Related Experiment Video
Updated: Jul 16, 2026


Thermochemical Studies of Ni(II) and Zn(II) Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
A DFT computational study of spin crossover in iron(III) and iron(II) tripodal imidazole complexes. A comparison of
Greg Brewer1, Myro Joy Olida, Ann M Schmiedekamp
1Department of Chemistry, Catholic University of America, Washington, DC 20064, USA.

Abstract:
B3LYP* functionals were used to model the sixteen iron(II) (1A, LS and 5T, HS) and iron(III) (2T, LS and 6A, HS) complexes of the 1 : 3 Schiff base condensate of tris(2-aminoethyl)amine and imidazole-4-carboxaldehyde, H3L1, and its deprotonated forms, [H2L1]1-, [HL1]2-, and [L1]3-. This ligand system is unusual in that [FeH3L1]3+, [FeH3L1]2+ and [FeL1]- all exhibit a spin crossover between 100-300 K. This makes these complexes ideal for a hybrid DFT computational approach and provides an opportunity to refine the value of the exact exchange admixture parameter, c3, and to predict properties of partially protonated complexes that are not experimentally available. The accepted value of 0.20 is larger than the value of approximately 0.13 that was found to best reproduce experimental data in terms of spin state predictions. With iron(III) B3LYP calculations showed that all of the complexes were low spin at 298 K with the exception of [FeH3L1]3+ which is spin crossover in agreement with experimental results. It was also shown for iron(III) that the ligand field increased as the number of protons decreased. In contrast all of the iron(II) complexes were close to the spin crossover region regardless of protonation state. Experimental structures are fairly well modeled by this system in regard to the key structural indicators of spin state, which are the bite and trans angles. The calculated iron to nitrogen atom distances are always larger in the high spin form than the low spin form but all iron to nitrogen bond distances are larger than the experimental values. In general non-bonded interactions are not well modeled by this methodology.
Related Concept Videos
 02:42
Valence Bond Theory
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human eye.
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:22
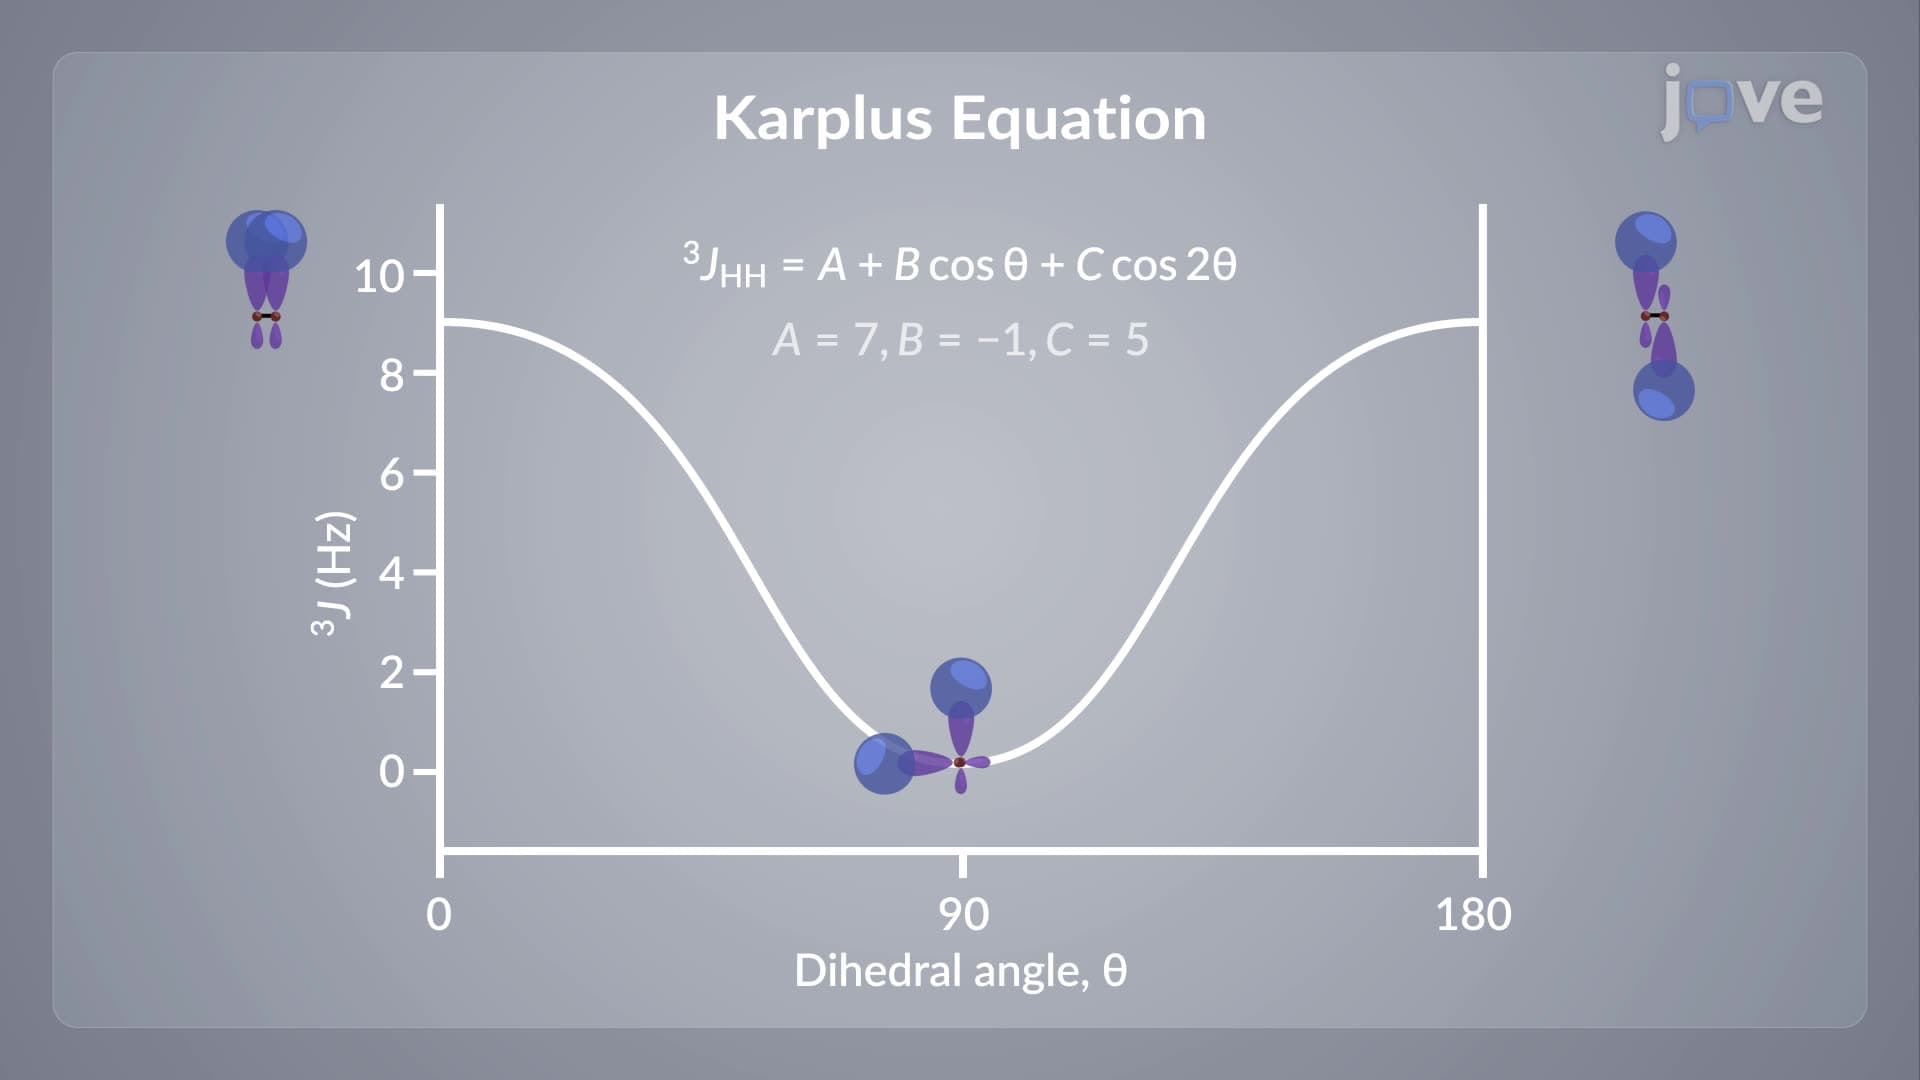
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...
 01:17
Spin–Spin Coupling: One-Bond Coupling