
Molecular and clinical characterization of a Moroccan Cog7 deficient patient
Bobby G Ng1, Christian Kranz, E E O Hagebeuk
1Department of Glycobiology and Carbohydrate Chemistry, Burnham Institute for Medical Research, La Jolla, CA 92037, USA.

Abstract:
Mutations in the N-linked glycosylation pathway cause rare autosomal recessive defects known as Congenital Disorders of Glycosylation (CDG). A previously reported mutation in the Conserved Oligomeric Golgi complex gene, COG7, defined a new subtype of CDG in a Tunisian family. The mutation disrupted the hetero-octomeric COG complex and altered both N- and O-linked glycosylation. Here we present clinical and biochemical data from a second family with the same mutation.
Related Concept Videos
 01:29
Alzheimer Disease l: Introduction
 01:23
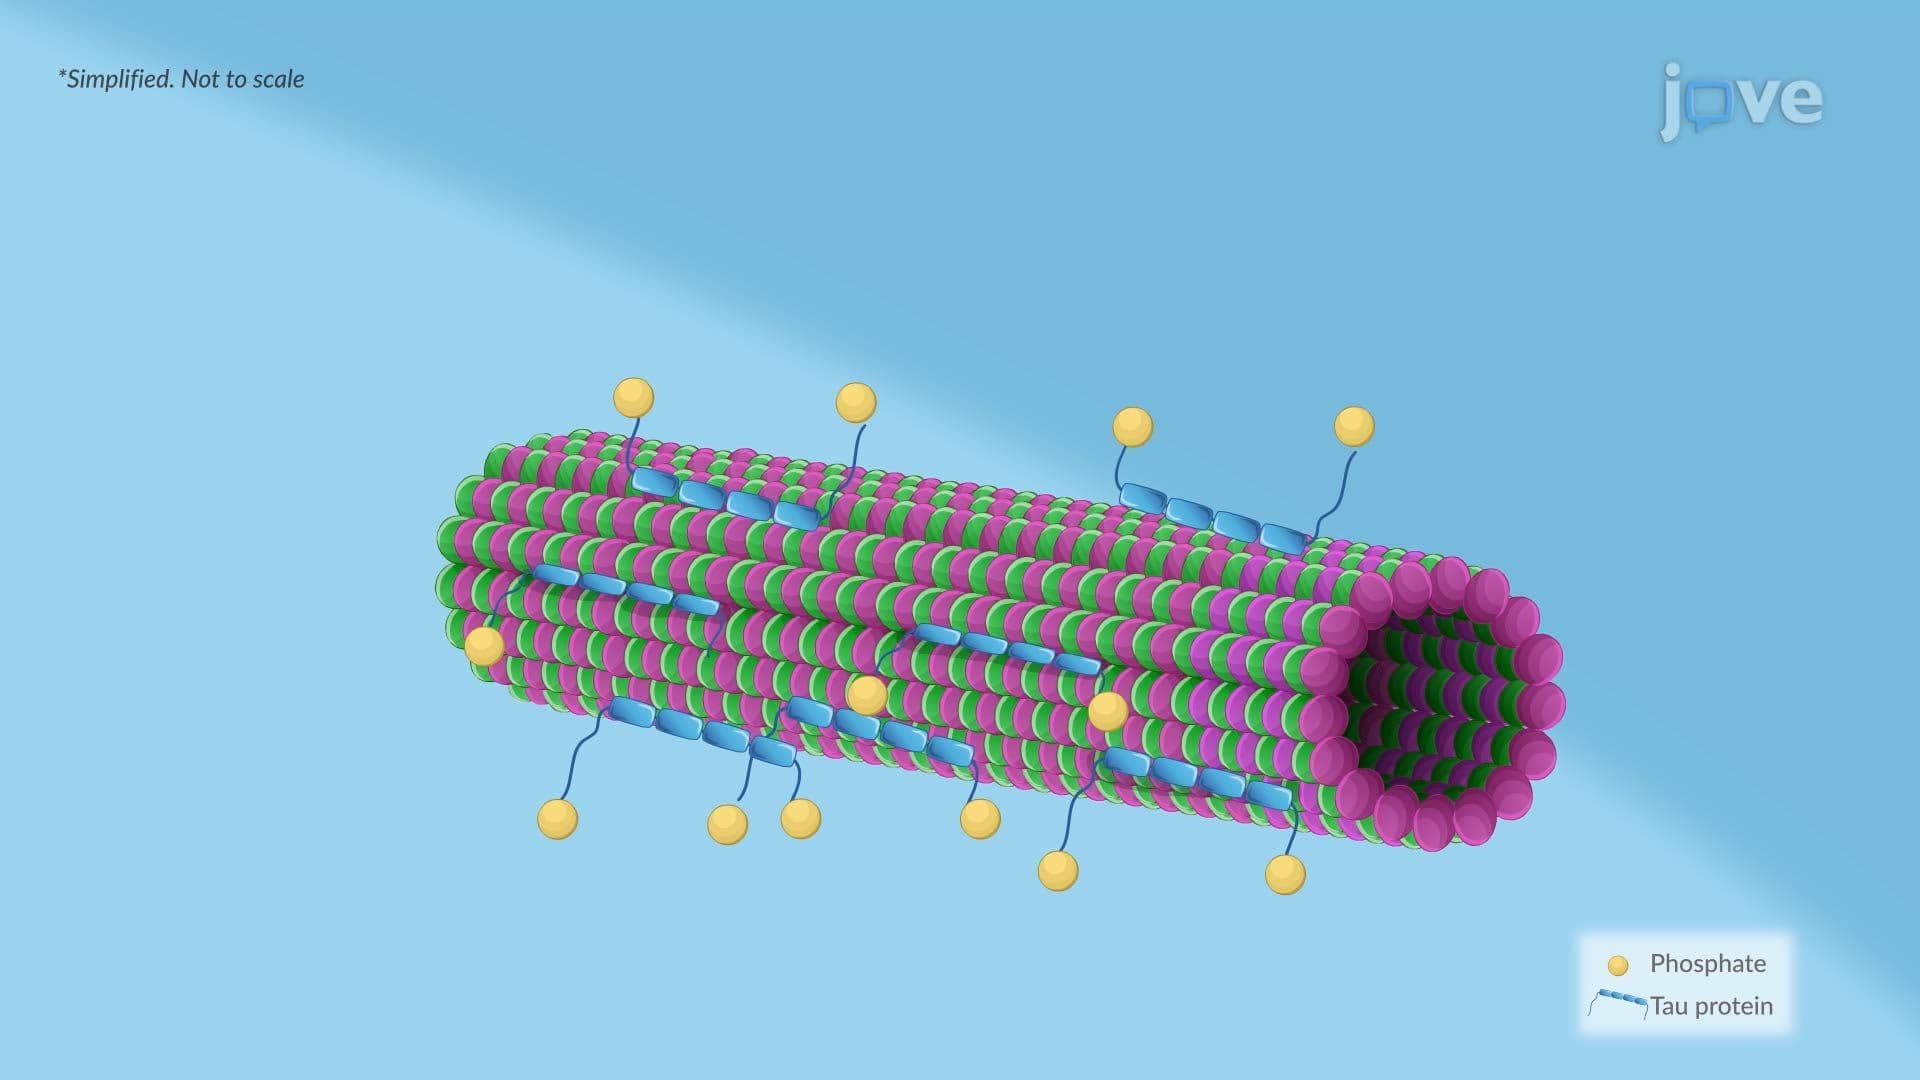
Alzheimer Disease ll: Pathophysiology
 01:21
Huntington Disease l: Introduction
 01:26
Alzheimer's Disease: Overview
The clinical diagnosis of AD hinges on the presence of memory and other cognitive impairments. Biomarkers, such as changes in Aβ and tau...
 01:24
Parkinson Disease ll: Pathophysiology
 01:20
Inborn Errors of Metabolism