Structural and functional characterization of binding sites in metallocarboxypeptidases based on Optimal Docking Area analysis
Related Experiment Video
Updated: Jul 15, 2026
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Daniel Fernández1, Josep Vendrell, Francesc X Avilés
1Institut de Biotecnologia i Biomedicina, Universitat Autònoma de Barcelona, 08193 Bellaterra, Spain.

Abstract:
The metallocarboxypeptidases (MCPs) belonging to the clan MC were studied by the Optimal Docking Area (ODA) method to evaluate protein-protein binding sites and to provide a basis for the identification of binding partners for this class of enzymes. The ODA method identifies surface patches with optimal desolvation energy based on the selection of low-energy docking regions, generated from a set of surface points around the protein. With few exceptions, the ODA method identified surface patches with a significant low-energy docking surface for all the MCPs with known three-dimensional structure. Overall, in 14 out of 24 cases, the detected ODA patches were correctly located (i.e. more than 50% of the predicted residues were in known protein-protein binding sites), yielding a global success rate of 58%. More specifically, the success rate increased up to 80% on the ODA patches detected for the catalytic domains of the M14A subfamily, independently on the partner. Interestingly, the ODA residues on the catalytic domain were correctly located in the interface with the N-terminal pro domain in all MCPs. The spatial distribution of the ODA patches for the different members of the family is in relation to the origin and function of the particular MCP, which allowed distinguishing between them. In good agreement with the experimentally characterized protein interfaces, the total average surface area of the theoretically derived ODA patches for the catalytic domain of MCPs is around 1700 A2 and their content in hydrophobic residues is about 40%. As a particular case, the average surface area of the ODA patches in MCPs of crop insect pests is about twice that of the MCPs of vertebrates, which might be related to their particular function. We recognized two binding regions for the catalytic domain of the MCPs, one of them accounting for nearly all the known intermolecular interactions made up by the enzymes. Protein inhibitors seem to have evolved to dock on this subset of ODA patches, evoking the binding mode of the N-terminal pro domains. The second binding region detected, for which no ligands have been identified so far, seems to be related to the acquisition/maintenance of the native structure of the peptidase. Overall, the ODA method has been successful in identifying low-energy docking areas in a set of structurally and functionally related proteins, suggesting that it can be easily extended to other families in the search for protein-protein binding sites and for their functional significance.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 00:49
Ligand Binding and Linkage
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis pathway,...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:04
Protein-protein Interfaces
 02:18
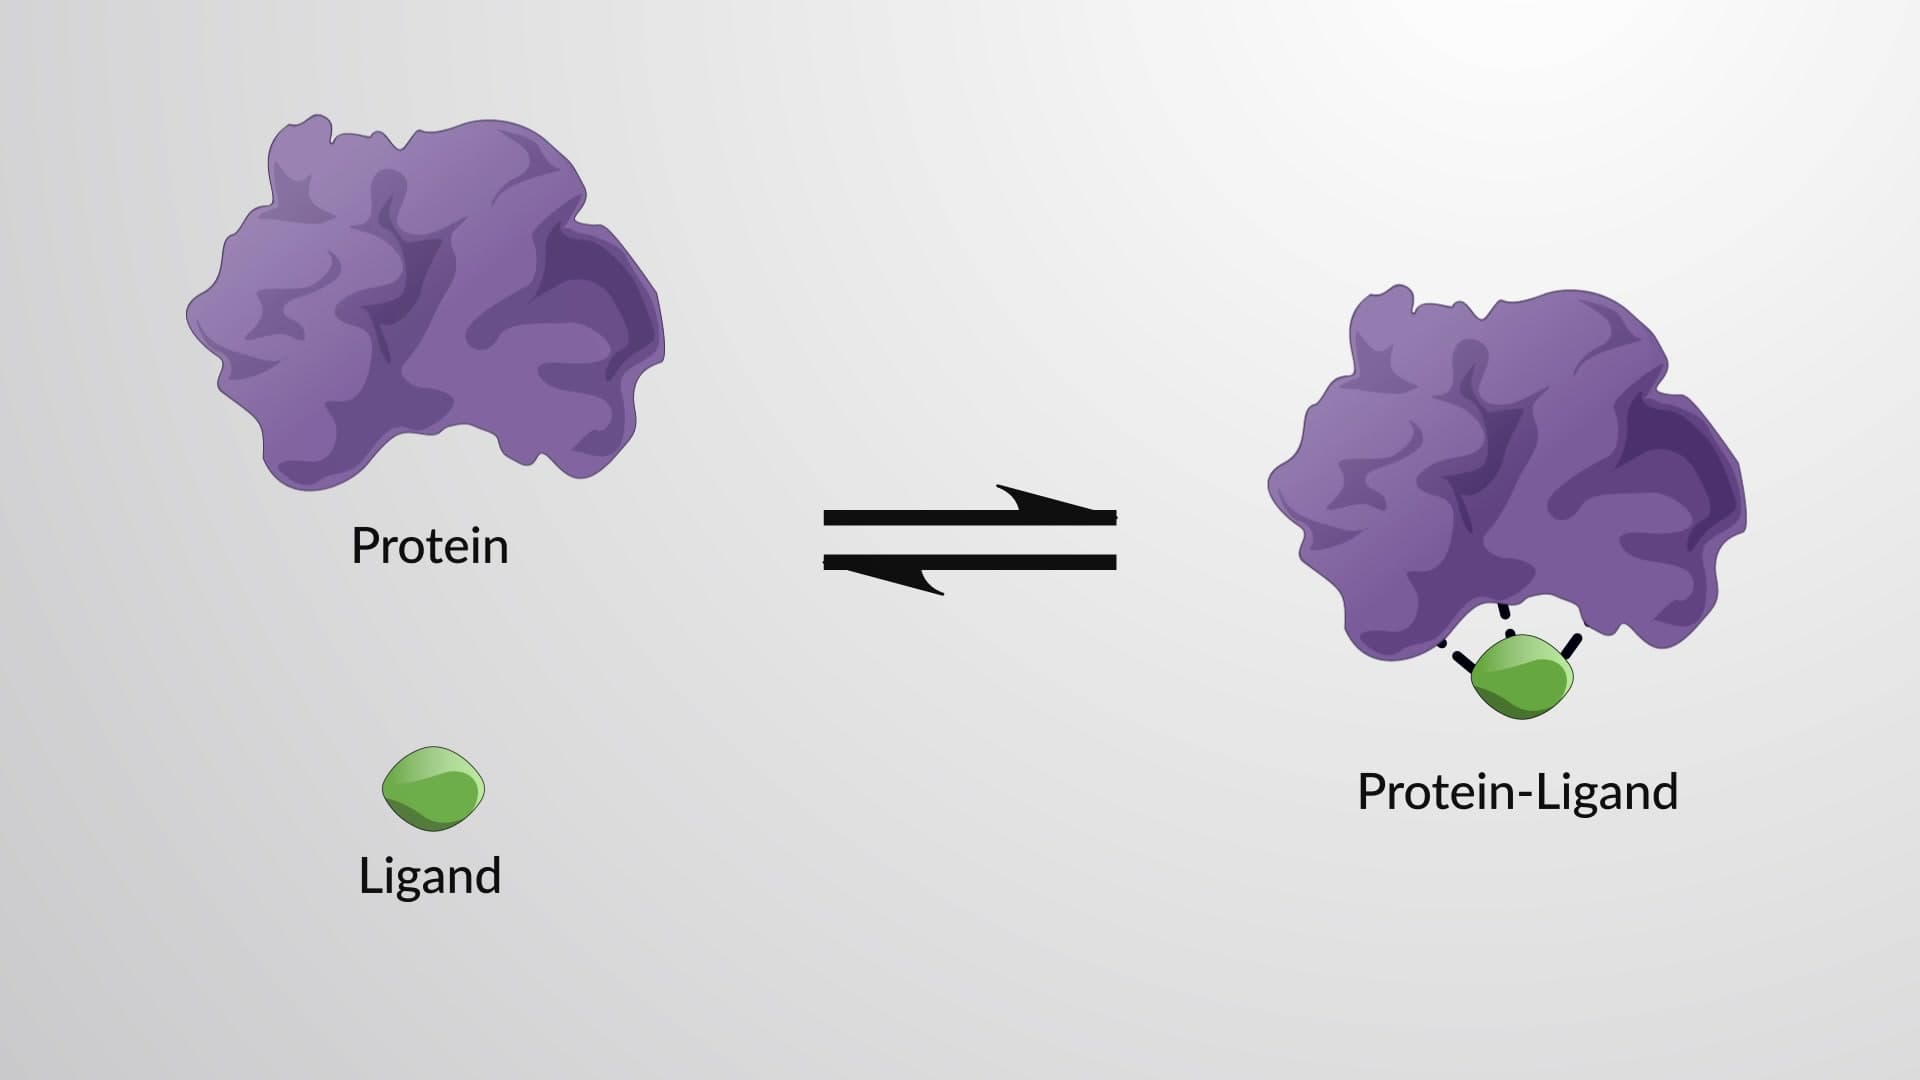
The Equilibrium Binding Constant and Binding Strength