Scoring function accuracy for membrane protein structure prediction
Related Experiment Video
Updated: Jul 15, 2026

A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
1Department of Chemistry, University of Rochester, Rochester, New York, USA.

Abstract:
We perform a systematic examination of the ability of several different high-resolution, atomic-detail scoring functions to discriminate native conformations of loops in membrane proteins from non-native but physically reasonable, or "decoy," conformations. Decoys constructed from changing a loop conformation while keeping the remainder of the protein fixed are a challenging test of energy function accuracy. Nevertheless, the best of the energy functions we examined recognized the native structure as lowest in energy around half the time, and consistently chose it as a low-energy structure. This suggests that the best of present energy functions, even without a representation of the lipid bilayer, are of sufficient accuracy to give reasonable confidence in predictions of membrane protein structure. We also constructed homology models for each structure, using other known structures in the same protein family as templates. Homology models were constructed using several scoring functions and modeling programs, but with a comparable sampling effort for each procedure. Our results indicate that the quality of sequence alignment is probably the most important factor in model accuracy for sequence identity from 20-40%; one can expect a reasonably accurate model for membrane proteins when sequence identity is greater than 30%, in agreement with previous studies. Most errors are localized in loop regions, which tend to be found outside the lipid bilayer. For the most discriminative energy functions, it appears that errors are most likely due to lack of sufficient sampling, although it should be stressed that present energy functions are still far from perfectly reliable.

Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:30
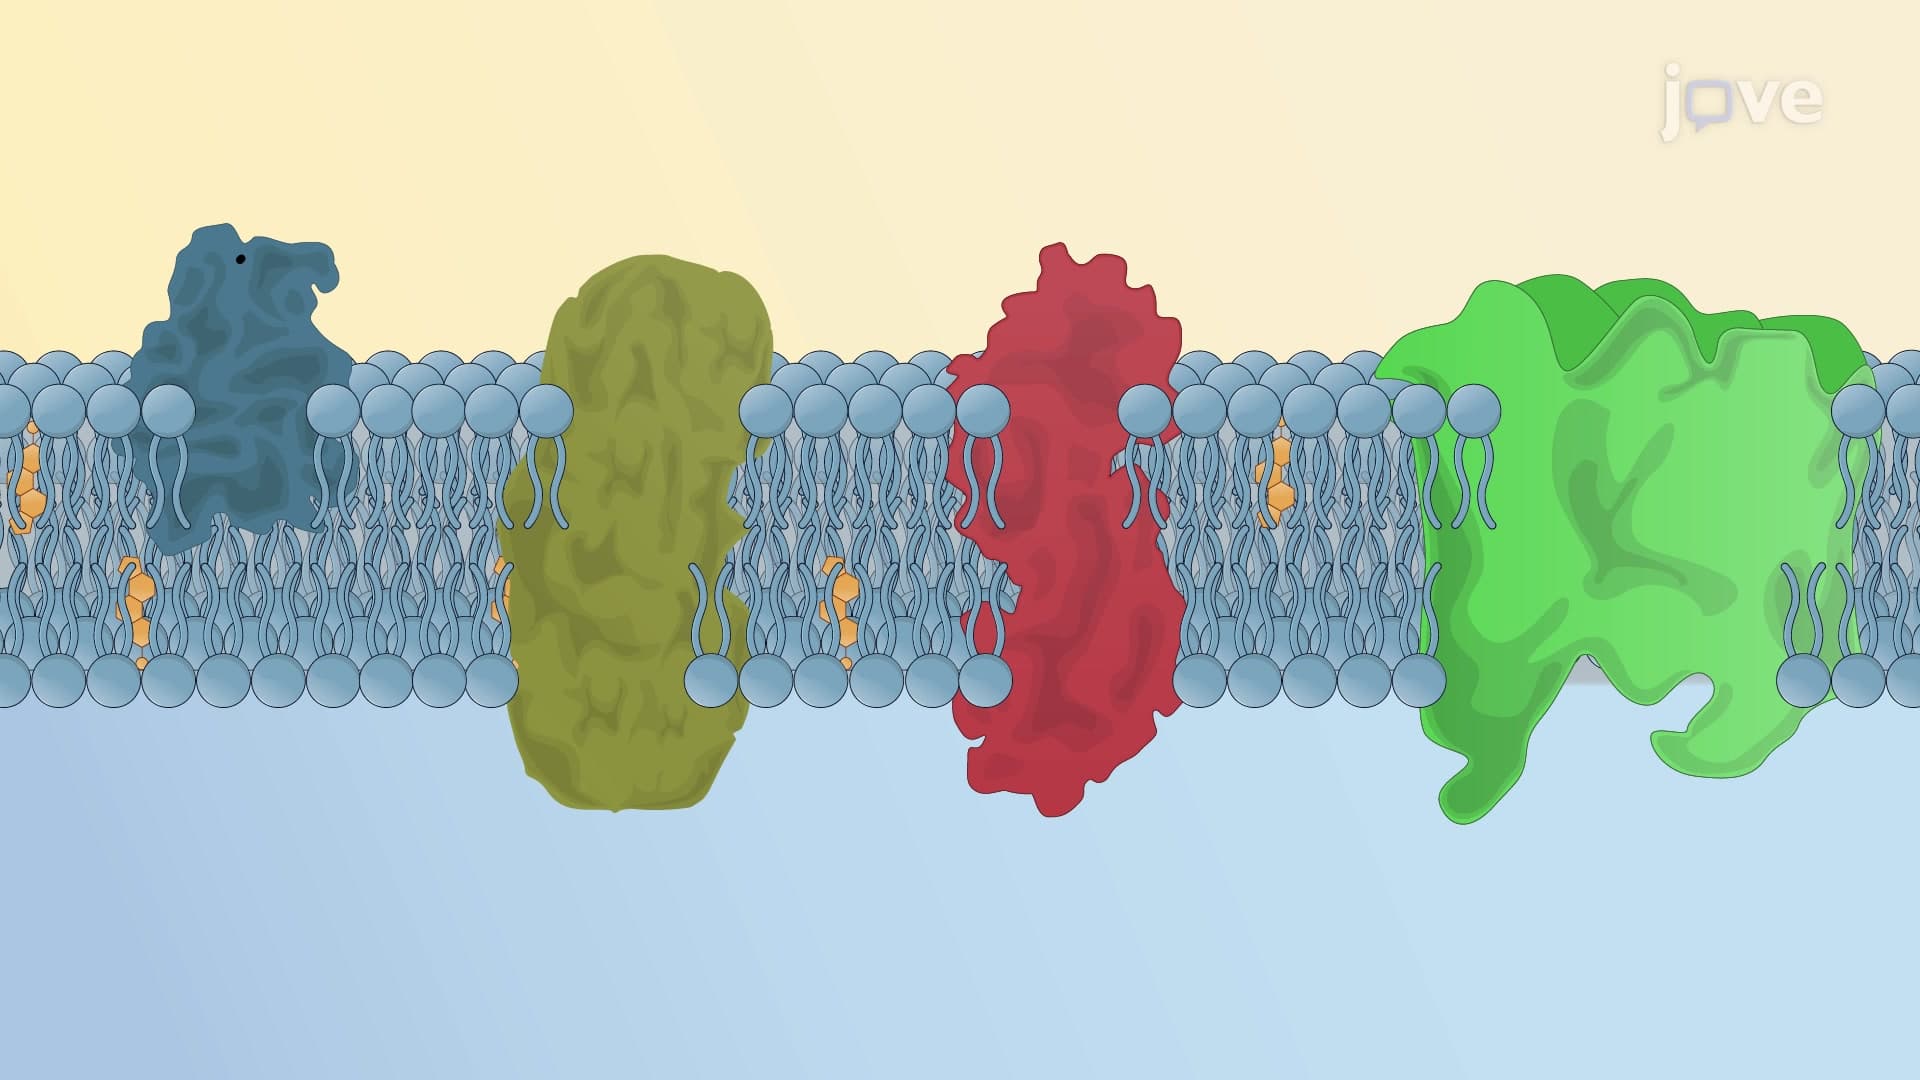
Membrane Proteins
01:30Membrane Proteins
 01:29
Protein Folding Quality Check in the RER
 01:25
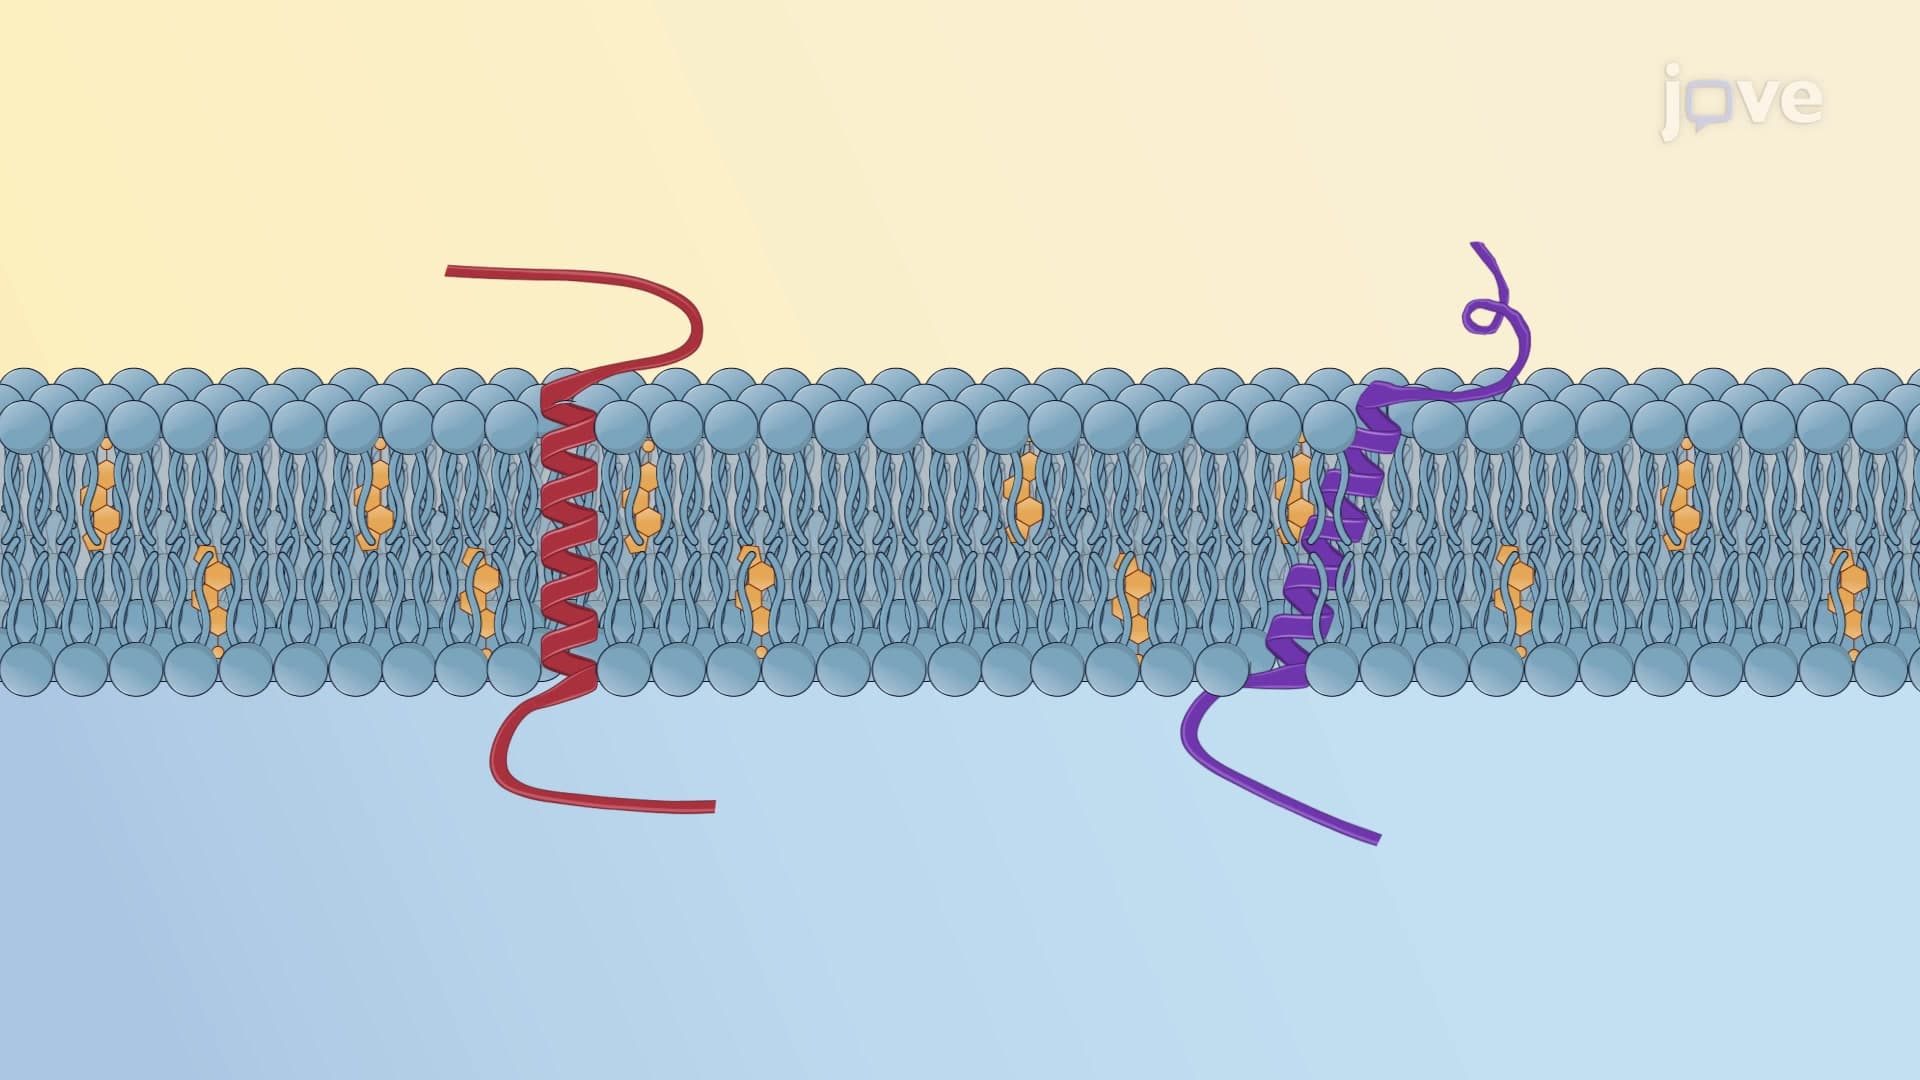
Single-pass Transmembrane Proteins
 01:09
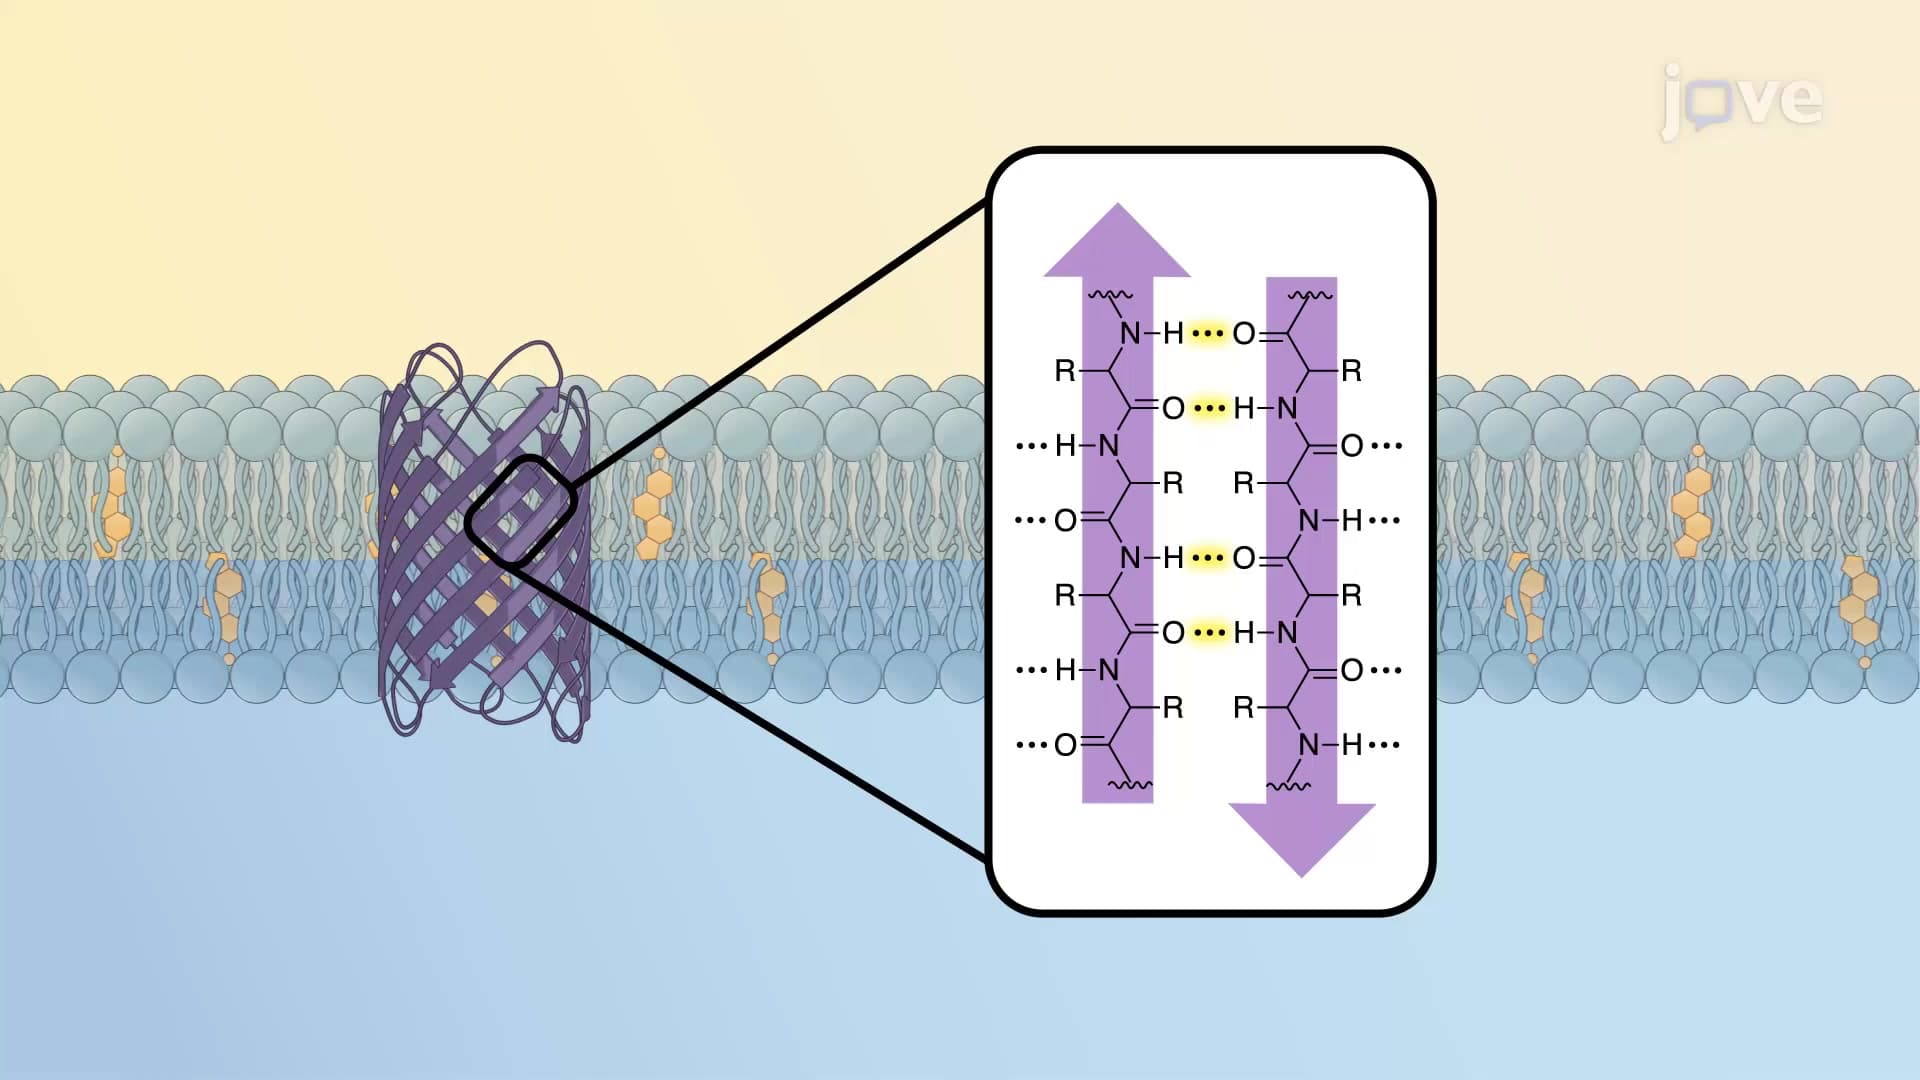
Multi-pass Transmembrane Proteins and β-barrels
α-Helix containing multi-pass transmembrane proteins
Multi-pass transmembrane proteins such as G-protein-linked receptors (GPCRs) and...