Mass-weighted molecular dynamics simulation of the protein-ligand complex of rhizopuspepsin and inhibitor
Related Experiment Video
Updated: Oct 21, 2026
Analyzing Protein Architectures and Protein-Ligand Complexes by Integrative Structural Mass Spectrometry
Published on: October 15, 2018
1Upjohn Research Laboratories, Kalamazoo, Michigan 49001.

Abstract:
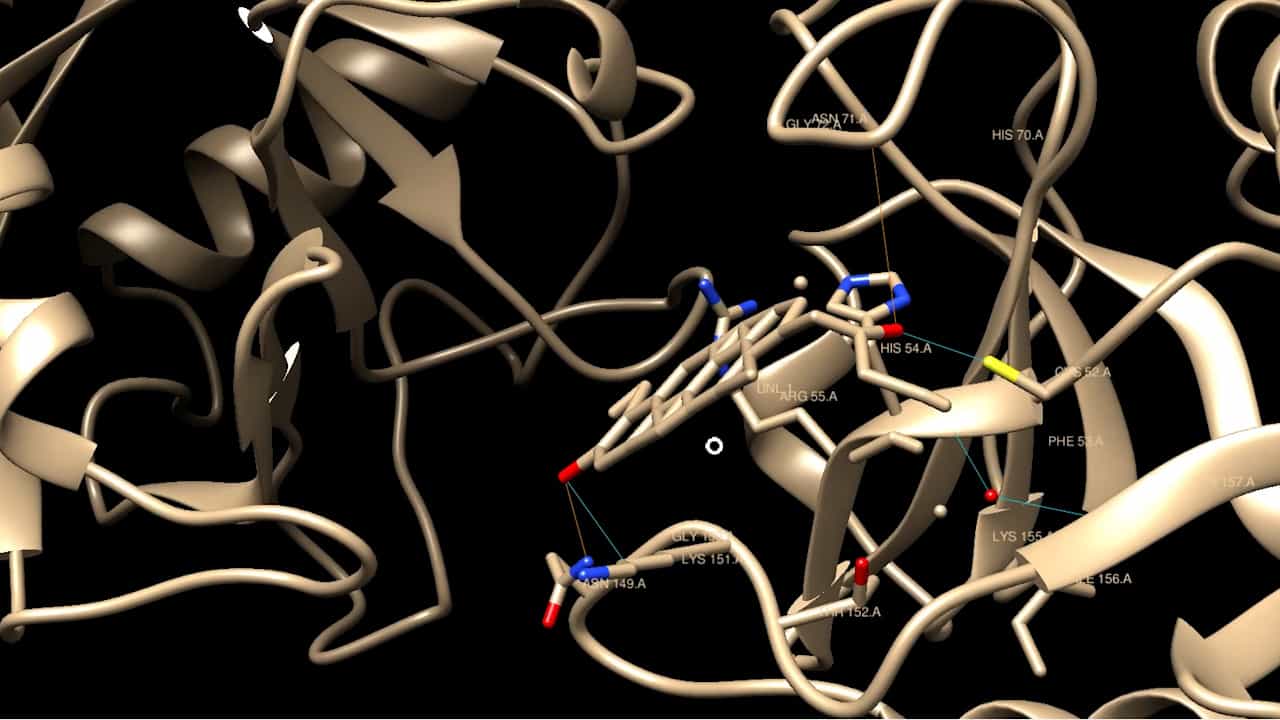
The mass-weighted molecular dynamics simulation method was developed previously for sampling the multidimensional conformational space of linear and cyclic polypeptides and studying their conformational flexibility. Herein results from molecular dynamics simulations of the protein-ligand complex of the aspartyl protease rhizopuspepsin and a polypeptide inhibitor are reported. The dihedral conformational space sampling for the linear peptide inhibitor in situ was found to be increased in the mass-weighted simulation as in other molecular systems previously studied. More significantly, the physical space of the enzyme binding pocket was also sampled efficiently in the simulations and multiple binding sites were identified for the inhibitor. These results suggest that it may be possible now to study, by computer simulations, the putative initial enzyme-inhibitor complex suggested experimentally from the time-dependent kinetics of enzyme inhibition by slow-binding inhibitors (Morrison, J. F., and C. T. Walsh. 1988. Adv. Enzymol. 61:201), and/or conformational substates in protein-ligand complexes suggested in the study of reassociation dynamics of myoglobin and carbon monoxide following photolysis (Austin, R. H., K. W. Beeson, L. Eisenstein, H. Frauenfelder, and I. C. Gunsalus. 1975. Biochemistry. 14:5355). Moreover, the intermediate binding steps and the molecular flexibility of the inhibitor shown in the MWMD simulation may have crucial roles in the ligand binding process.
More Related Videos
 10:01
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025