Related Experiment Video
Updated: Jul 15, 2026

Methods of Ex Situ and In Situ Investigations of Structural Transformations: The Case of Crystallization of Metallic Glasses
Published on: June 7, 2018
First-principles calculations of solid-state (17)O and (29)Si NMR spectra of Mg(2)SiO(4) polymorphs
Sharon E Ashbrook1, Laurent Le Pollès, Chris J Pickard
1School of Chemistry and EaStCHEM, University of St Andrews, North Haugh, St Andrews, UK KY16 9ST. sema@st-andrews.ac.uk

Abstract:
The nuclear magnetic resonance (NMR) shielding and electric field gradient (EFG) tensors of three polymorphs of Mg(2)SiO(4), forsterite (alpha-Mg(2)SiO(4)), wadsleyite (beta-Mg(2)SiO(4)) and ringwoodite (gamma-Mg(2)SiO(4)), have been calculated using a density functional theory (DFT) approach with a planewave basis set and pseudopotential approximation. These Mg(2)SiO(4) polymorphs are the principal components of the Earth down to depths of 660 km and have been proposed as the hosts of water in the Earth's upper mantle and transition zone. A comparison of our calculations with single-crystal spectroscopic data in the literature for the alpha-polymorph, forsterite, shows that both the magnitude and orientation of the shielding and EFG tensors for O and Si can be obtained with sufficient accuracy to distinguish subtle differences in atomic positions between published structures. We compare calculated (17)O MAS NMR quadrupolar powder lineshapes directly with experimental lineshapes and show that we are able to reproduce them within the precision with which the NMR parameters may be determined from multi-parameter fitting. The relatively small amounts of sample available for the beta- and gamma-polymorphs, arising from the high pressures required for synthesis, has hindered the extraction of NMR parameters in previous work. The application of DFT calculations to these high-pressure polymorphs confirms previous spectral assignments, and provides deeper insight into the empirical correlations and observations reported in the literature. These first-principles methods are highly promising for the determination of local bonding in more complex materials, such as the hydrated forms of Mg(2)SiO(4), by aiding analysis of their multinuclear NMR spectra.
More Related Videos
 06:37
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
 08:42
High-Sensitivity Nuclear Magnetic Resonance at Giga-Pascal Pressures: A New Tool for Probing Electronic and Chemical Properties of Condensed Matter under Extreme Conditions
Published on: October 10, 2014
Related Concept Videos
 02:54
Molecular and Ionic Solids
Molecular Solids
Molecular crystalline solids, such as ice, sucrose (table sugar), and iodine, are solids that are composed of neutral molecules as their constituent units. These molecules are held together by weak intermolecular forces such as London dispersion forces, dipole-dipole interactions, or hydrogen bonds, which...
 03:51
Molecular Orbital Theory II
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are slanted or...
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 01:27
Lattice Energies of Ionic Crystals
 01:15
NMR Spectroscopy: Chemical Shift Overview
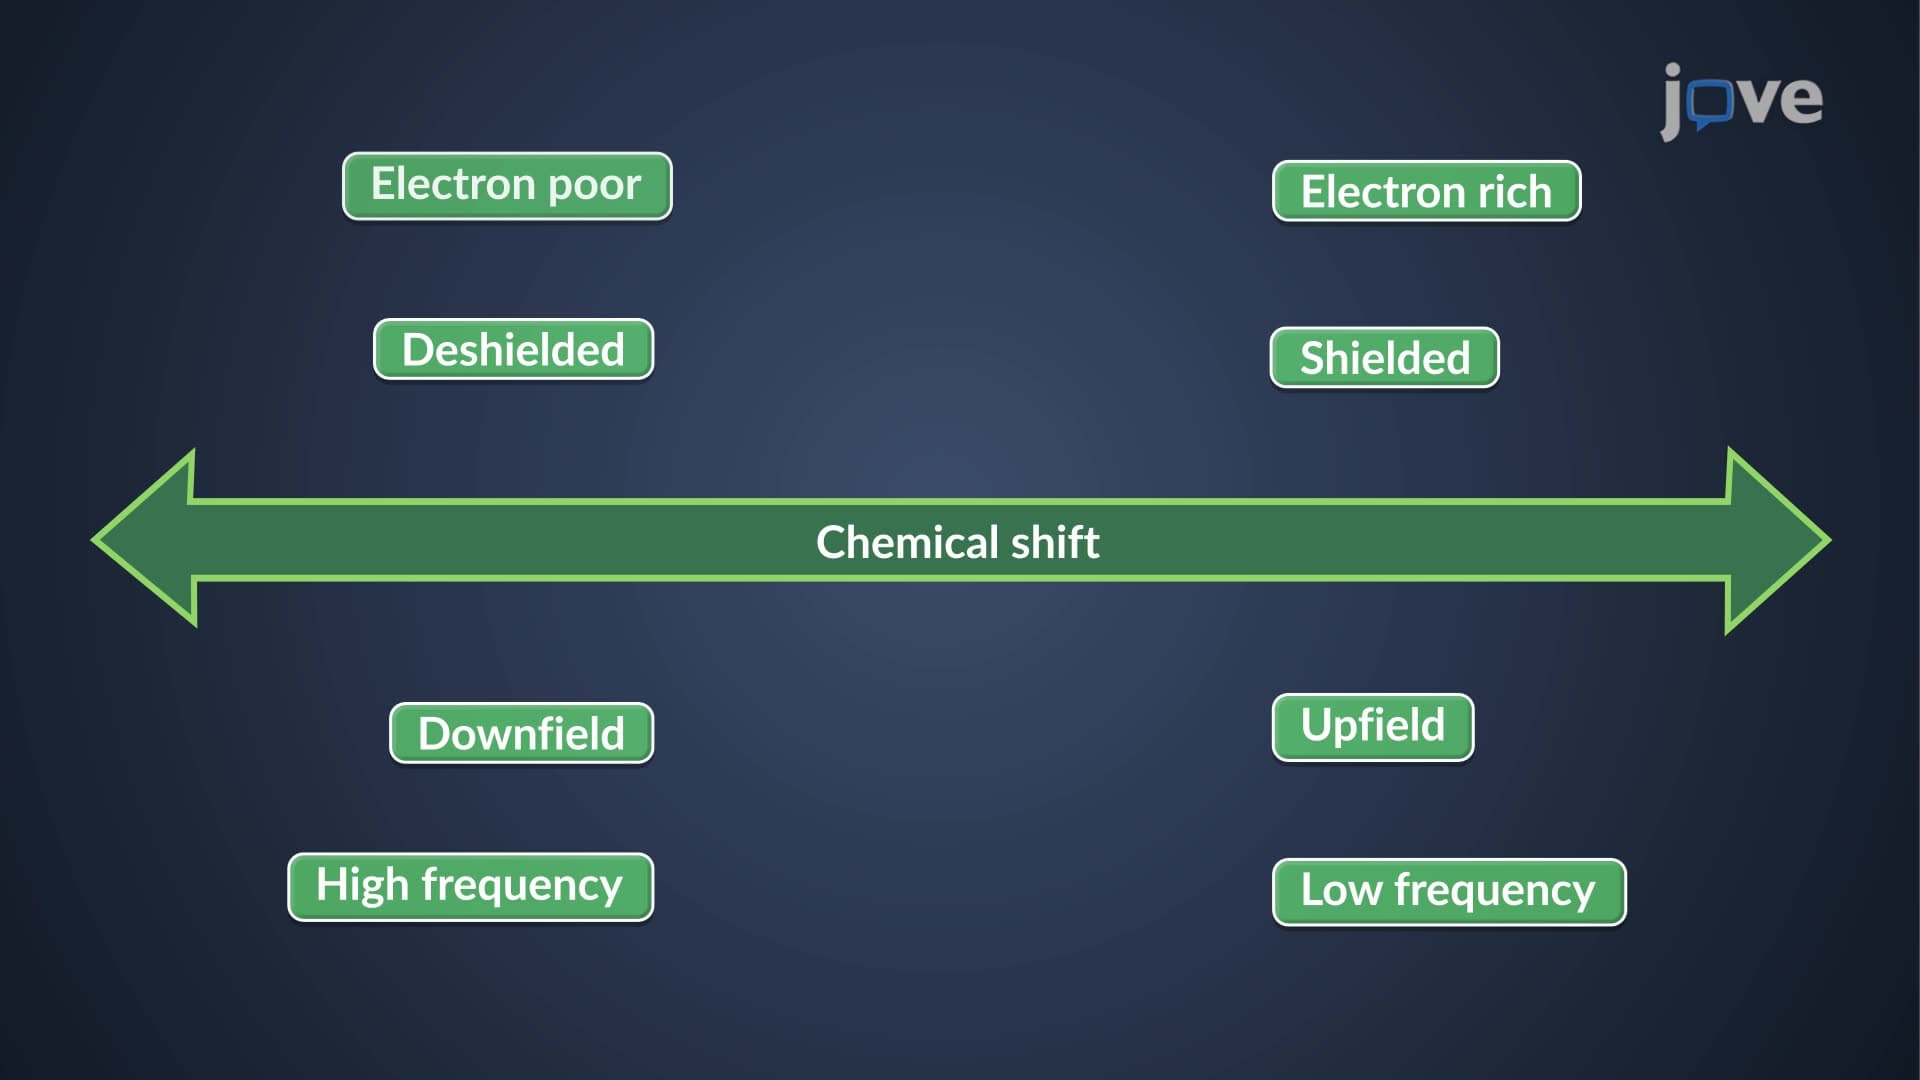
For instance, the proton...