Analytical excited state forces for the time-dependent density-functional tight-binding method
Related Experiment Video
Updated: Jul 14, 2026
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
D Heringer1, T A Niehaus, M Wanko
1General Electrics, Consumer & Industrial--Lighting, 1340 Budapest, Vaci ut 77, Hungary.

Abstract:
An analytical formulation for the geometrical derivatives of excitation energies within the time-dependent density-functional tight-binding (TD-DFTB) method is presented. The derivation is based on the auxiliary functional approach proposed in [Furche and Ahlrichs, J Chem Phys 2002, 117, 7433]. To validate the quality of the potential energy surfaces provided by the method, adiabatic excitation energies, excited state geometries, and harmonic vibrational frequencies were calculated for a test set of molecules in excited states of different symmetry and multiplicity. According to the results, the TD-DFTB scheme surpasses the performance of configuration interaction singles and the random phase approximation but has a lower quality than ab initio time-dependent density-functional theory. As a consequence of the special form of the approximations made in TD-DFTB, the scaling exponent of the method can be reduced to three, similar to the ground state. The low scaling prefactor and the satisfactory accuracy of the method makes TD-DFTB especially suitable for molecular dynamics simulations of dozens of atoms as well as for the computation of luminescence spectra of systems containing hundreds of atoms.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
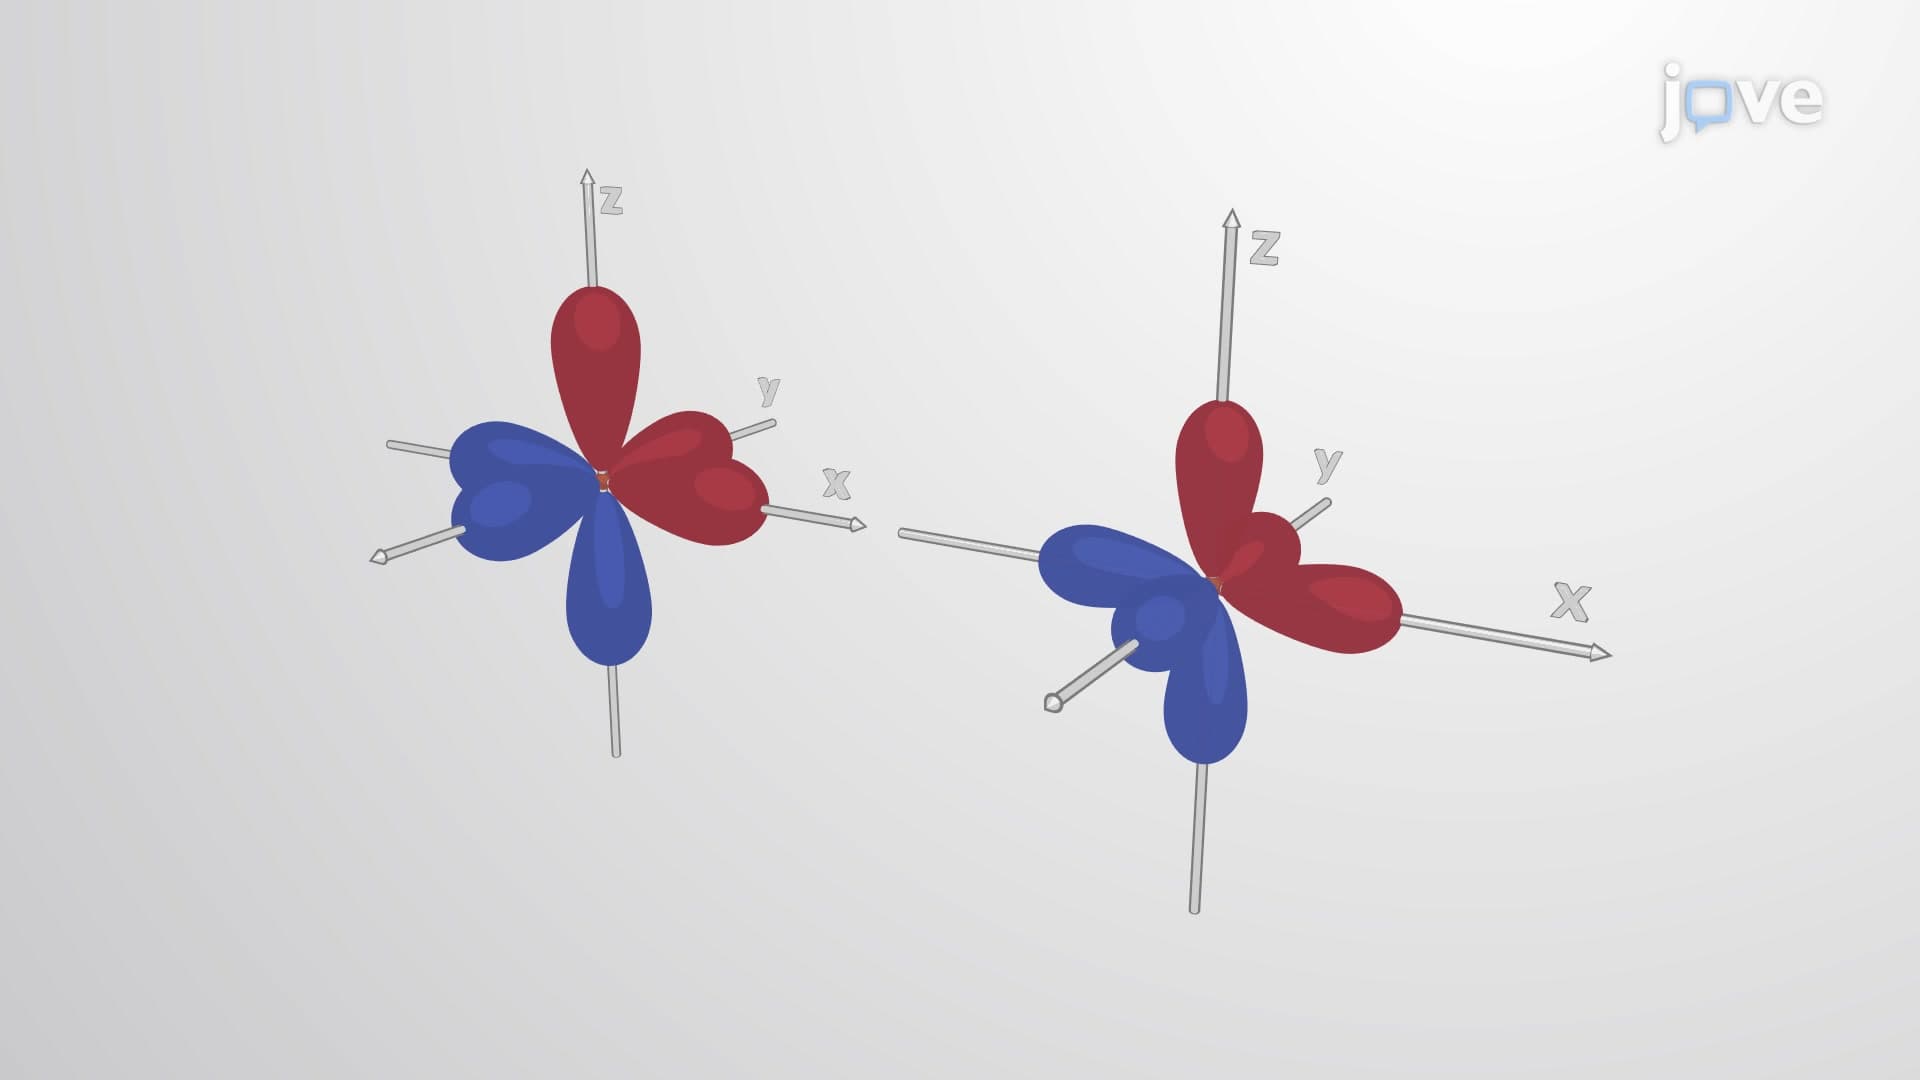
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 03:35
Hybridization of Atomic Orbitals II
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 02:35
Molecular Orbital Theory I
 03:24
Hybridization of Atomic Orbitals I
 03:51
Molecular Orbital Theory II