Related Experiment Video
Updated: Jul 13, 2026


Thermochemical Studies of Ni(II) and Zn(II) Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
A comparative study of semiempirical, ab initio, and DFT methods in evaluating metal-ligand bond strength, proton
Gilles Frison1, Gilles Ohanessian
1Laboratoire des Mécanismes Réactionnels, Département de Chimie, Ecole Polytechnique, CNRS, 91128 Palaiseau Cedex, France. gilles.frison@polytechnique.org

Abstract:
Although theoretical methods are now available which give very accurate results, often comparable to the experimental ones, modeling chemical or biological interesting systems often requires less demanding and less accurate theoretical methods, mainly due to computer limitations. Therefore, it is crucial to know the precision of such less reliable methods for relevant models and data. This has been done in this work for small zinc-active site models including O- (H(2)O and OH(-)) and N-donor (NH(3) and imidazole) ligands. Calculations using a number of quantum mechanical methods were carried out to determine their precision for geometries, coordination number relative stability, metal-ligand bond strengths, proton affinities, and interaction energies between first and second shell ligands. We have found that obtaining chemical accuracy can be as straightforward as HF geometry optimization with a double-zeta plus polarization basis followed by a B3LYP energy calculation with a triple-zeta quality basis set including diffuse and polarization functions. The use of levels as low as PM3 geometry optimization followed by a B3LYP single-point energy calculation with a double-zeta quality basis including polarization functions already yields useful trends in bond length, proton affinities or bond dissociation energies, provided that appropriate caution is taken with the optimized structures. The reliability of these levels of calculation has been successfully demonstrated for real biomimetic cases.
More Related Videos
 11:04
Ion Mobility-Mass Spectrometry Techniques for Determining the Structure and Mechanisms of Metal Ion Recognition and Redox Activity of Metal Binding Oligopeptides
Published on: September 7, 2019
 06:31
Preparation of SNS Cobalt(II) Pincer Model Complexes of Liver Alcohol Dehydrogenase
Published on: March 19, 2020
Related Concept Videos
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:09
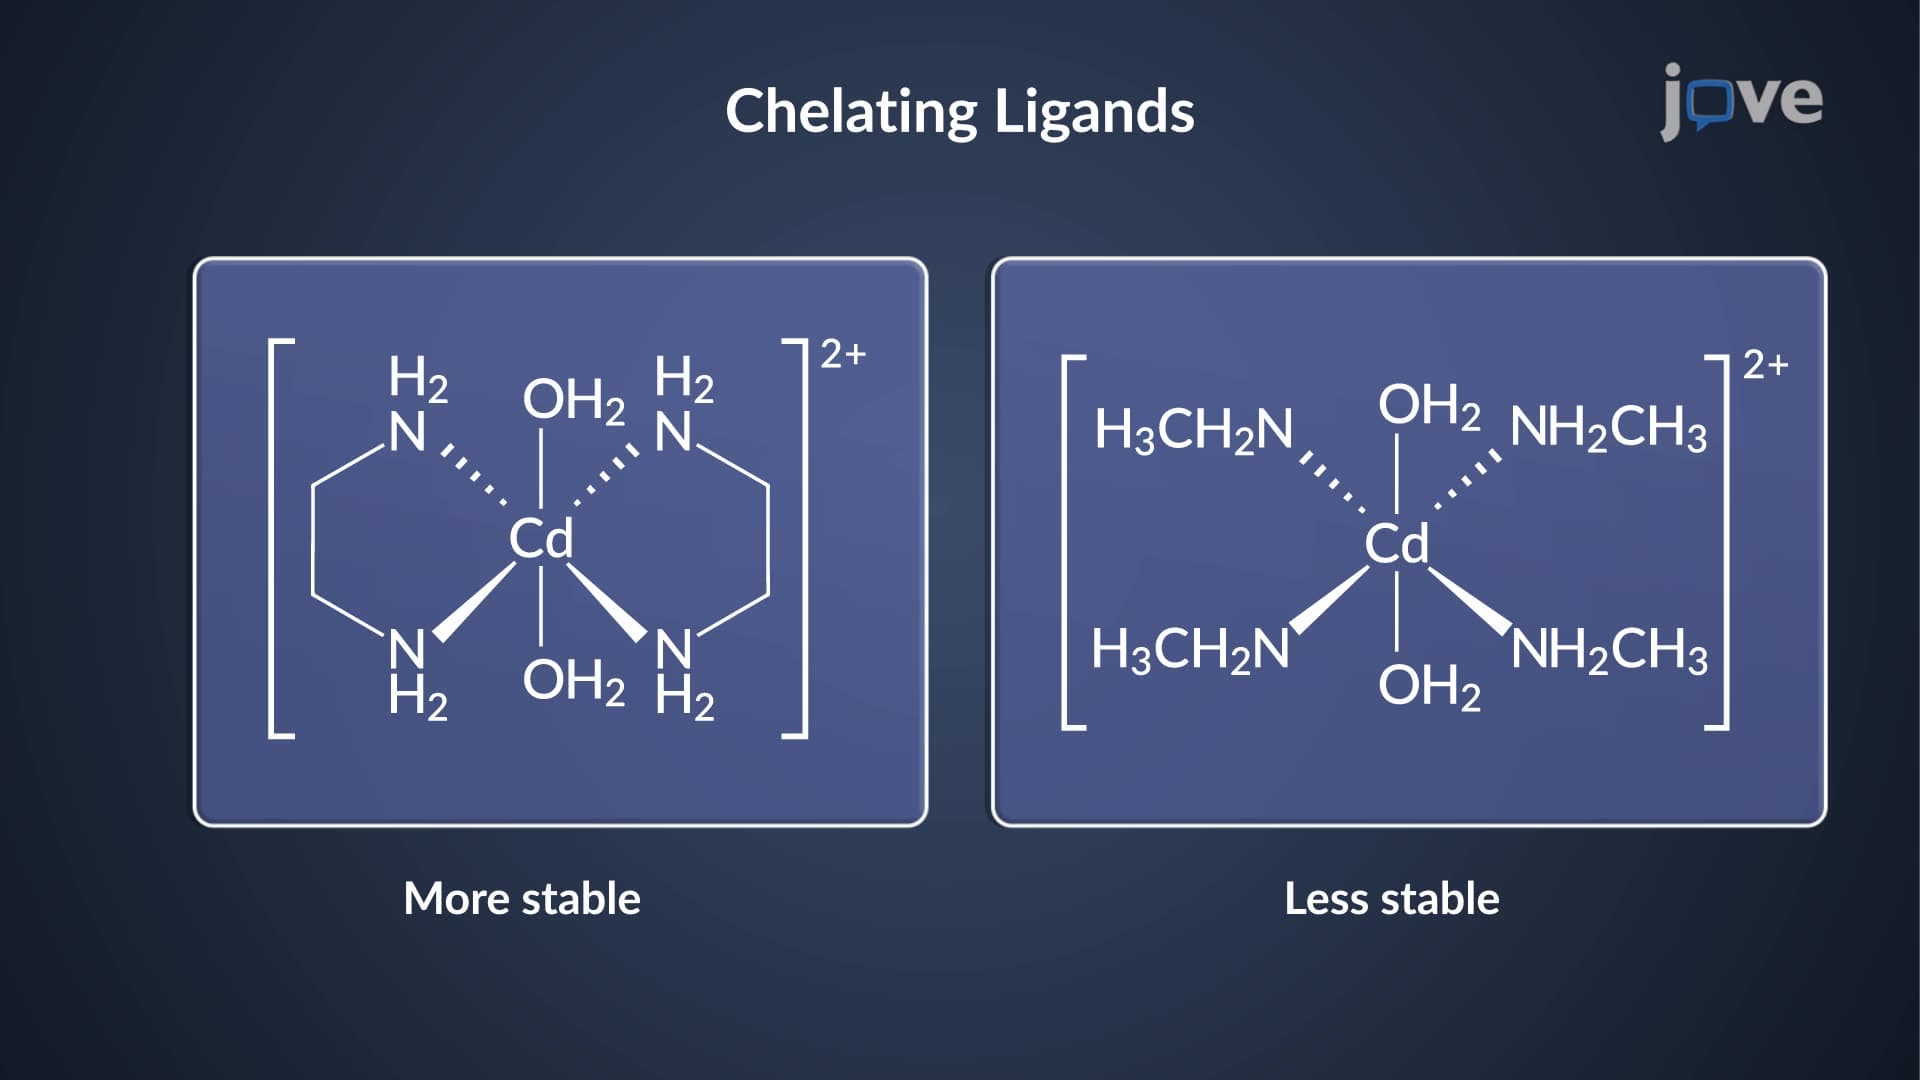
Complexation Equilibria: Factors Influencing Stability of Complexes
 02:42
Valence Bond Theory
 00:43
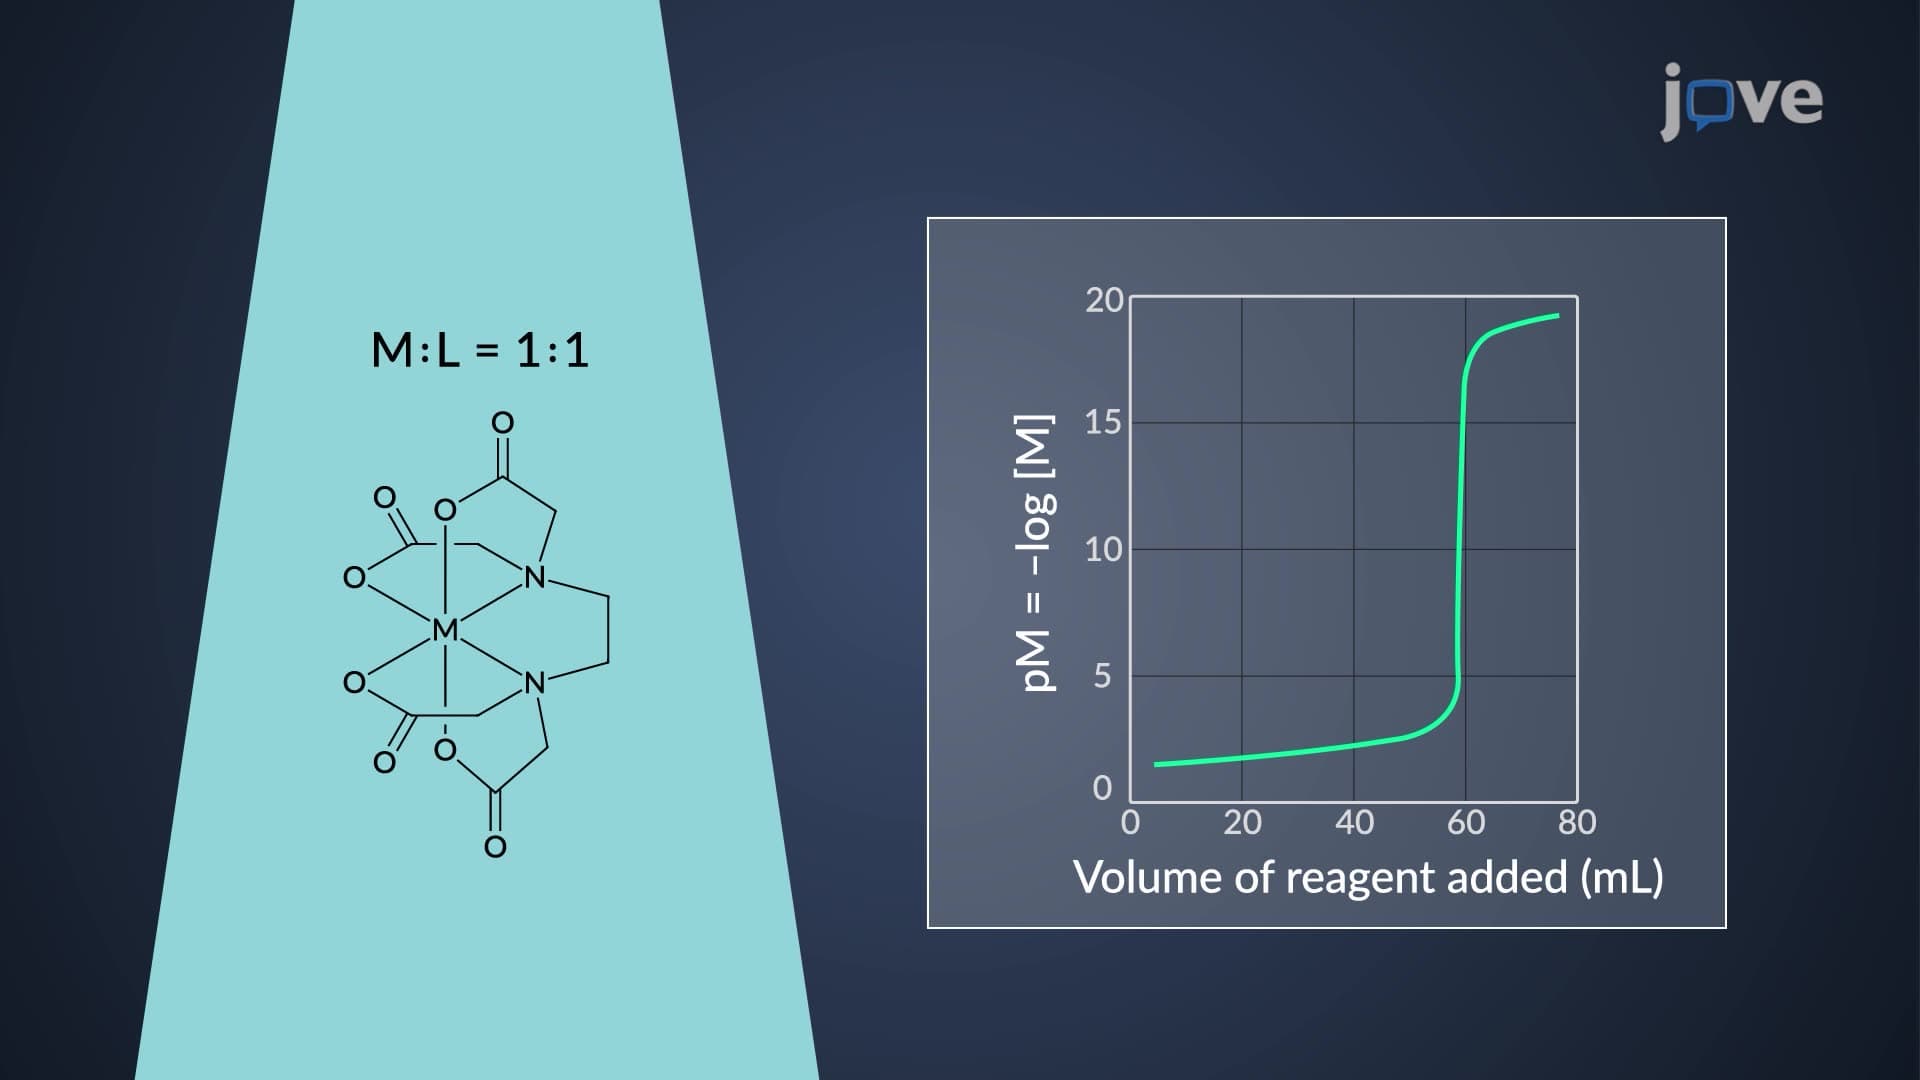
Complexometric Titration: Ligands
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...