Related Experiment Video
Updated: Jul 13, 2026

Catalytic Reactions at Amine-Stabilized and Ligand-Free Platinum Nanoparticles Supported on Titania During Hydrogenation of Alkenes and Aldehydes
Published on: June 24, 2022
Vibrational recognition of adsorption sites for CO on platinum and platinum-ruthenium surfaces
Ismaila Dabo1, Andrzej Wieckowski, Nicola Marzari
1Department of Materials Science and Engineering, Massachusetts Institute of Technology, 77 Massachusetts Avenue, Cambridge, Massachusetts 02139, USA. dabo_is@mit.edu

Abstract:
We have studied the vibrational properties of CO adsorbed on platinum and platinum-ruthenium surfaces using density-functional perturbation theory within the Perdew-Burke-Ernzerhof generalized-gradient approximation. The calculated C-O stretching frequencies are found to be in excellent agreement with spectroscopic measurements. The frequency shifts that take place when the surface is covered with ruthenium monolayers are also correctly predicted. This agreement for both shifts and absolute vibrational frequencies is made more remarkable by the frequent failure of local and semilocal exchange-correlation functionals in predicting the stability of the different adsorption sites for CO on transition metal surfaces. We have investigated the chemical origin of the C-O frequency shifts introducing an orbital-resolved analysis of the force and frequency density of states, and assessed the effect of donation and backdonation on the CO vibrational frequency using a GGA+molecular U approach. These findings rationalize and establish the accuracy of density-functional calculations in predicting absolute vibrational frequencies, notwithstanding the failure in determining relative adsorption energies, in the strong chemisorption regime.
More Related Videos
 11:38
In situ FTIR Spectroscopy as a Tool for Investigation of Gas/Solid Interaction: Water-Enhanced CO2 Adsorption in UiO-66 Metal-Organic Framework
Published on: February 1, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:25
Adsorption Isotherms II
 01:29
Adsorption Isotherms I
 01:28
Adsorption of Gases on Solids
 01:22
Heterogeneous Catalysis
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
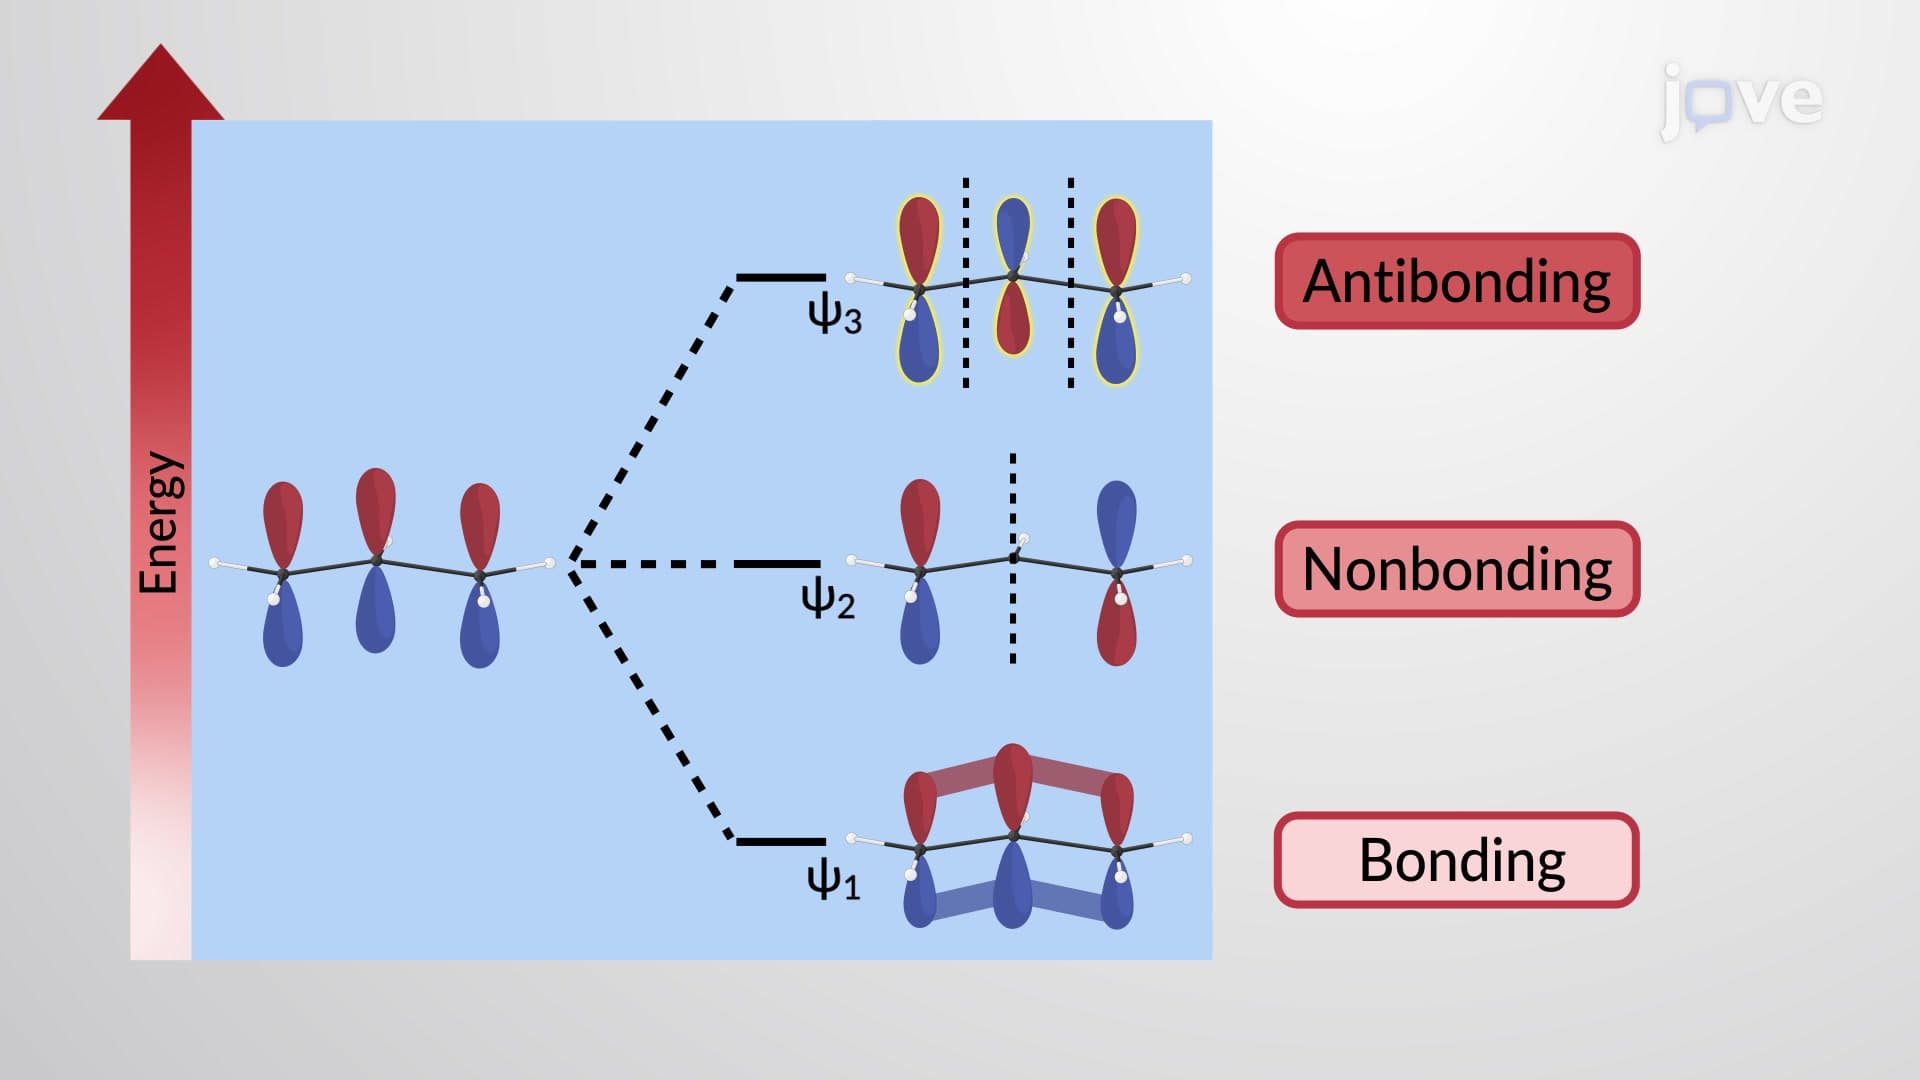 01:18
π Molecular Orbitals of the Allyl Cation and Anion