High-spin versus broken symmetry-Effect of DFT spin density representation on the geometries of three diiron (III) model compounds
Related Experiment Video
Updated: Jul 11, 2026
Visualizing Uniaxial-strain Manipulation of Antiferromagnetic Domains in Fe1+YTe Using a Spin-polarized Scanning Tunneling Microscope
Published on: March 24, 2019
1Department of Sciences and Technology, Universidad Metropolitana, P. O. Box 21150, San Juan, Puerto Rico 00928-1150, USA. binningrc@yahoo.com

Abstract:
Unrestricted density functional theory calculations have been conducted on three diiron(III) synthetic model compounds containing antiferromagnetically coupled high-spin (HS) irons for which crystallographic structures and Raman spectral data are available. Three density functionals have been employed: BPW91, PWC, and BOP. The study compares the effects on optimized geometries and harmonic vibrational frequencies of spin-paired (SP) low-spin, HS, and broken symmetry antiferromagnetically coupled singlet representations of the spin density distribution. The geometries around the diiron centers in the HS and broken symmetry (BS) representations are found to be similar, both markedly different from those arising from the SP representation. Small differences between the HS and BS results are seen in bond lengths, angles, Raman frequencies, and spin densities associated with oxo and peroxo bridges between the irons.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 07:24
Hyperspectral Imaging as a Tool to Study Optical Anisotropy in Lanthanide-Based Molecular Single Crystals
Published on: April 14, 2020
Related Concept Videos
 02:42
Valence Bond Theory
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human eye.
 01:22
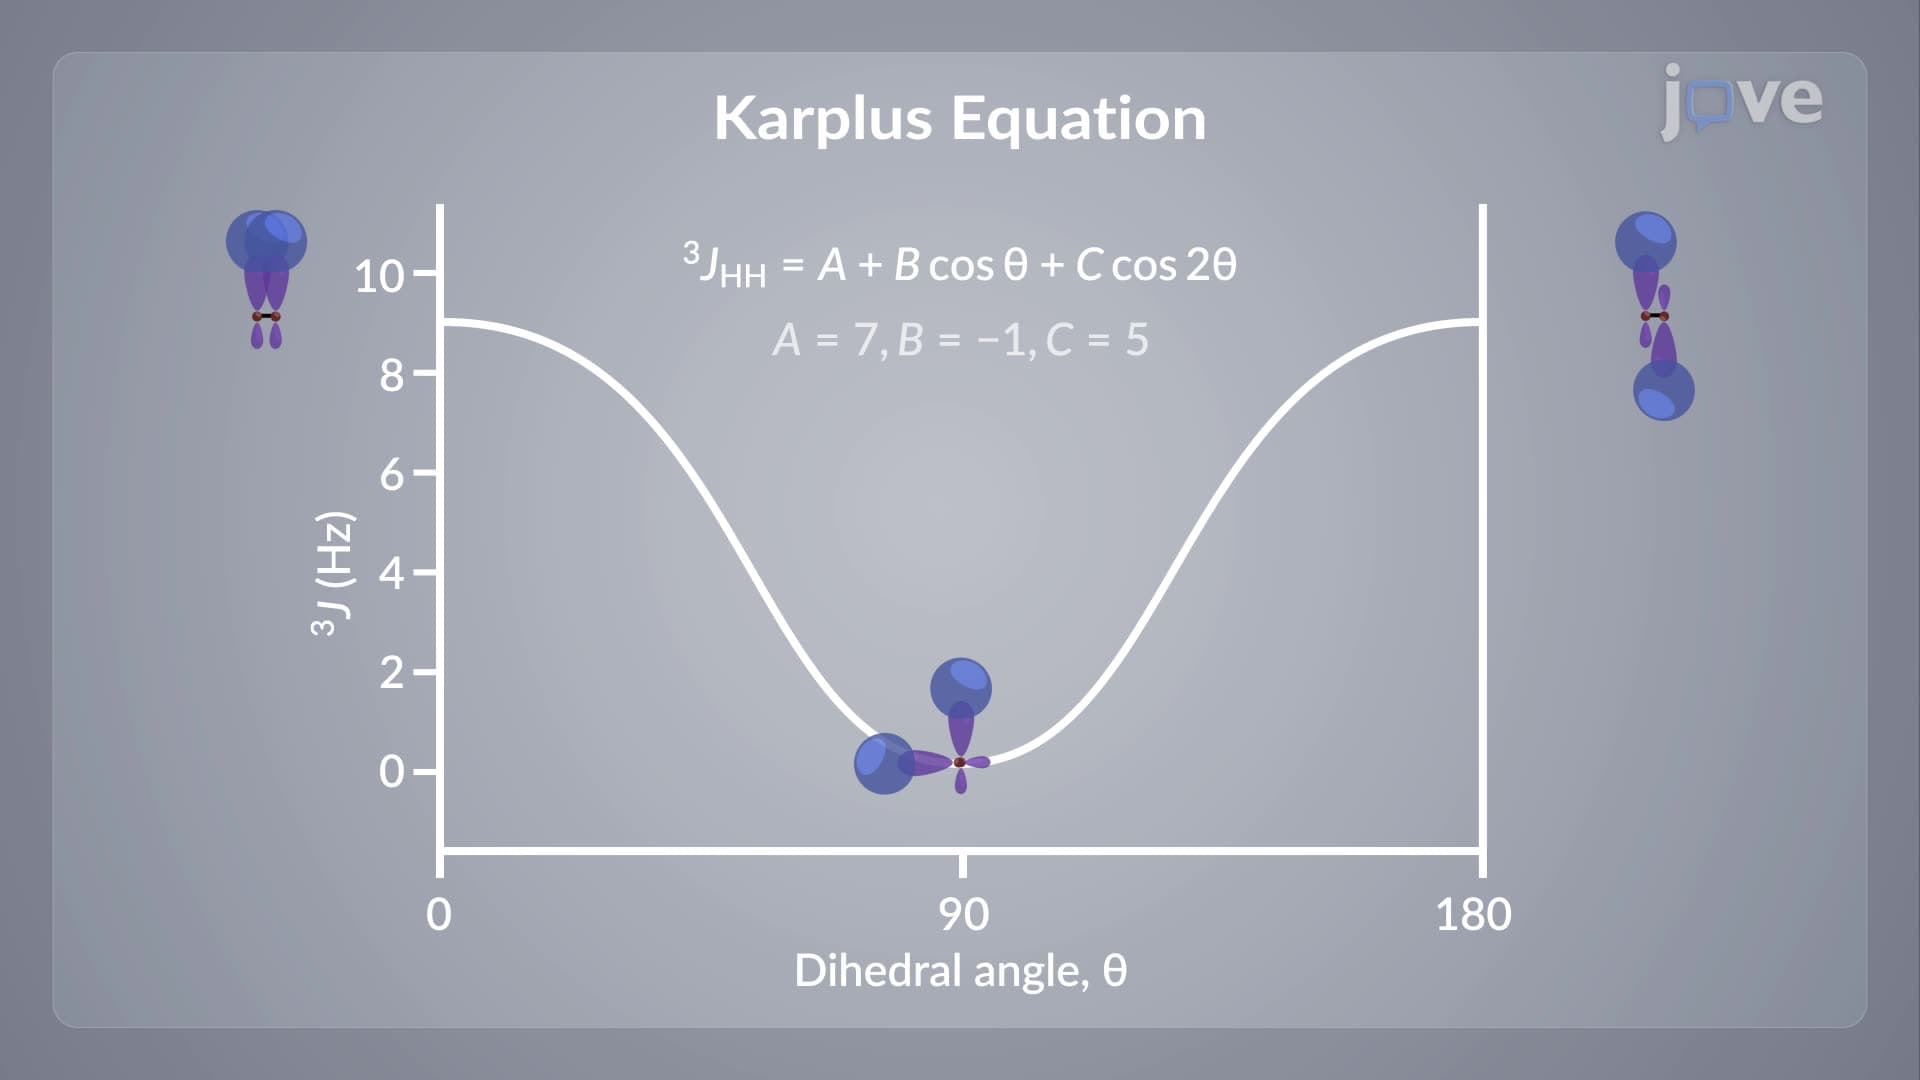
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...
 01:17
Spin–Spin Coupling: One-Bond Coupling
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...