Related Experiment Video
Updated: Jul 11, 2026


Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Quasiclassical trajectory calculations of the OH+NO2 association reaction on a global potential energy surface
Chao Chen1, Benjamin C Shepler, Bastiaan J Braams
1Department of Chemistry and Cherry L. Emerson Center for Scientific Computation, Emory University, Atlanta, Georgia 30322, USA.

Abstract:
We report a full-dimensional potential energy surface (PES) for the OH+NO(2) reaction based on fitting more than 55,000 energies obtained with density functional theory-B3LYP6-311G(d,p) calculations. The PES is invariant with respect to permutation of like nuclei and describes all isomers of HOONO, HONO(2), and the fragments OH+NO(2) and HO(2)+NO. Detailed comparison of the structures, energies, and harmonic frequencies of various stationary points on the PES are made with previous and present high-level ab initio calculations. Two hydrogen-bond complexes are found on the PES and confirmed by new ab initio CASPT2 calculations. Quasiclassical trajectory calculations of the cross sections for ground rovibrational OH+NO(2) association reactions to form HOONO and HONO(2) are done using this PES. The cross section to form HOONO is larger than the one to form HONO(2) at low collision energies but the reverse is found at higher energies. The enhancement of the HOONO complex at low collision energies is shown to be due, in large part, to the transient formation of a H-bond complex, which decays preferentially to HOONO. The association cross sections are used to obtain rate constants for formation of HOONO and HONO(2) for the ground rovibrational states in the high-pressure limit.

Related Concept Videos
 01:26
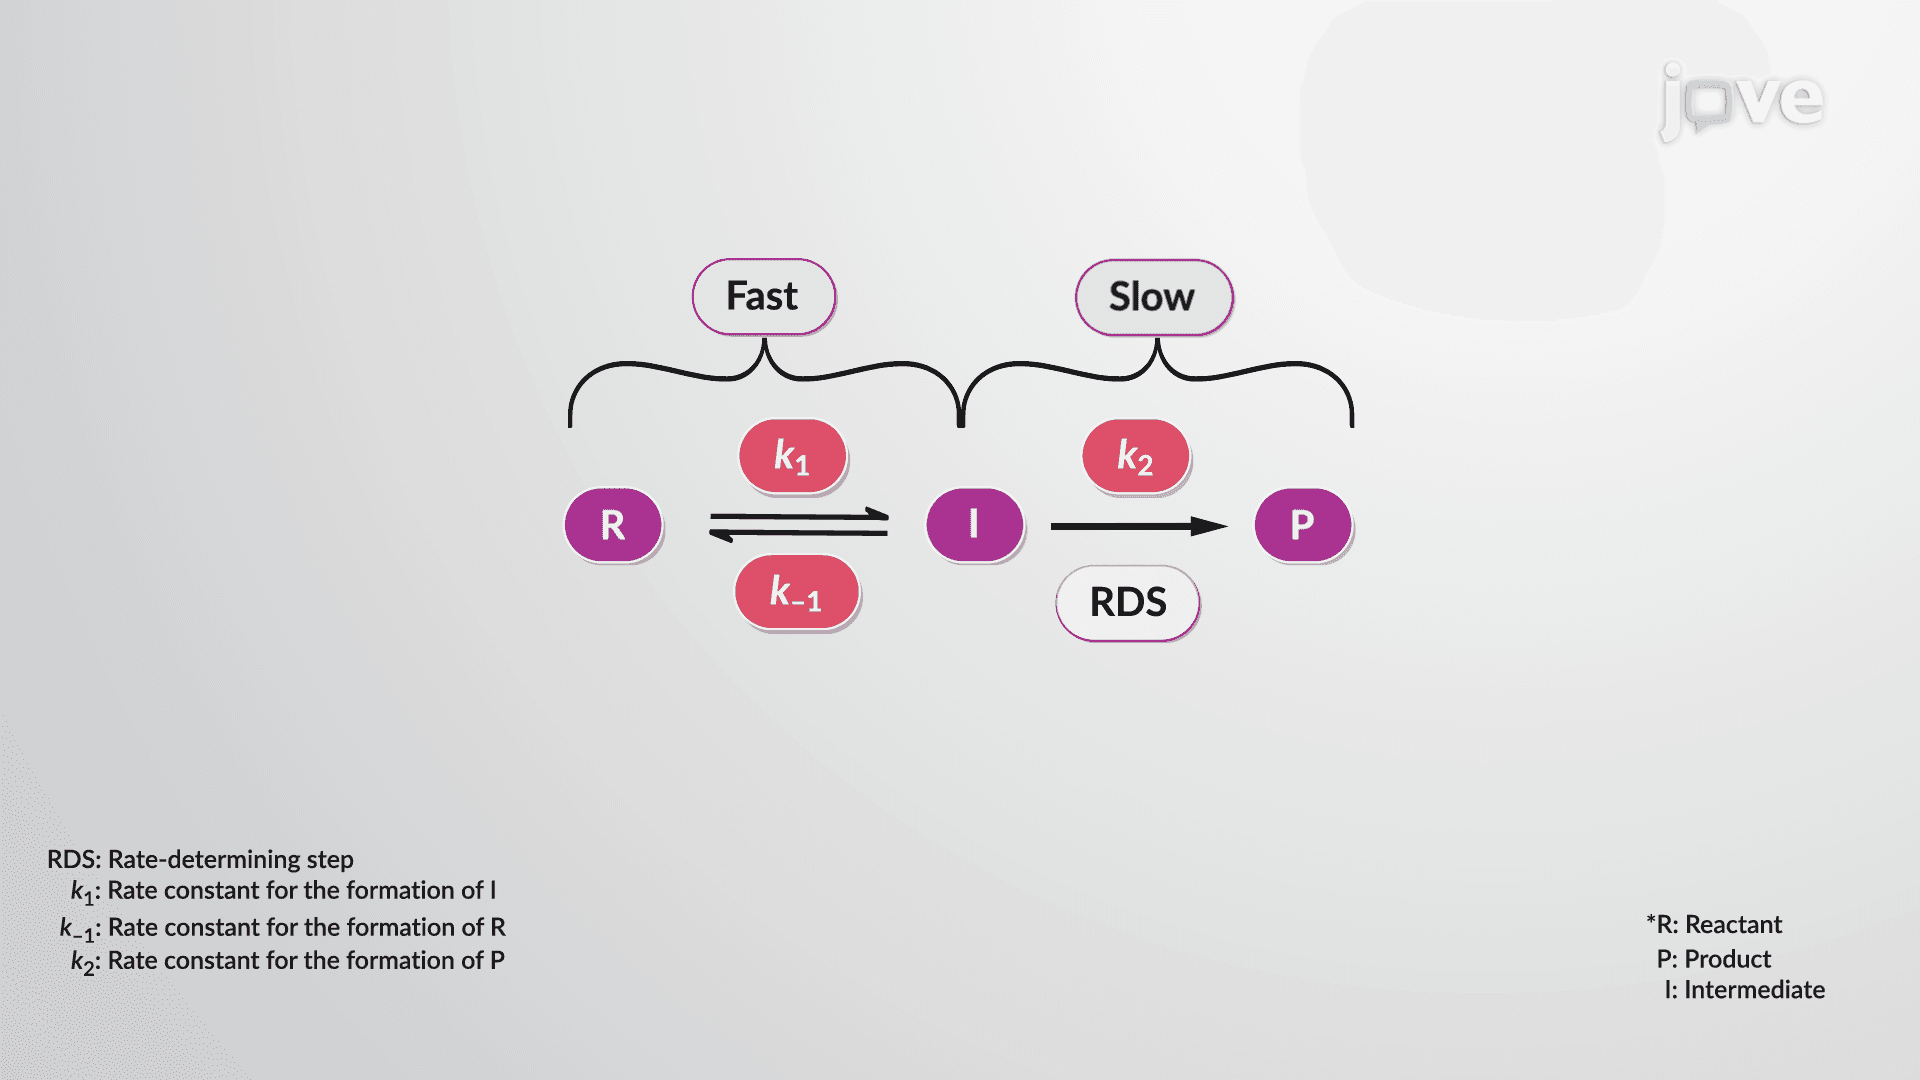
Reaction Mechanisms: The Steady-State Approximation
 03:51
Molecular Orbital Theory II
 02:34
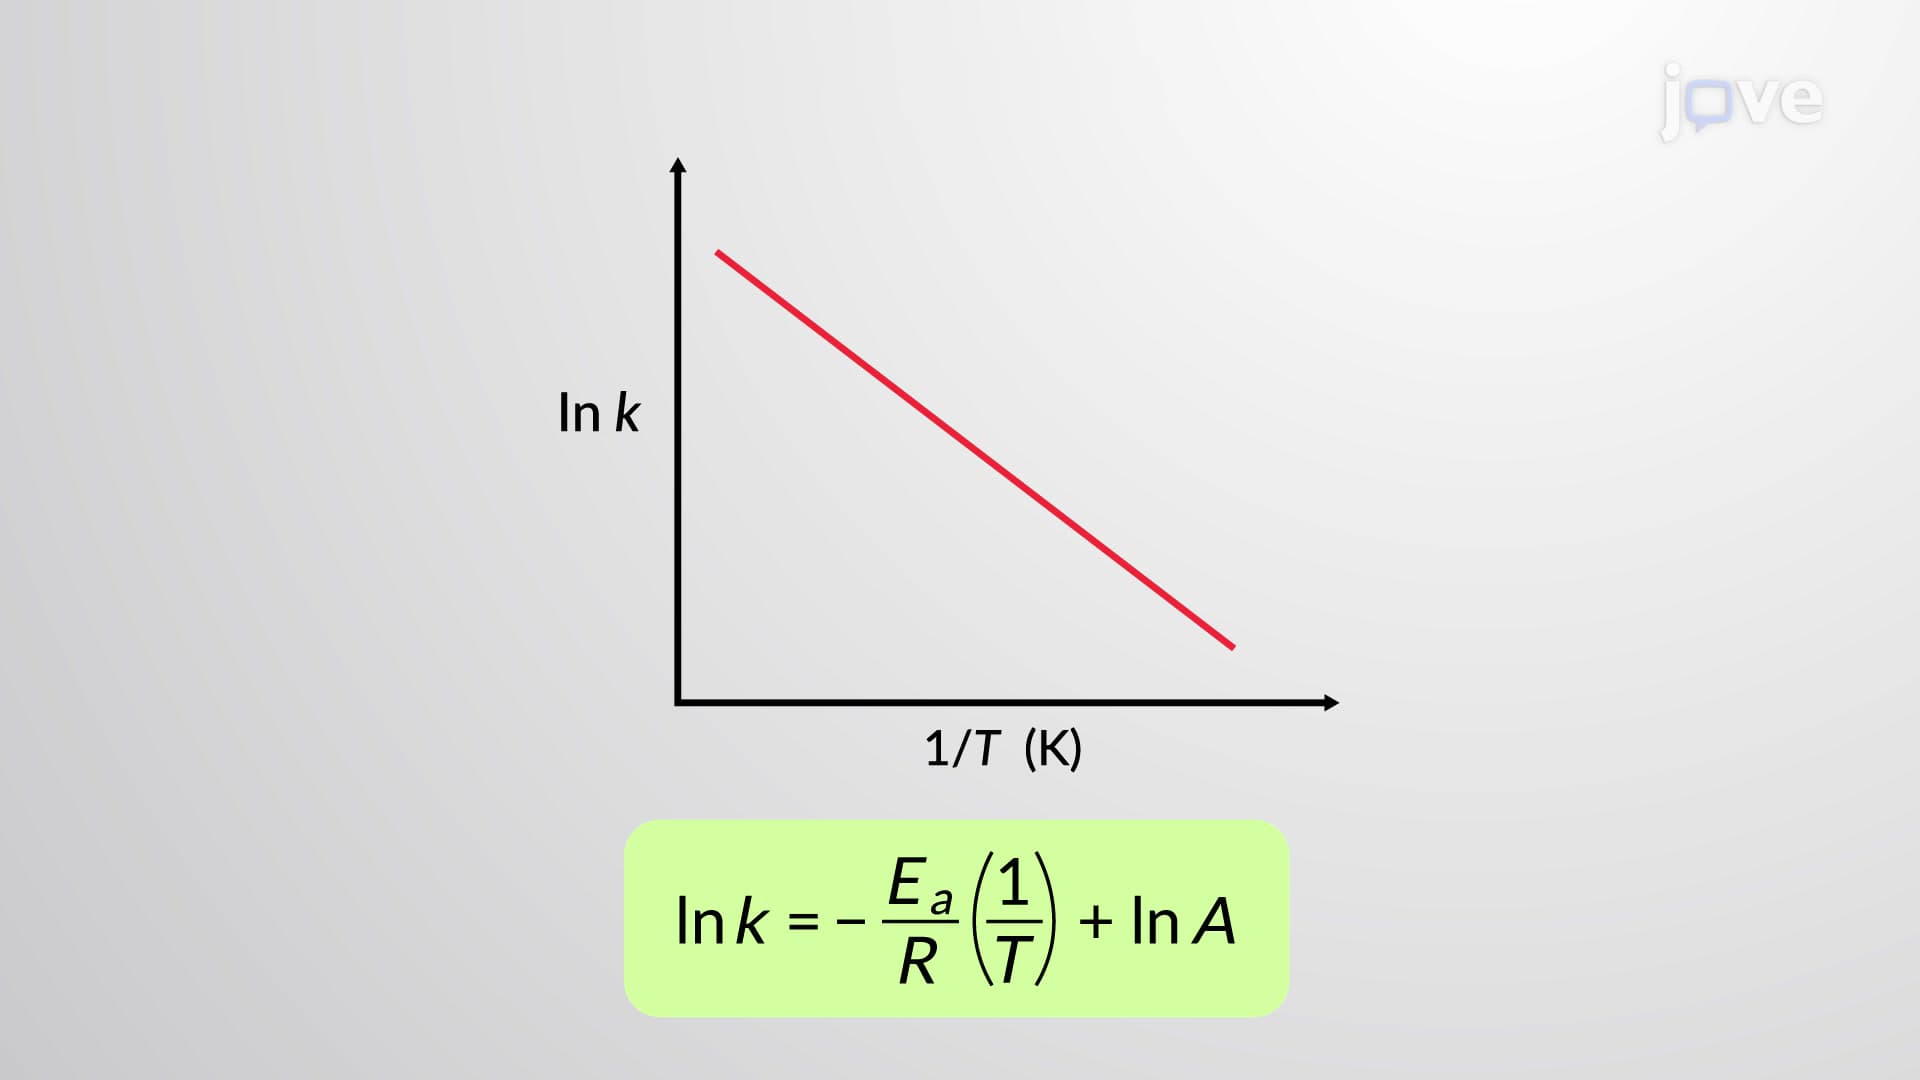
Arrhenius Plots
The Arrhenius equation can be used to...
 01:16
Potential-Energy Criterion for Equilibrium
 02:35
Molecular Orbital Theory I
 01:25
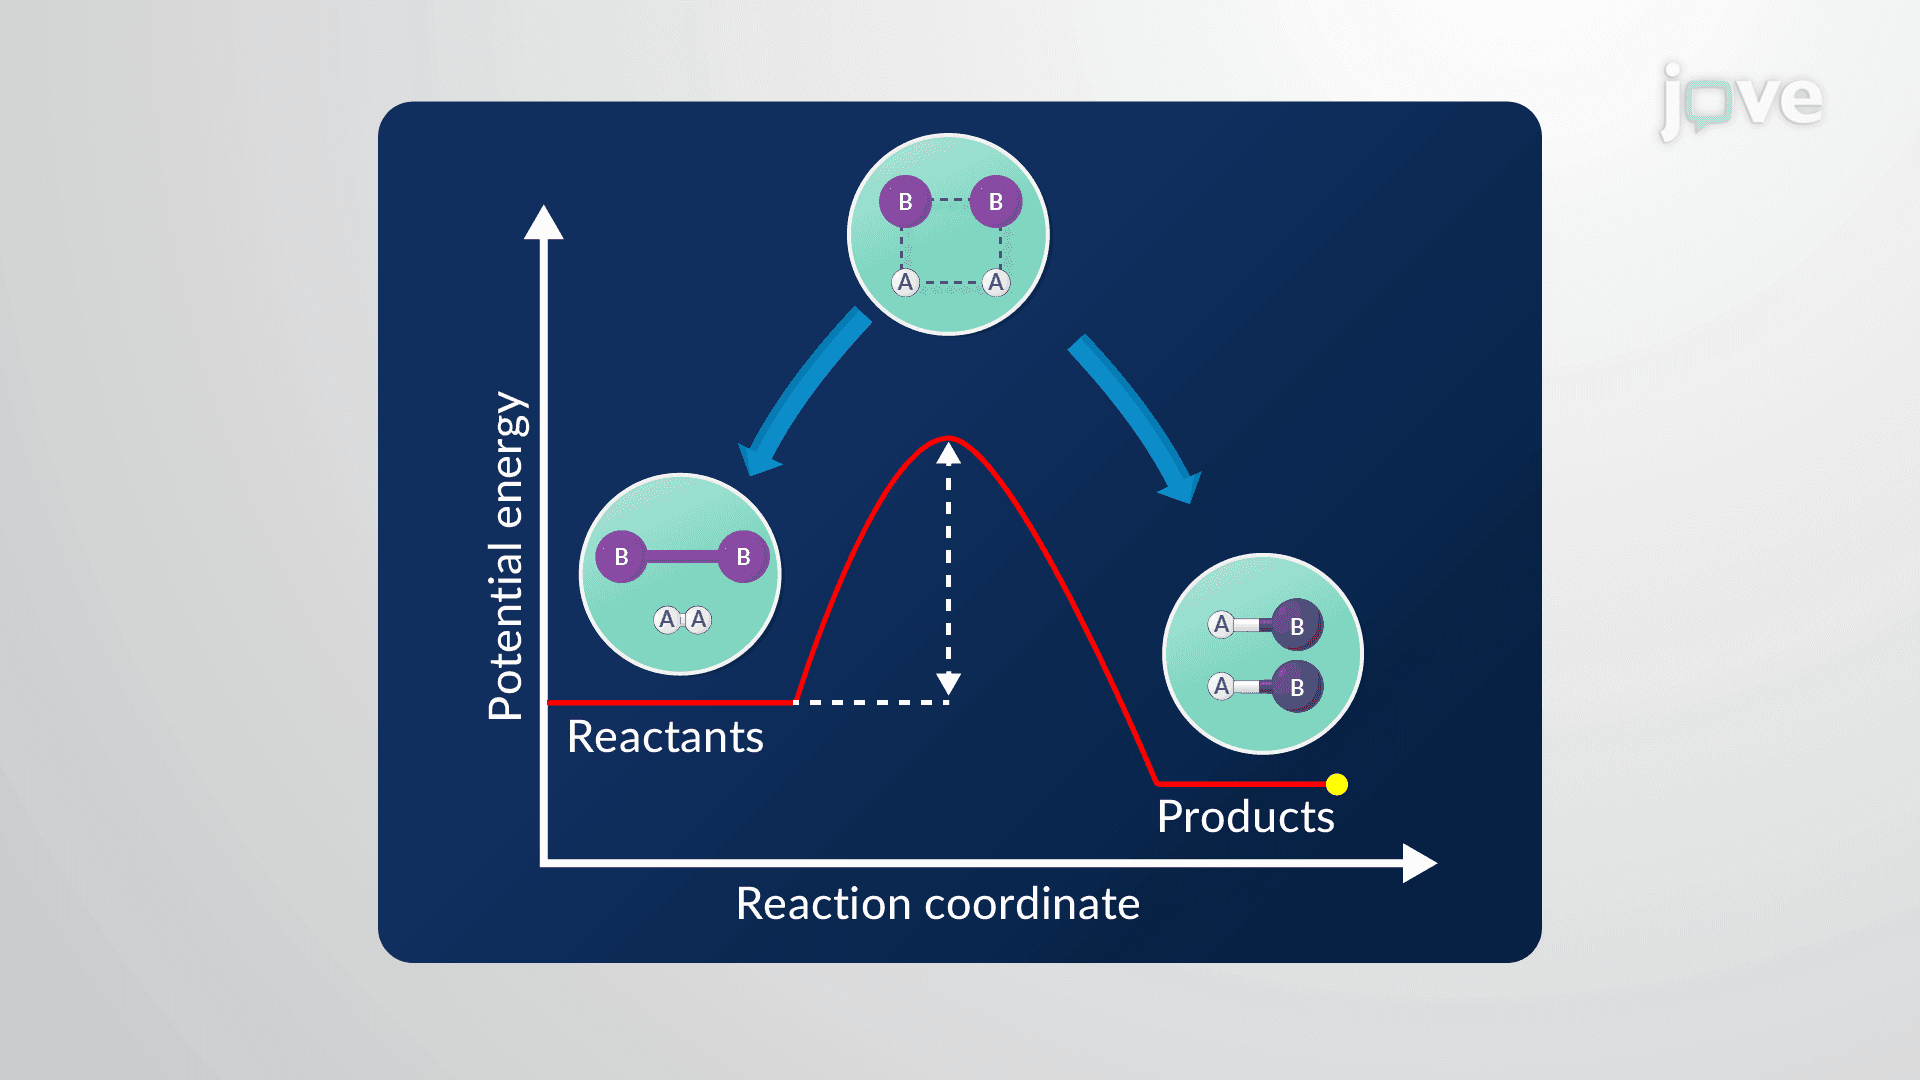
Transition State Theory