Related Experiment Video
Updated: Jul 10, 2026


Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
The origin of deficiency of the supermolecule second-order Moller-Plesset approach for evaluating interaction
Slawomir M Cybulski1, Marion L Lytle
1Department of Chemistry and Biochemistry, Miami University, Oxford, OH 45056, USA. cybulssm@muohio.edu

Abstract:
Calculations for the complex of thymine and adenine are used to show that the supermolecule second-order Moller-Plesset perturbation theory (MP2) approach for evaluating interaction energies fails in certain cases because of the behavior of one of its components: the uncoupled Hartree-Fock dispersion energy. A simple approach for correcting the MP2 supermolecule interaction energies is proposed. It focuses on correcting a relatively small difference between the MP2 and coupled cluster interaction energies, which is a very appealing feature of the new approach considering a benchmark role played by coupled cluster results.
More Related Videos


Related Concept Videos
 02:40
MO Theory and Covalent Bonding
 03:51
Molecular Orbital Theory II
 02:35
Molecular Orbital Theory I
 01:08
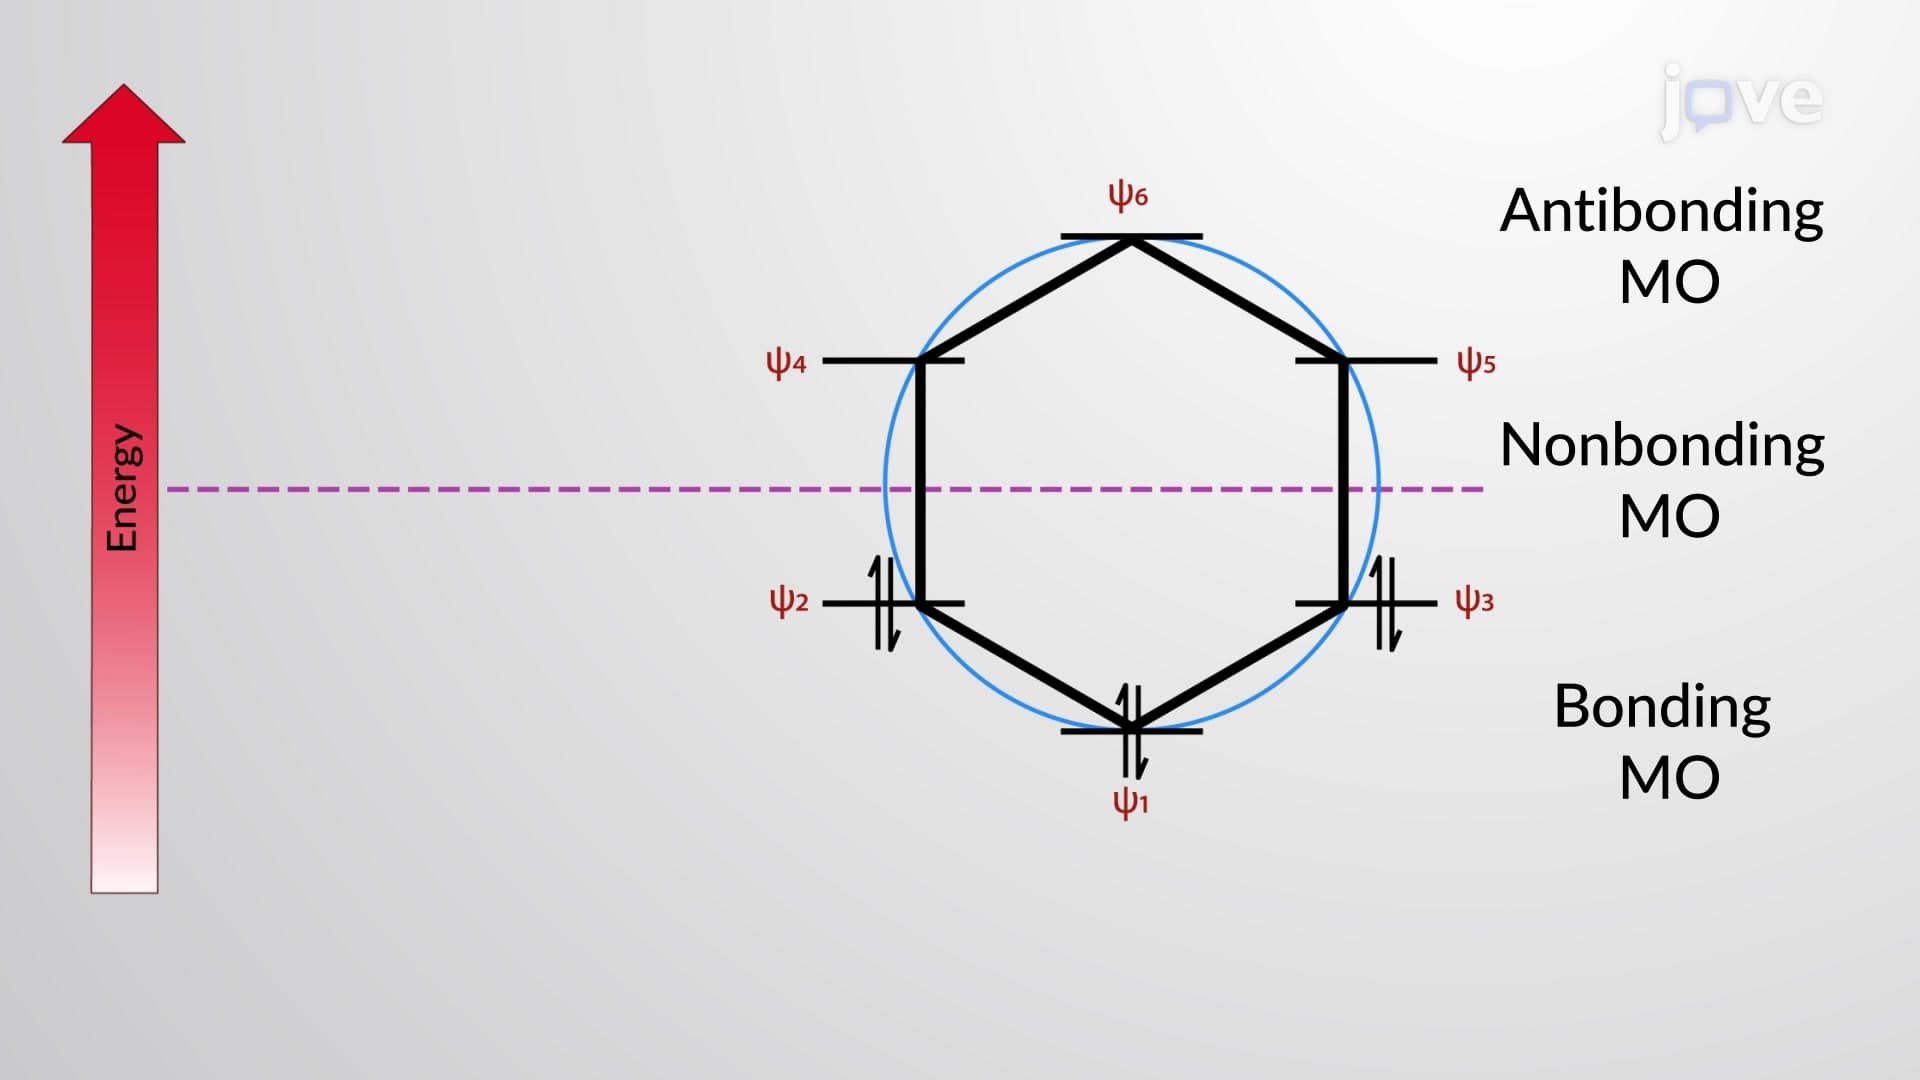
Hückel's Rule Diagram of π MOs: Frost Circle
A Frost circle is constructed by drawing a polygon whose number of edges is equal to the number of carbons of the given cyclic system, with one of the vertices pointing down. Then, a circle is drawn enclosing the polygon so that...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...