Related Experiment Video
Updated: Jul 10, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Energies of ions in water and nanopores within density functional theory
1Sandia National Laboratories, MS 1415, Albuquerque, New Mexico 87185, USA. kleung@sandia.gov

Abstract:
Accurate calculations of electrostatic potentials and treatment of substrate polarizability are critical for predicting the permeation of ions inside water-filled nanopores. The ab initio molecular dynamics method, based on density functional theory (DFT), accounts for the polarizability of materials, water, and solutes, and it should be the method of choice for predicting accurate electrostatic energies of ions. In practice, DFT coupled with the use of periodic boundary conditions in a charged system leads to large energy shifts. Results obtained using different DFT packages may vary because of the way pseudopotentials and long-range electrostatics are implemented. Using maximally localized Wannier functions, we apply robust corrections that yield relatively unambiguous ion energies in select molecular and aqueous systems and inside carbon nanotubes. Large binding energies are predicted for ions in metallic carbon nanotube arrays, while Na+ and Cl- energies are found to exhibit asymmetry in water that is smaller than but comparable with those computed using nonpolarizable water force fields.
More Related Videos

Related Concept Videos
 03:13
Intermolecular Forces
 01:23
Theory of Strong Electrolytes
 03:21
The Energies of Atomic Orbitals
 02:42
Aqueous Solutions and Heats of Hydration
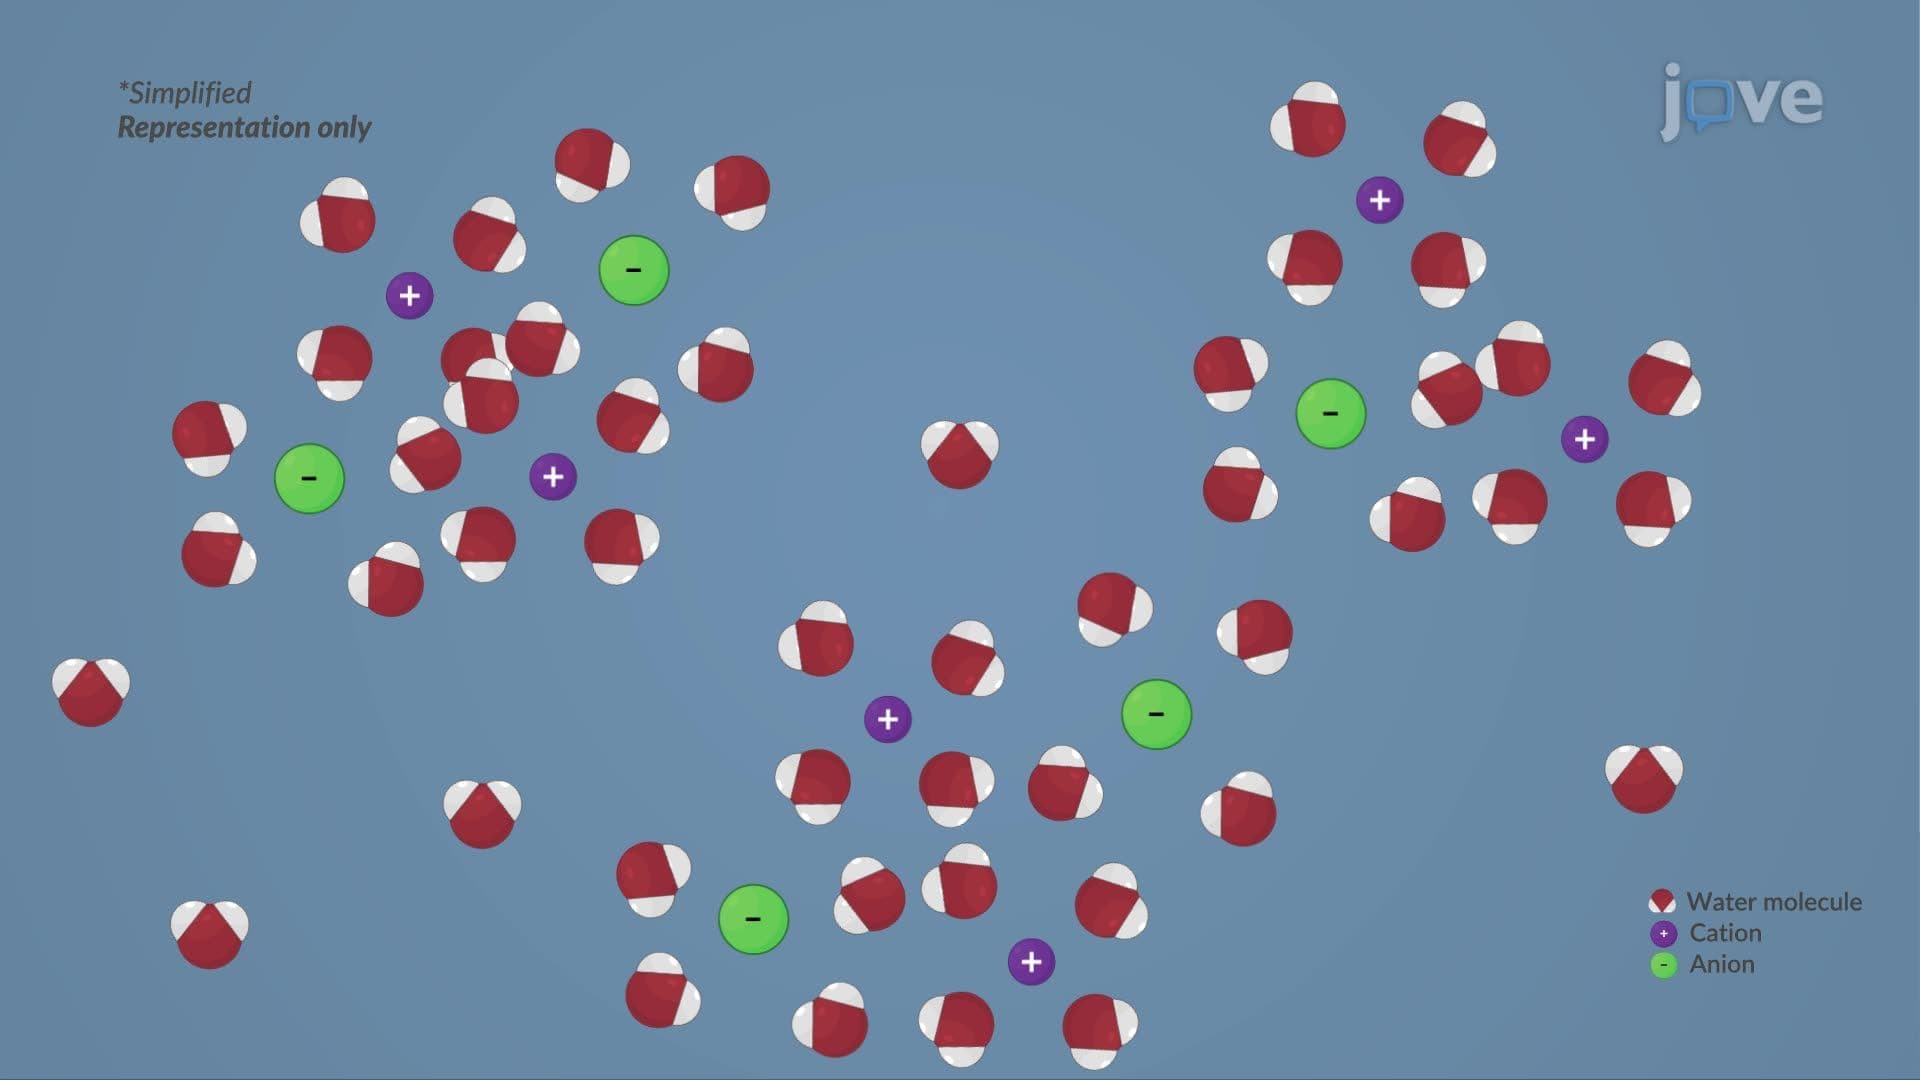
When ionic compounds dissolve in water, the ions in the solid separate and disperse uniformly throughout the solution because water molecules surround and solvate the ions, reducing the strong electrostatic forces between them. This process...
 02:45
The Quantum-Mechanical Model of an Atom
 02:54
Trends in Lattice Energy: Ion Size and Charge