Identifying evolutionary trees and substitution parameters for the general Markov model with invariable sites
Related Experiment Video
Updated: Jul 10, 2026
Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
Elizabeth S Allman1, John A Rhodes
1Department of Mathematics and Statistics, University of Alaska Fairbanks, Fairbanks, AK 99775, USA. e.allman@uaf.edu

Abstract:
The general Markov plus invariable sites (GM+I) model of biological sequence evolution is a two-class model in which an unknown proportion of sites are not allowed to change, while the remainder undergo substitutions according to a Markov process on a tree. For statistical use it is important to know if the model is identifiable; can both the tree topology and the numerical parameters be determined from a joint distribution describing sequences only at the leaves of the tree? We establish that for generic parameters both the tree and all numerical parameter values can be recovered, up to clearly understood issues of 'label swapping'. The method of analysis is algebraic, using phylogenetic invariants to study the variety defined by the model. Simple rational formulas, expressed in terms of determinantal ratios, are found for recovering numerical parameters describing the invariable sites.
More Related Videos
Related Concept Videos
 03:21
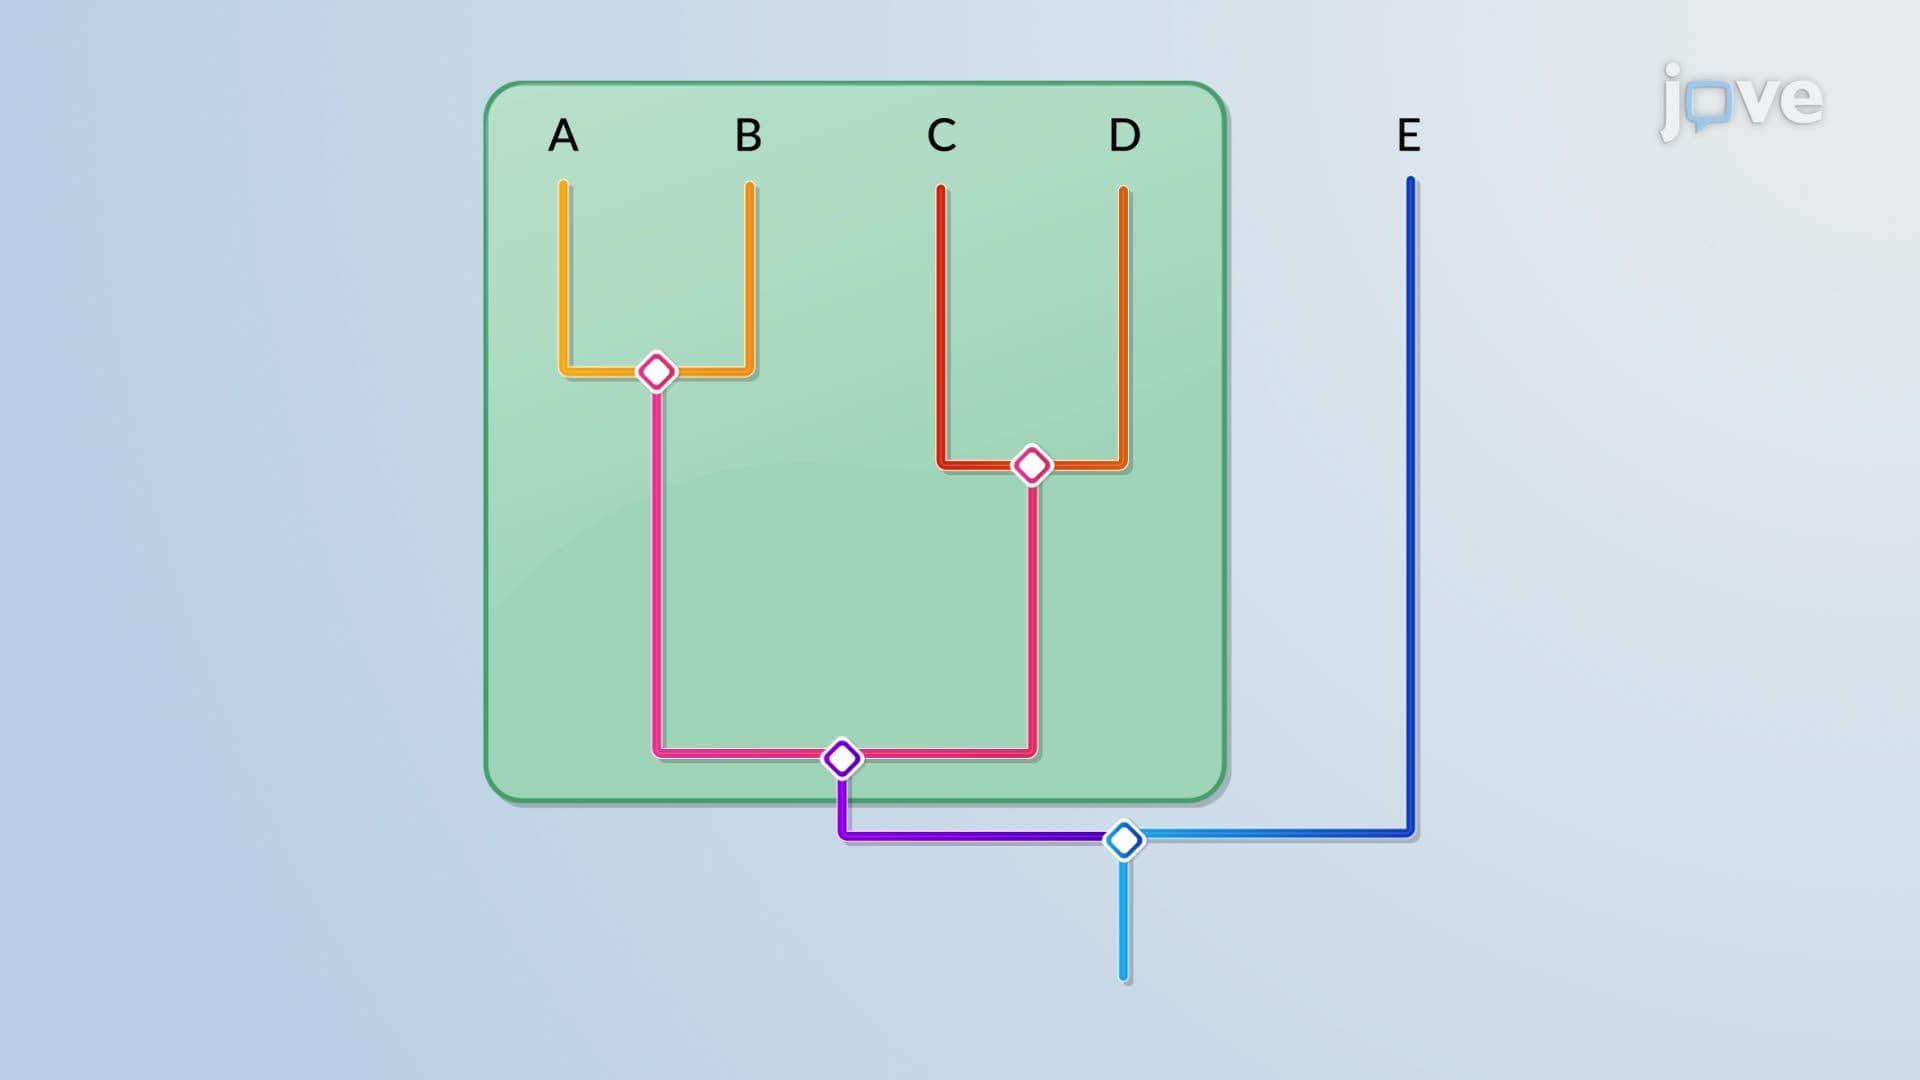
Phylogenetic Trees
03:21Phylogenetic Trees
 01:28
Microbial Phylogeny
 02:54
Evolutionary Relationships through Genome Comparisons
 01:19
Survival Tree
Building a Survival Tree
Constructing a survival tree begins...
 02:13
Overview of Transposition and Recombination