Related Experiment Video
Updated: Jul 9, 2026

Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
Phylogenetic analysis using complete signature information of whole genomes and clustered Neighbour-Joining method
Xiaomeng Wu1, Xiu-Feng Wan, Gang Wu
1Bioinformatics Research Group, Department of Computing Science, University of Alberta-Edmonton, Alberta, Canada. xiaomeng@cs.ualberta.ca

Abstract:
A new method called Complete Composition Vector (CCV), which is a collection of Composition Vectors (CV), is described to infer evolutionary relationships between species using their complete genomic sequences. Such a method bypasses the complexity of performing multiple sequence alignments and avoids the ambiguity of choosing individual genes for species tree construction. It is expected to effectively retain the rich evolutionary information contained in the whole genomic sequence. The method was applied to infer the evolutionary footprints for several datasets that have been previously studied. The final phylogenies were built by an improved clustered Neighbour-Joining method. The generated phylogenetic trees are highly consistent with taxonomy hierarchy and previous studies, with some biologically interesting disagreements.
Related Concept Videos
 02:54
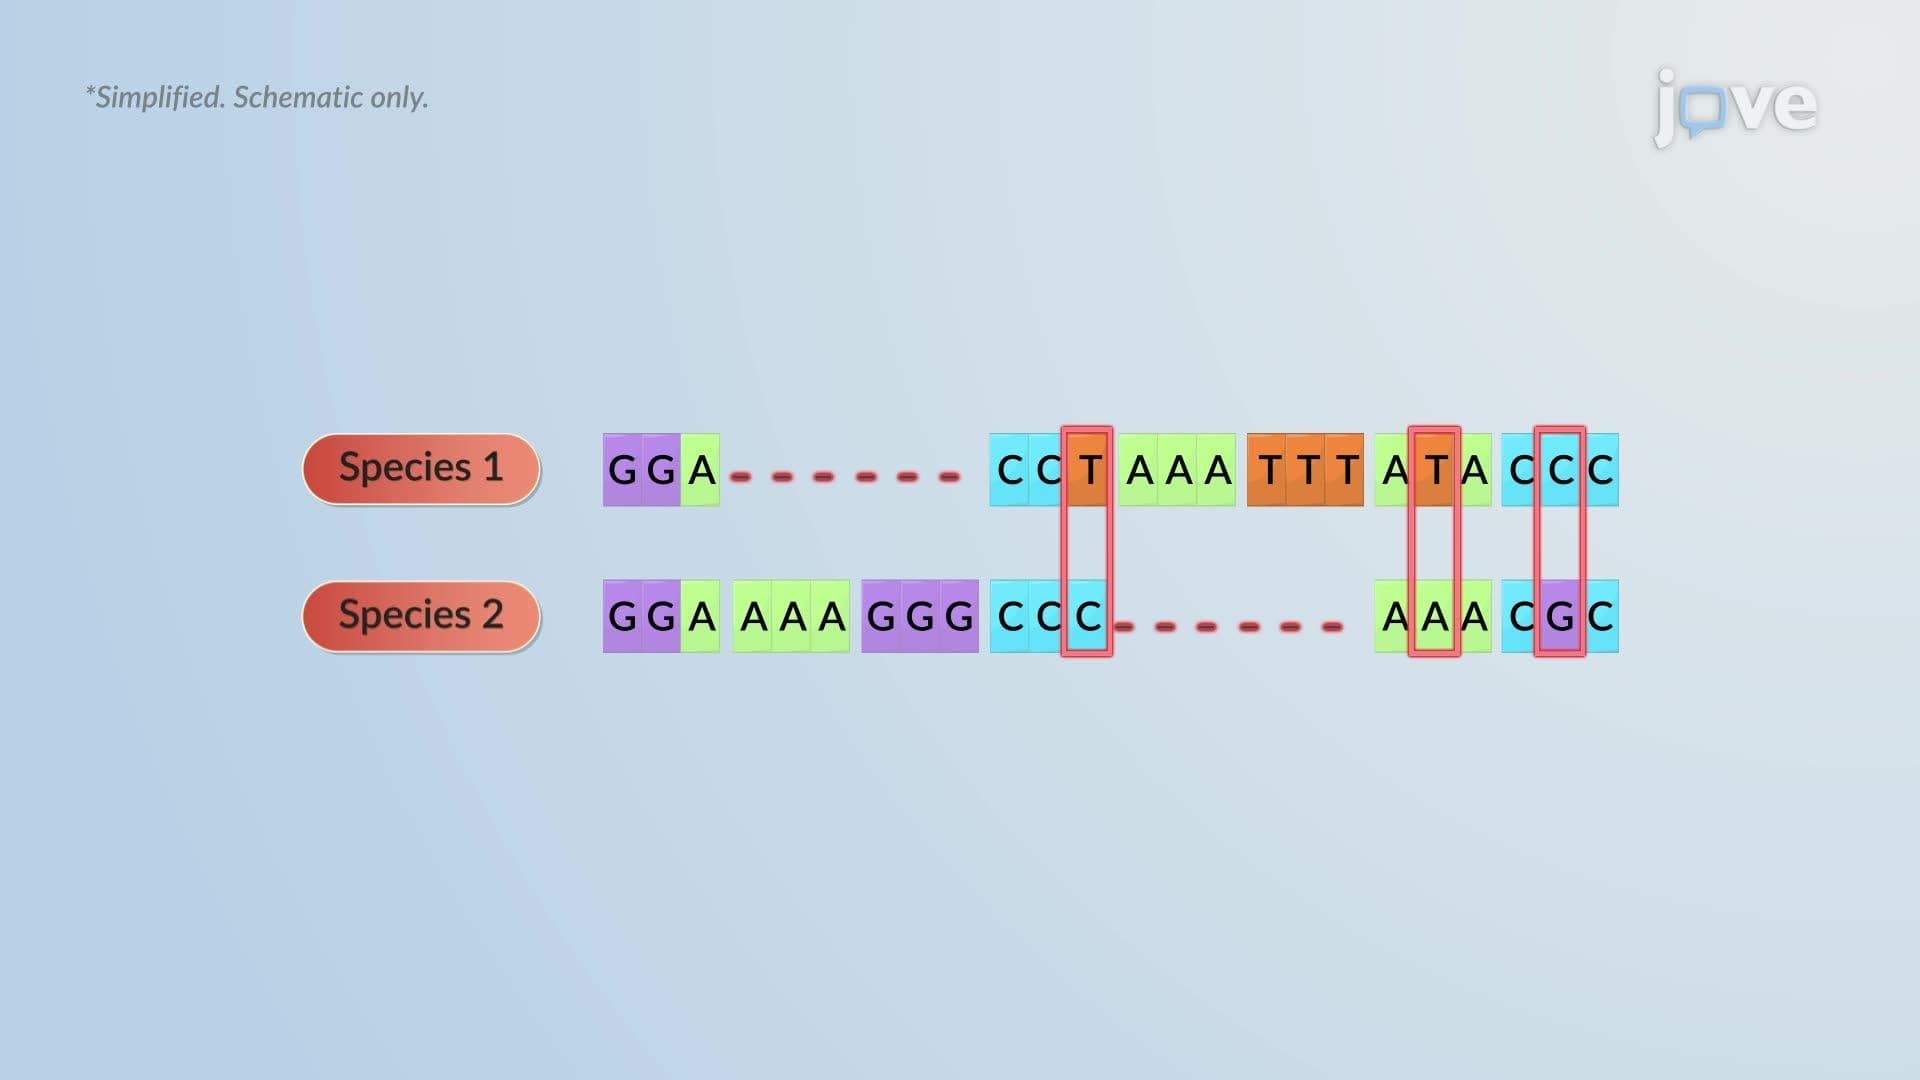
Evolutionary Relationships through Genome Comparisons
 01:29
Modern Molecular Taxonomy
 01:28
Microbial Phylogeny
 02:02
Genomics
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...
 01:20
Applications of Molecular Taxonomy