Related Experiment Video
Updated: Jul 9, 2026

Synthesizing Amino Acids Modified with Reactive Carbonyls in Silico to Assess Structural Effects Using Molecular Dynamics Simulations
Published on: April 26, 2024
Using quantum mechanics to improve estimates of amino acid side chain rotamer energies
P Douglas Renfrew1, Glenn L Butterfoss, Brian Kuhlman
1Department of Biochemistry and Biophysics, University of North Carolina at Chapel Hill, Chapel Hill, North Carolina 27599-7260, USA.

Abstract:
Amino acid side chains adopt a discrete set of favorable conformations typically referred to as rotamers. The relative energies of rotamers partially determine which side chain conformations are more often observed in protein structures and accurate estimates of these energies are important for predicting protein structure and designing new proteins. Protein modelers typically calculate side chain rotamer energies by using molecular mechanics (MM) potentials or by converting rotamer probabilities from the protein database (PDB) into relative free energies. One limitation of the knowledge-based energies is that rotamer preferences observed in the PDB can reflect internal side chain energies as well as longer-range interactions with the rest of the protein. Here, we test an alternative approach for calculating rotamer energies. We use three different quantum mechanics (QM) methods (second order Møller-Plesset (MP2), density functional theory (DFT) energy calculation using the B3LYP functional, and Hartree-Fock) to calculate the energy of amino acid rotamers in a dipeptide model system, and then use these pre-calculated values in side chain placement simulations. Energies were calculated for over 36,000 different conformations of leucine, isoleucine, and valine dipeptides with backbone torsion angles from the helical and strand regions of the Ramachandran plot. In a subset of cases these energies differ significantly from those calculated with standard molecular mechanics potentials or those derived from PDB statistics. We find that in these cases the energies from the QM methods result in more accurate placement of amino acid side chains in structure prediction tests.

Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 02:27
Predicting Molecular Geometry
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 02:07
Improving Translational Accuracy
 01:02
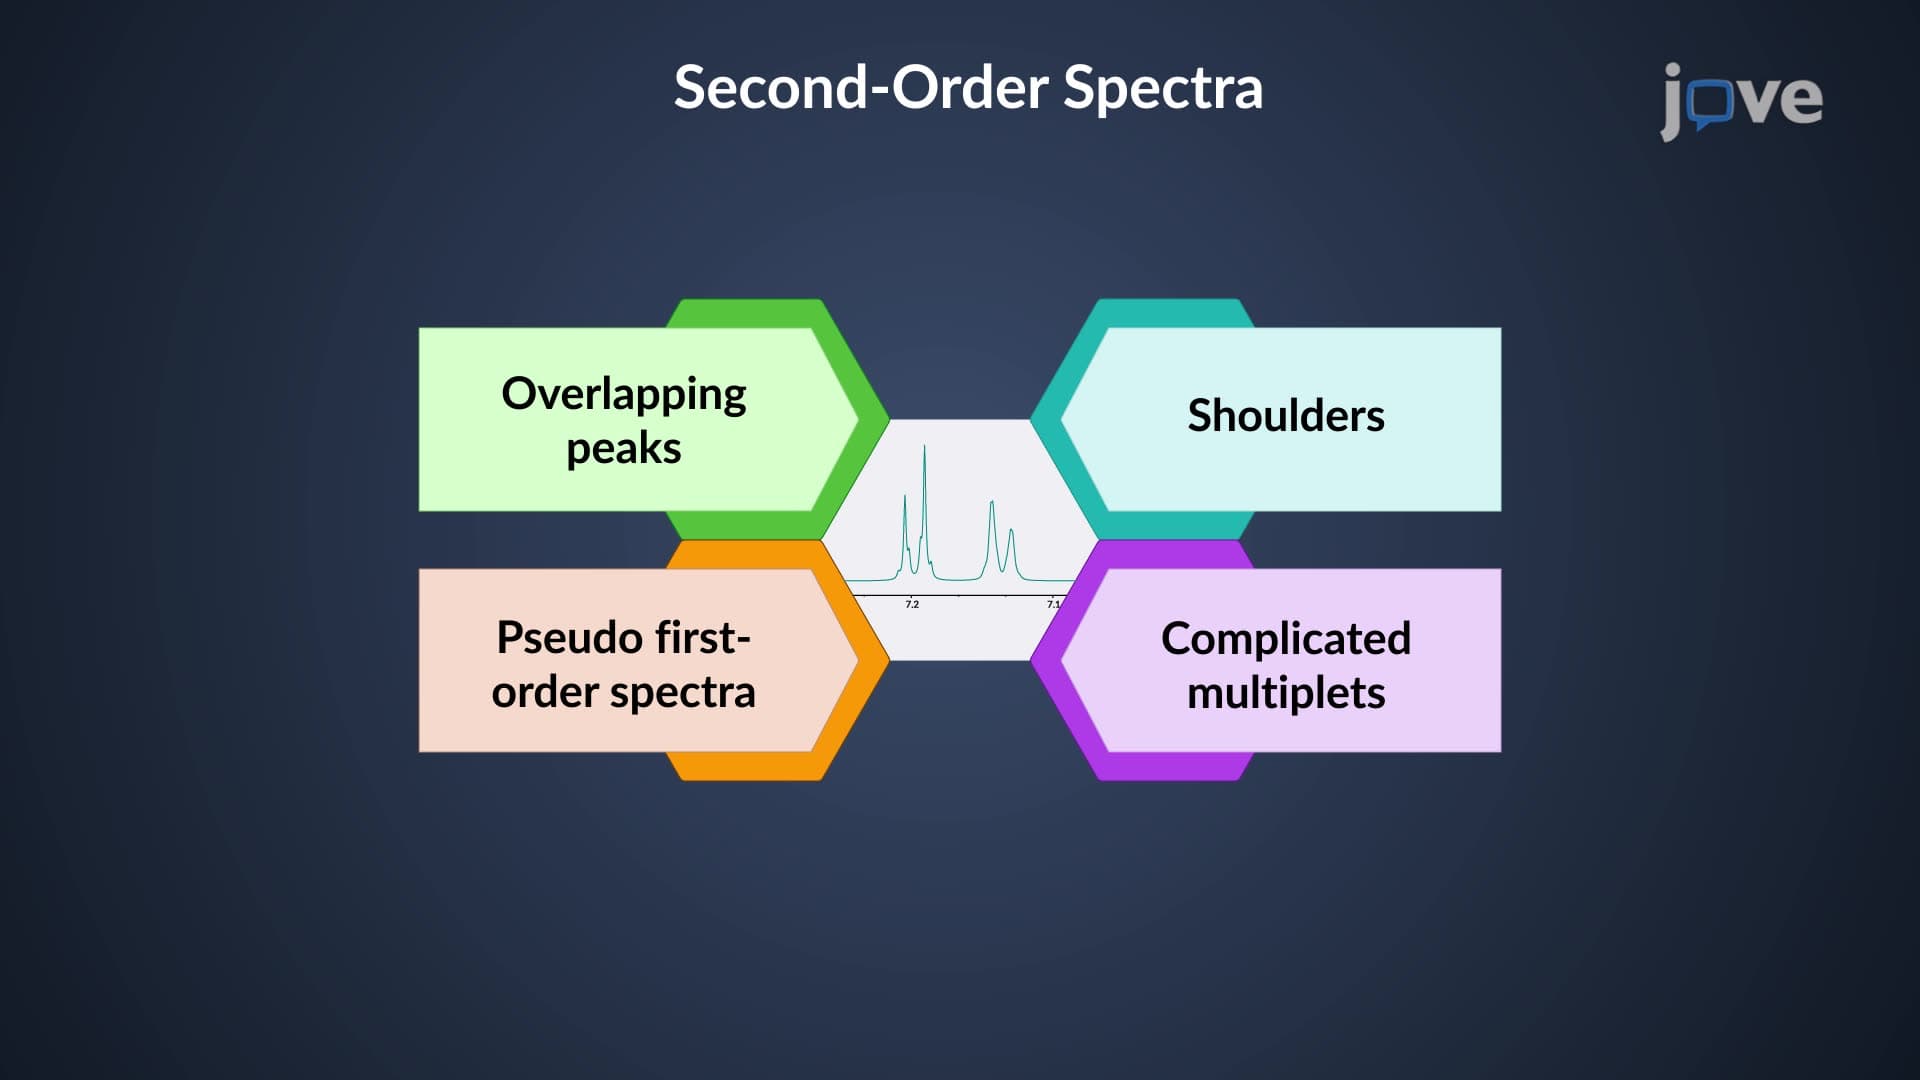
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are slanted or...