Related Experiment Video
Updated: Jul 8, 2026
Reverse Yeast Two-hybrid System to Identify Mammalian Nuclear Receptor Residues that Interact with Ligands and/or Antagonists
Published on: November 15, 2013
Reduction of lipophilicity at the lipophilic domain of RXR agonists enables production of subtype preference:
Kayo Takamatsu1, Atsushi Takano, Nobumasa Yakushiji
1Okayama University Graduate School of Medicine, Dentistry and Pharmaceutical Sciences, 1-1-1, Tsushima-Naka, Okayama 700-8530, Japan.

Abstract:
Retinoid X receptor agonists (RXR agonists, rexinoids) are interesting candidates for the treatment of cancers such as tamoxifen-resistant breast cancer and taxol-resistant lung cancer. However, well-known RXR agonists possess a strong lipophilic character. In addition, although RXR has three subtypes, no subtype-selective RXR agonists are known. Thus we aimed to produce less-lipophilic and subtype-selective RXR agonists. By designing sulfonamide-type RXR agonists, 4-[N-methanesulfonyl-N-(5,5,8,8-tetramethyl-5,6,7,8-tetrahydro-2-naphthyl)amino]benzoic acid (8 a) was found to prefer RXRalpha over RXRbeta and RXRgamma, although the potency is less than the potencies of well-known RXR pan-agonists. Moreover, our results suggest that the reduction of lipophilicity at the hydrophobic interaction region of RXR agonists enables production of RXR subtype preference. Our finding will be useful for the creation of more potent and less-lipophilic subtype-selective RXR agonists aimed at the reduction of undesirable side effects.
Insights
Researchers developed novel Retinoid X receptor (RXR) agonists that are less lipophilic and selective for the RXR-alpha subtype. This finding may lead to more effective cancer treatments with fewer side effects.
Area of Science:
- Medicinal Chemistry
- Molecular Biology
- Oncology
Background:
- Retinoid X receptor (RXR) agonists are potential cancer therapeutics, including for tamoxifen-resistant breast and taxol-resistant lung cancers.
- Existing RXR agonists are highly lipophilic and lack subtype selectivity, limiting their therapeutic potential and increasing side effects.
- RXR receptors have three subtypes (RXRalpha, RXRbeta, RXRgamma), and subtype-selective agonists are currently unknown.
Purpose of the Study:
- To design and synthesize less-lipophilic and subtype-selective RXR agonists.
- To investigate the relationship between lipophilicity and RXR subtype preference.
Main Methods:
- Design and synthesis of novel sulfonamide-type RXR agonists.
- Evaluation of RXR subtype selectivity (RXRalpha, RXRbeta, RXRgamma) for the synthesized compounds.
Main Results:
- A novel sulfonamide-type RXR agonist, compound 8a, demonstrated preference for RXRalpha over RXRbeta and RXRgamma.
- The potency of compound 8a was lower than that of known RXR pan-agonists.
- Reducing lipophilicity in the hydrophobic interaction region of RXR agonists appears to enable subtype preference.
Conclusions:
- The development of less-lipophilic, RXR-alpha-selective agonists is feasible.
- Findings suggest a strategy for creating more potent and less-lipophilic subtype-selective RXR agonists.
- This research could lead to cancer therapies with reduced undesirable side effects.
Related Concept Videos
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:25
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous ligand's action.
 01:31
Transducer Mechanism: Nuclear Receptors
About 48 different soluble family members of nuclear receptors are identified that can be divided into two main classes:
 01:27
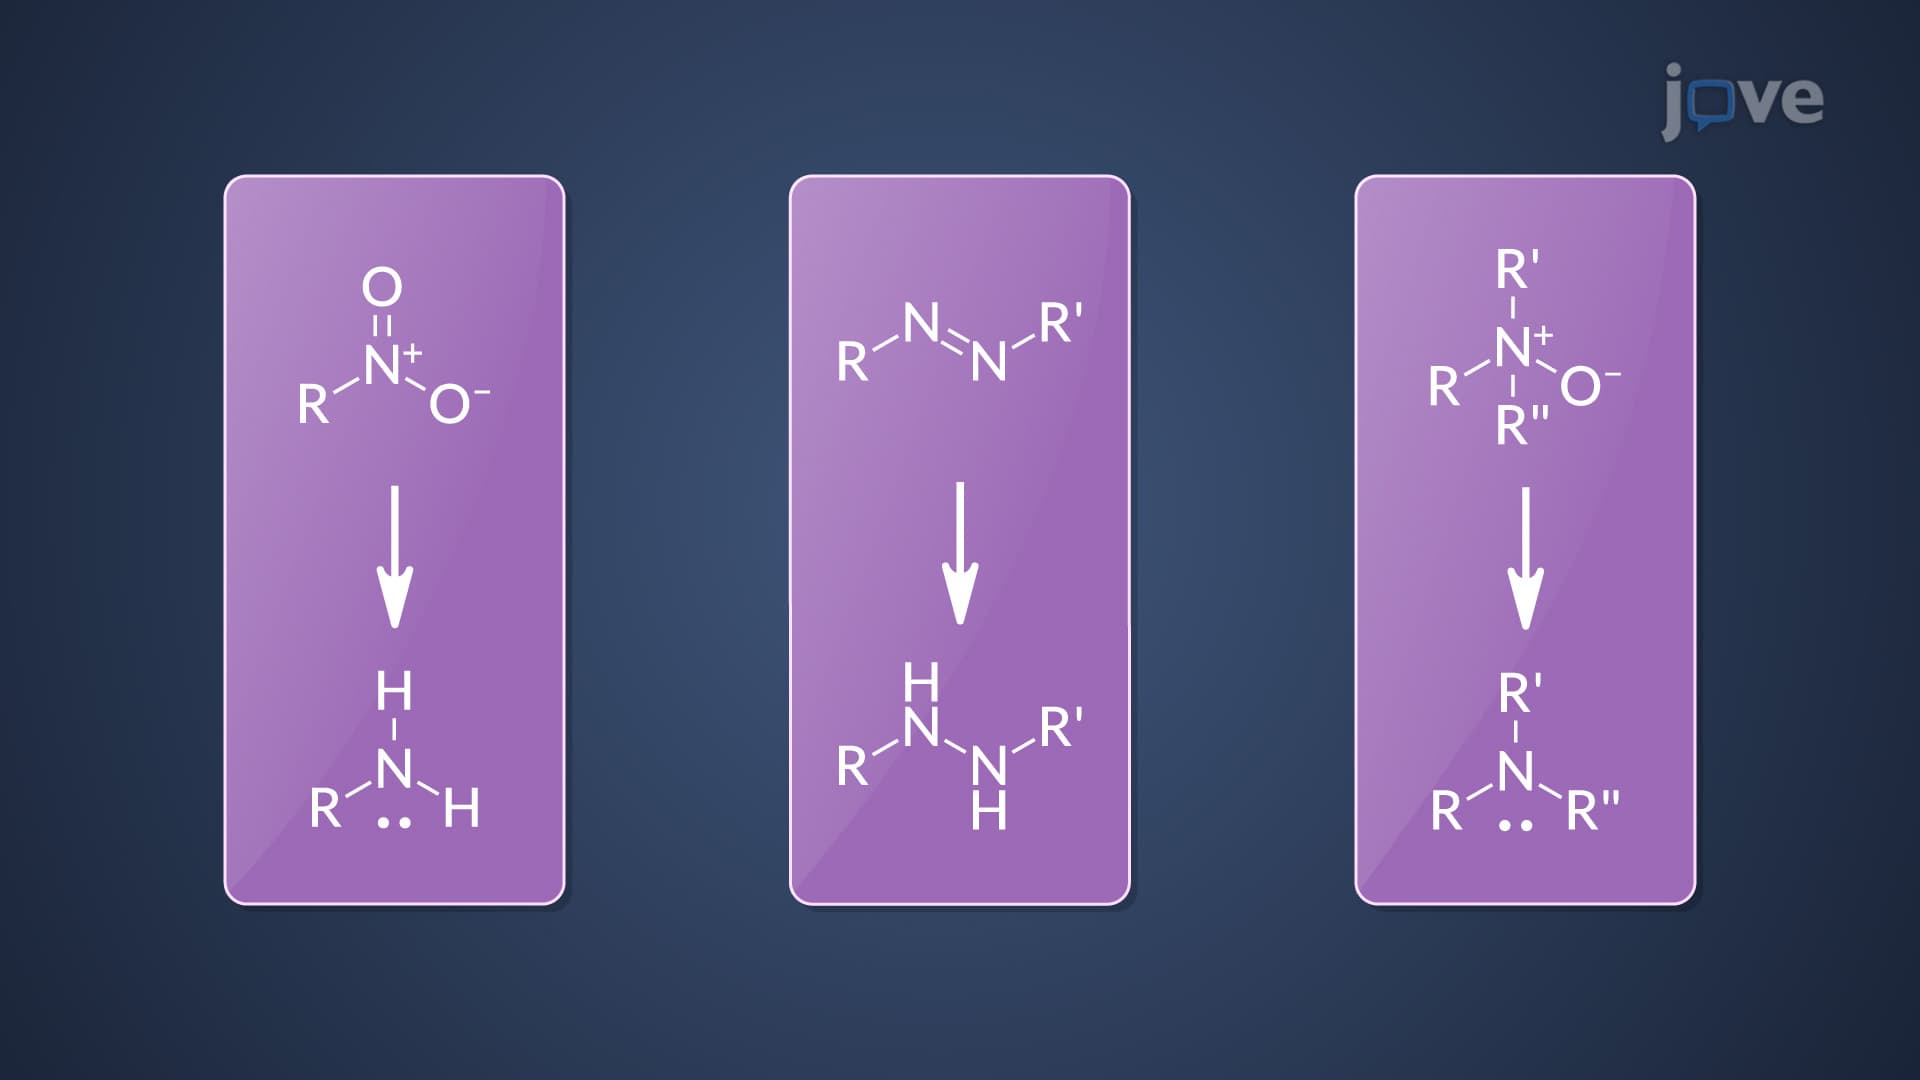
Phase I Reactions: Reductive Reactions
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:17
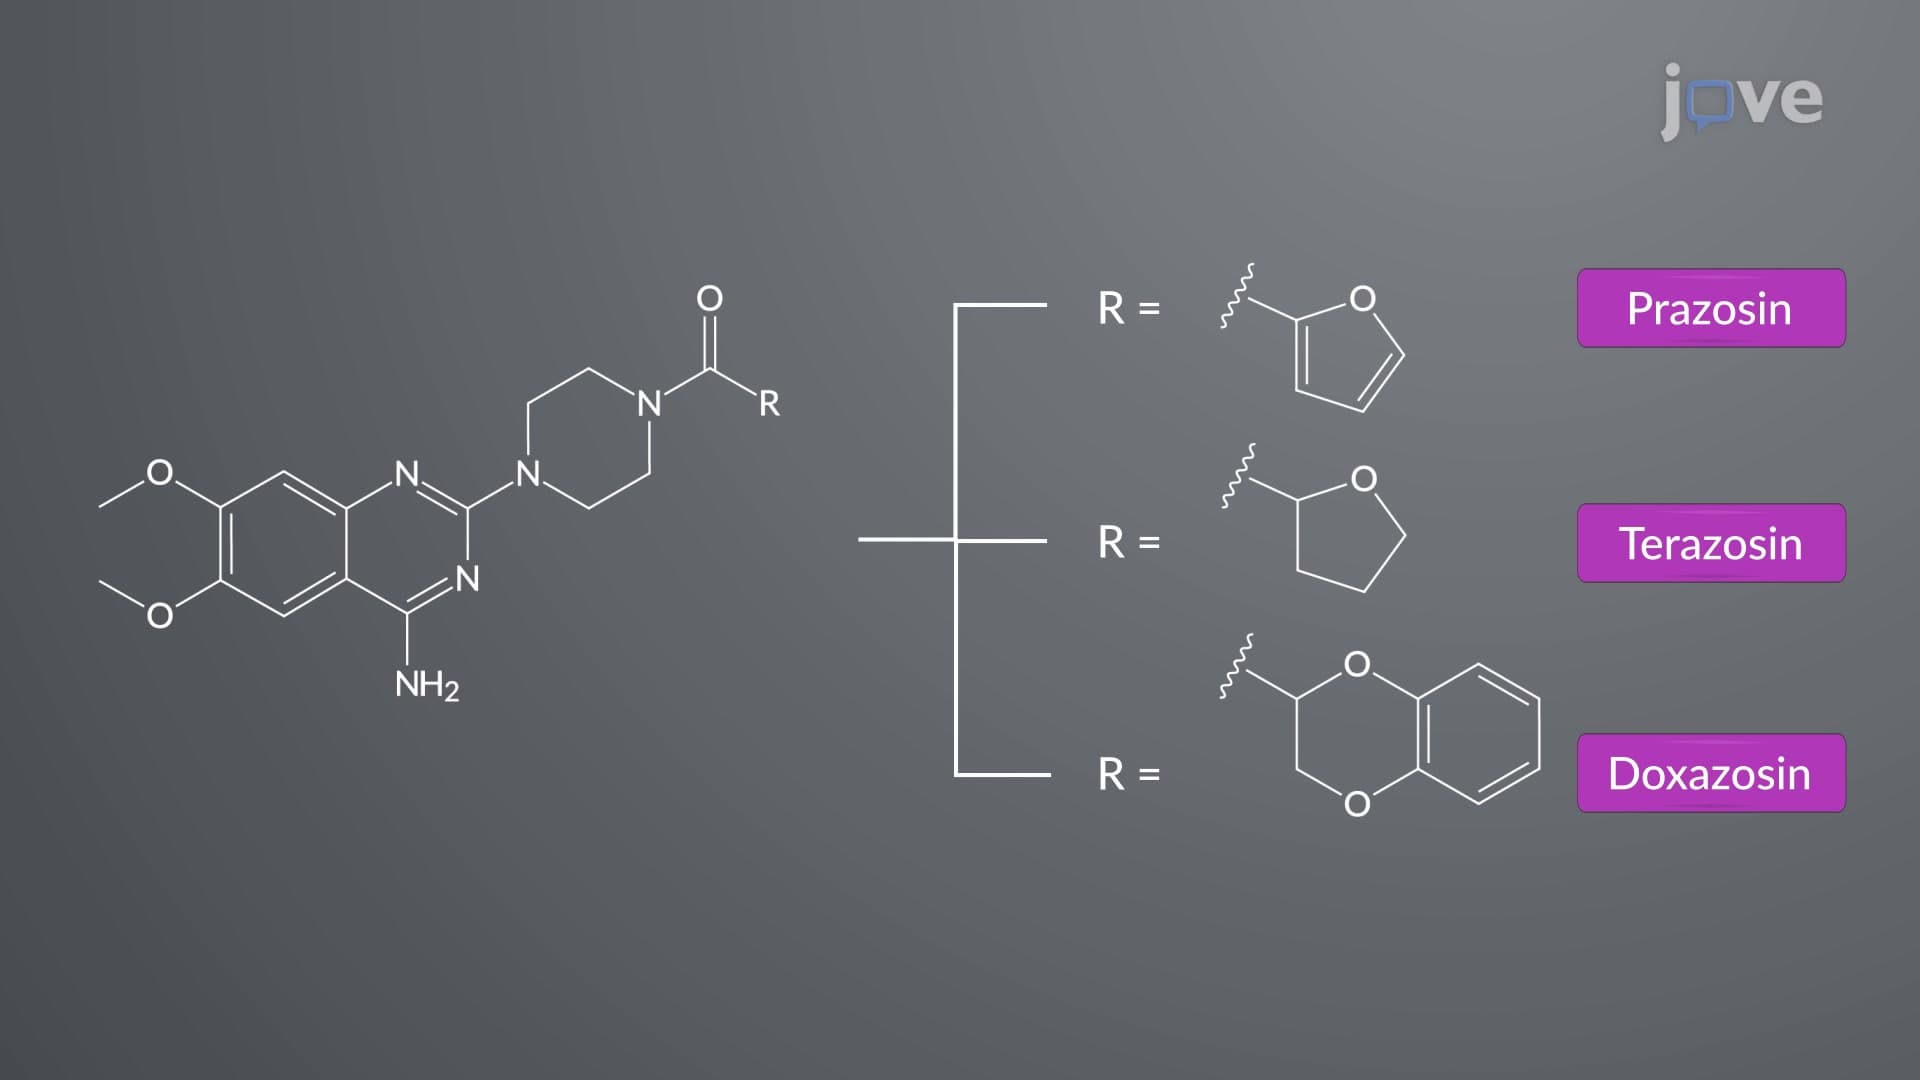
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...