Related Experiment Video
Updated: Jul 8, 2026
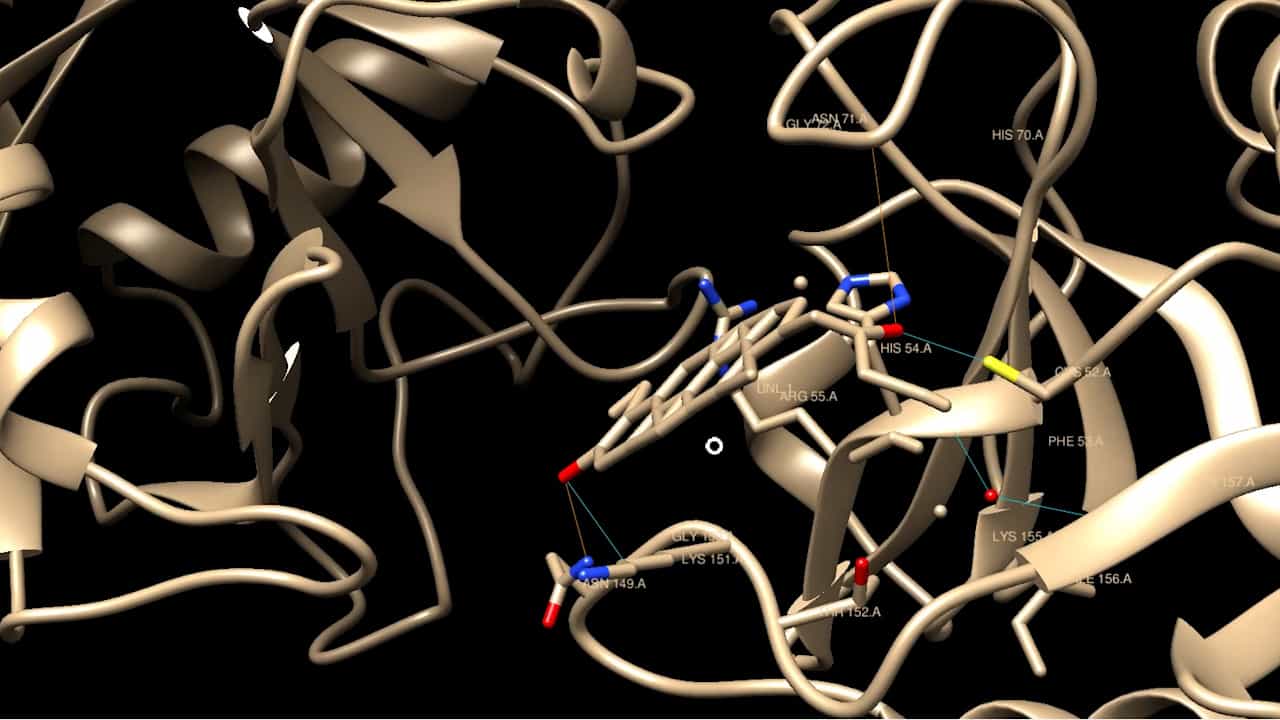

Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Maximum common binding modes (MCBM): consensus docking scoring using multiple ligand information and interaction
Steffen Renner1, Swetlana Derksen, Sebastian Radestock
1Chemical R&D, Merz Pharmaceuticals GmbH, Eckenheimer Landstrasse 100, D-60318 Frankfurt am Main, Germany. steffen.renner@mpi-dortmund.mpg.de

Abstract:
Improving the scoring functions for small molecule-protein docking is a highly challenging task in current computational drug design. Here we present a novel consensus scoring concept for the prediction of binding modes for multiple known active ligands. Similar ligands are generally believed to bind to their receptor in a similar fashion. The presumption of our approach was that the true binding modes of similar ligands should be more similar to each other compared to false positive binding modes. The number of conserved (consensus) interactions between similar ligands was used as a docking score. Patterns of interactions were modeled using ligand receptor interaction fingerprints. Our approach was evaluated for four different data sets of known cocrystal structures (CDK-2, dihydrofolate reductase, HIV-1 protease, and thrombin). Docking poses were generated with FlexX and rescored by our approach. For comparison the CScore scoring functions from Sybyl were used, and consensus scores were calculated thereof. Our approach performed better than individual scoring functions and was comparable to consensus scoring. Analysis of the distribution of docking poses by self-organizing maps (SOM) and interaction fingerprints confirmed that clusters of docking poses composed of multiple ligands were preferentially observed near the native binding mode. Being conceptually unrelated to commonly used docking scoring functions our approach provides a powerful method to complement and improve computational docking experiments.
More Related Videos
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
 14:34
A Bilingual Computational Workflow for Identifying Potential PLK1 Inhibitors in American Sign Language and English
Published on: April 3, 2026
Related Concept Videos
 02:18
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 00:49
Ligand Binding and Linkage