Related Experiment Video
Updated: Jul 2, 2026

Quantitative Analysis of Dietary Vitamin A Metabolites in Murine Ocular and Non-Ocular Tissues Using High-Performance Liquid Chromatography
Published on: December 27, 2024
Structural and Thermodynamic Discrimination between Agonists and Antagonists of Retinoic Acid Receptor γ and the
Teresa Żołek1, Szymon Sutuła2, Andrzej Kutner3
1Department of Organic and Physical Chemistry, Faculty of Pharmacy, Medical University of Warsaw, 1 Banacha, Warsaw 02-097, Poland.

Abstract:
Synthetic ligands of retinoic acid receptor γ (RARγ) and vitamin D receptor (VDR) can act as agonists or antagonists by reshaping receptor conformation and cofactor recruitment, yet the structural and energetic determinants of this selectivity remain incompletely defined. Here, we present an integrated structural and thermodynamic analysis combining conformational characterization, molecular dynamics simulations, and molecular mechanics Poisson-Boltzmann surface area calculations to compare representative agonists and antagonists of human RARγ and VDR. Binding free energies and effective enthalpic contributions were evaluated for selected ligands, whereas the apparent entropic term, estimated indirectly as -TΔSapp = ΔGbind - ΔHeff, was used only as a qualitative internal descriptor. The thermodynamic profiles indicate that effective enthalpic stabilization is a more informative comparative descriptor than binding free energy alone, with clear agonist-antagonist separation for RARγ and a directional, less statistically resolved trend for VDR. These energetic patterns were interpreted alongside ligand-dependent stabilization of helix 12 and receptor-ligand interaction networks. Newly determined MicroED structures of AGN194310 and AGN205728 were incorporated as ligand-specific conformational references, improving structural definition without being treated as direct evidence of receptor-bound bioactive conformations. Comparative analysis of RARγ and VDR suggests that functional selectivity is not governed solely by affinity but emerges from the interplay among ligand-binding energetics, interaction networks, and receptor conformational dynamics. These results provide a retrospective mechanistic benchmark for interpreting agonist- and antagonist-associated behavior in nuclear receptors and may guide future studies on functionally selective modulators.
More Related Videos
 08:18
Methodology for Studying Interactions of Vitamin A Membrane Receptors and Opsin Protein with their Ligands in Generating the Retinylidene Protein
Published on: October 4, 2024
 08:33
Optimized Automated Analysis of Live Neuronal Mitochondria Homeostasis Modulation by Isoform-Specific Retinoic Acid Receptors
Published on: July 28, 2023
Related Concept Videos
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with one...
 01:25
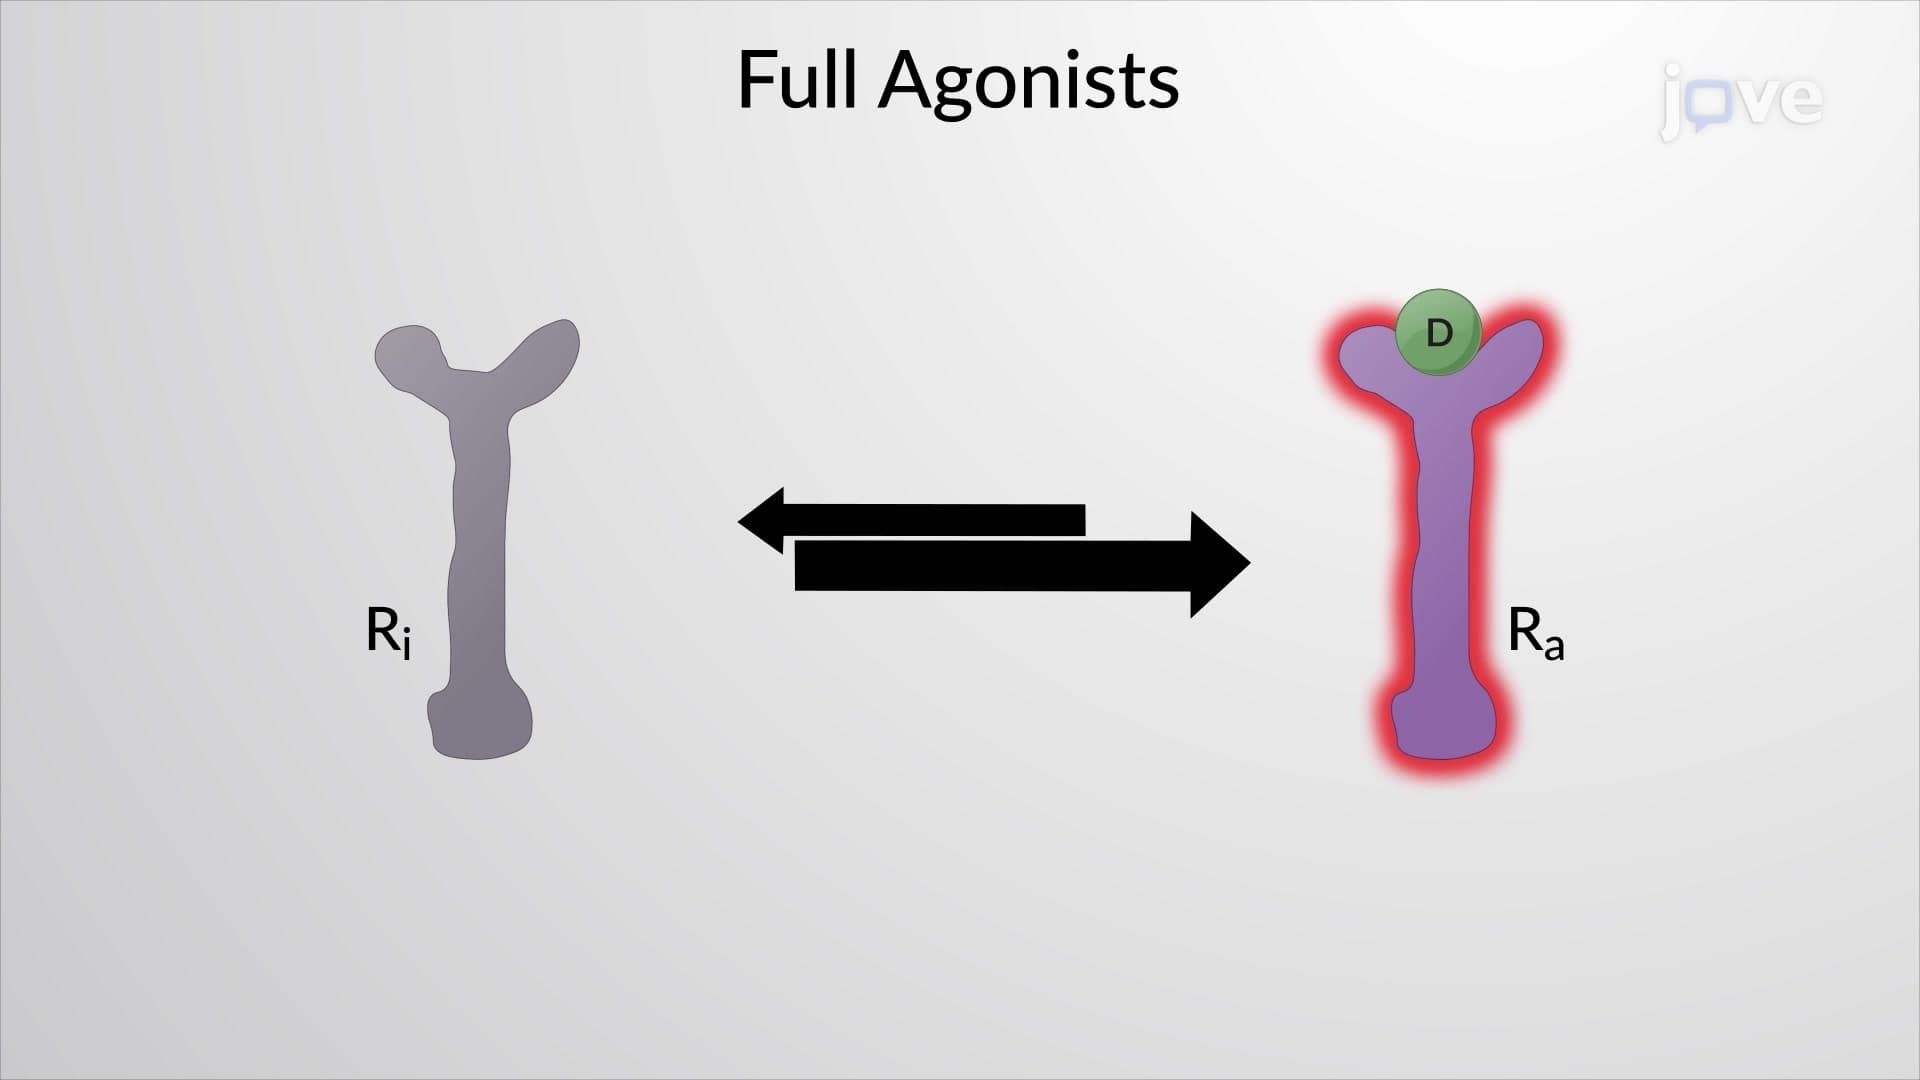
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous ligand's action.
 01:31
Transducer Mechanism: Nuclear Receptors
About 48 different soluble family members of nuclear receptors are identified that can be divided into two main classes:
 01:29
Drug-Receptor Interactions
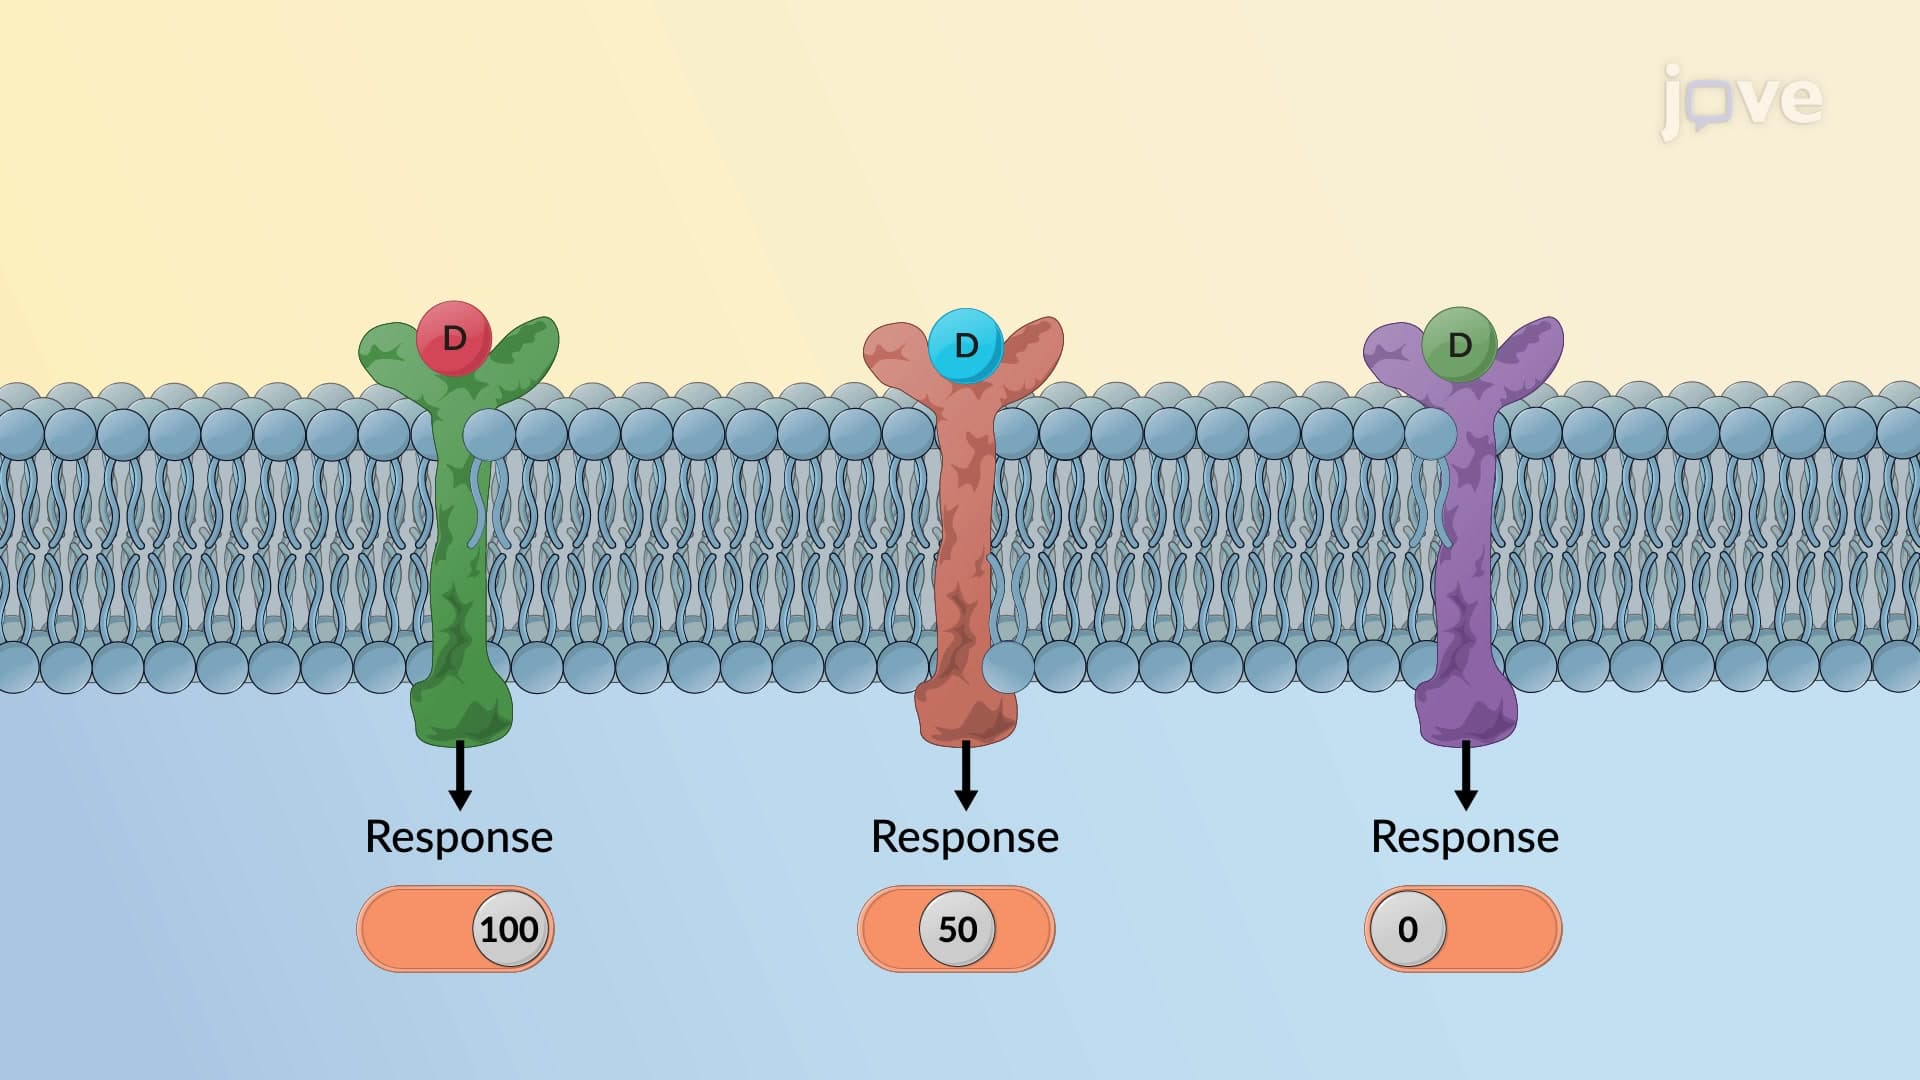
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue.
 01:28
Drug-Receptor Interaction: Antagonist
Antagonists can be classified as competitive or noncompetitive based on their...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...