Related Experiment Video
Updated: Jul 6, 2026
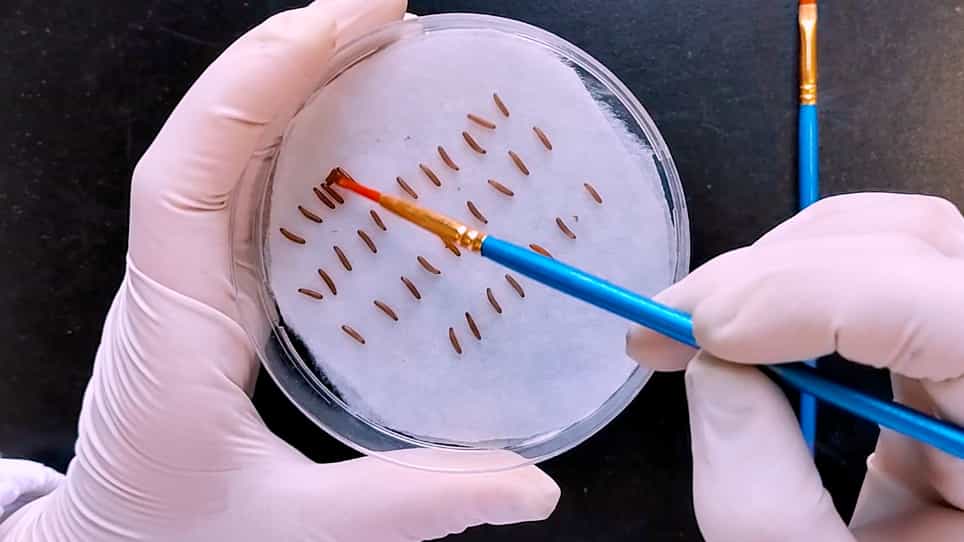

Construction of Homozygous Mutants of Migratory Locust Using CRISPR/Cas9 Technology
Published on: March 16, 2022
Haplotype reconstruction for scnp DNA: a consensus vote approach with extensive sequence data from populations of the
Zu-Shi Huang1, Ya-Jie Ji, De-Xing Zhang
1State Key Laboratory of Integrated Management of Pest Insects and Rodents, Institute of Zoology, Chinese Academy of Sciences, Beijing 100101, China.

Abstract:
Single copy nuclear polymorphic (scnp) DNA is potentially a powerful molecular marker for evolutionary studies of populations. However, a practical obstacle to its employment is the general problem of haplotype determination due to the common occurrence of heterozygosity in diploid organisms. We explore here a 'consensus vote' (CV) approach to this question, combining statistical haplotype reconstruction and experimental verification using as an example an indel-free scnp DNA marker from the flanking region of a microsatellite locus of the migratory locust. The raw data comprise 251-bp sequences from 526 locust individuals (1052 chromosomes), with 71 (28.3%) polymorphic nucleotide sites (including seven triallelic sites) and 141 distinct genotypes (with frequencies ranging from 0.2 to 25.5%). Six representative statistical haplotype reconstruction algorithms are employed in our CV approach, including one parsimony method, two expectation-maximization (EM) methods and three Bayesian methods. The phases of 116 ambiguous individuals inferred by this approach are verified by molecular cloning experiments. We demonstrate the effectiveness of the CV approach compared to inferences based on individual statistical algorithms. First, it has the unique power to partition the inferrals into a reliable group and an uncertain group, thereby allowing the identification of the inferrals with greater uncertainty (12.7% of the total sample in this case). This considerably reduces subsequent efforts of experimental verification. Second, this approach is capable of handling genotype data pooled from many geographical populations, thus tolerating heterogeneity of genetic diversity among populations. Third, the performance of the CV approach is not influenced by the number of heterozygous sites in the ambiguous genotypes. Therefore, the CV approach is potentially a reliable strategy for effective haplotype determination of nuclear DNA markers. Our results also show that rare variations and rare inferrals tend to be more vulnerable to inference error, and hence deserve extra surveillance.
Related Concept Videos
 02:54
Evolutionary Relationships through Genome Comparisons
 01:05
Single Nucleotide Polymorphisms-SNPs
 02:51
Multi-species Conserved Sequences
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved DNA...
 02:53
Conservative Site-specific Recombination and Phase Variation
The recognition sites for Cre recombinase called LoxP...
 02:26
Comparing Copy Number Variations and SNPs
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...