Related Experiment Video
Updated: Jul 6, 2026
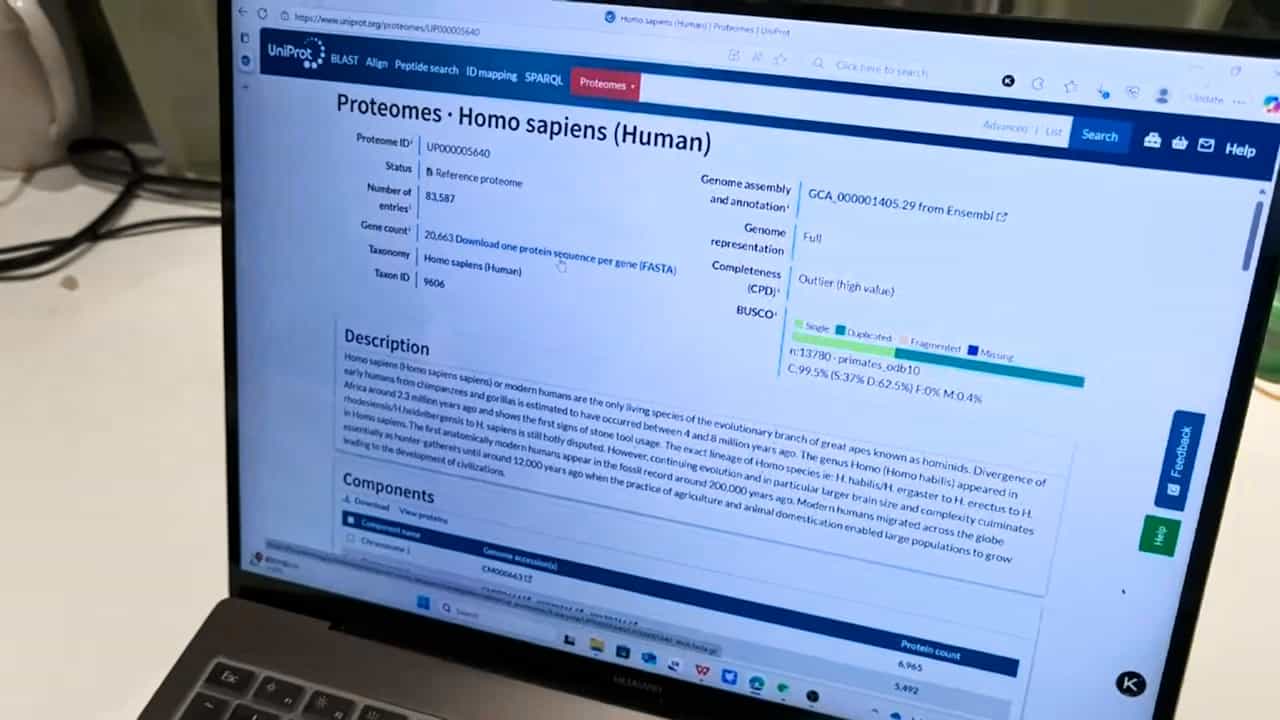

Navigating the Mass Spectrometry-Based Proteomic Data Using Free Computational Tools
Published on: August 19, 2025
A noise model for mass spectrometry based proteomics
Peicheng Du1, Gustavo Stolovitzky, Peter Horvatovich
1IBM Computational Biology Center, P.O. Box 218, Yorktown Heights, NY 10598, USA. pdu@us.ibm.com

Motivation:
Mass spectrometry data are subjected to considerable noise. Good noise models are required for proper detection and quantification of peptides. We have characterized noise in both quadrupole time-of-flight (Q-TOF) and ion trap data, and have constructed models for the noise.
Results:
We find that the noise in Q-TOF data from Applied Biosystems QSTAR fits well to a combination of multinomial and Poisson model with detector dead-time correction. In comparison, ion trap noise from Agilent MSD-Trap-SL is larger than the Q-TOF noise and is proportional to Poisson noise. We then demonstrate that the noise model can be used to improve deisotoping for peptide detection, by estimating appropriate cutoffs of the goodness of fit parameter at prescribed error rates. The noise models also have implications in noise reduction, retention time alignment and significance testing for biomarker discovery.
Related Concept Videos
 01:19
Mass Spectrometry: Overview
 01:16
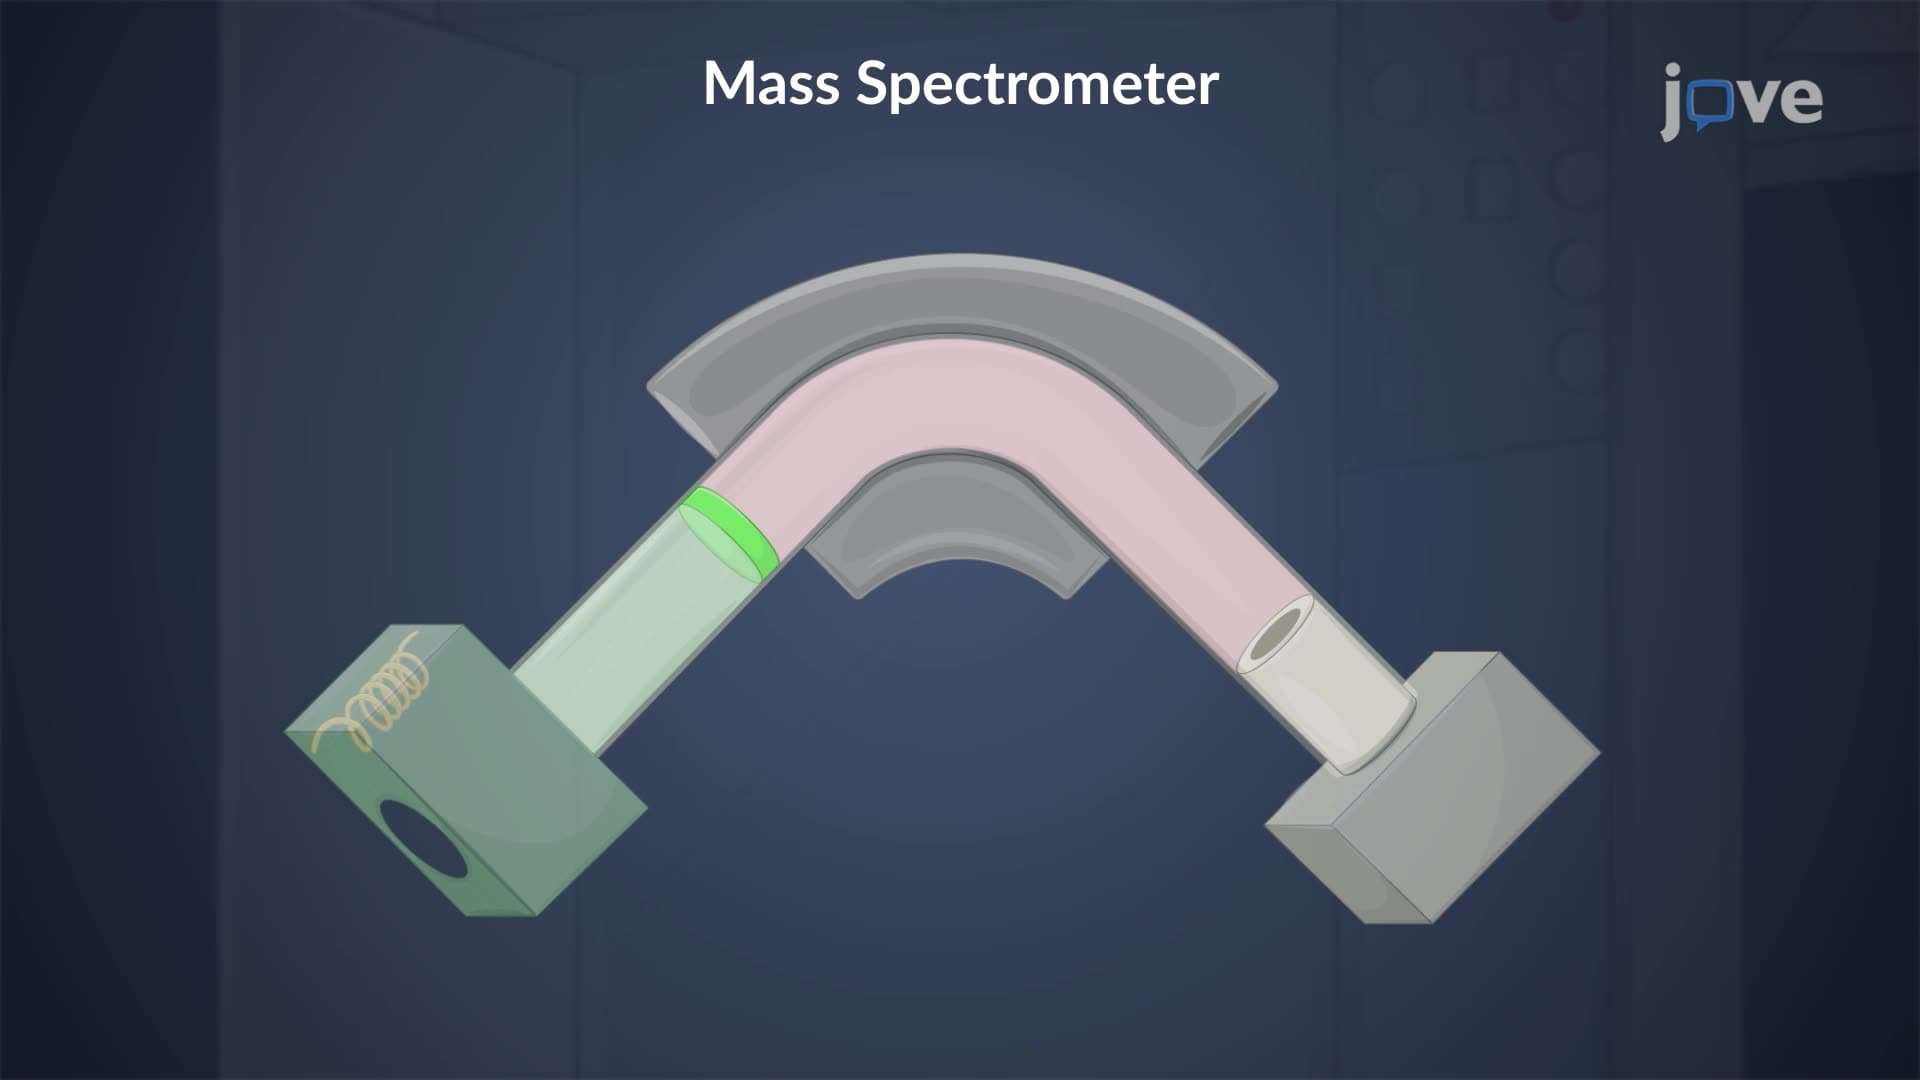
Mass Spectrometers
 01:21
Tandem Mass Spectrometry
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
 01:19
MALDI-TOF Mass Spectrometry
 01:24
Mass Spectrum: Interpretation