Related Experiment Video
Updated: Sep 19, 2026
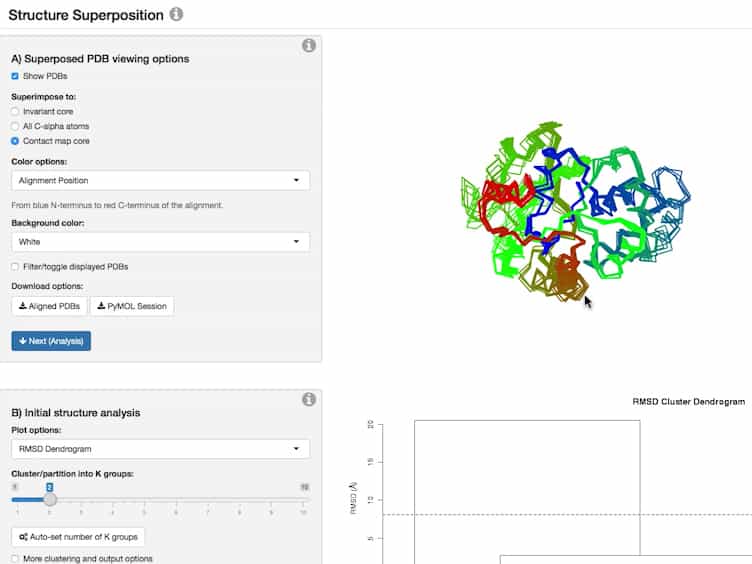

Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Conformational dynamics of fuzzy interfaces in disordered protein complexes: mapping key residues and binding modes
María Laura Fernández1,2,3, María Clara Bastien3, Franco G Tavolaro3
1Departamento de Física, Facultad de Ciencias Exactas y Naturales, Universidad de Buenos Aires, Ciudad Universitaria, Buenos Aires, Argentina.

Abstract:
Understanding the dynamic behavior of fuzzy complexes formed by intrinsically disordered proteins (IDPs) is challenging due to their conformational flexibility. Nuclear Magnetic Resonance (NMR) structural bundles satisfy experimental constraints and inherently provide an overview of alternative conformations in solution. However, they assign equal weight to every sampled model, obscuring the temporal prevalence and dynamics of inter-residue contacts. In this study, we performed dual 1-microsecond (µs) Molecular Dynamics (MD) simulations for eight biologically relevant fuzzy complexes, utilizing the most distant conformers from deposited NMR models, as starting configurations. Through structural clustering and contact-persistence analysis, we classified the complexes into two distinct dynamic categories. The first group comprises complexes whose trajectories remain consistent with experimental data. Notably, large-scale motions such as relative chain rotations observed during simulations are already reflected as distinct orientations within the starting NMR conformers. Conversely, the second group comprises complexes that exhibit interaction modes absent in the structural NMR ensembles. These include alternative stable interfaces arising from chain reorientation, as well as highly dynamic interfaces characterized by rapidly exchanging contacts lacking any persistent stabilizing core. Furthermore, the microsecond trajectories captured intermittent dissociation and re-association events where the proteins sampled transient contacts through alternative regions. Overall, leveraging NMR models as a baseline for MD clustering and contact weighting, provides a systematic bioinformatics analysis to solve the atomistic details of flexible interfaces. Beyond categorizing diverse binding behaviors, this approach systematically refines the interaction surfaces across all studied cases, uncovers dynamic features that escape detection in ensemble-averaged NMR structures. Furthermore, it maps key residues to guide further experimental approaches, such as mutational studies or therapeutic targeting, among other applications.
More Related Videos
 09:25
NMR 15N Relaxation Experiments for the Investigation of Picosecond to Nanoseconds Structural Dynamics of Proteins
Published on: November 1, 2024
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:04
Protein-protein Interfaces
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence.
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 02:18
Intrinsically Disordered Proteins