Related Experiment Video
Updated: Jul 5, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Potential energy curve for isomerization of N2H2 and C2H4 using the improved virtual orbital multireference
Rajat K Chaudhuri1, Karl F Freed, Sudip Chattopadhyay
1Indian Institute of Astrophysics, Bangalore 560 034, India. rkchaudh@iiap.res.in

Abstract:
Multireference Møller-Plesset (MRMP) perturbation theory [K. Hirao, Chem. Phys. Lett. 190, 374 (1992)] is modified to use improved virtual orbitals (IVOs) and is applied to study ground state potential energy curves for isomerization and dissociation of the N2H2 and C2H4 molecules. In contrast to traditional MRMP or multistate multiconfiguration quasidegenerate perturbation theory where the reference functions are obtained from (often difficult to converge) state averaged multiconfiguration self-consistent field methods, our reference functions are represented in terms of computationally efficient IVOs. For convenience in comparisons with other methods, a first order complete active space configuration interaction (CASCI) calculation with the IVOs is followed by the use of the IVOs in MRMP to incorporate residual electron correlation effects. The potential energy curves calculated from the IVO-MRMP method are compared with computations using state-of-the-art coupled cluster singles and doubles (CCSD) methods and variants thereof to assess the efficacy of the IVO-MRMP scheme. The present study clearly demonstrates that unlike the CCSD and its variants, the IVO-MRMP approach provides smooth and reliable ground state potential energy curves for isomerization of these systems. Although the rigorously size-extensive completely renormalized CC theory with noniterative triples corrections (CR-CC(2,3)) likewise provides relatively smooth curves, the CR-CC(2,3) calculations overestimate the cis-trans barrier height for N2H2. The ground state spectroscopic constants predicted by the IVO-CASCI method agree well with experiment and with other highly correlated ab initio methods.
More Related Videos
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
 10:52
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 03:51
Molecular Orbital Theory II
 02:18
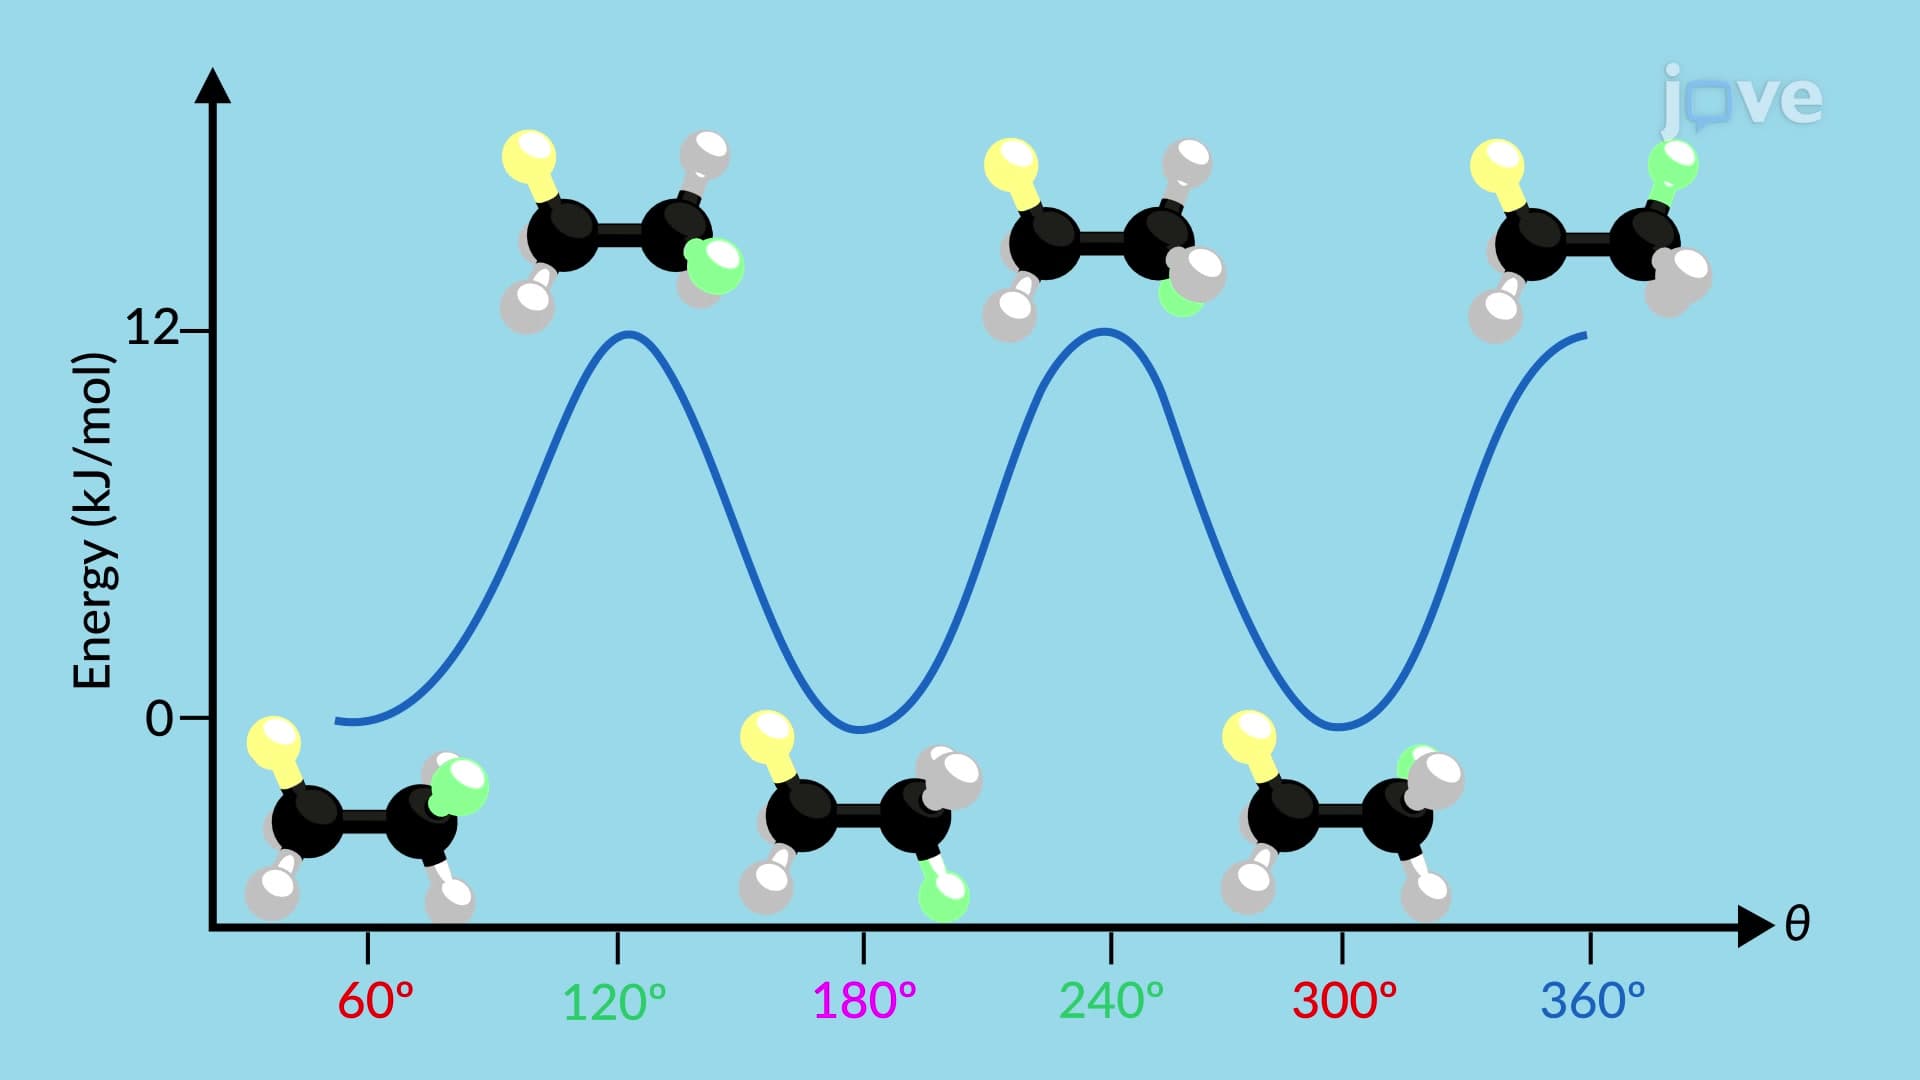
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered ethane, the...
 02:40
MO Theory and Covalent Bonding
 01:18
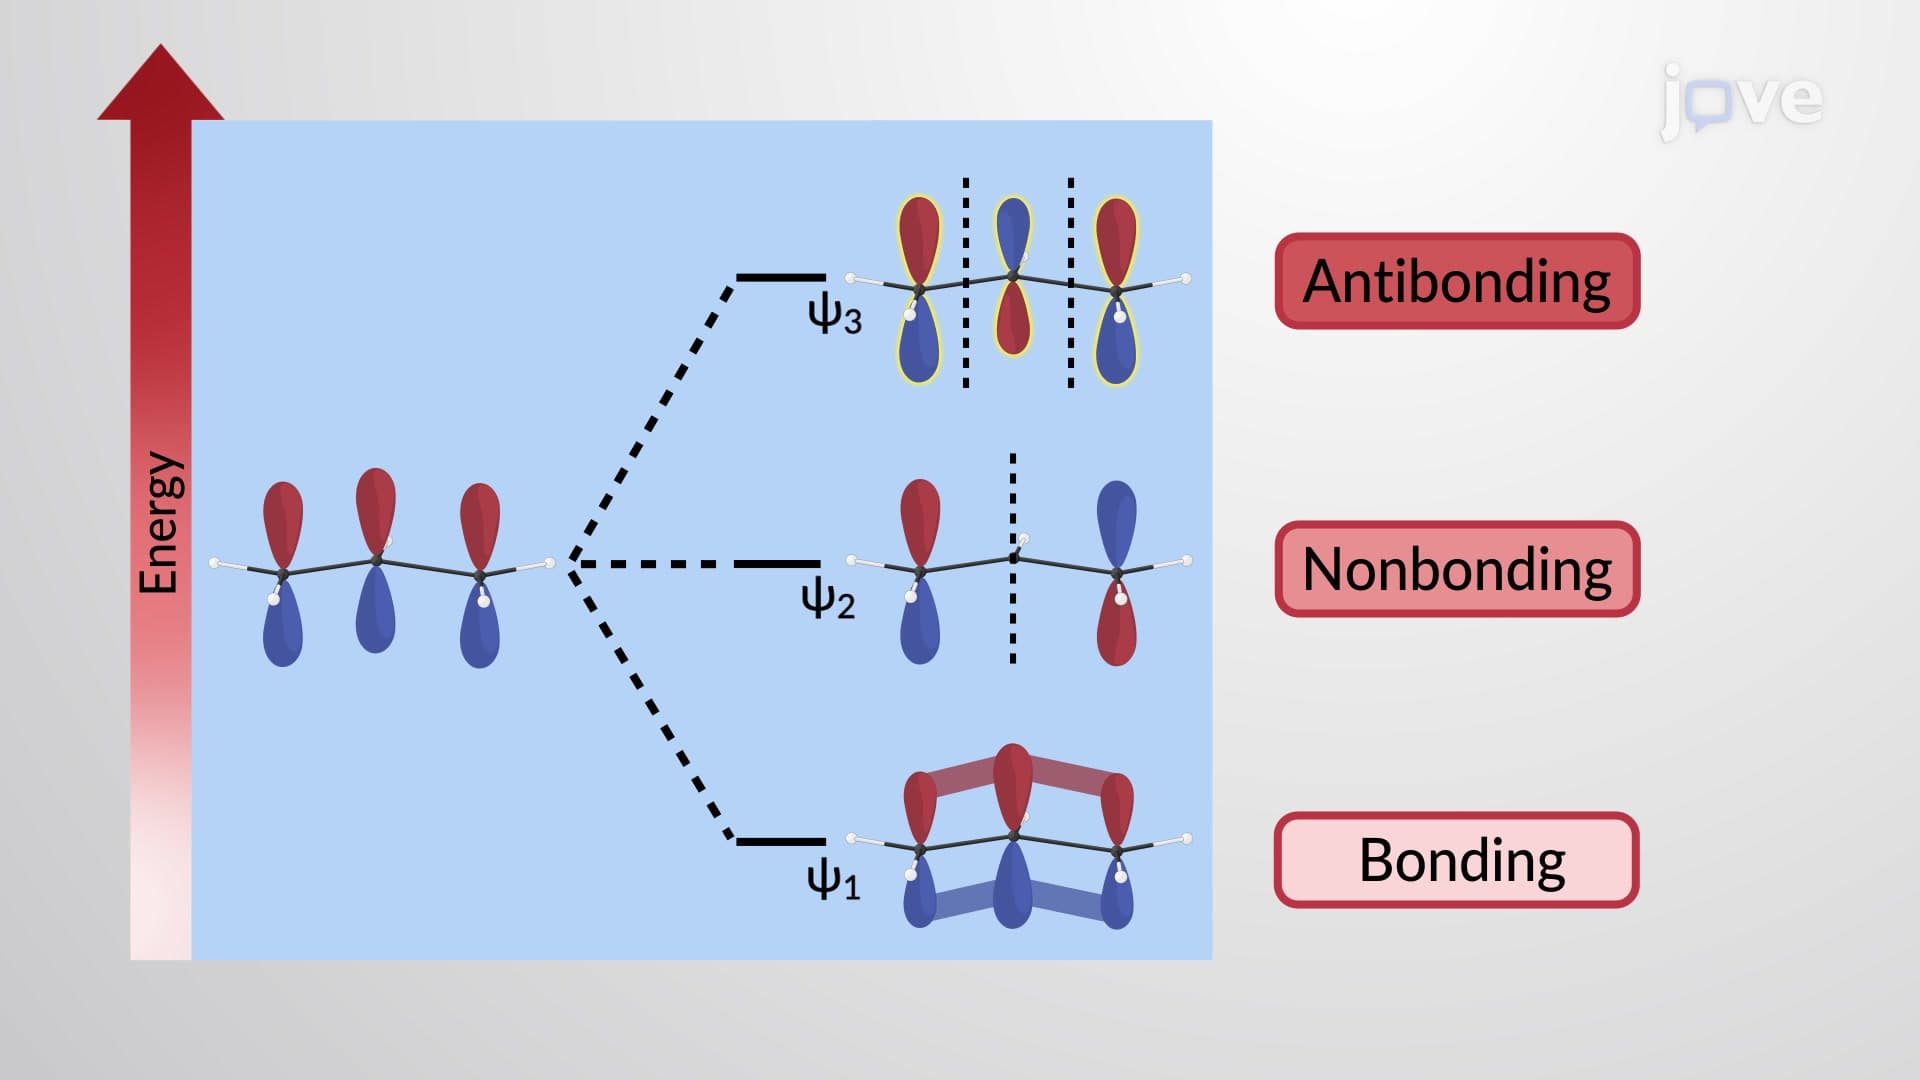
π Molecular Orbitals of the Allyl Cation and Anion