Related Experiment Video
Updated: Jul 5, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
An atomic orbital-based reformulation of energy gradients in second-order Møller-Plesset perturbation theory
Sabine Schweizer1, Bernd Doser, Christian Ochsenfeld
1Theoretische Chemie, Auf der Morgenstelle 8, Universität Tübingen, Tübingen, Germany.

Abstract:
A fully atomic orbital (AO)-based reformulation of second-order Møller-Plesset perturbation theory (MP2) energy gradients is introduced, which provides the basis for reducing the computational scaling with the molecular size from the fifth power to linear. Our formulation avoids any transformation between the AO and the molecular orbital (MO) basis and employs pseudodensity matrices similar to the AO-MP2 energy expressions within the Laplace scheme for energies. The explicit computation of perturbed one-particle density matrices emerging in the new AO-based gradient expression is avoided by reformulating the Z-vector method of Handy and Schaefer [J. Chem. Phys. 81, 5031 (1984)] within a density matrix-based scheme.
More Related Videos


Related Concept Videos
 02:35
Molecular Orbital Theory I
 03:51
Molecular Orbital Theory II
 01:18
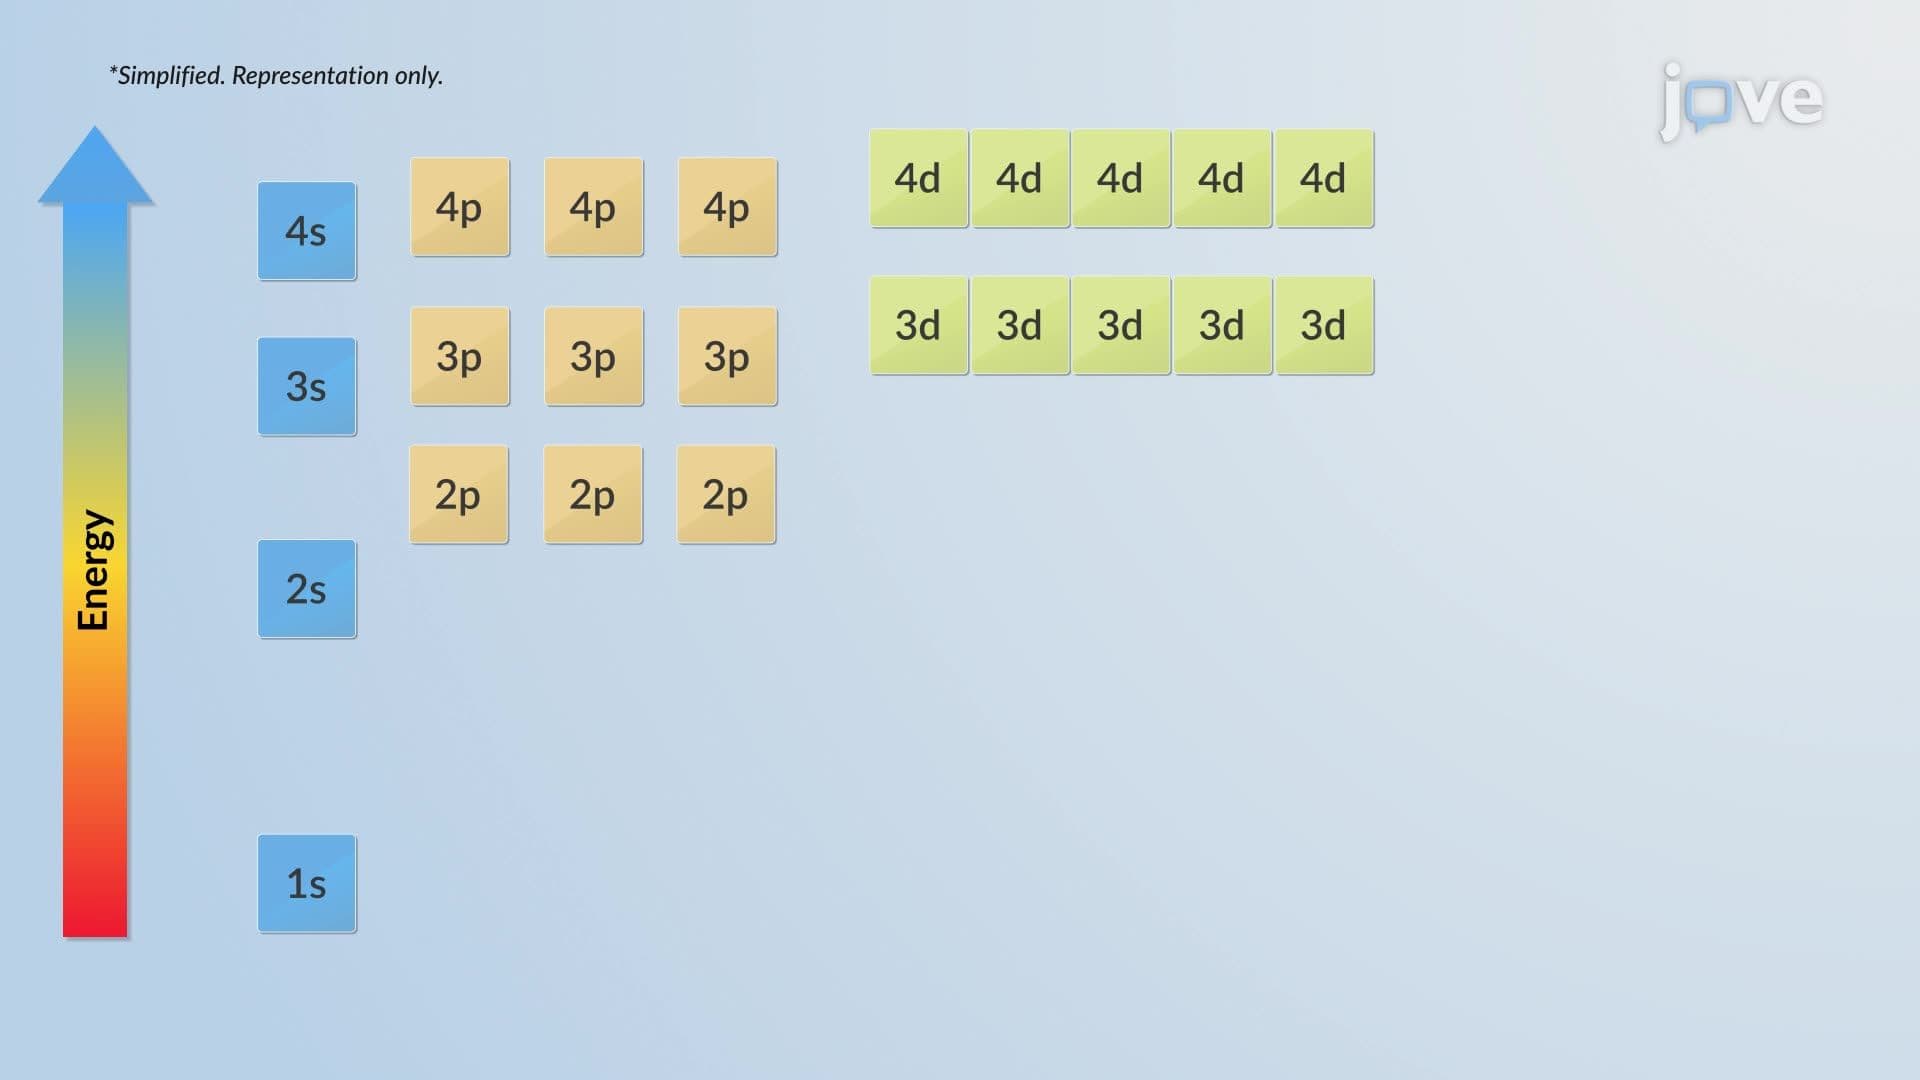
Electron Orbital Model
 03:21
The Energies of Atomic Orbitals
 02:40
MO Theory and Covalent Bonding
 02:44
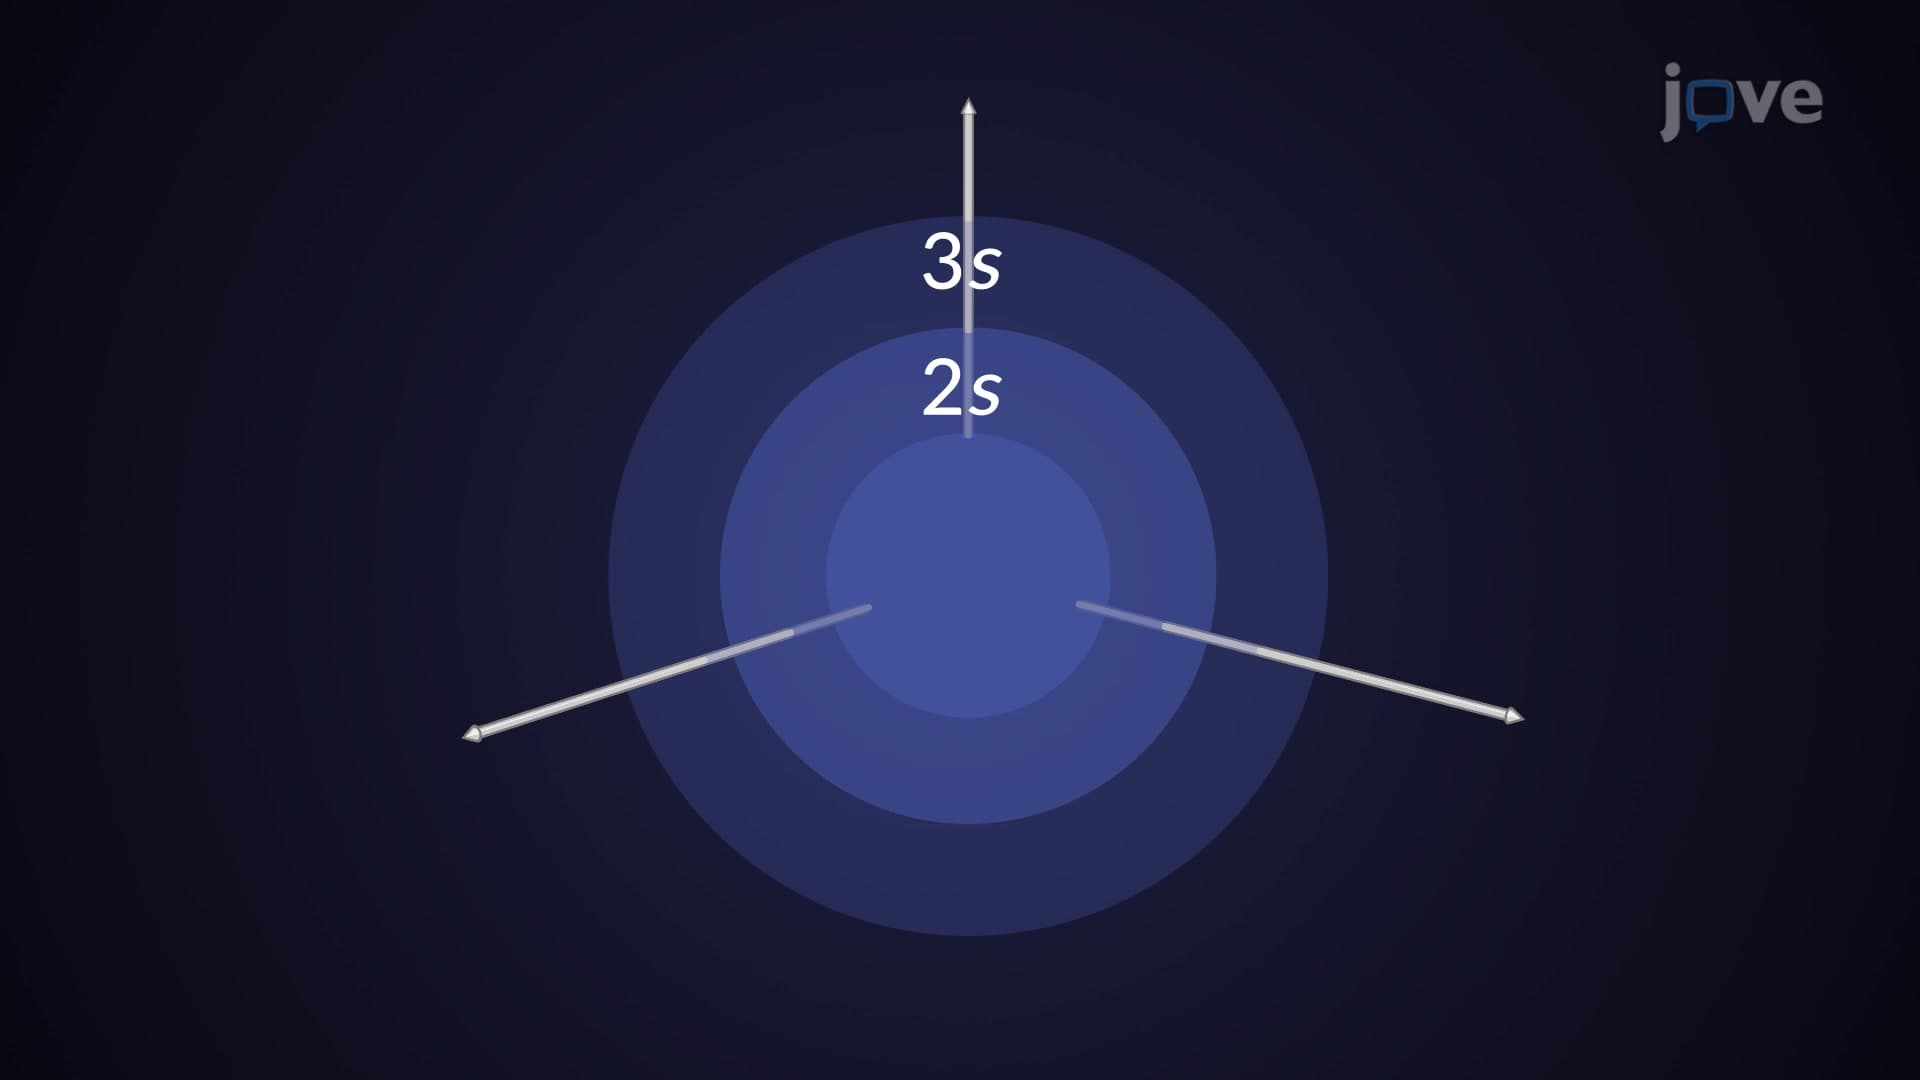
Atomic Orbitals