Related Experiment Video
Updated: Jul 5, 2026

One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
Spectrum of medium-chain acyl-CoA dehydrogenase deficiency detected by newborn screening
Ho-Wen Hsu1, Thomas H Zytkovicz, Anne Marie Comeau
1New England Newborn Screening Program, University of Massachusetts Medical School, 305 South St, Jamaica Plain, MA 02130, USA. ho-wen.hsu@umassmed.edu

Objective:
Our goal was to describe the clinical spectrum of medium-chain acyl-CoA dehydrogenase deficiency detected by routine newborn screening and assess factors associated with elevations of octanoylcarnitine in newborns and characteristics associated with adverse clinical consequences of medium-chain acyl-CoA dehydrogenase deficiency.
Methods:
The first 47 medium-chain acyl-CoA dehydrogenase deficiency cases detected by the New England Newborn Screening Program were classified according to initial and follow-up octanoylcarnitine values, octanoylcarnitine-decanoylcarnitine ratios, medium-chain acyl-CoA dehydrogenase genotype, follow-up biochemical parameters, and feeding by breast milk or formula.
Results:
All 20 patients who were homozygous for 985A-->G had high initial octanoylcarnitine values (7.0-36.8 microM) and octanoylcarnitine-decanoylcarnitine ratios (7.0-14.5), whereas the 27 patients with 0 to 1 copy of 985A-->G exhibited a wide range of octanoylcarnitine values (0.5-28.6 microM) and octanoylcarnitine-decanoylcarnitine ratios (0.8-12.7). Initial newborn octanoylcarnitine values decreased by days 5 to 8, but the octanoylcarnitine-decanoylcarnitine ratio generally remained stable. Among 985A-->G homozygotes, breastfed newborns had higher initial octanoylcarnitine values than newborns who received formula. Adverse events occurred in 5 children, 4 985A-->G homozygotes and 1 compound heterozygote with a very high initial octanoylcarnitine: 2 survived severe neonatal hypoglycemia, 1 survived a severe hypoglycemic episode at 15 months of age, and 2 died as a result of medium-chain acyl-CoA dehydrogenase deficiency at ages 11 and 33 months.
Conclusion:
Newborn screening for medium-chain acyl-CoA dehydrogenase deficiency has detected cases with a wide range of genotypes and biochemical abnormalities. Although most children do well, adverse outcomes have not been entirely avoided. Assessment of potential risk and determination of appropriate treatment remain a challenge.
More Related Videos
 06:53
Visualization of Mitochondrial Respiratory Function using Cytochrome C Oxidase / Succinate Dehydrogenase (COX/SDH) Double-labeling Histochemistry
Published on: November 23, 2011
 06:48
Fingerprinting Cardiolipin in Leukocytes by Mass Spectrometry for a Rapid Diagnosis of Barth Syndrome
Published on: March 23, 2022
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 02:59
Animal Mitochondrial Genetics
 01:28
Overview of Fatty Acid Metabolism
Fatty acids are catabolized in a process called beta-oxidation, which takes place in the matrix of the mitochondria and converts their fatty acid chains into two-carbon units of acetyl groups. The acetyl...
 01:15
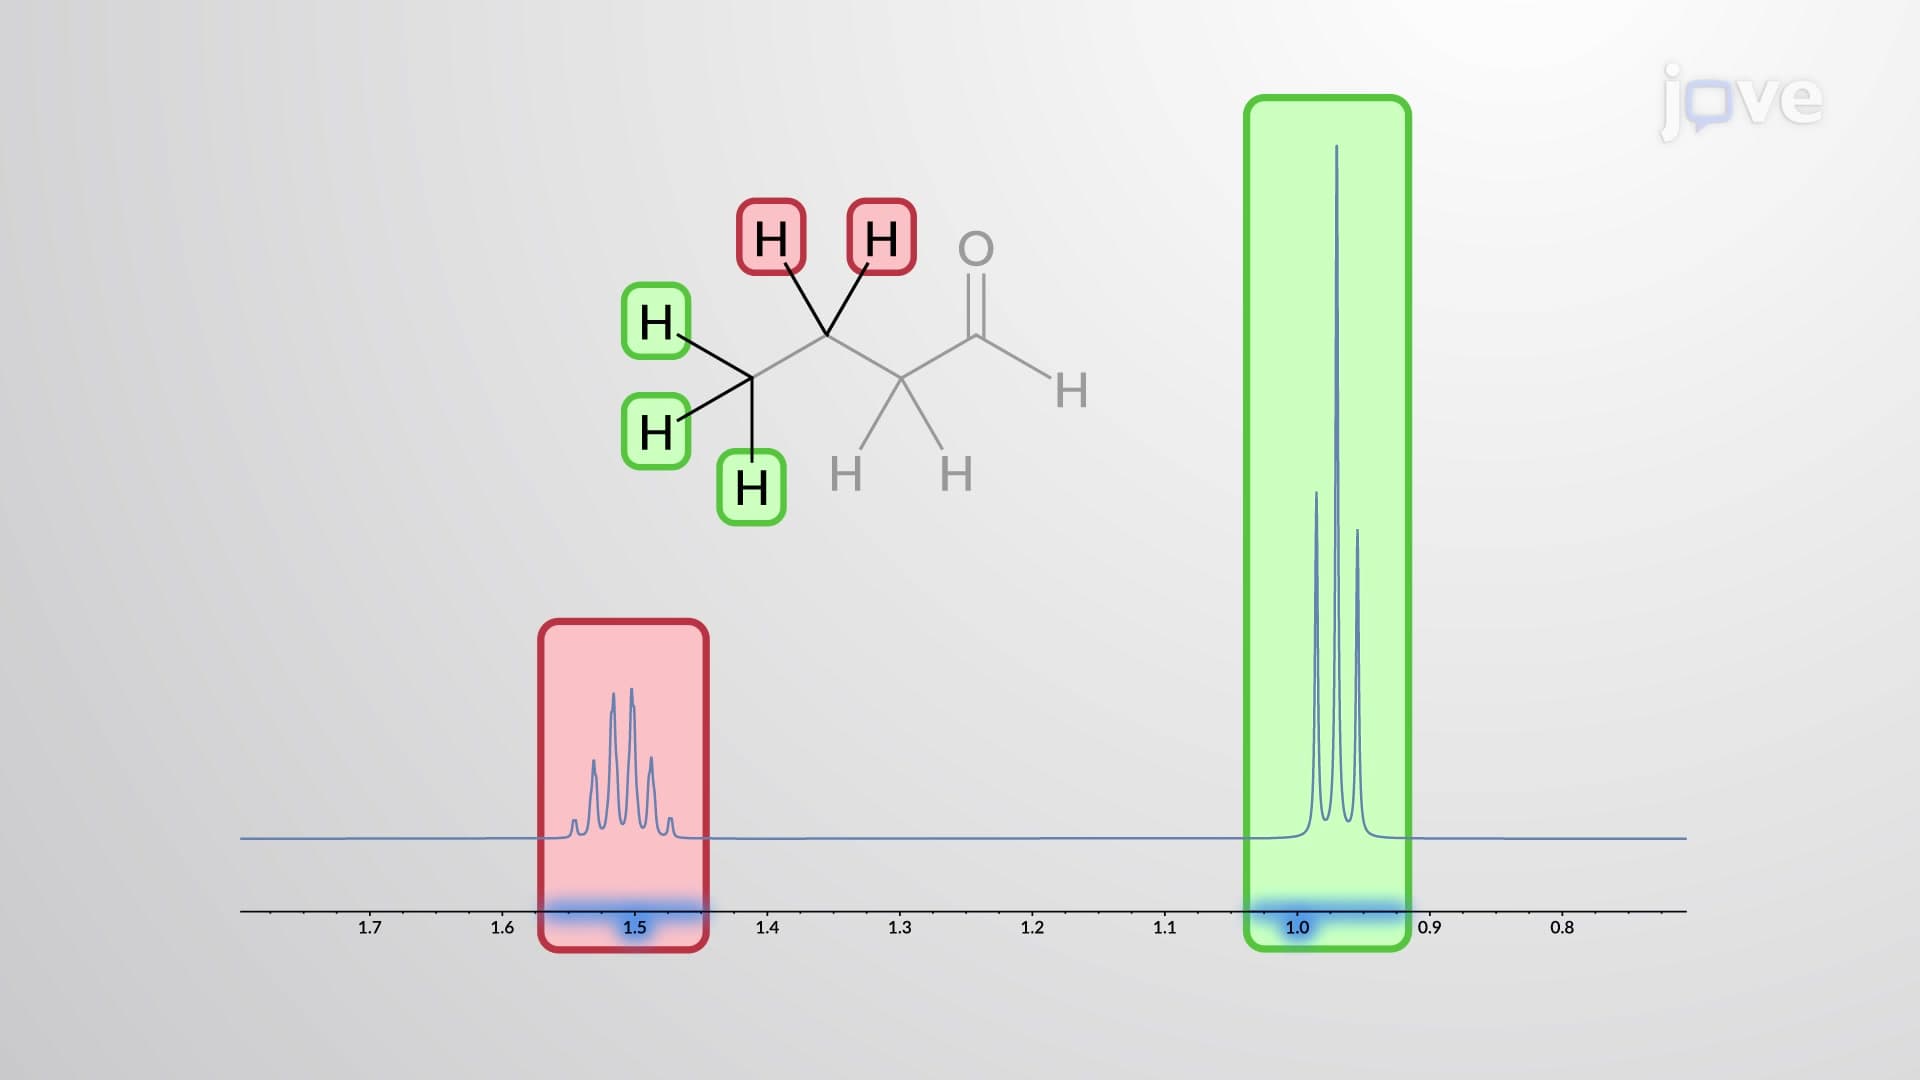
NMR Spectroscopy and Mass Spectrometry of Aldehydes and Ketones