Related Experiment Video
Updated: Sep 12, 2026


Fingerprinting Cardiolipin in Leukocytes by Mass Spectrometry for a Rapid Diagnosis of Barth Syndrome
Published on: March 23, 2022
Monocarboxylate transporter 1 deficiency: a rare case report and systematic review of genetically confirmed cases
Youssef Magdy Khalifa1,2, Mohanad Yaser Assem1,2, Ziad Fathi Ahmed1
1Faculty of medicine, Mansoura University, Daqahlia, Egypt.

Abstract:
Monocarboxylate transporter 1 (MCT1) deficiency is an ultra-rare autosomal recessive metabolic disorder caused by pathogenic variants in the SLC16A1 gene, which encodes a proton-linked transporter of lactate, pyruvate, and ketone bodies. Few cases have been genetically confirmed worldwide, and diagnosis is often delayed due to nonspecific clinical features and the absence of characteristic metabolic markers. We report the case of a 5-year-old Egyptian boy with recurrent episodes of ketoacidosis and neurological symptoms, who was confirmed by whole-exome sequencing to have homozygous SLC16A1 pathogenic variants. Clinical data, biochemical findings, treatment course, and follow-up were documented in accordance with CARE guidelines. To contextualize this case, we conducted a systematic review of the literature following PRISMA 2020 guidelines, identifying genetically confirmed cases of MCT1 deficiency and summarizing clinical, genetic, and therapeutic insights. The patient presented with high anion gap metabolic acidosis (pH 6.8, bicarbonate 5.2 mmol/L) and normal glucose, lactate, and ammonia levels, consistent with impaired monocarboxylate transport. He developed severe rebound hypokalemia (serum potassium 1.7 mmol/L) during bicarbonate therapy, necessitating intensive correction. He recovered with supportive management, including careful fluid and electrolyte replacement. Our systematic review identified seven eligible studies, encompassing eight patients across six countries. Age of onset ranged from 2 months to 6 years, with clinical presentations including recurrent vomiting, lethargy, seizures, and metabolic crises. While outcomes varied, early recognition and supportive management were associated with more favorable neurodevelopmental trajectories. This study highlights the diagnostic and therapeutic challenges of MCT1 deficiency. The case illustrates not only the typical metabolic phenotype but also treatment-related complications such as hypokalemia. Findings from the systematic review emphasize the importance of genetic testing for early recognition, multidisciplinary management, and the need for greater clinical awareness of this emerging metabolic disorder. Expanding patient registries and investigating genotype-phenotype correlations are essential to improve outcomes in affected individuals.
Related Concept Videos
 01:27
Glucose Transporters
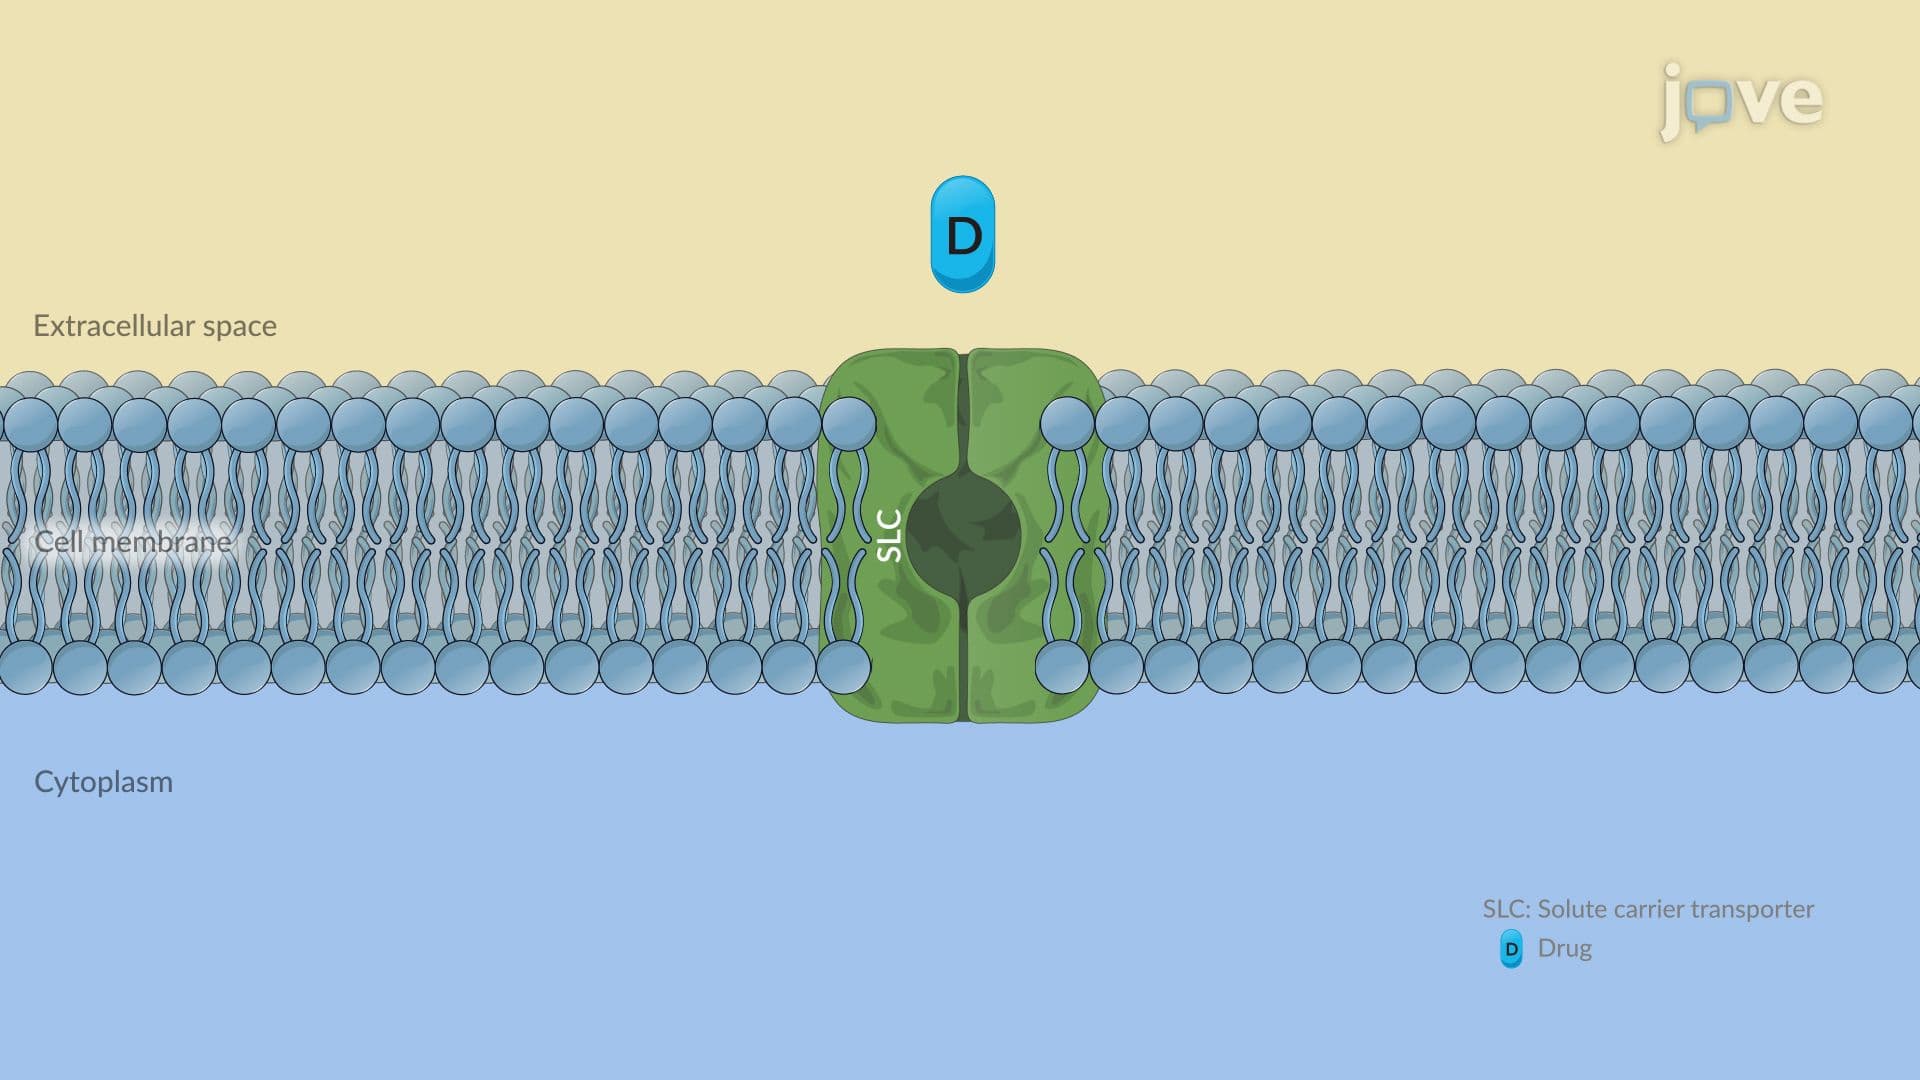
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes:
 01:16
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters
 01:20
Inborn Errors of Metabolism
 01:44
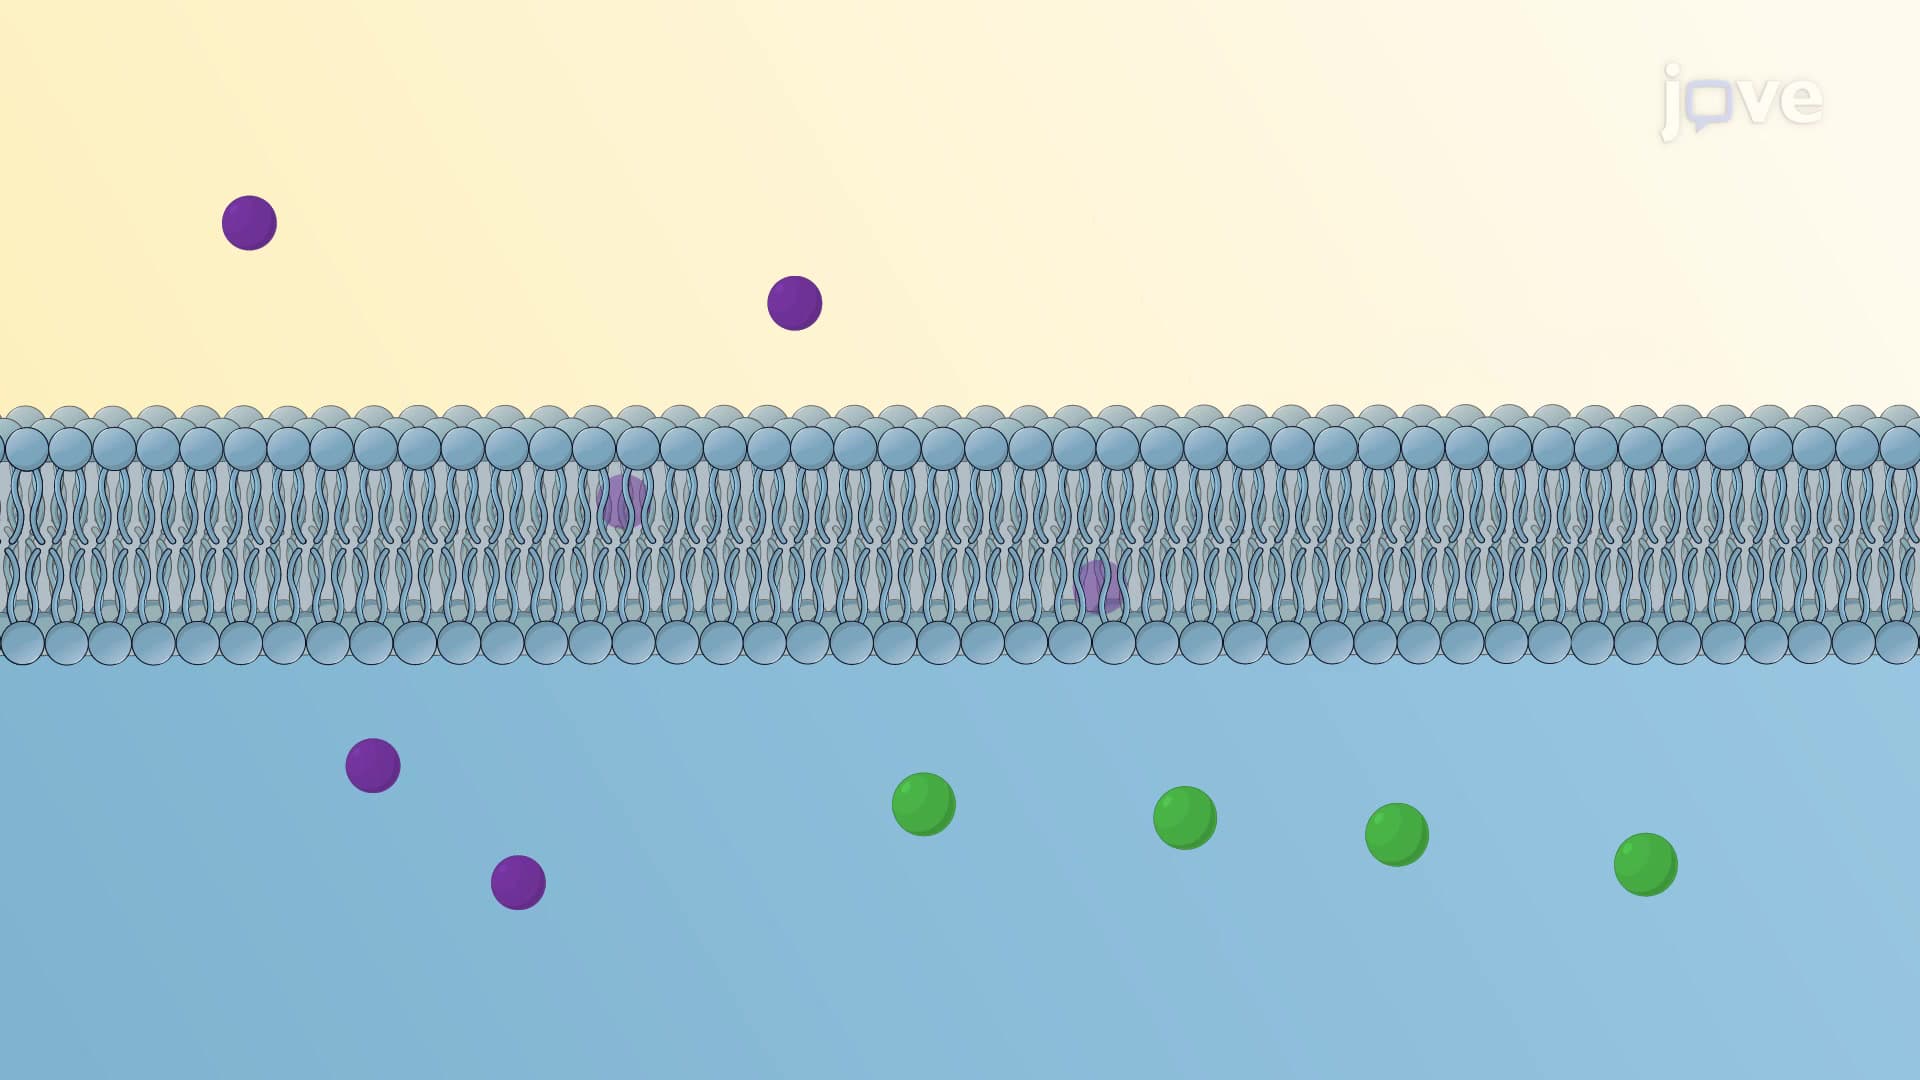
The Significance of Membrane Transport
Transporters facilitate either an active or passive movement of solutes. They can allow a single-molecule transport down its...
 01:31
Membrane Transporters
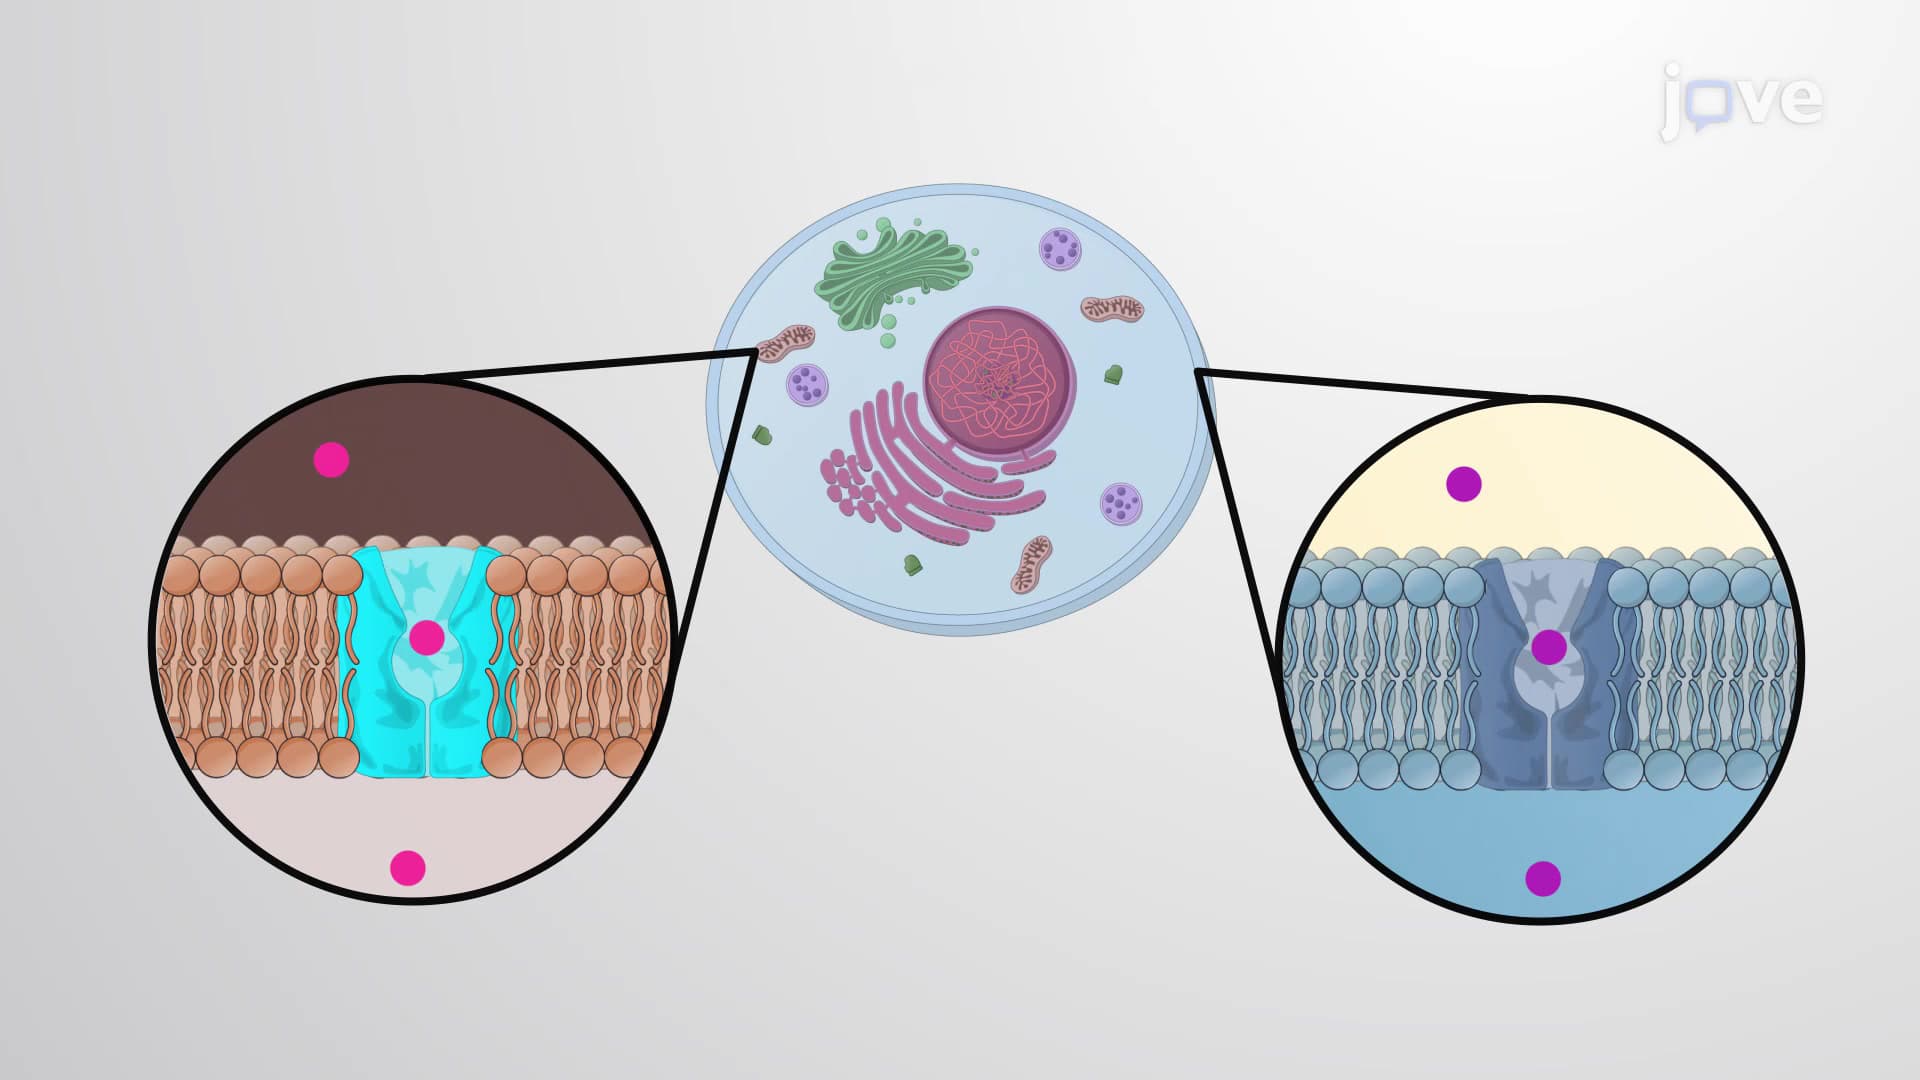
Transporters are mainly composed of alpha-helices, built from bundles of ten or more helices traversing the plasma membrane. The solute-binding sites are located midway, where some of the helices are broken or distorted, making space for the binding site through...
 01:31
ABC Transporters: Exporter