Related Experiment Video
Updated: Jul 4, 2026

Biosynthesis of a Flavonol from a Flavanone by Establishing a One-pot Bienzymatic Cascade
Published on: August 14, 2019
Pharmacophore mapping of flavone derivatives for aromatase inhibition
Shuchi Nagar1, Md Ataul Islam, Suvadra Das
1Department of Chemical Technology, University of Calcutta, 92, A.P.C. Road, Kolkata, 700009, India.

Abstract:
Aromatase, which catalyses the final step in the steroidogenesis pathway of estrogen, has been target for the design of inhibitor in the treatment of hormone dependent breast cancer for postmenopausal women. The extensive SAR studies performed in the last 30 years to search for potent, selective and less toxic compounds, have led to the development of second and third generation of non-steroidal aromatase inhibitors (AI). Besides the development of synthetic compounds, several naturally occurring and synthetic flavonoids, which are ubiquitous natural phenolic compounds and mediate the host of biological activities, are found to demonstrate inhibitory effects on aromatase. The present study explores the pharmacophores, i.e., the structural requirements of flavones (Fig. 1) for inhibition of aromatase activity, using quantitative structure activity relationship (QSAR) and space modeling approaches. The classical QSAR studies generate the model (R (2) = 0.924, Q (2) = 0.895, s = 0.233) that shows the importance of aromatic rings A and C, along with substitutional requirements in meta and para positions of ring C for the activity. 3D QSAR of Comparative Molecular Field Analysis (CoMFA, R (2) = 0.996, R(2)(cv) = 0.791) and Comparative Molecular Similarity Analysis (CoMSIA, R (2) = 0.992, R(2)(cv) = 0.806) studies show contour maps of steric and hydrophobic properties and contribution of acceptor and donor of the molecule, suggesting the presence of steric hindrance due to ring C and R''-substituent, bulky hydrophobic substitution in ring A, along with acceptors at positions 11, and alpha and gamma of imidazole ring, and donor in ring C favor the inhibitory activity. Further space modeling (CATALYST) study (R = 0.941, Delta( cost ) = 96.96, rmsd = 0.876) adjudge the presence of hydrogen bond acceptor (keto functional group), hydrophobic (ring A) and aromatic rings (steric hindrance) along with critical distance among features are important for the inhibitory activity.
More Related Videos
Related Concept Videos
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:11
Pharmacogenetics of Drug Targets: β₂-Adrenergic Receptors, Apo E, Thymidylate Synthase
 01:17
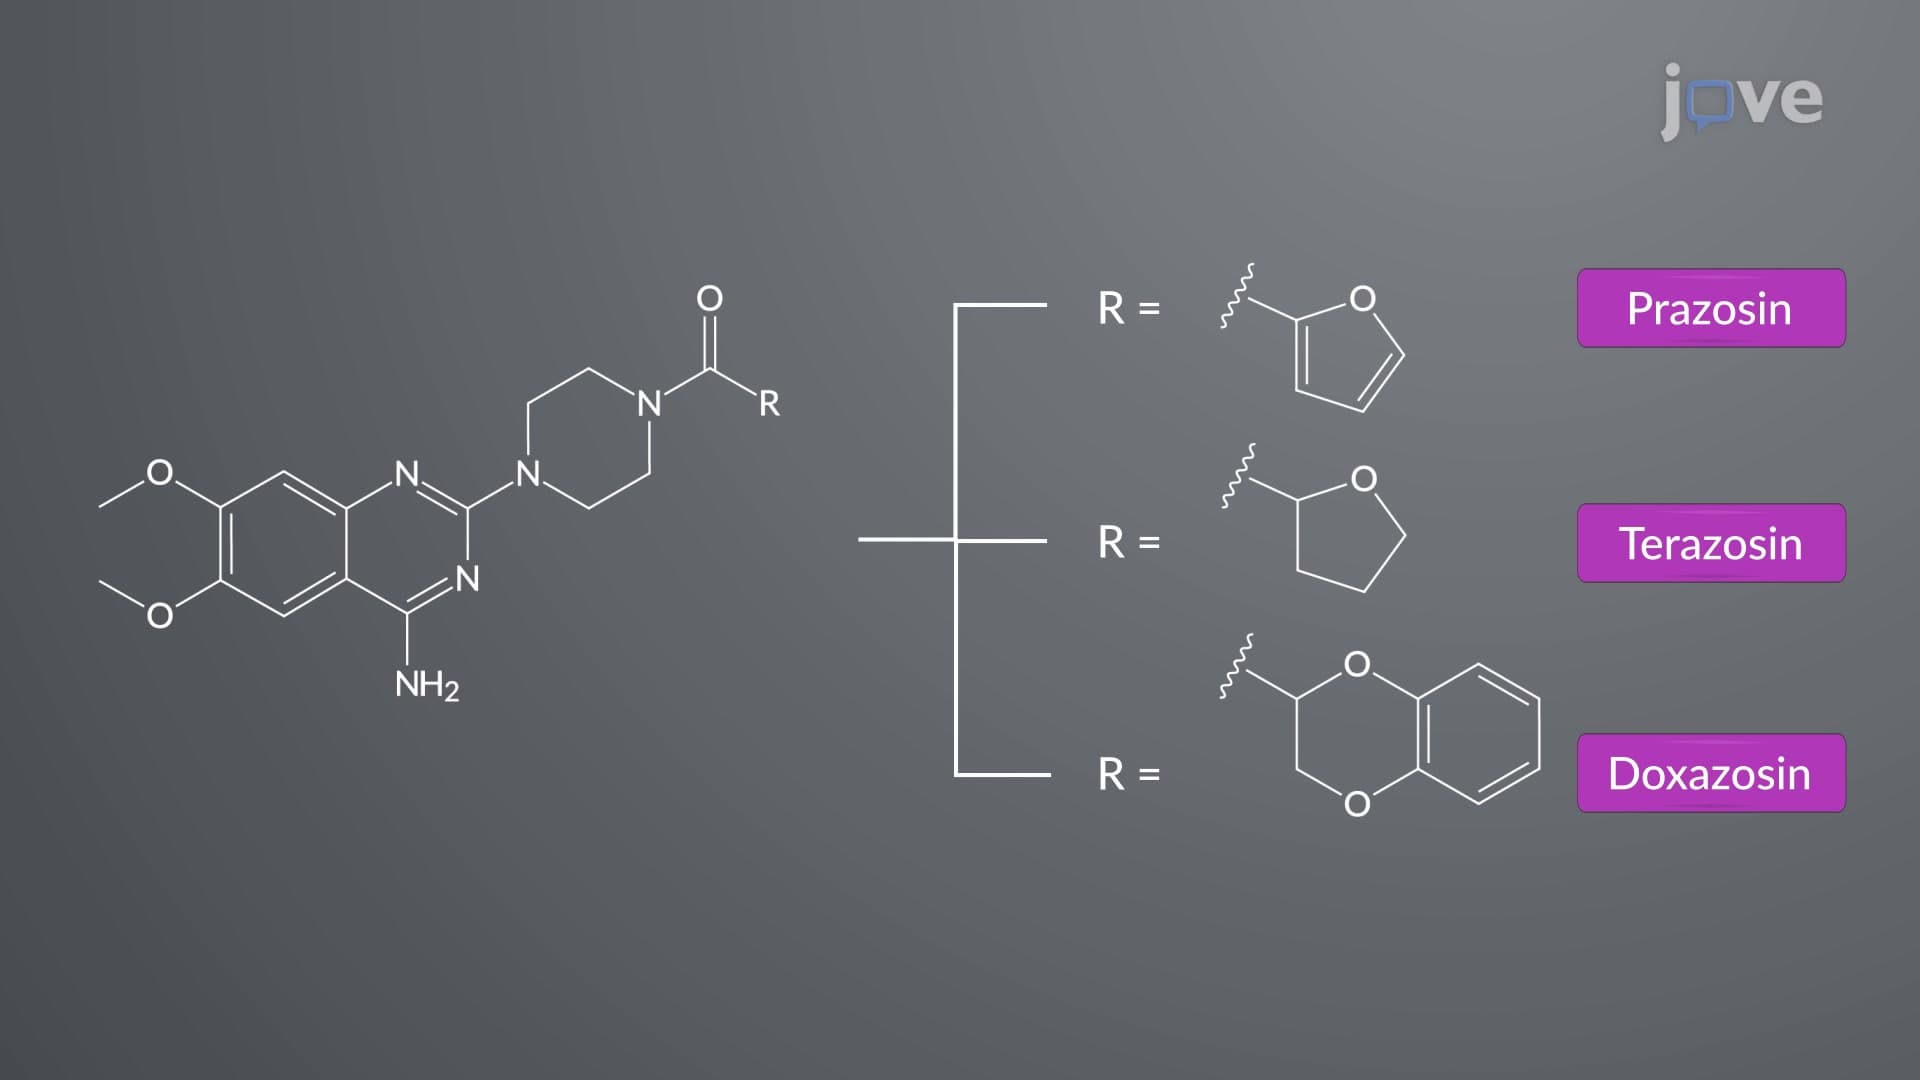
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:22
Direct-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
The direct-acting...
 01:29
Indirect-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
Reversible inhibitors display short to medium durations of action. Short-acting agents include simple alcohols with...