Related Experiment Video
Updated: Jul 4, 2026
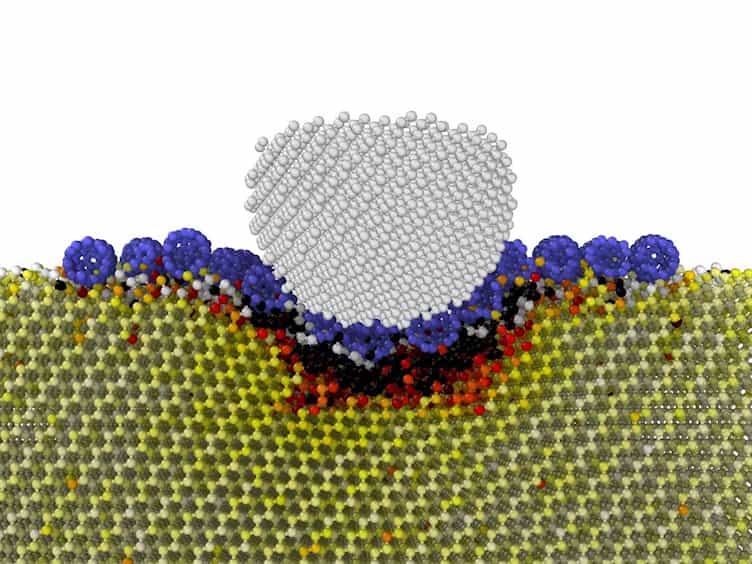

Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Full-dimensional quantum calculations of ground-state tunneling splitting of malonaldehyde using an accurate ab
Yimin Wang1, Bastiaan J Braams, Joel M Bowman
1Cherry L. Emerson Center for Scientific Computation and Department of Chemistry, Emory University, Atlanta, Georgia 30322, USA.

Abstract:
Quantum calculations of the ground vibrational state tunneling splitting of H-atom and D-atom transfer in malonaldehyde are performed on a full-dimensional ab initio potential energy surface (PES). The PES is a fit to 11 147 near basis-set-limit frozen-core CCSD(T) electronic energies. This surface properly describes the invariance of the potential with respect to all permutations of identical atoms. The saddle-point barrier for the H-atom transfer on the PES is 4.1 kcalmol, in excellent agreement with the reported ab initio value. Model one-dimensional and "exact" full-dimensional calculations of the splitting for H- and D-atom transfer are done using this PES. The tunneling splittings in full dimensionality are calculated using the unbiased "fixed-node" diffusion Monte Carlo (DMC) method in Cartesian and saddle-point normal coordinates. The ground-state tunneling splitting is found to be 21.6 cm(-1) in Cartesian coordinates and 22.6 cm(-1) in normal coordinates, with an uncertainty of 2-3 cm(-1). This splitting is also calculated based on a model which makes use of the exact single-well zero-point energy (ZPE) obtained with the MULTIMODE code and DMC ZPE and this calculation gives a tunneling splitting of 21-22 cm(-1). The corresponding computed splittings for the D-atom transfer are 3.0, 3.1, and 2-3 cm(-1). These calculated tunneling splittings agree with each other to within less than the standard uncertainties obtained with the DMC method used, which are between 2 and 3 cm(-1), and agree well with the experimental values of 21.6 and 2.9 cm(-1) for the H and D transfer, respectively.


Related Concept Videos
 01:18
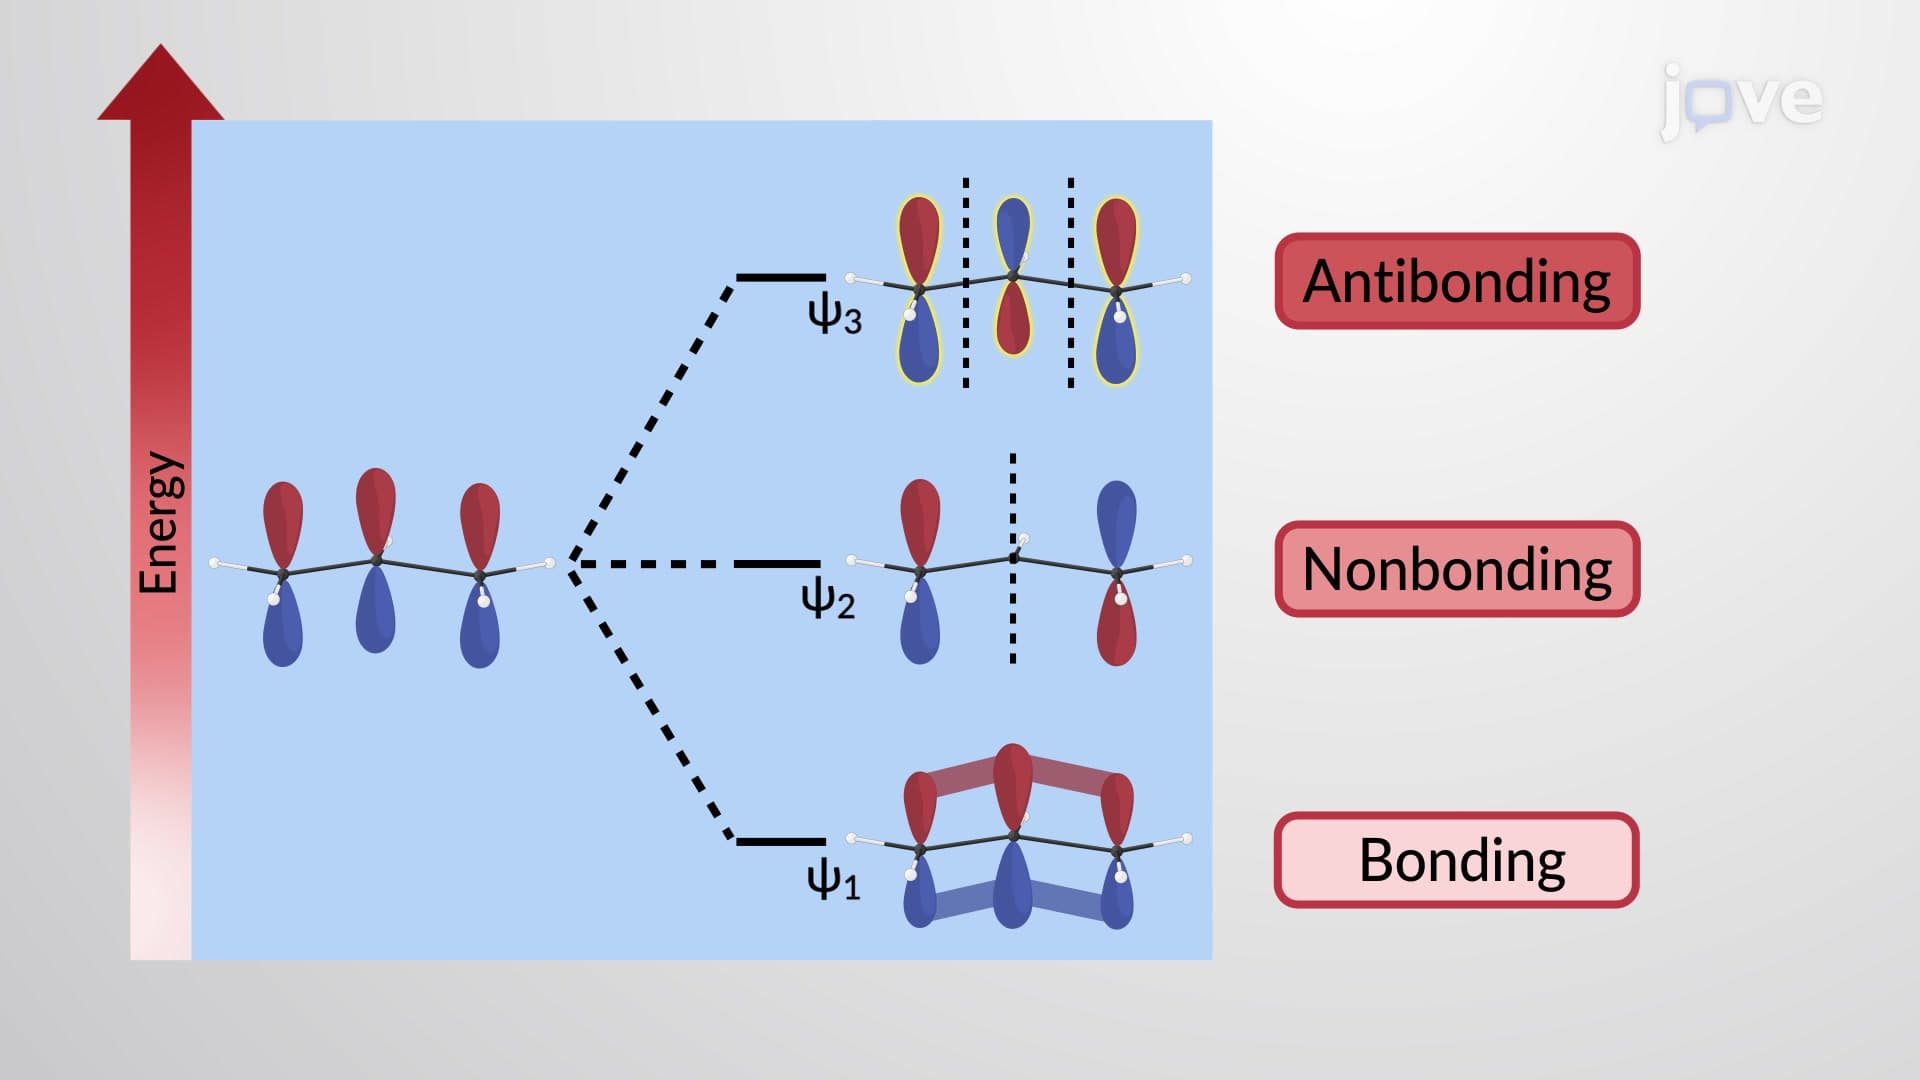
π Molecular Orbitals of the Allyl Cation and Anion
 03:51
Molecular Orbital Theory II
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:29
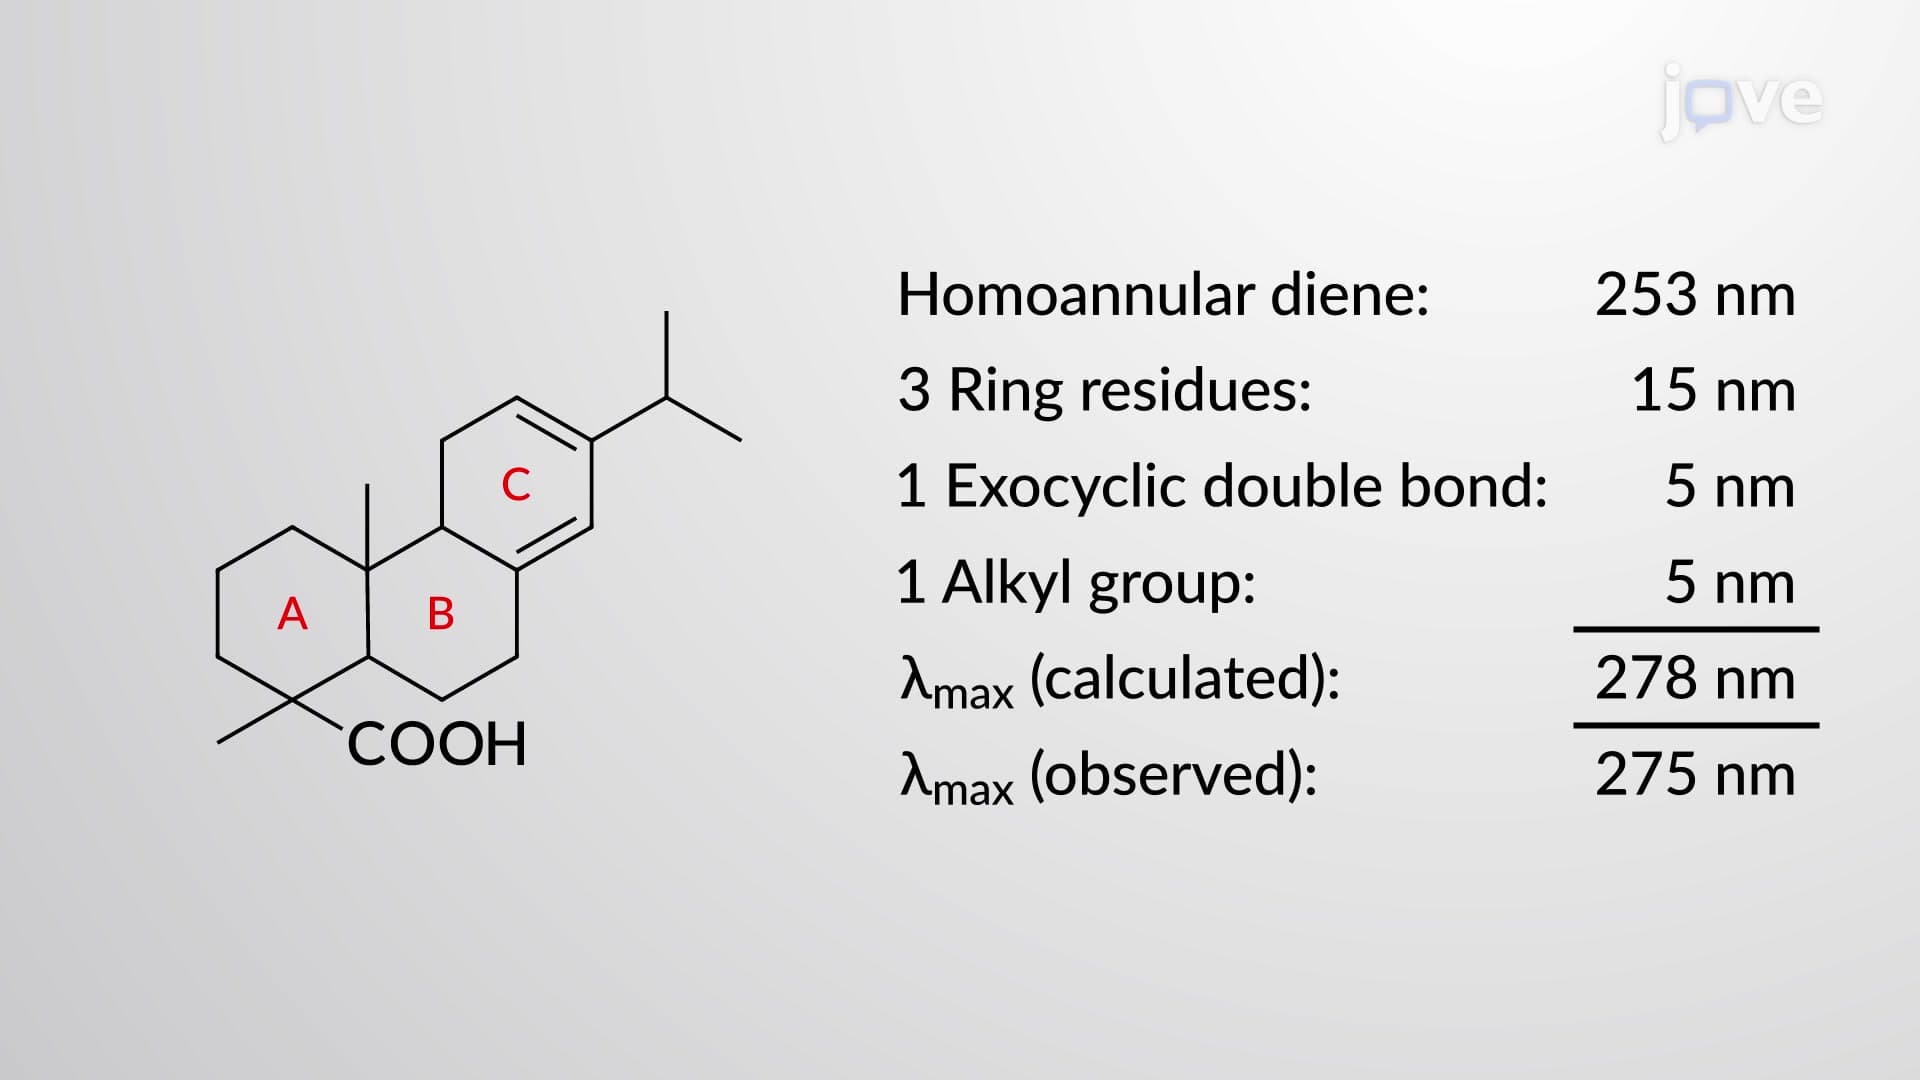
UV–Vis Spectroscopy: Woodward–Fieser Rules
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:10
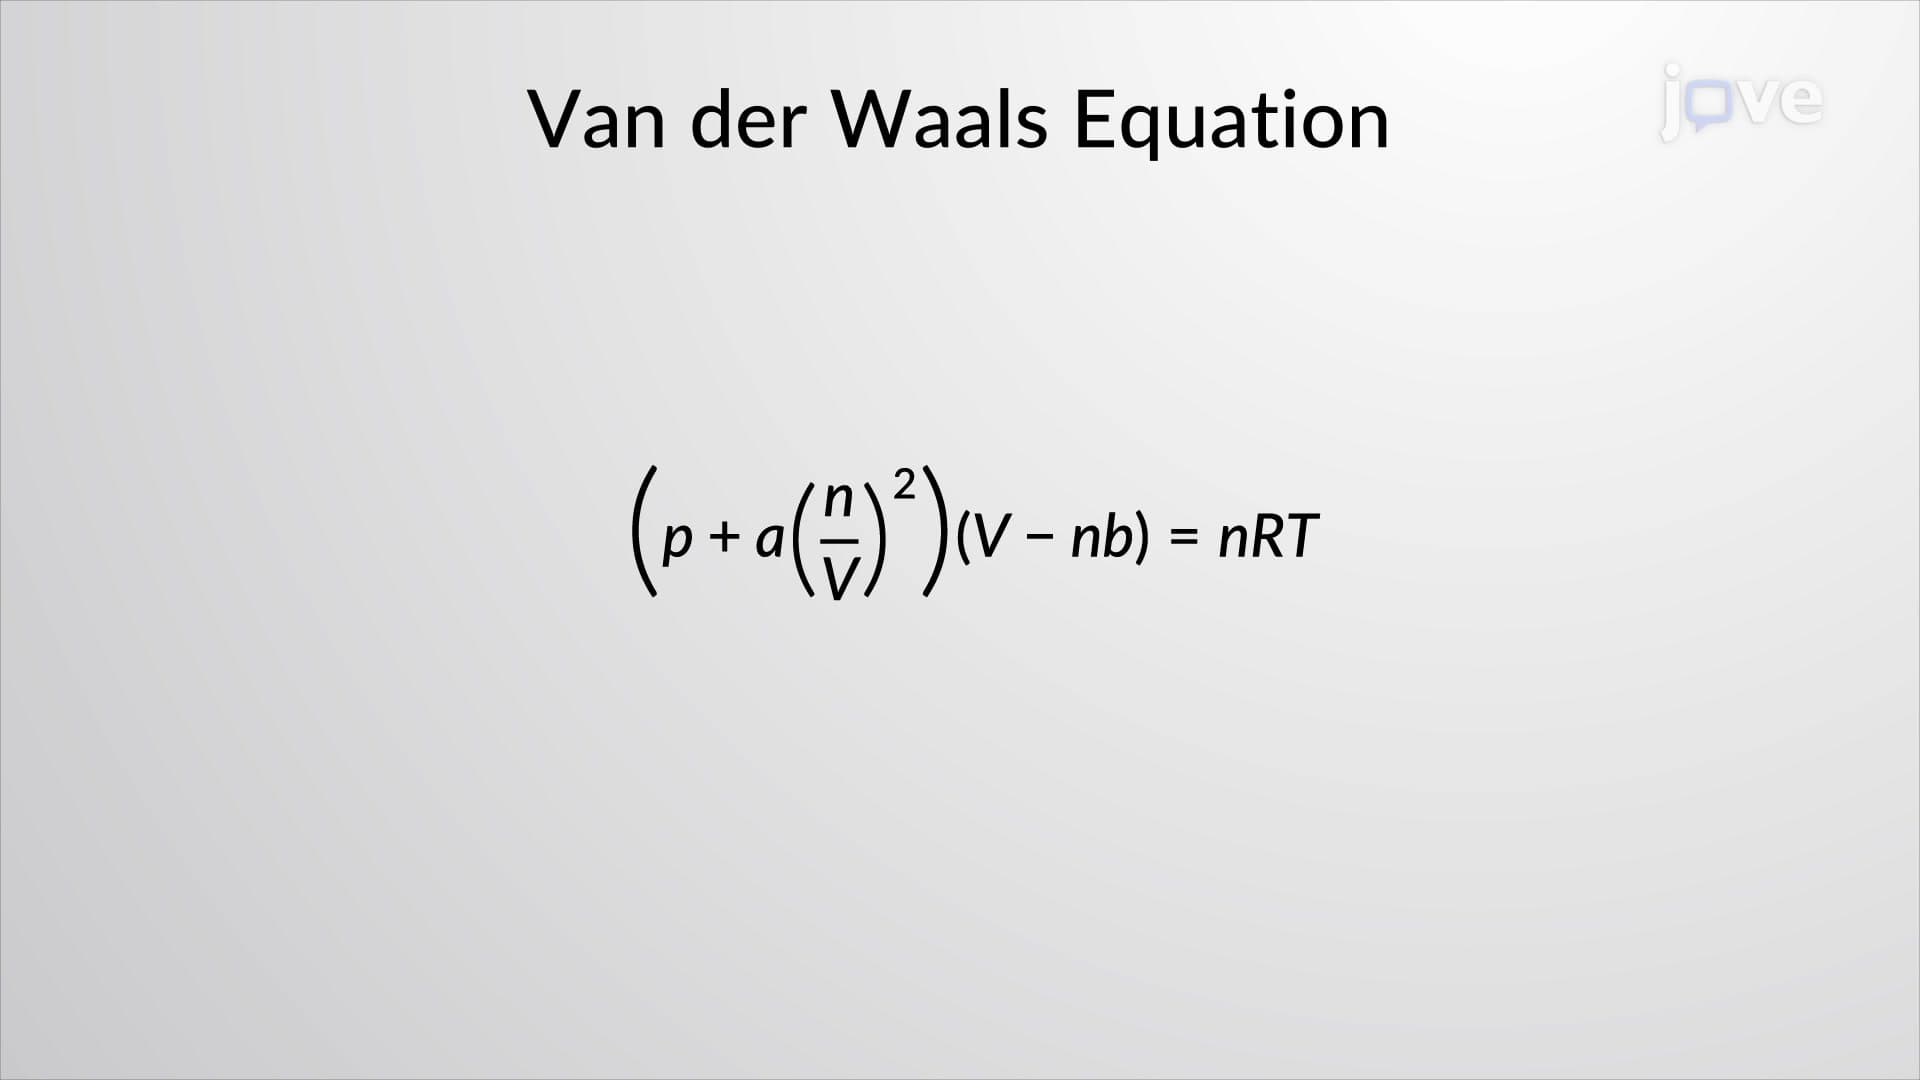
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...