Related Experiment Video
Updated: Jul 3, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Simple coupled-cluster singles and doubles method with perturbative inclusion of triples and explicitly correlated
Edward F Valeev1, T Daniel Crawford
1Department of Chemistry, Virginia Tech, Blacksburg, Virginia 24061, USA. evaleev@vt.edu

Abstract:
To approach the complete basis set limit of the "gold-standard" coupled-cluster singles and doubles plus perturbative triples [CCSD(T)] method, we extend the recently proposed perturbative explicitly correlated coupled-cluster singles and doubles method, CCSD(2)(R12) [E. F. Valeev, Phys. Chem. Chem. Phys. 8, 106 (2008)], to account for the effect of connected three-electron correlations. The natural choice of the zeroth-order Hamiltonian produces a perturbation expansion with rigorously separable second-order energy corrections due to the explicitly correlated geminals and conventional triple and higher excitations. The resulting CCSD(T)(R12) energy is defined as a sum of the standard CCSD(T) energy and an amplitude-dependent geminal correction. The method is technically very simple: Its implementation requires no modification of the standard CCSD(T) program and the formal cost of the geminal correction is small. We investigate the performance of the open-shell version of the CCSD(T)(R12) method as a possible replacement of the standard complete-basis-set CCSD(T) energies in the high accuracy extrapolated ab initio thermochemistry model of Stanton et al. [J. Chem. Phys. 121, 11599 (2004)]. Correlation contributions to the heat of formation computed with the new method in an aug-cc-pCVXZ basis set have mean absolute basis set errors of 2.8 and 1.0 kJmol when X is T and Q, respectively. The corresponding errors of the standard CCSD(T) method are 9.1, 4.0, and 2.1 kJmol when X=T, Q, and 5. Simple two-point basis set extrapolations of standard CCSD(T) energies perform better than the explicitly correlated method for absolute correlation energies and atomization energies, but no such advantage found when computing heats of formation. A simple Schwenke-type two-point extrapolation of the CCSD(T)(R12)aug-cc-pCVXZ energies with X=T,Q yields the most accurate heats of formation found in this work, in error on average by 0.5 kJmol and at most by 1.7 kJmol.

Related Concept Videos
 01:20
Spin–Spin Coupling: Two-Bond Coupling (Geminal Coupling)
The central atom need not be NMR-active because its electrons are affected by the electron polarization of the spin-active atoms. However, spin information is transmitted less effectively than in one-bond coupling, and 2J values are usually weaker than 1J values. The energy of...
 01:22
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
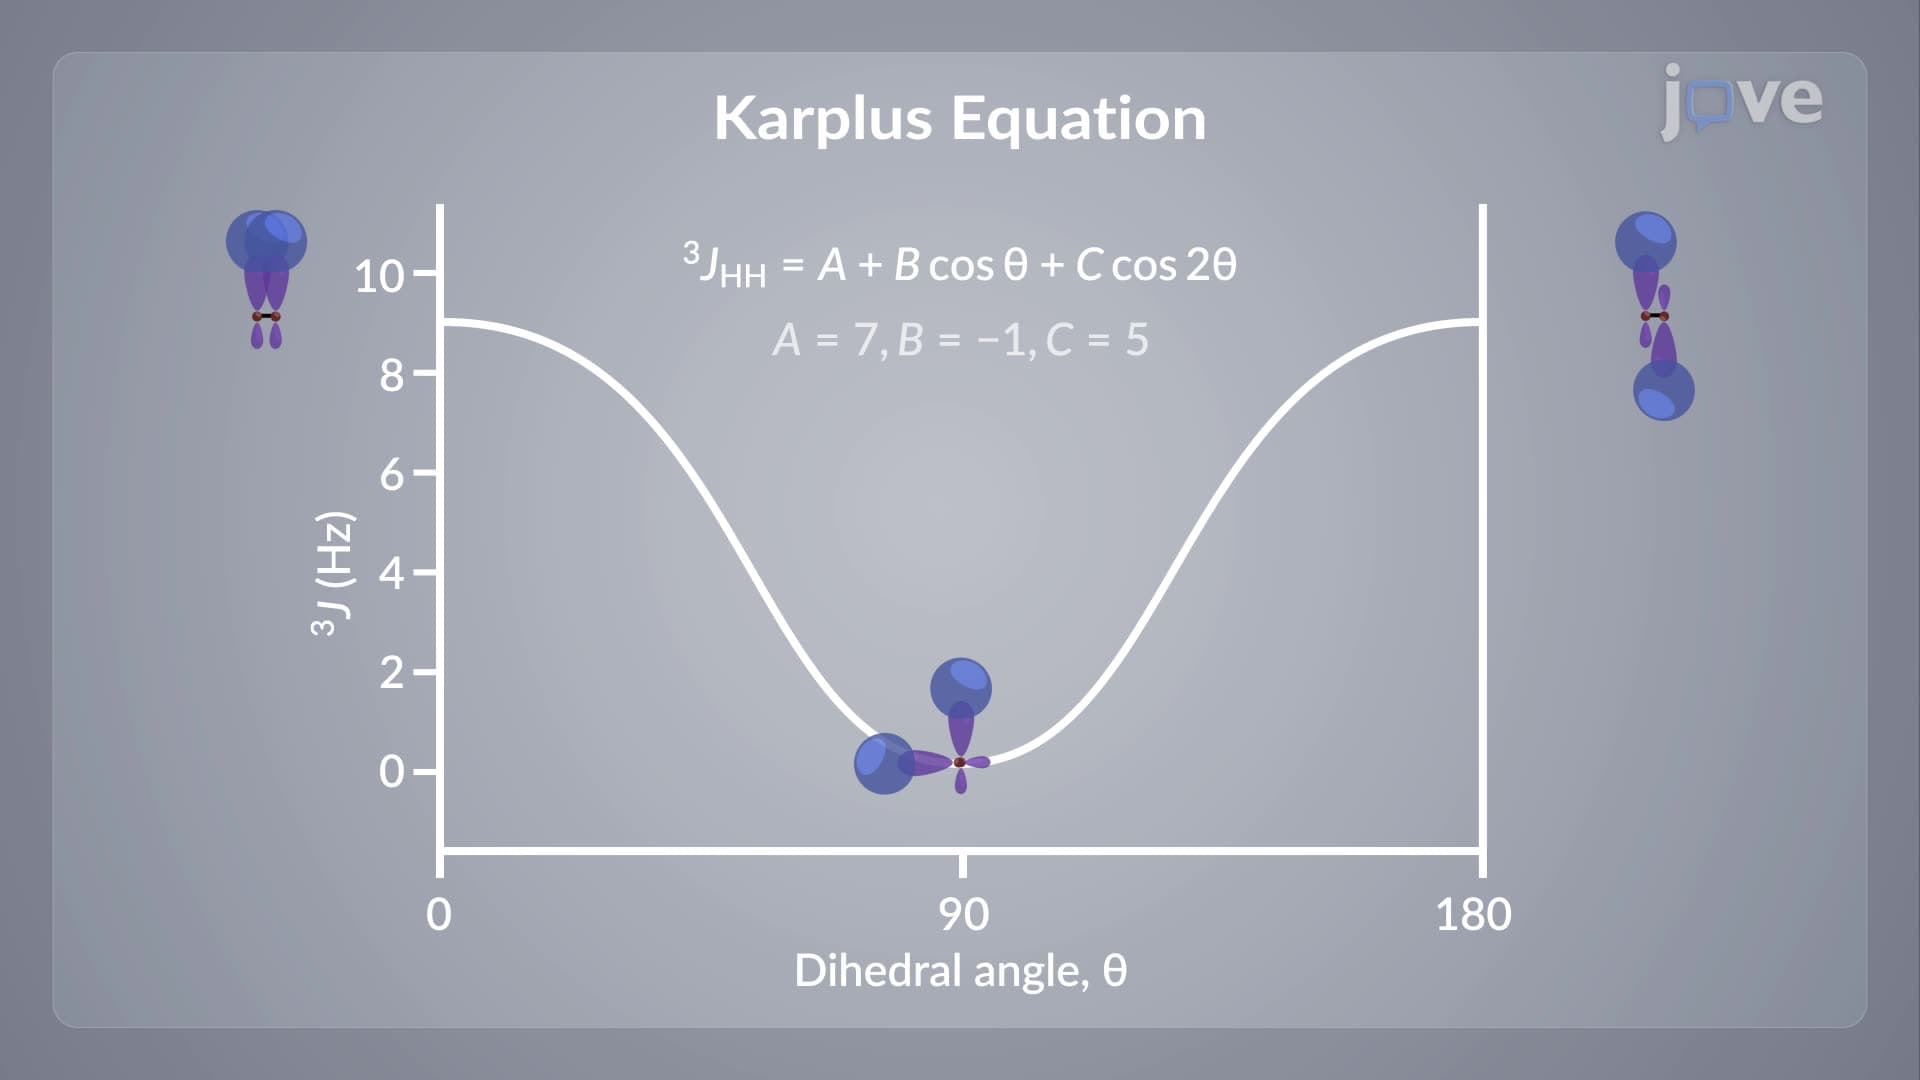
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene π orbitals.
 01:12
Reduced Mass Coordinates: Isolated Two-body Problem
 01:17
Spin–Spin Coupling: One-Bond Coupling
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...