Related Experiment Video
Updated: Jul 3, 2026
Versatile CO2 Transformations into Complex Products: A One-pot Two-step Strategy
Published on: November 9, 2019
Structural characterization of a 140 degrees domain movement in the two-step reaction catalyzed by
Albert S Reger1, Rui Wu, Debra Dunaway-Mariano
1Hauptman-Woodward Medical Research Institute, Buffalo, New York 14203, USA.

Abstract:
Members of the adenylate-forming family of enzymes play a role in the metabolism of halogenated aromatics and of short, medium, and long chain fatty acids, as well as in the biosynthesis of menaquinone, peptide antibiotics, and peptide siderophores. This family includes a subfamily of acyl- and aryl-CoA ligases that catalyze thioester synthesis through two half-reactions. A carboxylate substrate first reacts with ATP to form an acyl-adenylate. Subsequent to the release of the product PP i, the enzyme binds CoA, which attacks the activated acyl group to displace AMP. Structural and functional studies on different family members suggest that these enzymes alternate between two conformations during catalysis of the two half-reactions. Specifically, after the initial adenylation step, the C-terminal domain rotates by approximately 140 degrees to adopt a second conformation for thioester formation. Previously, we determined the structure of 4-chlorobenzoate:CoA ligase (CBL) in the adenylate forming conformation bound to 4-chlorobenzoate. We have determined two new crystal structures. We have determined the structure of CBL in the original adenylate-forming conformation, bound to the adenylate intermediate. Additionally, we have used a novel product analogue, 4-chlorophenacyl-CoA, to trap the enzyme in the thioester-forming conformation and determined this structure in a new crystal form. This work identifies a novel binding pocket for the CoA nucleotide. The structures presented herein provide the foundation for biochemical analyses presented in the accompanying manuscript in this issue [Wu et al. (2008) Biochemistry 47, 8026-8039]. The complete characterization of this enzyme allows us to provide an explanation for the use of the domain alternation strategy by these enzymes.
Related Concept Videos
 01:21
[3,3] Sigmatropic Rearrangement of 1,5-Dienes: Cope Rearrangement
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration
 01:11
ortho–para-Directing Activators: –CH3, –OH, –NH2, –OCH3
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
 01:15
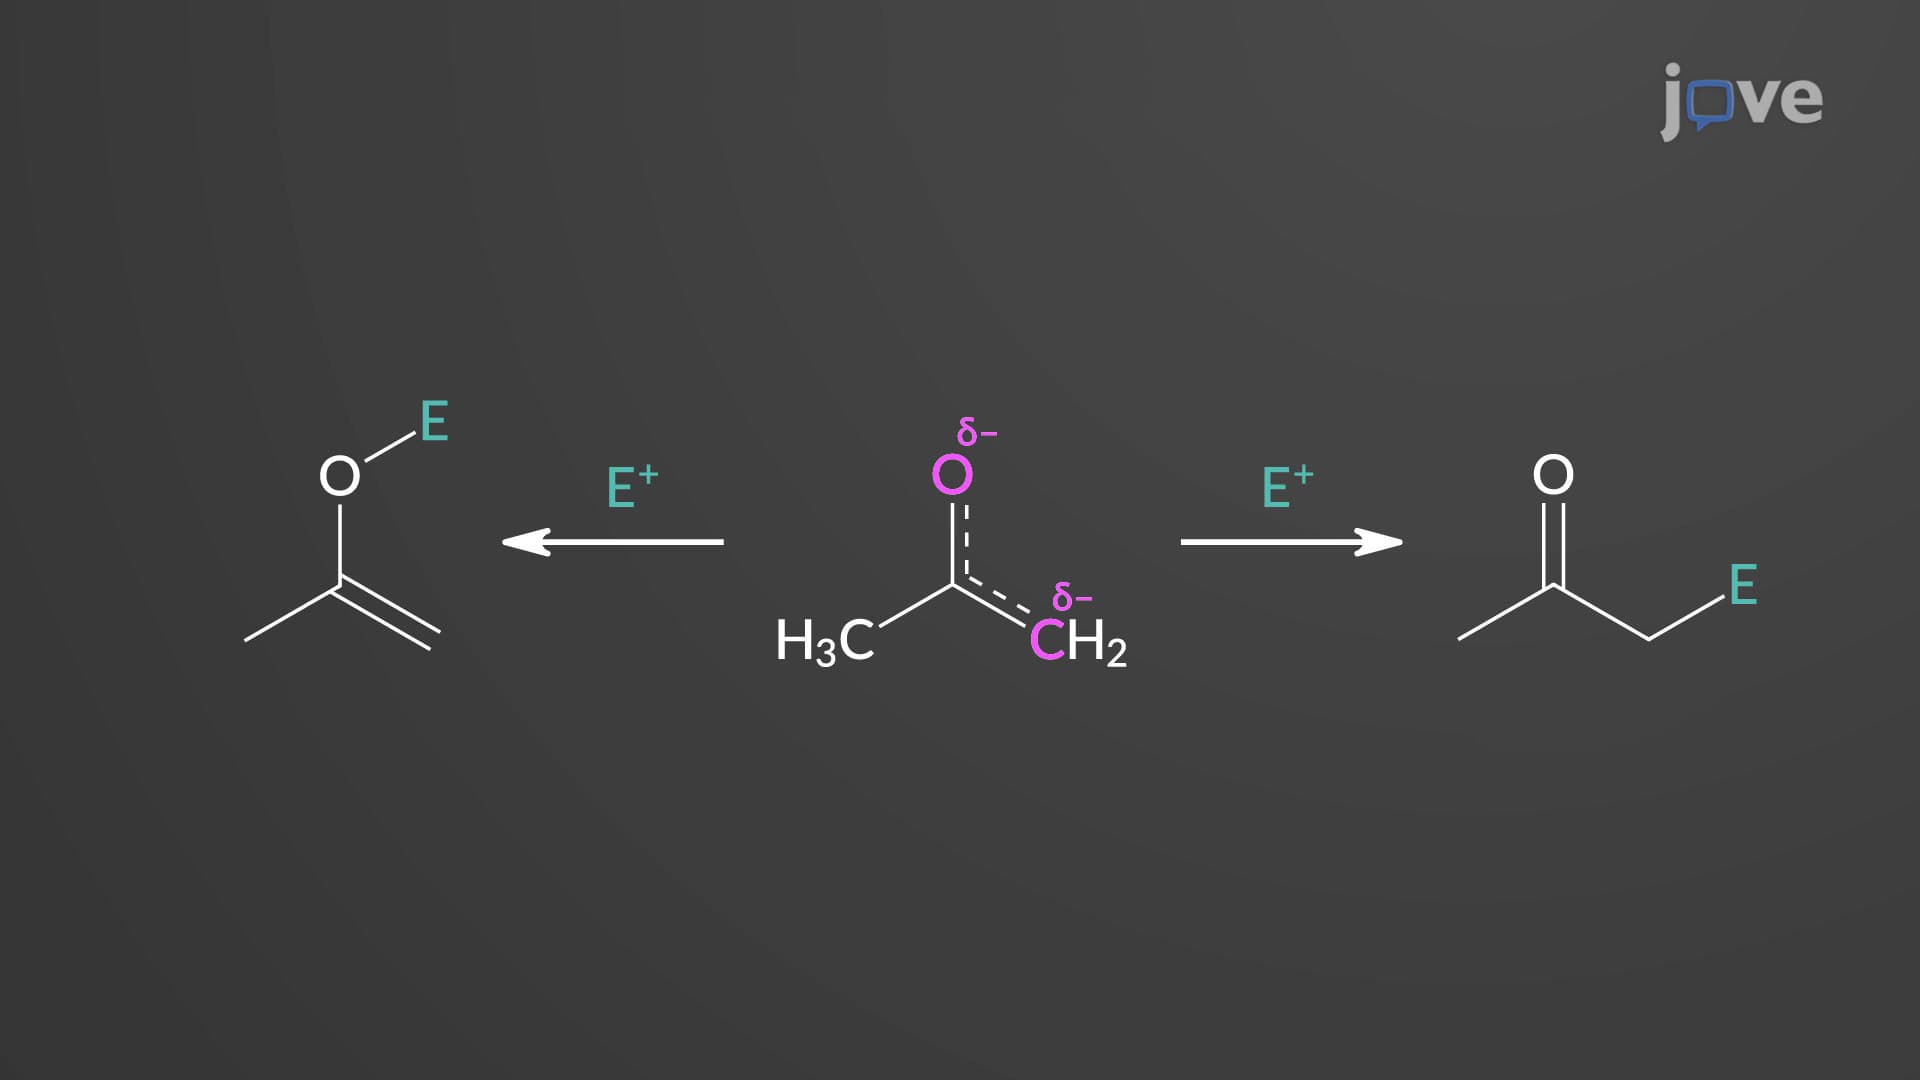
Enolate Mechanism Conventions
C-attack...
 01:29
Diels–Alder Reaction Forming Bridged Bicyclic Products: Stereochemistry