Related Experiment Video
Updated: Jun 30, 2026

Protein WISDOM: A Workbench for In silico De novo Design of BioMolecules
Published on: July 25, 2013
Calculation of absolute ligand binding free energy to a ribosome-targeting protein as a function of solvent model
1Computational Sciences and Engineering Branch, U.S. Army Research Laboratory, Aberdeen Proving Ground, Maryland 21005, USA. michael.lee@amedd.army.mil

Abstract:
A comparative analysis is provided of the effect of different solvent models on the calculation of a potential of mean force (PMF) for determining the absolute binding affinity of the small molecule inhibitor pteroic acid bound to ricin toxin A-chain (RTA). Solvent models include the distance-dependent dielectric constant, several different generalized Born (GB) approximations, and a hybrid explicit/GB-based implicit solvent model. We found that the simpler approximation of dielectric screening and a GB model, with Born radii fitted to a switching-window dielectric-boundary surface Poisson solvent model, severely overpredicted the binding affinity as compared to the experimental value, estimated to range from -4.4 to -6.0 kcal/mol. In contrast, GB models that are parametrized to fit the Lee-Richards molecular surface performed much better, predicting binding free energy within 1-3 kcal/mol of experimental estimates. However, the predicted free-energy profiles of these GB models displayed alternative binding modes not observed in the crystal structure. Finally, the most rigorous and computationally costly approach in this work, which used a hybrid explicit/implicit solvent model, correctly determined a binding funnel in the PMF near the crystallographic bound state and predicted an absolute binding affinity that was 2 kcal/mol more favorable than the estimated experimental binding affinity.
More Related Videos

Related Concept Videos
 02:18
The Equilibrium Binding Constant and Binding Strength
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:22
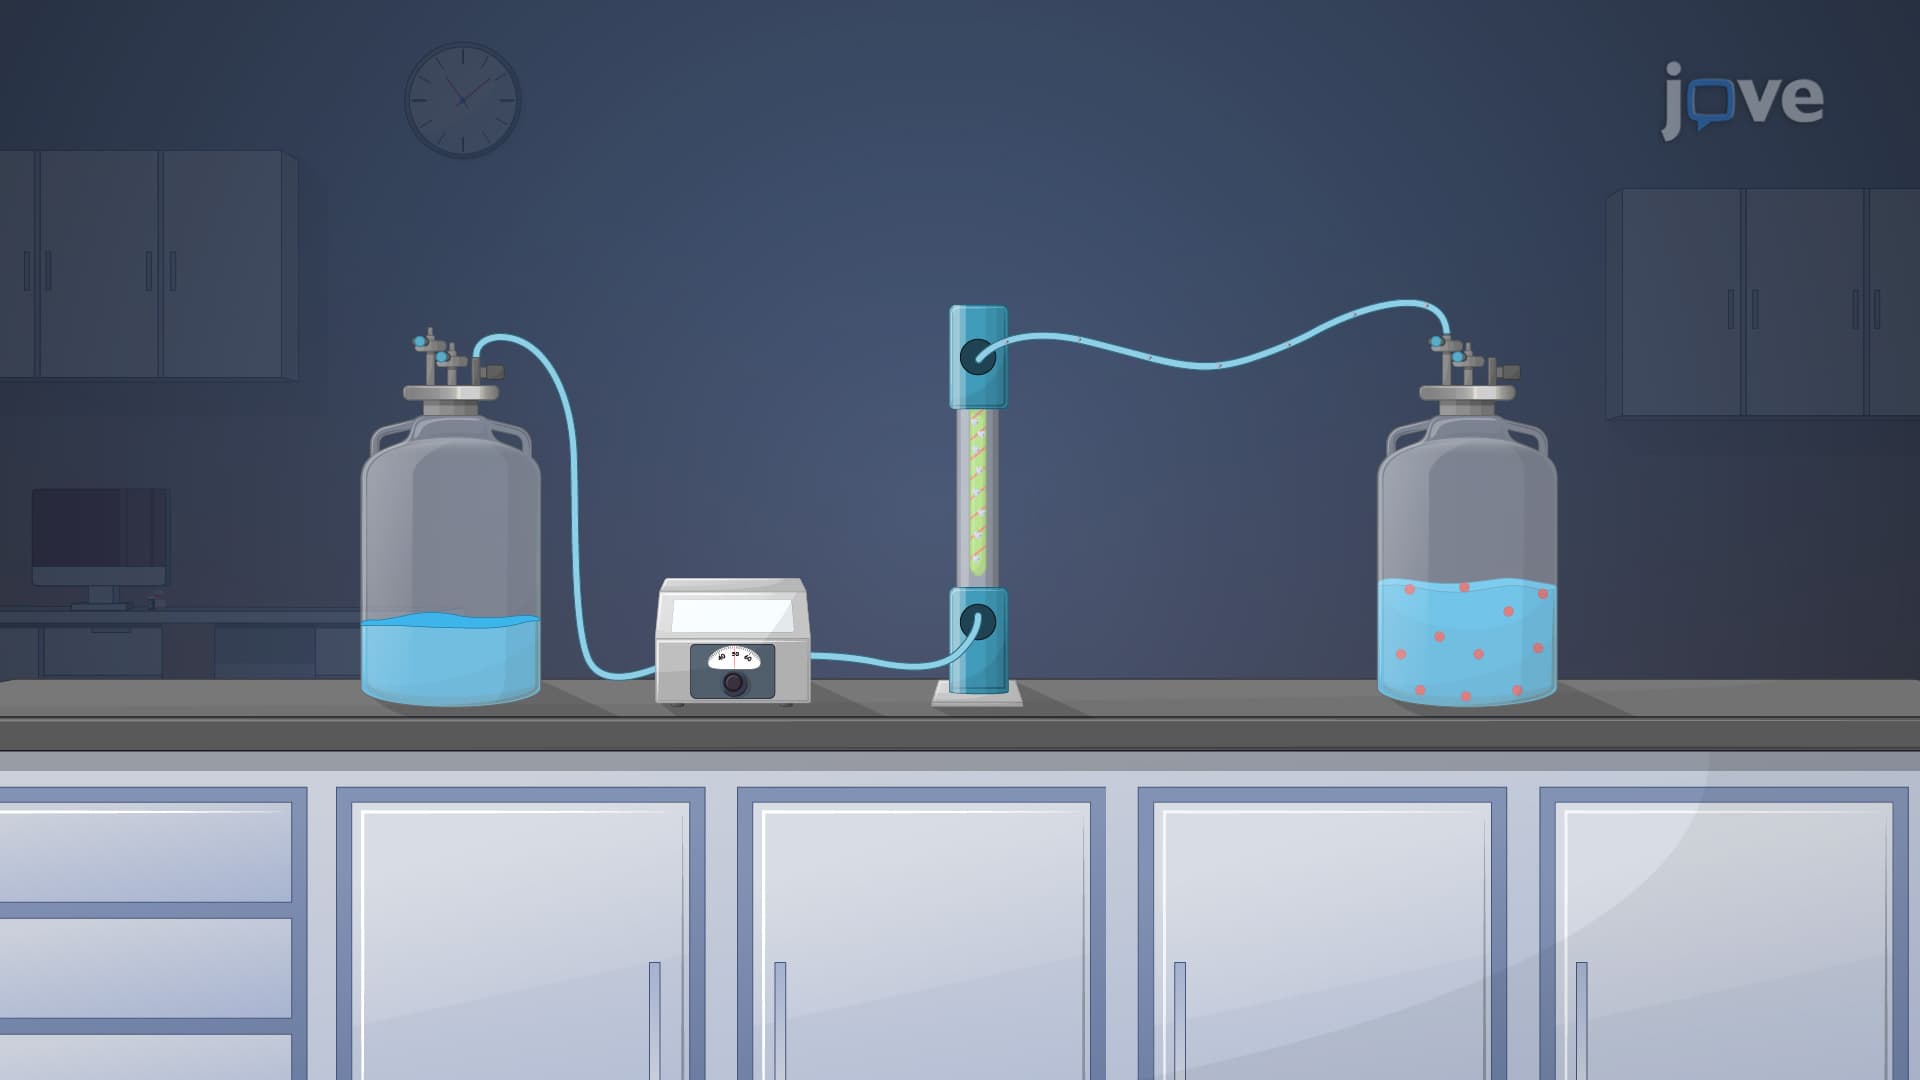
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...