Related Experiment Video
Updated: Jun 29, 2026

Isolating Free Carbenes, their Mixed Dimers and Organic Radicals
Published on: April 19, 2019
Molecular structure and benzene ring deformation of three cyanobenzenes from gas-phase electron diffraction and
Anna Rita Campanelli1, Aldo Domenicano, Fabio Ramondo
1Department of Chemistry, University of Rome "La Sapienza", I-00185 Rome, Italy.

Abstract:
The molecular structures of cyanobenzene, p-dicyanobenzene, and 1,2,4,5-tetracyanobenzene have been accurately determined by gas-phase electron diffraction and ab initio/DFT MO calculations. The equilibrium structures of these molecules are planar, but their average geometries in the gaseous phase are nonplanar because of large-amplitude vibrational motions of the substituents out of the plane of the benzene ring. The use of nonplanar models in electron diffraction analysis is necessary to yield ring angles consistent with the results of MO calculations. The angular deformation of the benzene ring in the three molecules is found to be much smaller than obtained from previous electron diffraction studies, as well as from microwave spectroscopy studies of cyanobenzene. While the deformation of the ring CC bonds and CCC angles in p-dicyanobenzene is well interpreted as arising from the superposition of independent effects from each substituent, considerable deviation from additivity occurs in 1,2,4,5-tetracyanobenzene. The changes in the ring geometry and C ipso-C cyano bond lengths in this molecule indicate an enhanced ability of the cyano group to withdraw pi-electrons from the benzene ring, compared with cyanobenzene and p-dicyanobenzene. In particular, gas-phase electron diffraction and MP2 or B3LYP calculations show a small but consistent increase in the mean length of the ring CC bonds for each cyano group and a further increase in 1,2,4,5-tetracyanobenzene. Comparison with accurate results from X-ray and neutron crystallography indicates that in p-dicyanobenzene the internal ring angle at the place of substitution opens slightly as the molecule is frozen in the crystal. The small geometrical change, about 0.6 degrees , is shown to be real and to originate from intermolecular C identical withN...HC interactions in the solid state.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 10:35
Novel Techniques for Observing Structural Dynamics of Photoresponsive Liquid Crystals
Published on: May 29, 2018
Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 01:07
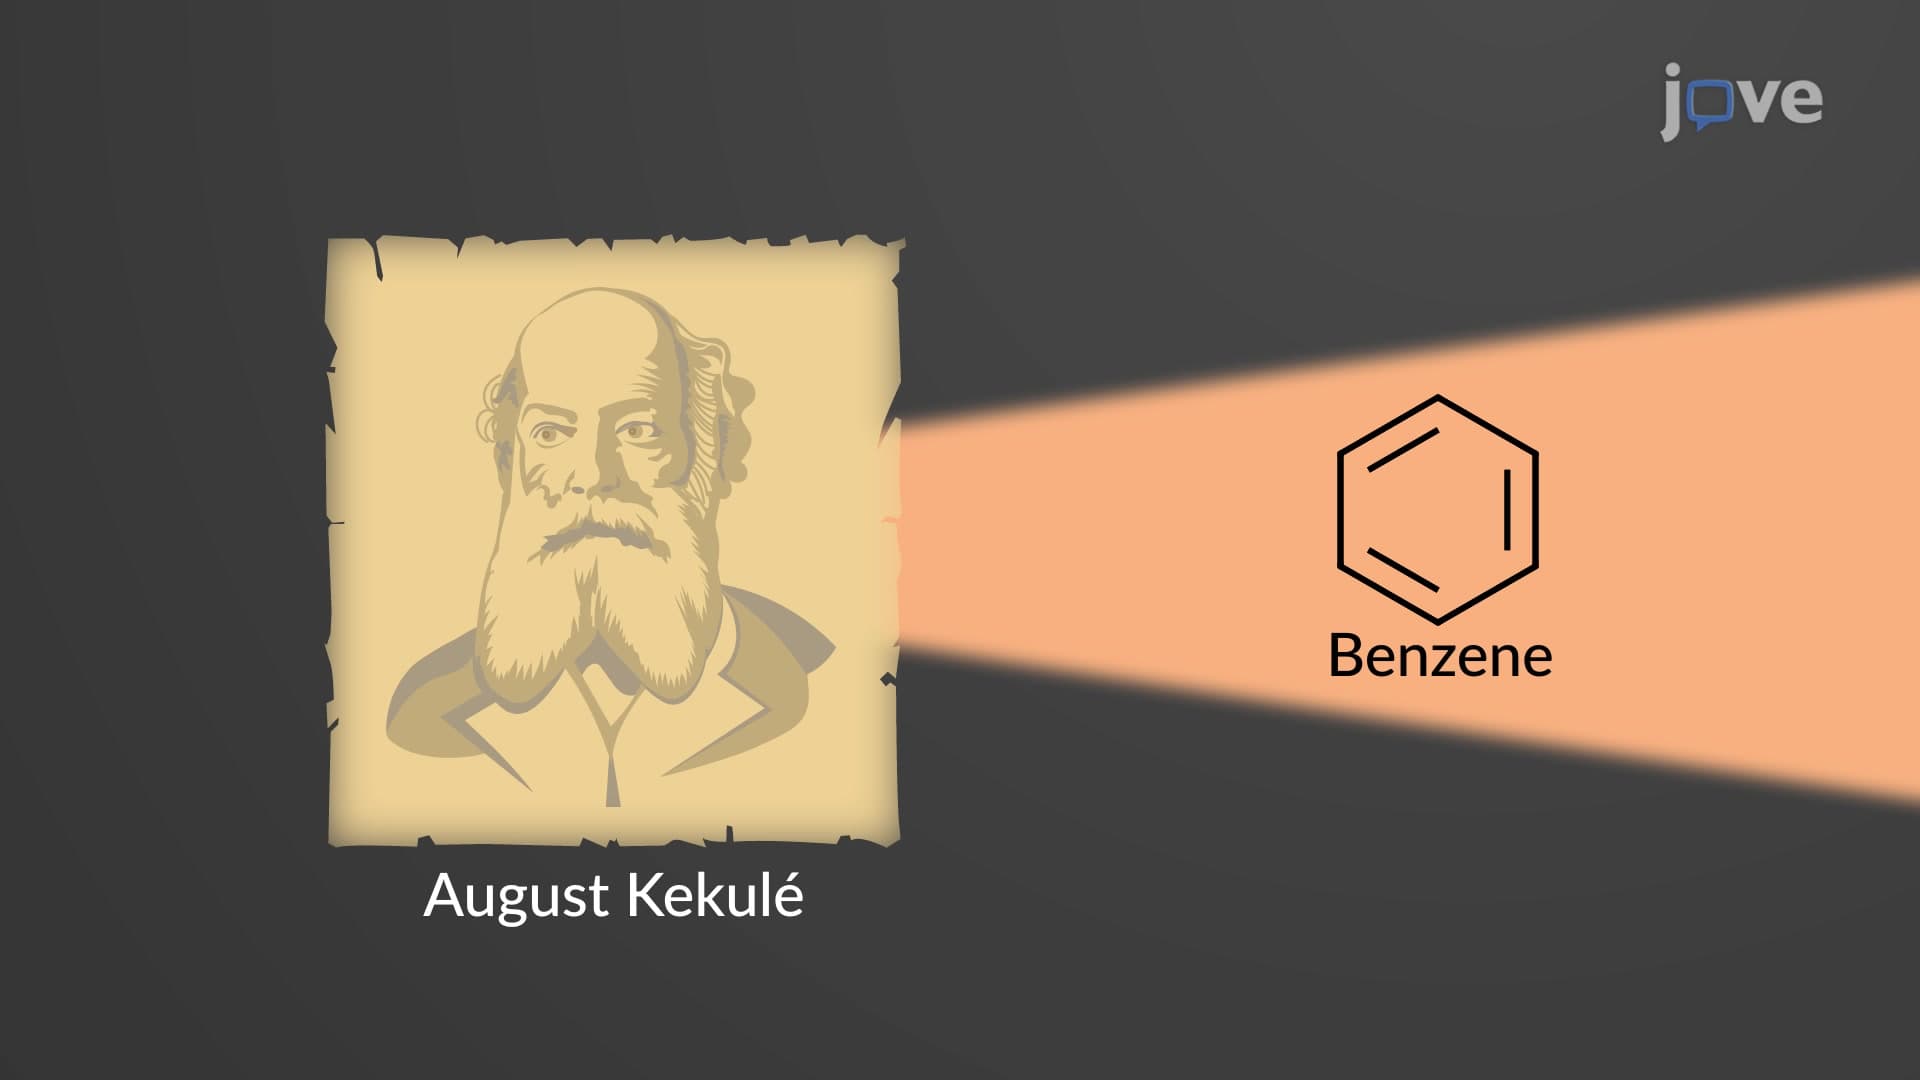
Structure of Benzene: Kekulé Model
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.
 02:29
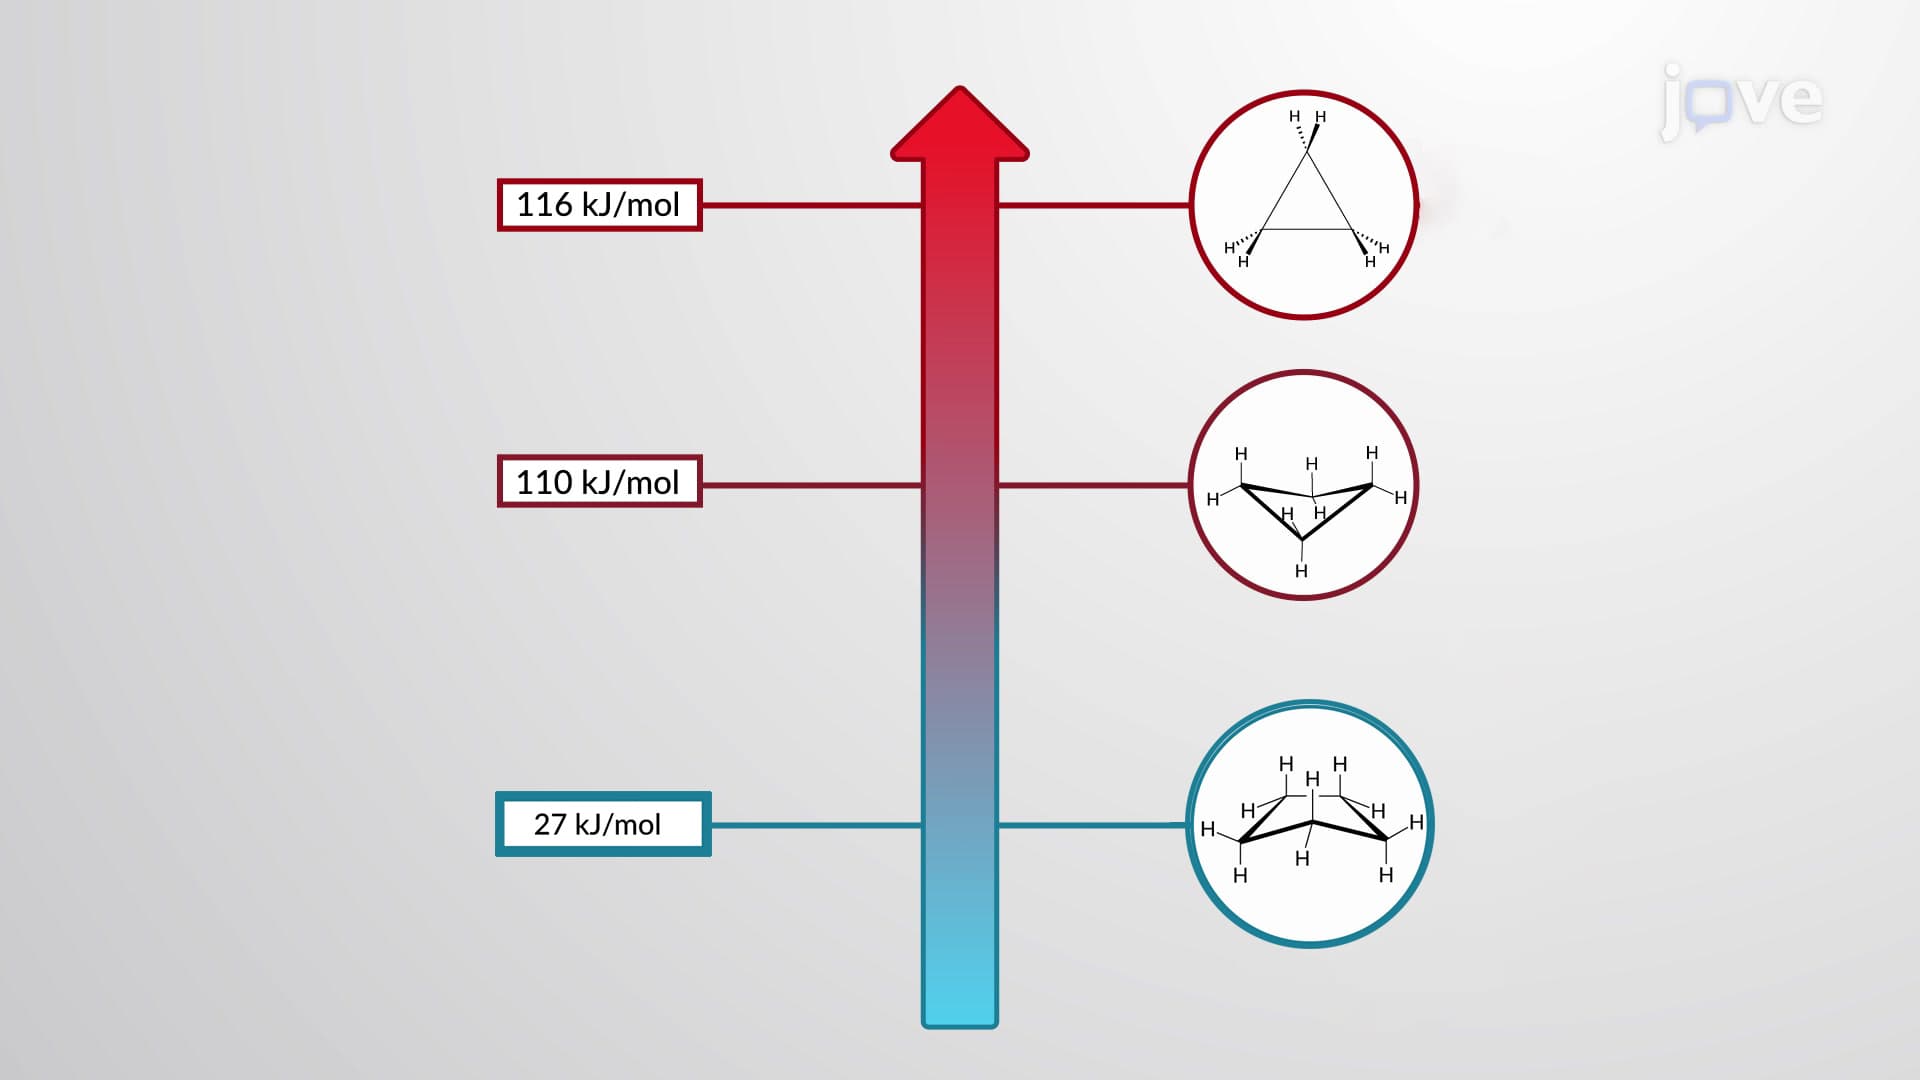
Conformations of Cycloalkanes
 01:37
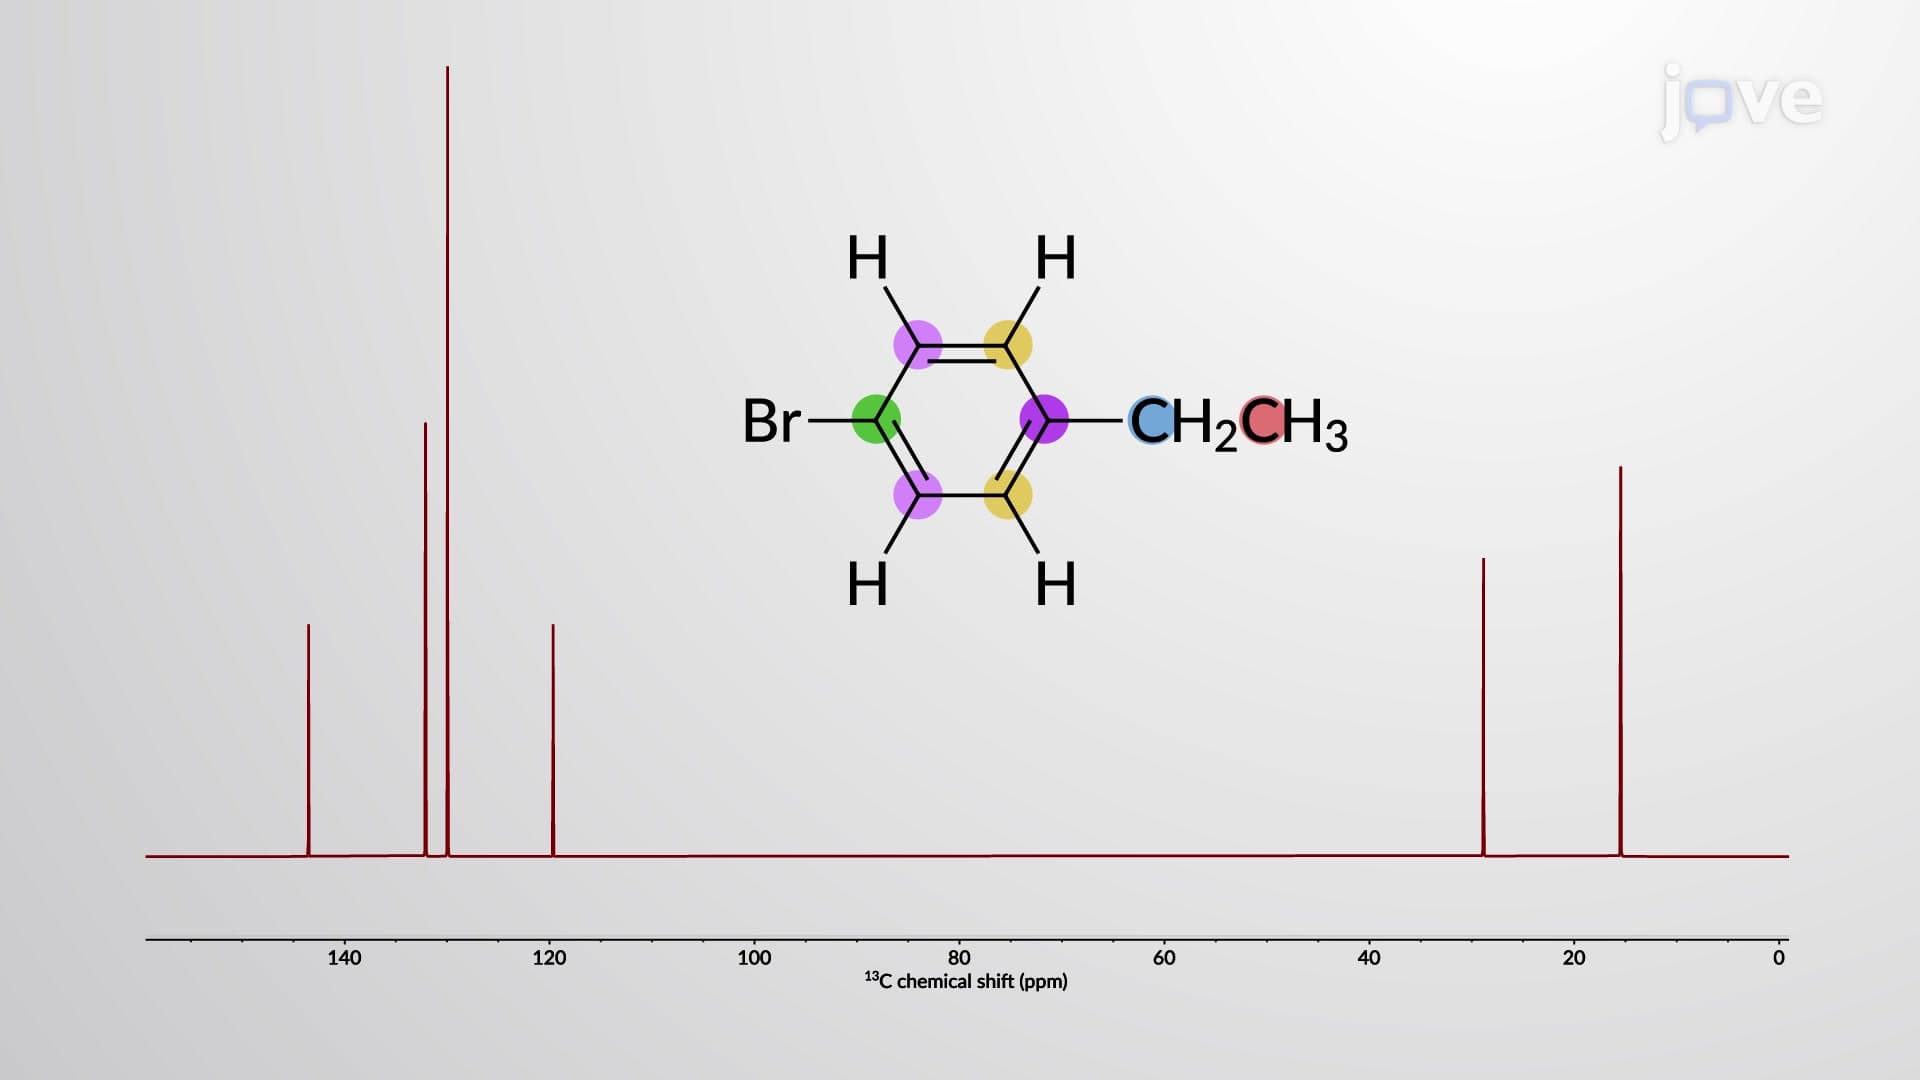
NMR Spectroscopy of Benzene Derivatives
 02:27
Predicting Molecular Geometry
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal tetrahedral value,...