Related Experiment Video
Updated: Oct 11, 2026

Time-resolved Photophysical Characterization of Triplet-harvesting Organic Compounds at an Oxygen-free Environment Using an iCCD Camera
Published on: December 27, 2018
Shared-reference analysis of pathway-dependent excited-state responses in donor-side and terminal-side substituted
1Department of Chemistry, College of Science, King Saud University, P.O. Box 2455, Riyadh 11451, Saudi Arabia.

Abstract:
Distinct substitution pathways within a conjugated chromophore may produce comparable changes in conventional energetic or spectral descriptors without necessarily producing comparable underlying excited-state electronic responses. Here, donor-side and terminal-side substitution pathways in a phenyl-thiophene framework were evaluated independently relative to the same molecular reference using a common gas-phase DFT/TD-DFT protocol. Ground-state structures and frontier orbitals were obtained at the B3LYP/6-311++G(d,p) level, while vertical singlet excitations were evaluated using TD-CAM-B3LYP/6-311++G(d,p). The lowest singlet state, S1, and the state with the largest oscillator strength within the common S1-S10 manifold, Sbright, were treated separately and analyzed using canonical orbital contributions, natural transition orbitals, quantitative real-space hole-electron descriptors, and interfragment charge-transfer (IFCT) analysis. Because the two pathways contain chemically different substituent sets, the comparison is interpreted in terms of pathway-dependent responses rather than as an independent separation of substitution-position and substituent-identity effects. The two predefined substitution pathways exhibit distinct electronic and excited-state response patterns within the examined molecular set. In the donor-side series, D0H and D1-OCH3 exhibit S1 ≠ Sbright, with the lowest and brightest states showing distinct hole-electron organization and qualitatively different fragment-resolved redistribution. In contrast, D2-NH2 and the shared reference J retain S1 = Sbright. All terminal-side derivatives preserve S1 = Sbright within the calculated S₁-S₁₀ manifold while exhibiting substituent-dependent spatial and fragment-resolved responses. T2-NO2 is the only terminal-side derivative to show a calculated red shift relative to J, with its S1 excitation energy lower by 0.310 eV, together with the largest hole-electron centroid distance among the terminal-side derivatives and pronounced redistribution toward its terminal NO2 region. Targeted TD-ωB97X-D calculations for J, D1-OCH3, and T2-NO2 preserve the principal state assignments and the relative energetic directions examined for D1-OCH3 and T2-NO2, while showing functional sensitivity in the absolute excitation properties. Together, the NTO, real-space, and IFCT analyses show that transition-density pattern, spatial hole-electron organization, and interfragment redistribution provide complementary information and should not be treated as interchangeable descriptors. The shared-reference, state-resolved comparison therefore identifies pathway-dependent electronic responses within the investigated molecular set and shows that similarity in energetic or spectral descriptors alone does not establish similarity in the underlying excited-state electronic organization.
Related Concept Videos
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 01:26
Thermal and Photochemical Electrocyclic Reactions: Overview
 01:29
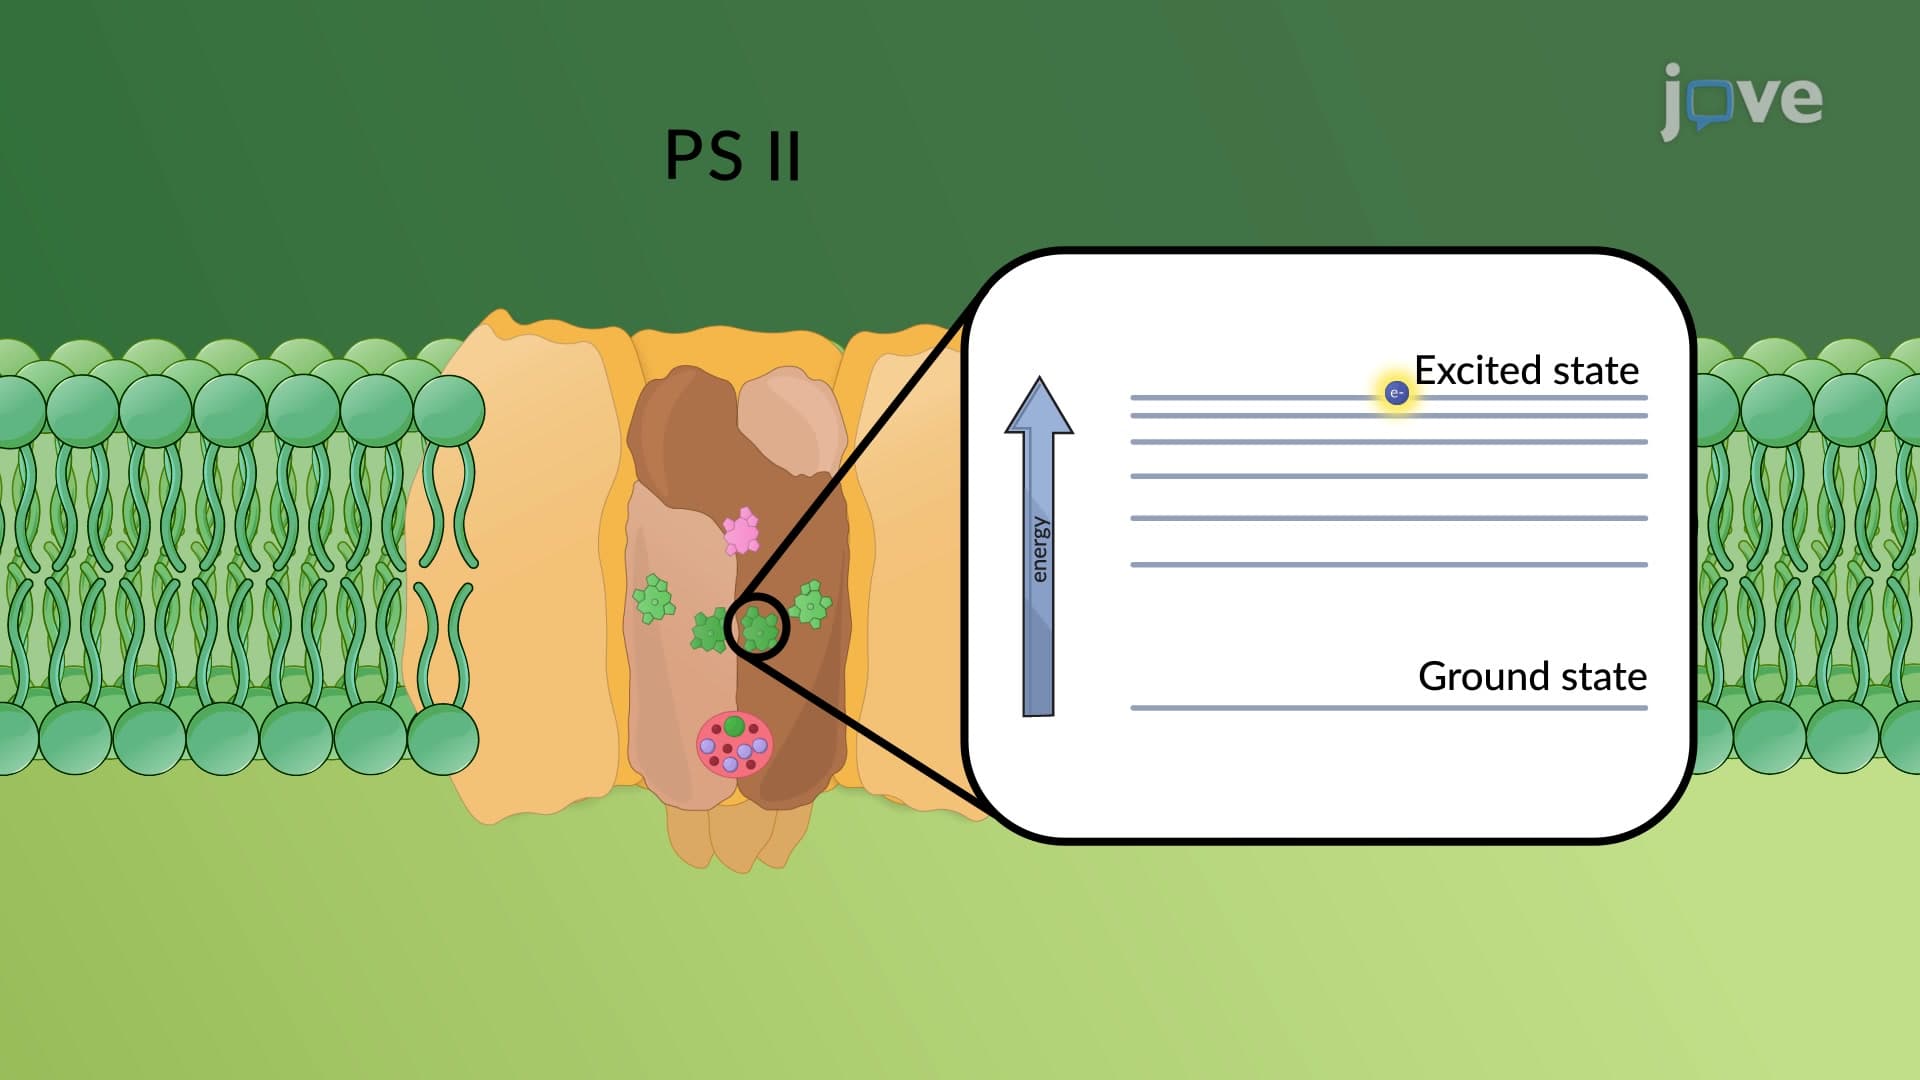
The Photochemical Reaction Center
 01:22
Photoreceptors and Visual Pathways
 01:12
Cycloaddition Reactions: MO Requirements for Photochemical Activation
 01:17
Thermal Electrocyclic Reactions: Stereochemistry
Selection Rules: Thermal Activation
Conjugated systems containing an even number of π-electron pairs undergo a conrotatory ring closure. For example, thermal electrocyclization of (2E,4E)-2,4-hexadiene, a conjugated diene containing two π-electron pairs, gives trans-3,4-dimethylcyclobutene.