Related Experiment Video
Updated: Jun 29, 2026

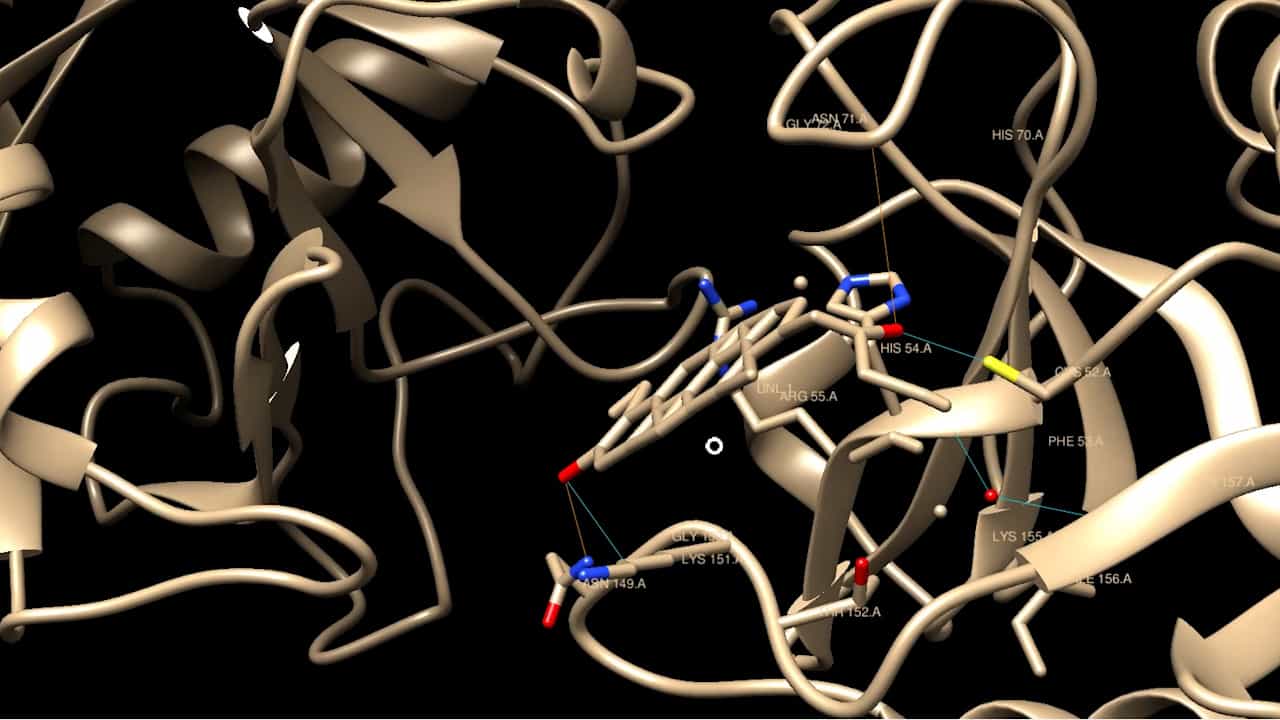
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Protein docking by the underestimation of free energy funnels in the space of encounter complexes
Yang Shen1, Ioannis Ch Paschalidis, Pirooz Vakili
1Department of Biomedical Engineering, BioMolecular Engineering Research Center, Boston University, Boston, Massachusetts, United States of America.

Abstract:
Similarly to protein folding, the association of two proteins is driven by a free energy funnel, determined by favorable interactions in some neighborhood of the native state. We describe a docking method based on stochastic global minimization of funnel-shaped energy functions in the space of rigid body motions (SE(3)) while accounting for flexibility of the interface side chains. The method, called semi-definite programming-based underestimation (SDU), employs a general quadratic function to underestimate a set of local energy minima and uses the resulting underestimator to bias further sampling. While SDU effectively minimizes functions with funnel-shaped basins, its application to docking in the rotational and translational space SE(3) is not straightforward due to the geometry of that space. We introduce a strategy that uses separate independent variables for side-chain optimization, center-to-center distance of the two proteins, and five angular descriptors of the relative orientations of the molecules. The removal of the center-to-center distance turns out to vastly improve the efficiency of the search, because the five-dimensional space now exhibits a well-behaved energy surface suitable for underestimation. This algorithm explores the free energy surface spanned by encounter complexes that correspond to local free energy minima and shows similarity to the model of macromolecular association that proceeds through a series of collisions. Results for standard protein docking benchmarks establish that in this space the free energy landscape is a funnel in a reasonably broad neighborhood of the native state and that the SDU strategy can generate docking predictions with less than 5 A ligand interface C(alpha) root-mean-square deviation while achieving an approximately 20-fold efficiency gain compared to Monte Carlo methods.
Related Concept Videos
 02:04
Protein-protein Interfaces
02:04Protein-Protein Interfaces
 02:35
Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
02:35Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...