Related Experiment Video
Updated: Jun 27, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Polynomial-time quantum algorithm for the simulation of chemical dynamics
Ivan Kassal1, Stephen P Jordan, Peter J Love
1Department of Chemistry and Chemical Biology, Harvard University, Cambridge, MA 02138, USA.

Abstract:
The computational cost of exact methods for quantum simulation using classical computers grows exponentially with system size. As a consequence, these techniques can be applied only to small systems. By contrast, we demonstrate that quantum computers could exactly simulate chemical reactions in polynomial time. Our algorithm uses the split-operator approach and explicitly simulates all electron-nuclear and interelectronic interactions in quadratic time. Surprisingly, this treatment is not only more accurate than the Born-Oppenheimer approximation but faster and more efficient as well, for all reactions with more than about four atoms. This is the case even though the entire electronic wave function is propagated on a grid with appropriately short time steps. Although the preparation and measurement of arbitrary states on a quantum computer is inefficient, here we demonstrate how to prepare states of chemical interest efficiently. We also show how to efficiently obtain chemically relevant observables, such as state-to-state transition probabilities and thermal reaction rates. Quantum computers using these techniques could outperform current classical computers with 100 qubits.

Related Concept Videos
 02:45
The Quantum-Mechanical Model of an Atom
 01:26
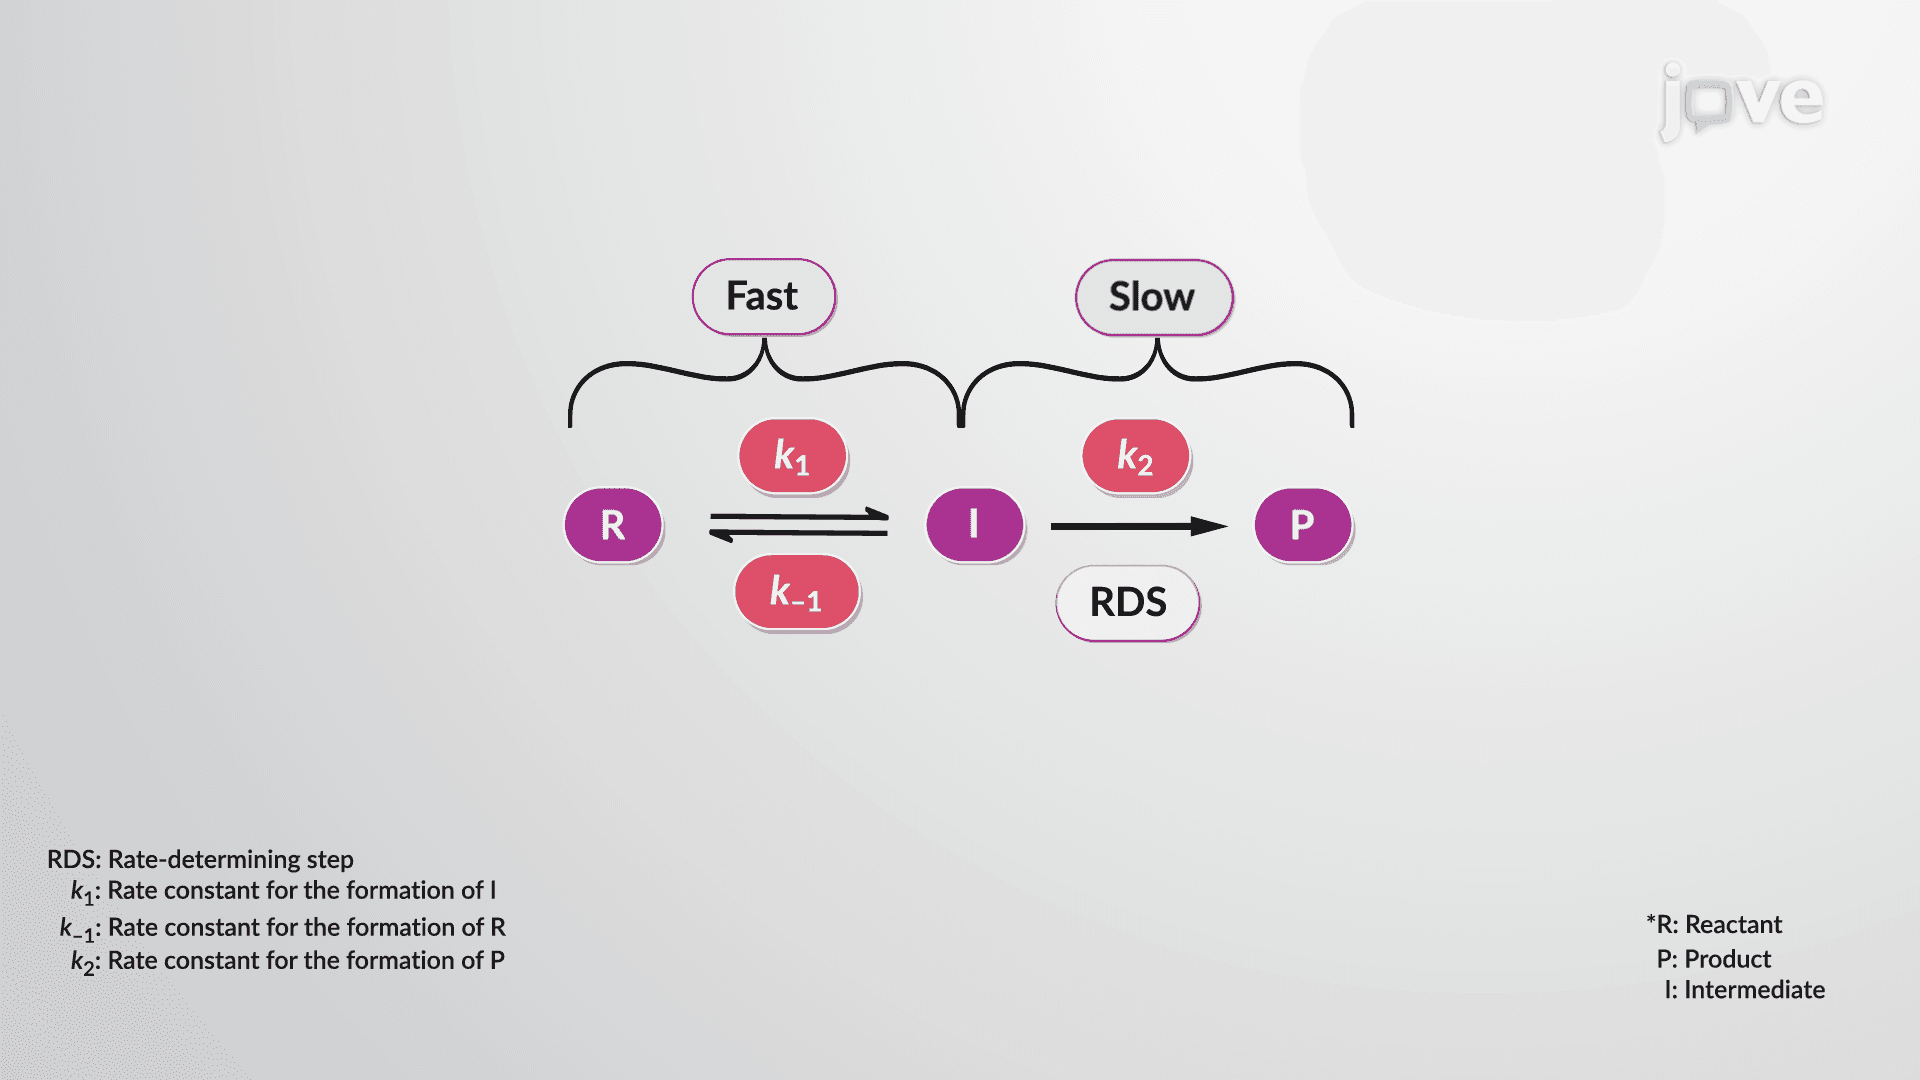
Reaction Mechanisms: The Steady-State Approximation
 01:23
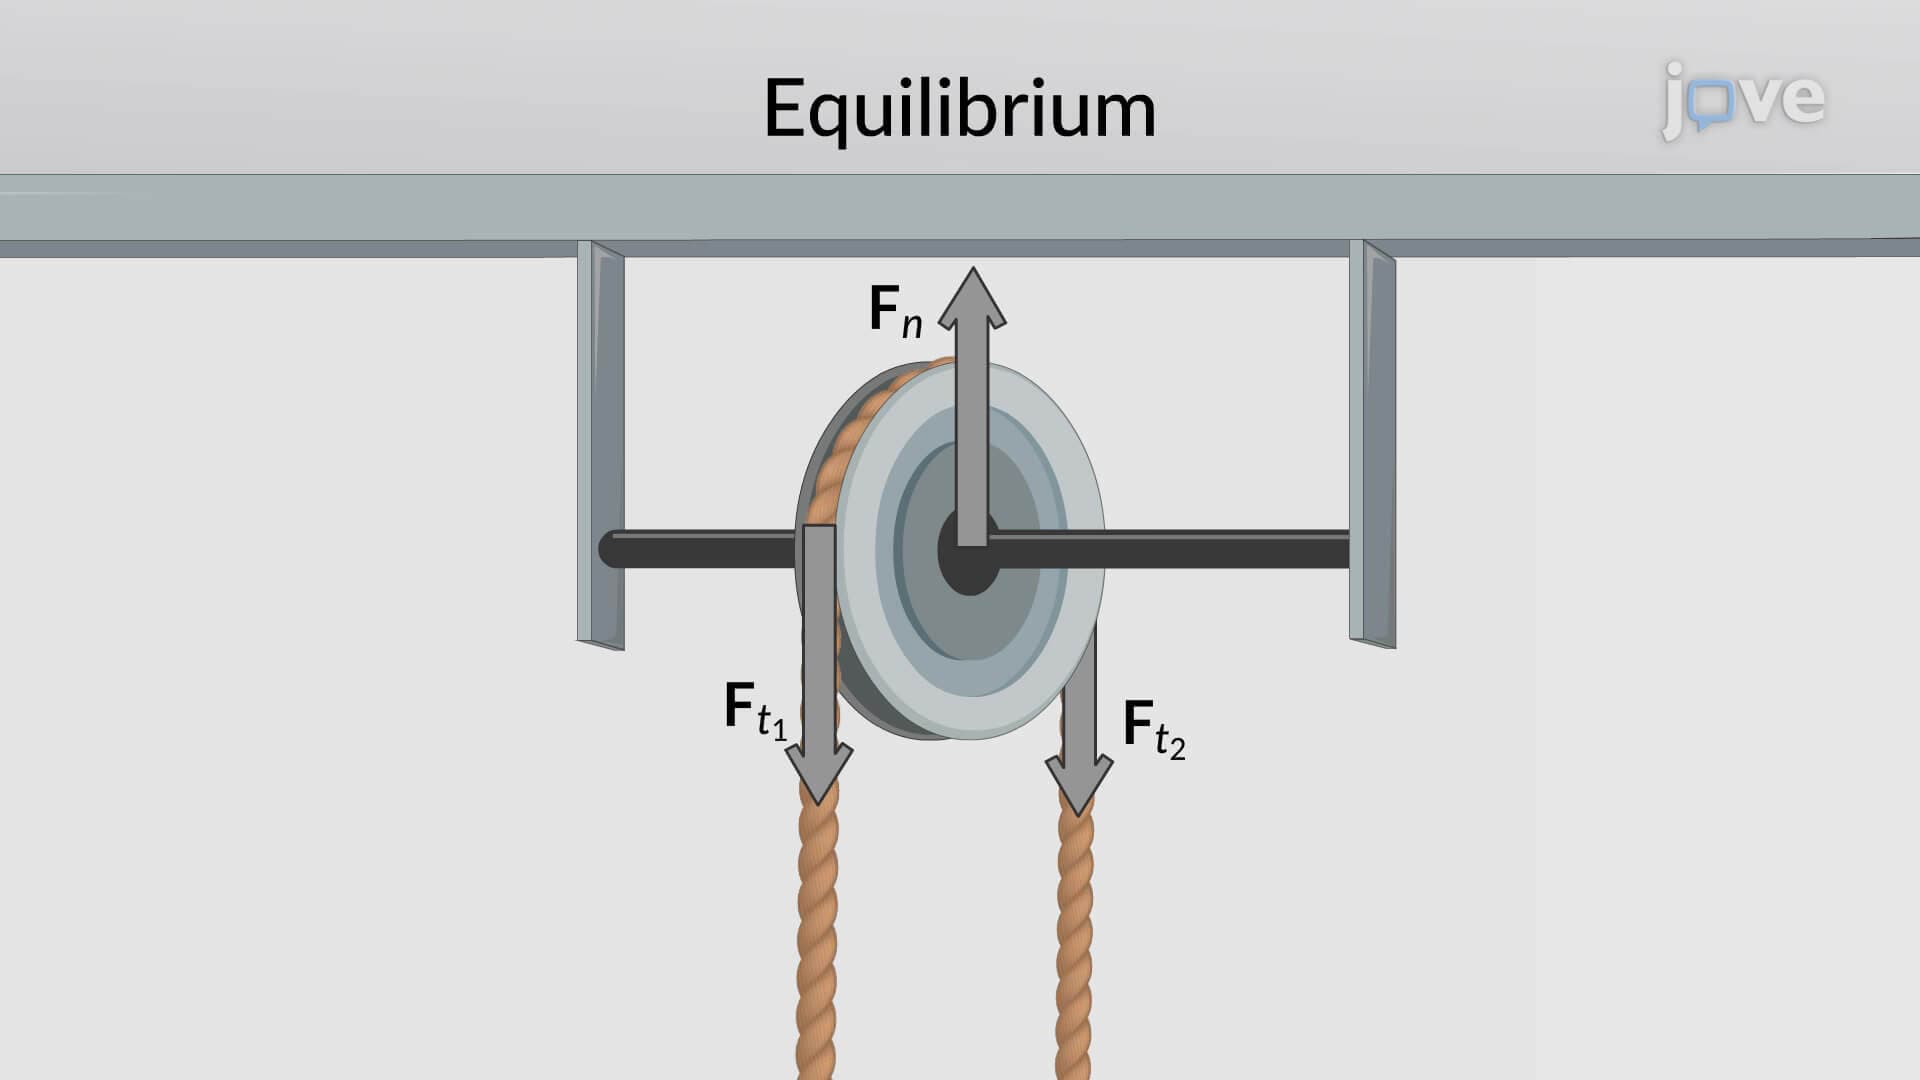
Equilibrium Conditions for a Particle
To understand the concept of equilibrium, let us first consider the forces acting on an object. When different forces act on an object, they can...
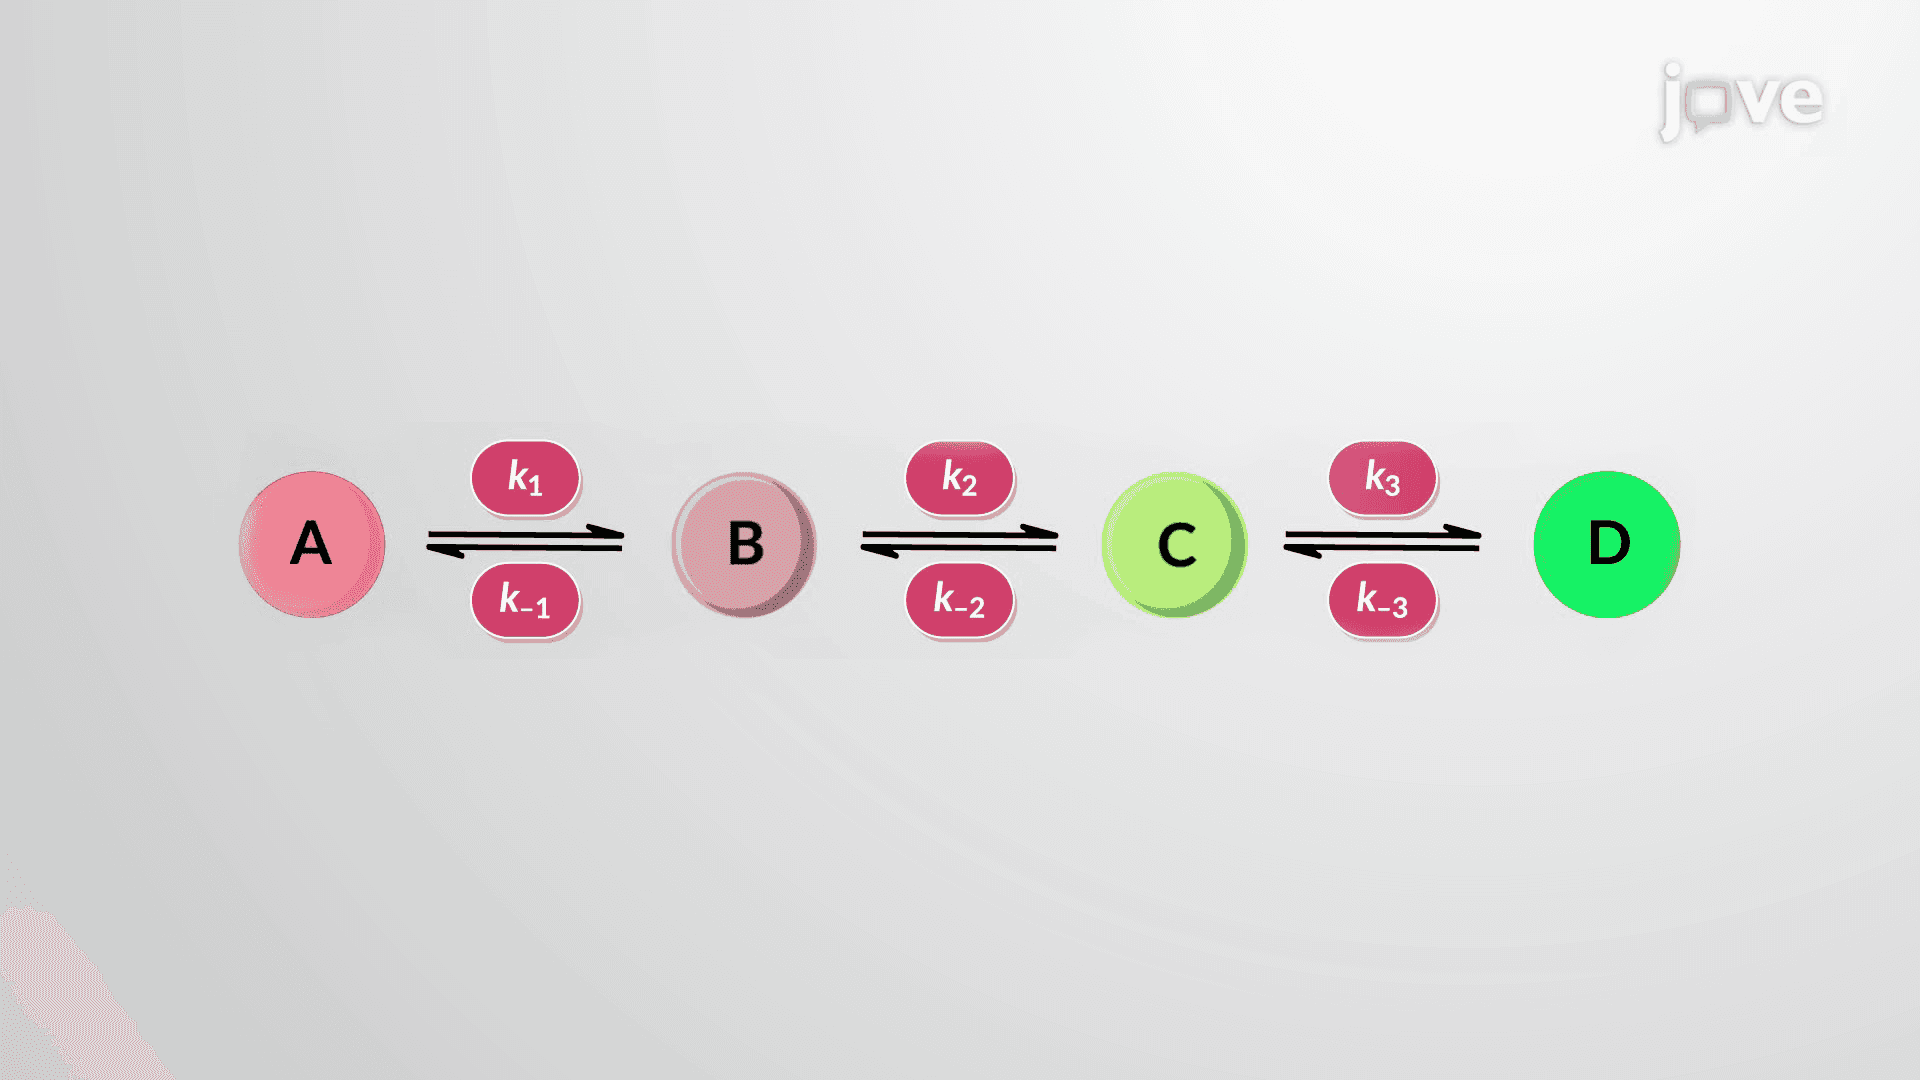 01:29
Reaction Mechanisms: Rate-limiting Step Approximation
 01:27
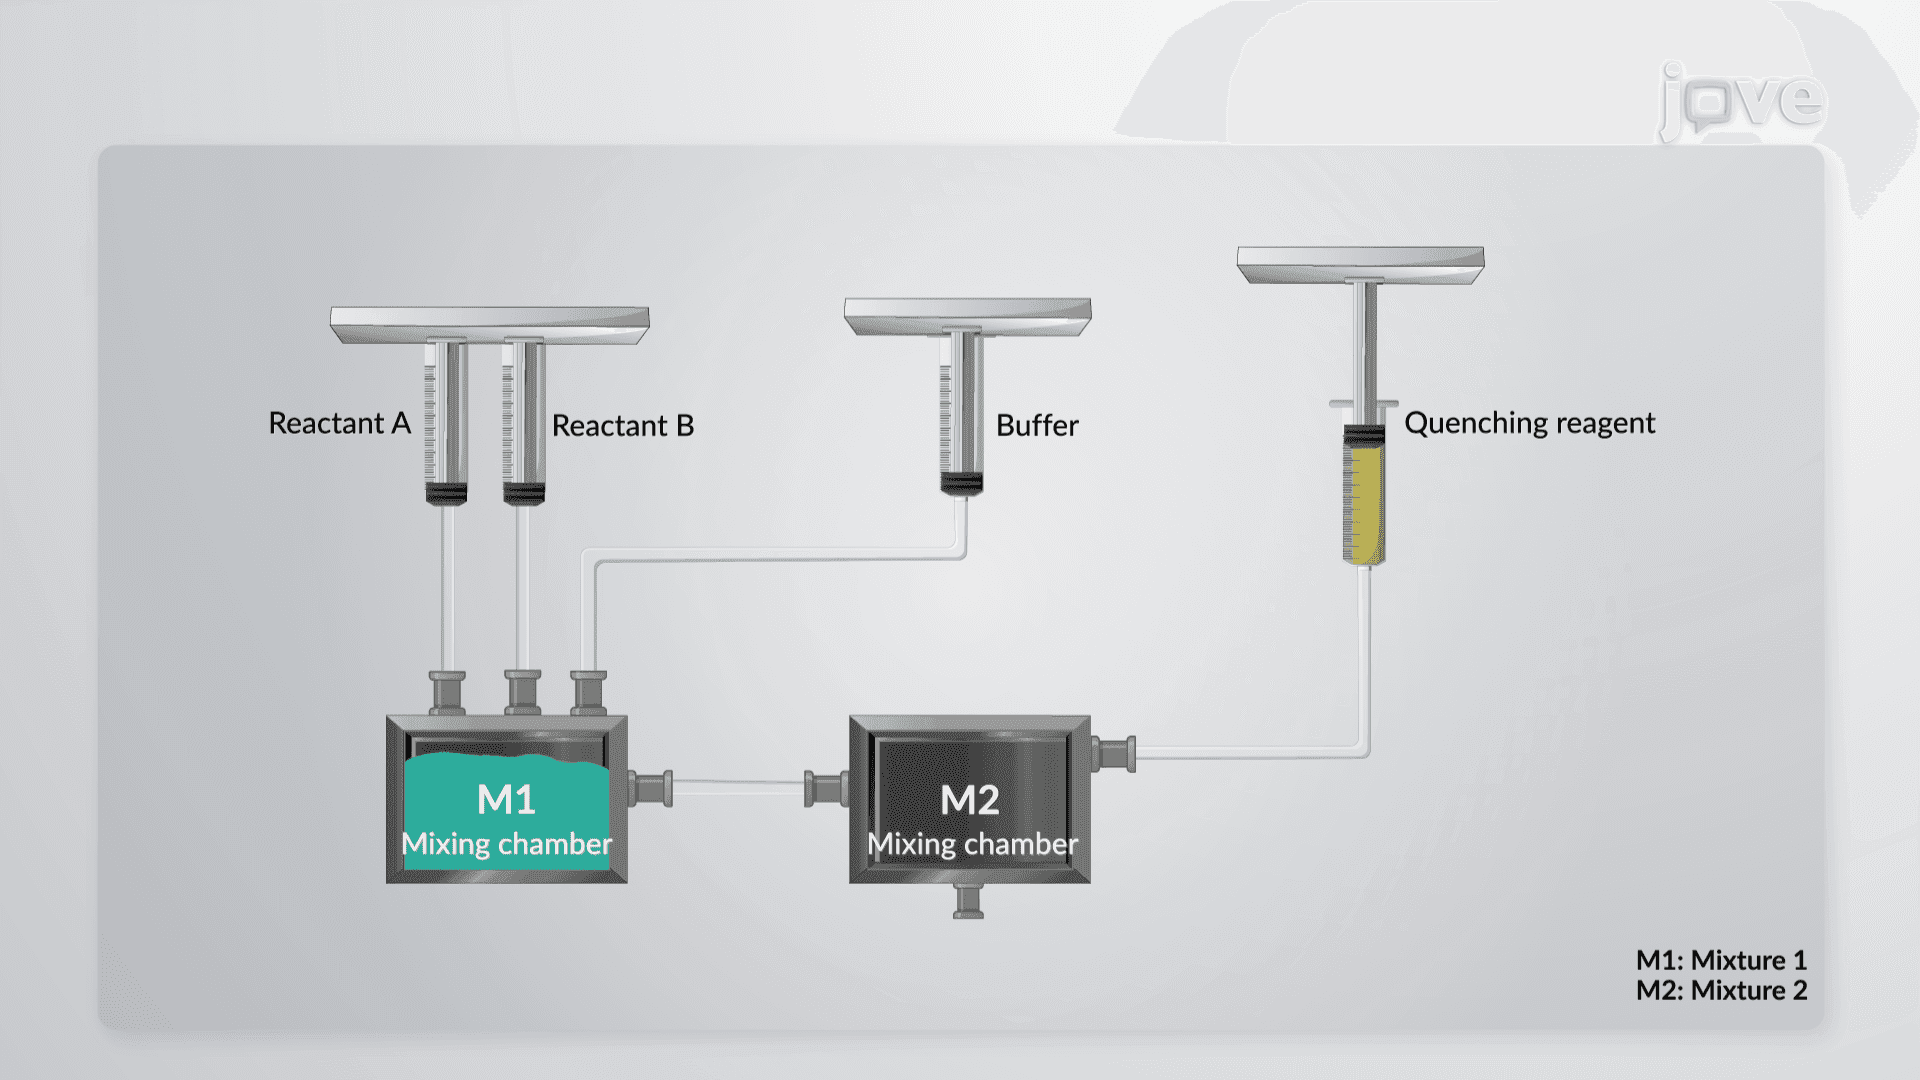
Fast Reactions
 02:32
The de Broglie Wavelength