Related Experiment Video
Updated: Jun 27, 2026

Isolating Free Carbenes, their Mixed Dimers and Organic Radicals
Published on: April 19, 2019
A benchmark theoretical study of the electron affinities of benzene and linear acenes
B Hajgató1, M S Deleuze, D J Tozer
1Research Group of Theoretical Chemistry, Department SBG, Hasselt University, Agoralaan Gebouw D, B-3590 Diepenbeek, Belgium.

Abstract:
A benchmark theoretical determination of the electron affinities of benzene and linear oligoacenes ranging from naphthalene to hexacene is presented, using the principles of a focal point analysis. These energy differences have been obtained from a series of single-point calculations at the Hartree-Fock, second-, third-, and partial fourth-order Moller-Plesset (MP2, MP3, and MP4SDQ) levels and from coupled cluster calculations including single and double excitations (CCSD) as well as perturbative estimates of connected triple excitations [CCSD(T)], using basis sets of improving quality, containing up to 1386, 1350, 1824, 1992, 1630, and 1910 basis functions in the computations, respectively. Studies of the convergence properties of these energy differences as a function of the size of the basis set and order attained in electronic correlation enable a determination of the vertical electron affinities of the four larger terms of the oligoacene (C(2+4n)H(2+2n)) series within chemical accuracy (0.04 eV). According to our best estimates, these amount to +0.28, +0.82, +1.21, and +1.47 eV when n=3, 4, 5, and 6. Adiabatic electron affinities have been further calculated by incorporating corrections for zero-point vibrational energies and for geometrical relaxations. The same procedure was applied to determine the vertical electron affinities of benzene and naphthalene, which are found to be markedly negative ( approximately -1.53 and approximately -0.48 eV, respectively). Highly quantitative insights into experiments employing electron transmission spectroscopy on these compounds were also amenable from such an approach, provided diffuse atomic functions are deliberately removed from the basis set, in order to enforce confinement in the molecular region and enable a determination of pseudoadiabatic electron affinities (with respect to the timescale of nuclear motions). Comparison was made with calculations employing density functional theory and especially designed models that exploit the integer discontinuity in the potential or incorporate a potential wall in the unrestricted Kohn-Sham orbital equation for the anion.
Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 01:37
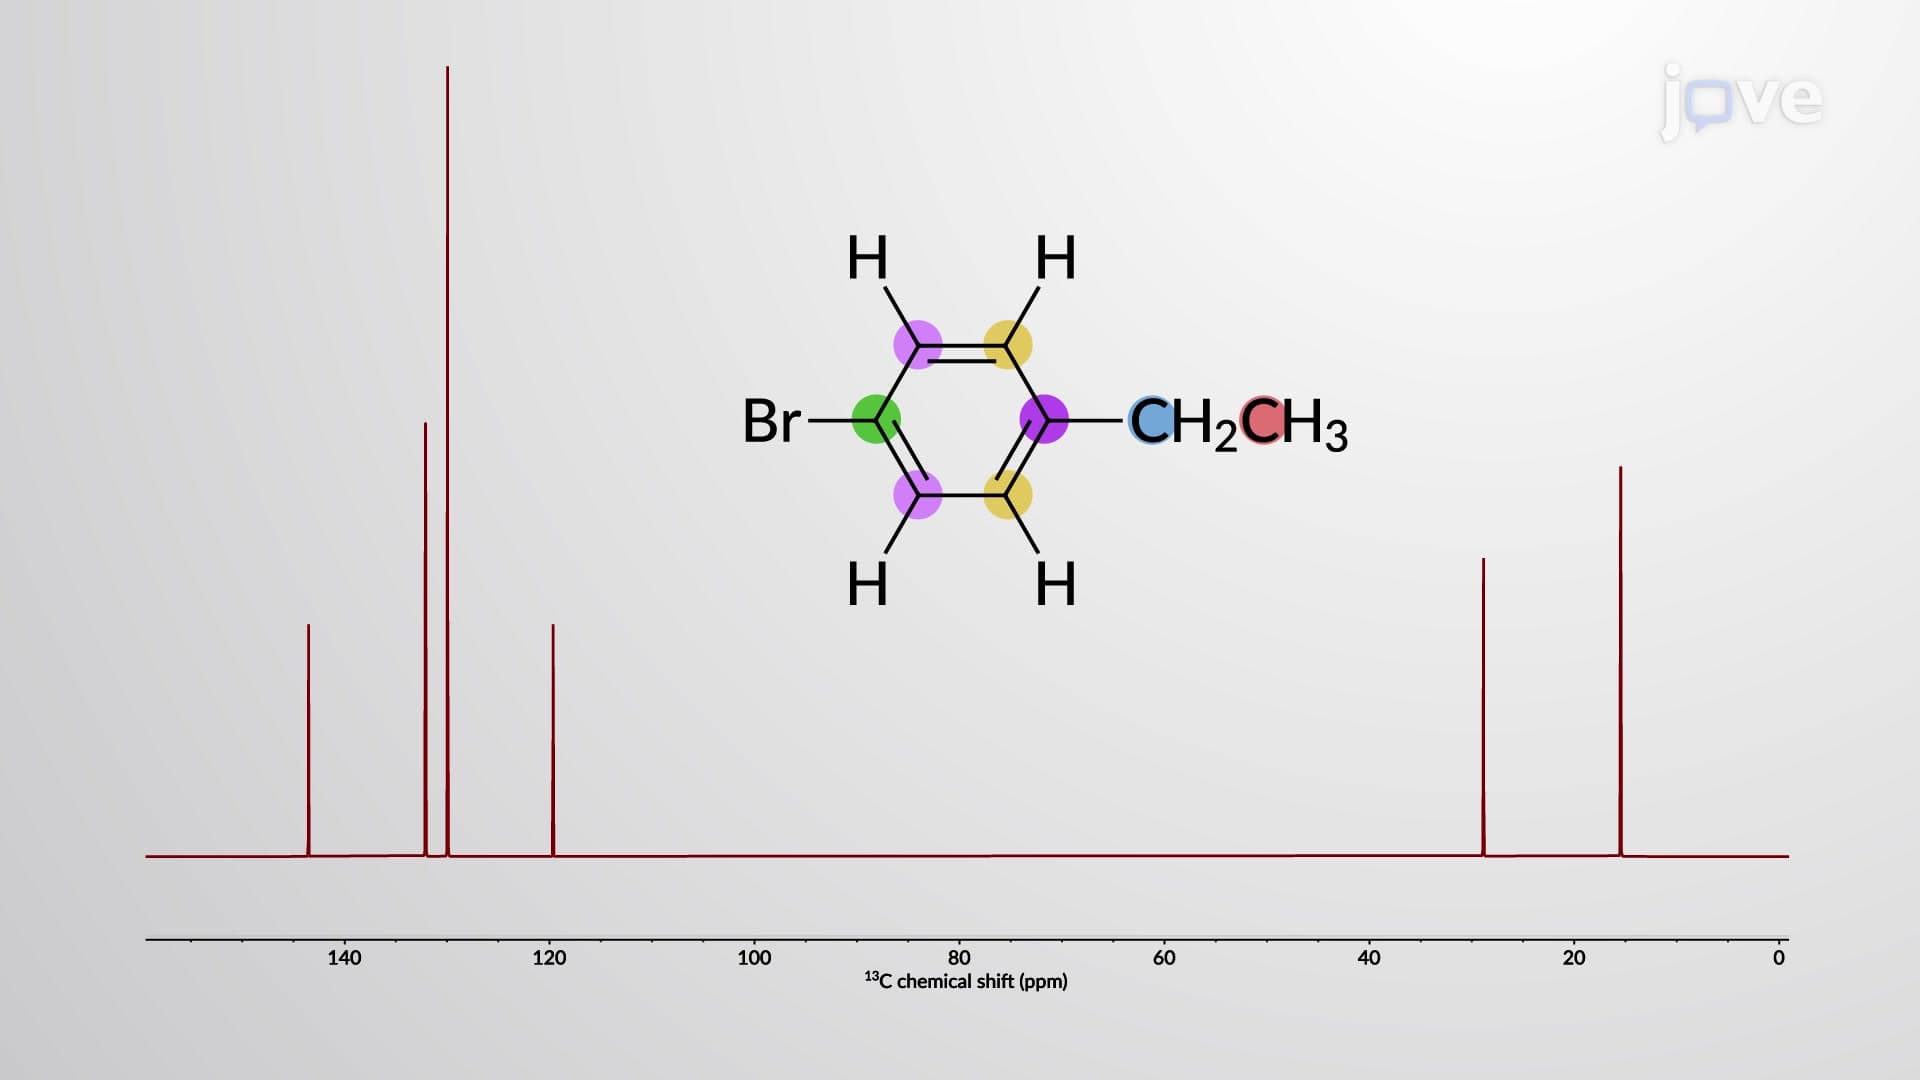
NMR Spectroscopy of Benzene Derivatives
 01:07
Structure of Benzene: Kekulé Model
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.
 02:42
Acidity of 1-Alkynes
The acidic strength of hydrocarbons follows the order: Alkynes > Alkenes > Alkanes. The strength of an acid is commonly expressed in units of pKa — the lower the pKa, the stronger the acid. Among the hydrocarbons, terminal alkynes have lower pKa values and are, therefore, more acidic. For example, the pKa values for ethane, ethene, and acetylene are 51, 44, and 25, respectively, as shown here.
 02:37
Structure and Physical Properties of Alkynes
In nature, compounds containing both carbon and hydrogen are known as "hydrocarbons". Aliphatic hydrocarbons are compounds whose molecules contain saturated single bonds (i.e., alkanes) or unsaturated double or triple bonds. Alkenes contain carbon–carbon double bonds and have a structural formula CnH2n. Unsaturated hydrocarbons containing carbon–carbon triple bonds are called "alkynes" and are structurally represented by the formula CnH2n-2.
The simplest alkyne is ethyne, or...
 01:18
Benzene to 1,4-Cyclohexadiene: Birch Reduction Mechanism