Comparison of methods for estimating the nucleotide substitution matrix
Related Experiment Video
Updated: Jun 27, 2026
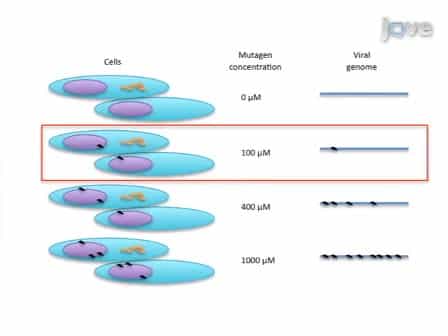
Isolation of Fidelity Variants of RNA Viruses and Characterization of Virus Mutation Frequency
Published on: June 16, 2011
Maribeth Oscamou1, Daniel McDonald, Von Bing Yap
1Department of Applied Mathematics, University of Colorado, Boulder, CO, USA. Maribeth.Oscamou@colorado.edu

Background:
The nucleotide substitution rate matrix is a key parameter of molecular evolution. Several methods for inferring this parameter have been proposed, with different mathematical bases. These methods include counting sequence differences and taking the log of the resulting probability matrices, methods based on Markov triples, and maximum likelihood methods that infer the substitution probabilities that lead to the most likely model of evolution. However, the speed and accuracy of these methods has not been compared.
Results:
Different methods differ in performance by orders of magnitude (ranging from 1 ms to 10 s per matrix), but differences in accuracy of rate matrix reconstruction appear to be relatively small. Encouragingly, relatively simple and fast methods can provide results at least as accurate as far more complex and computationally intensive methods, especially when the sequences to be compared are relatively short.
Conclusion:
Based on the conditions tested, we recommend the use of method of Gojobori et al. (1982) for long sequences (> 600 nucleotides), and the method of Goldman et al. (1996) for shorter sequences (< 600 nucleotides). The method of Barry and Hartigan (1987) can provide somewhat more accuracy, measured as the Euclidean distance between the true and inferred matrices, on long sequences (> 2000 nucleotides) at the expense of substantially longer computation time. The availability of methods that are both fast and accurate will allow us to gain a global picture of change in the nucleotide substitution rate matrix on a genomewide scale across the tree of life.
Related Concept Videos
 02:26
Comparing Copy Number Variations and SNPs
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
 02:54
Evolutionary Relationships through Genome Comparisons
 02:52
Predicting Products: Substitution vs. Elimination
The following factors can influence the mechanisms competing against each other:
 02:16
Comparing Mitochondrial, Chloroplast, and Prokaryotic Genomes