Related Experiment Video
Updated: Jun 26, 2026
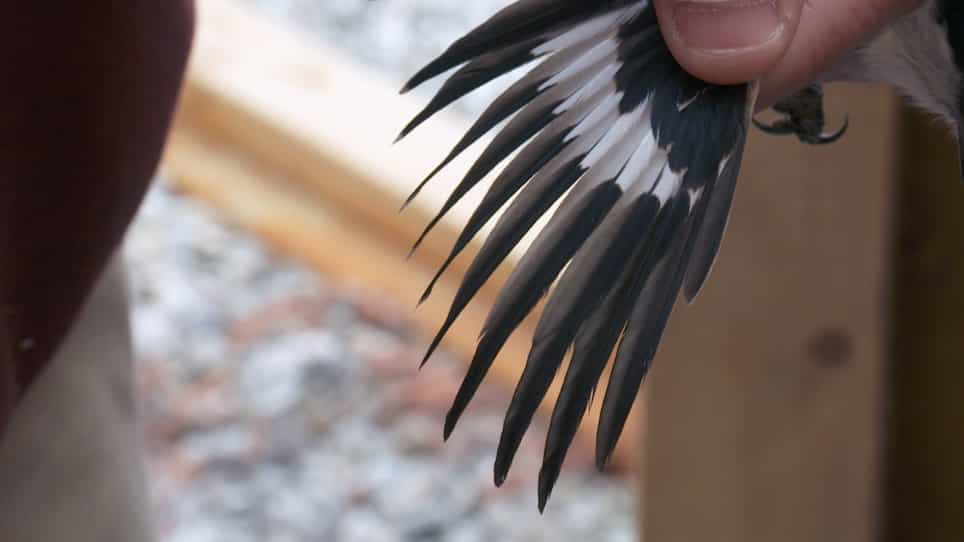

Visually Sexing Loggerhead Shrike (Lanius Ludovicianus) Using Plumage Coloration and Pattern
Published on: March 8, 2020
Patchy distributions belie morphological and genetic homogeneity in rosy-finches
Sergei V Drovetski1, Robert M Zink, Nicolle A Mode
1CIBIO, Centro de Investigação em Biodiversidade e Recursos Genéticos, Campus Agrário de Vairão, Rua Padre Armando Quintas, Crasto 4485-661 Vairão, Portugal. svd@mail.icav.up.pt

Abstract:
DNA sequence data often appear to contradict low-level avian taxonomy, which is usually based on patterns of external phenotypic similarity. We examined such an apparent contradiction in the Nearctic rosy-finches. On the basis of several phenotypic characters the finches were divided into three species congeneric with three Asian species. When Nearctic taxa were analyzed in a principal components analysis, 66.9% of phenotypic variation was explained by differences between the Bering Sea and continental populations, sexual dimorphism and a latitudinal cline. Our phylogenetic analysis of mitochondrial ND2 sequences revealed four clades among six species of rosy-finches. Three clades corresponded to three Asian species. The fourth clade included all three Nearctic species. Their haplotypes were not reciprocally monophyletic and the combined genetic variability of all Nearctic taxa was lower than in two of their Asian congeners. A Z-specific intron (ACO1I9) and an autosomal coding locus (MC1R) provided little additional phylogenetic information, most likely because of the longer coalescence times relative to ND2. Phylogeographic analyses of ND2 data revealed significant gene flow among neighboring localities regardless of their taxonomic assignment. Our analyses showed that DNA and phenotypic data are not in conflict, but rather complement each other, and together help clarify species limits. Our data are consistent with a single species in North America, not three.
Related Concept Videos
 01:21
Testing a Claim about Mean: Unknown Population SD
Estimating a population mean requires the samples to be approximately normally distributed. The data should be collected from the randomly selected samples having no sampling bias. There is no specific requirement for sample size. But if the sample size is less than 30, and we don't know the population standard deviation, a different approach is used; instead...
 01:49
Hardy-Weinberg Principle
 02:55
The Evidence for Evolution
 02:39
Gene Flow
 01:25
What is Population Genetics?
 02:16
Genetics of Speciation