Related Experiment Video
Updated: Jun 26, 2026

Facile Preparation of Ultrafine Aluminum Hydroxide Particles with or without Mesoporous MCM-41 in Ambient Environments
Published on: May 11, 2017
Uranyl interaction with the hydrated (001) basal face of gibbsite: a combined theoretical and spectroscopic study
Edouard Veilly1, Jérôme Roques, Marie-Camille Jodin-Caumon
1Institut de Physique Nucleaire d'Orsay, Universite Paris-Sud 11, CNRS UMR 8608, 15 Rue Georges Clemenceau, Bâtiment 100, 91406 Orsay Cedex, France.

Abstract:
The sorption of uranyl cations and water molecules on the basal (001) face of gibbsite was studied by combining vibrational and fluorescence spectroscopies together with density functional theory (DFT) computations. Both the calculated and experimental values of O-H bond lengths for the gibbsite bulk are in good agreement. In the second part, water sorption with this surface was studied to take into account the influence of hydration with respect to the uranyl adsorption. The computed water configurations agreed with previously published molecular dynamics studies. The uranyl adsorption in acidic media was followed by time-resolved laser-induced fluorescence spectroscopy and Raman spectrometry measurements. The existence of only one kind of adsorption site for the uranyl cation was then indicated in good agreement with the DFT calculations. The computation of the uranyl adsorption has been performed by means of a bidentate interaction with two surface oxygen atoms. The optimized structures displayed strong hydrogen bonds between the surface and the -yl oxygen of uranyl. The uranium-surface bond strength depends on the protonation state of the surface oxygen atoms. The calculated U-O(surface) bond lengths range between 2.1-2.2 and 2.6-2.7 A for the nonprotonated and protonated surface O atoms, respectively.
More Related Videos
 12:05
U2O5 Film Preparation via UO2 Deposition by Direct Current Sputtering and Successive Oxidation and Reduction with Atomic Oxygen and Atomic Hydrogen
Published on: February 21, 2019
 11:38
In situ FTIR Spectroscopy as a Tool for Investigation of Gas/Solid Interaction: Water-Enhanced CO2 Adsorption in UiO-66 Metal-Organic Framework
Published on: February 1, 2020
Related Concept Videos
 03:24
Hybridization of Atomic Orbitals I
 02:42
Valence Bond Theory
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:29
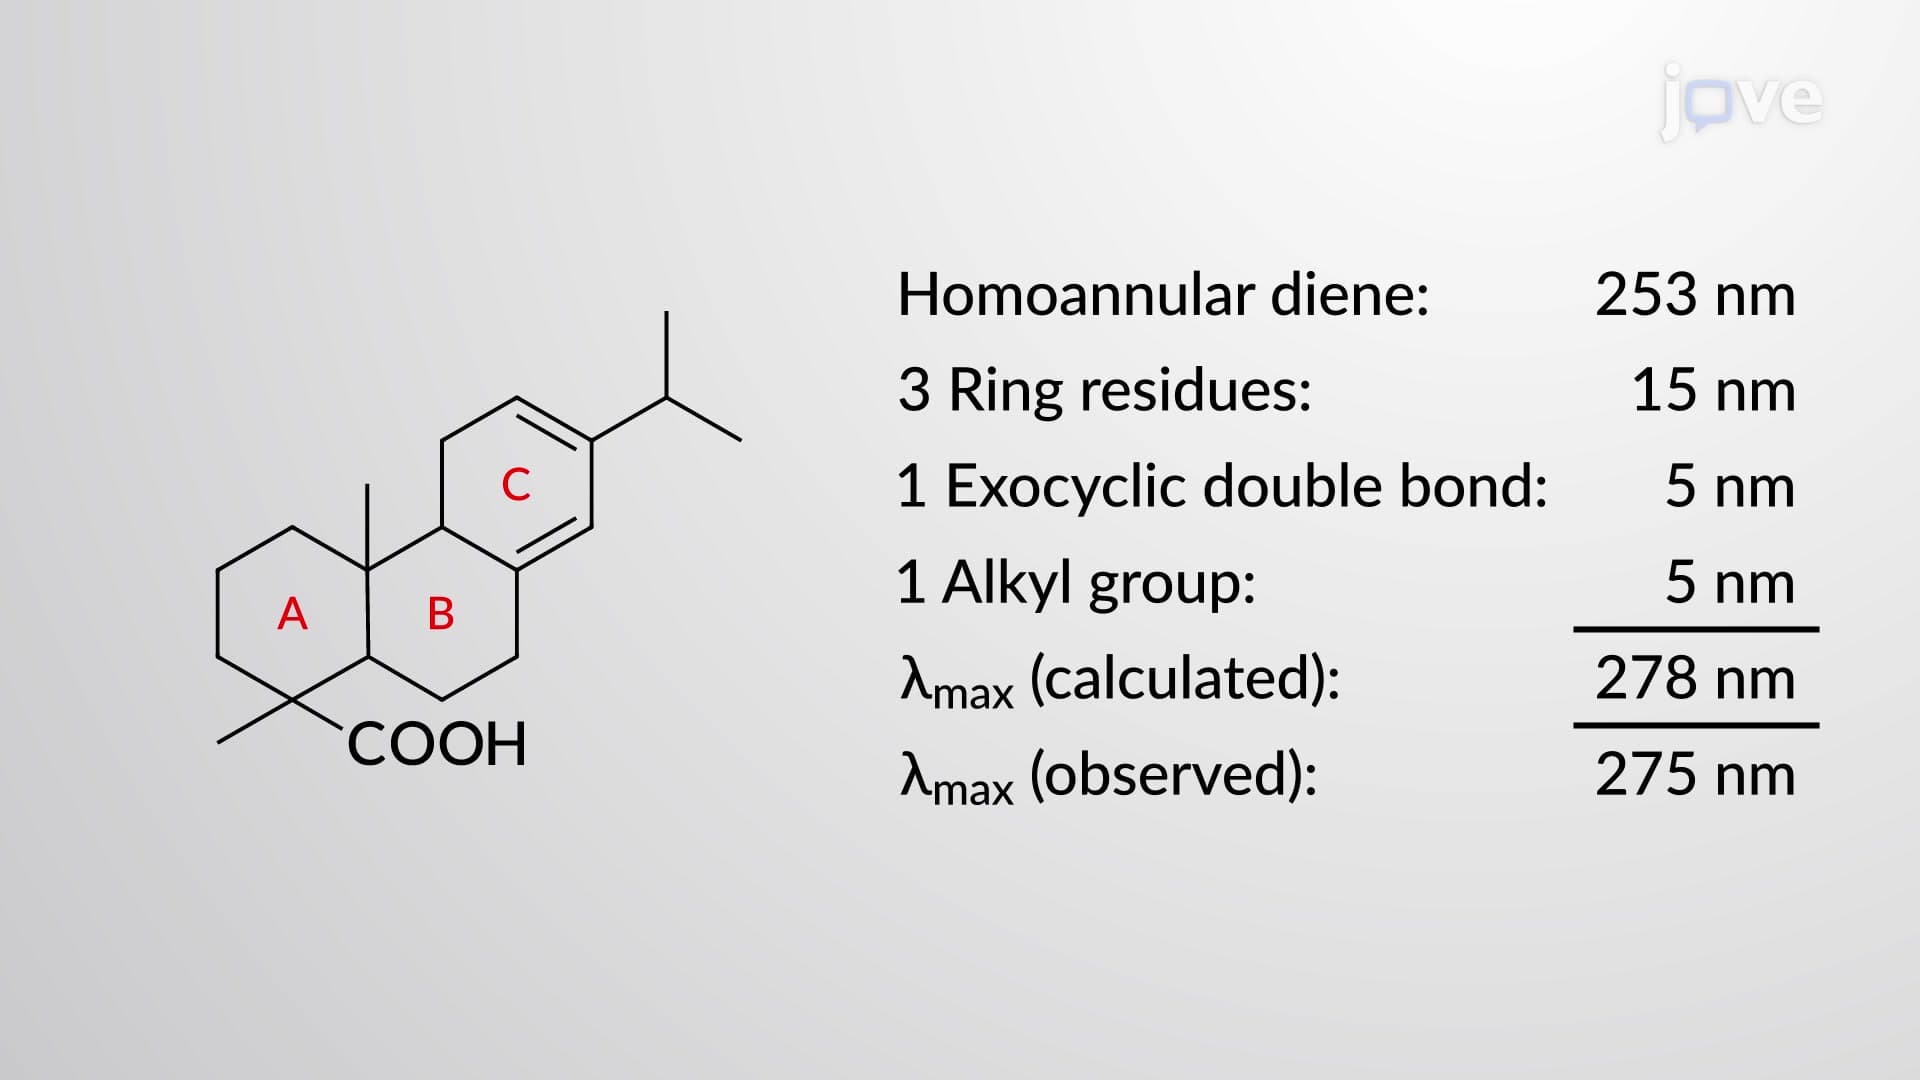
UV–Vis Spectroscopy: Woodward–Fieser Rules
 03:35
Hybridization of Atomic Orbitals II
 01:23
IR Spectrum Peak Broadening: Hydrogen Bonding
However, the extent of hydrogen bonding influences the observed stretching frequency and band broadening. Intermolecular or intramolecular hydrogen bonding...