Related Experiment Video
Updated: Jun 26, 2026

A Rapid and Facile Pipeline for Generating Genomic Point Mutants in C. elegans Using CRISPR/Cas9 Ribonucleoproteins
Published on: April 30, 2018
An ALS-linked mutant SOD1 produces a locomotor defect associated with aggregation and synaptic dysfunction when
Jiou Wang1, George W Farr, David H Hall
1Howard Hughes Medical Institute, Yale School of Medicine, New Haven, Connecticut, United States of America.

Abstract:
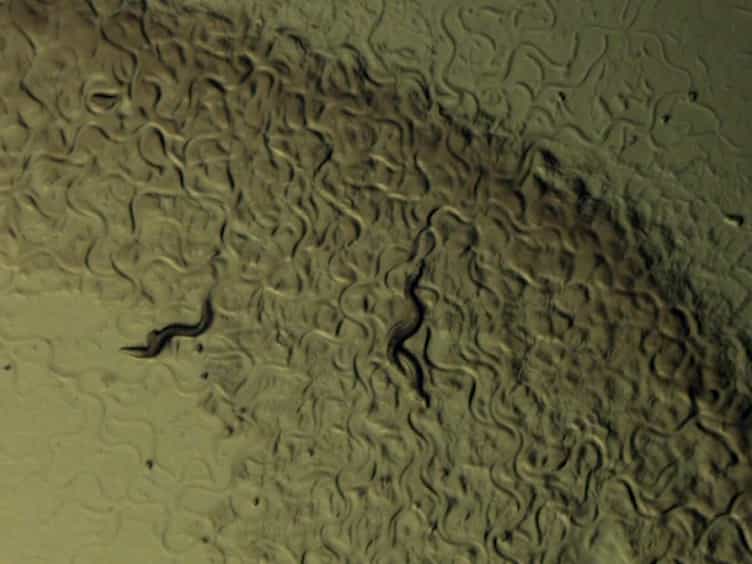
The nature of toxic effects exerted on neurons by misfolded proteins, occurring in a number of neurodegenerative diseases, is poorly understood. One approach to this problem is to measure effects when such proteins are expressed in heterologous neurons. We report on effects of an ALS-associated, misfolding-prone mutant human SOD1, G85R, when expressed in the neurons of Caenorhabditis elegans. Stable mutant transgenic animals, but not wild-type human SOD1 transgenics, exhibited a strong locomotor defect associated with the presence, specifically in mutant animals, of both soluble oligomers and insoluble aggregates of G85R protein. A whole-genome RNAi screen identified chaperones and other components whose deficiency increased aggregation and further diminished locomotion. The nature of the locomotor defect was investigated. Mutant animals were resistant to paralysis by the cholinesterase inhibitor aldicarb, while exhibiting normal sensitivity to the cholinergic agonist levamisole and normal muscle morphology. When fluorescently labeled presynaptic components were examined in the dorsal nerve cord, decreased numbers of puncta corresponding to neuromuscular junctions were observed in mutant animals and brightness was also diminished. At the EM level, mutant animals exhibited a reduced number of synaptic vesicles. Neurotoxicity in this system thus appears to be mediated by misfolded SOD1 and is exerted on synaptic vesicle biogenesis and/or trafficking.