Theoretical study of the effect of structural modifications on the hyperpolarizabilities of indigo derivatives

Prasanta K Nandi1, Nabamita Panja, Tapan K Ghanty

  • 1Department of Chemistry, Bengal Engineering and Science University, Shibpur, Howrah 711 103, India. nandi_pk@yahoo.co.in

Related Concept Videos

Inductive Effects on Chemical Shift: Overview01:27

Inductive Effects on Chemical Shift: Overview

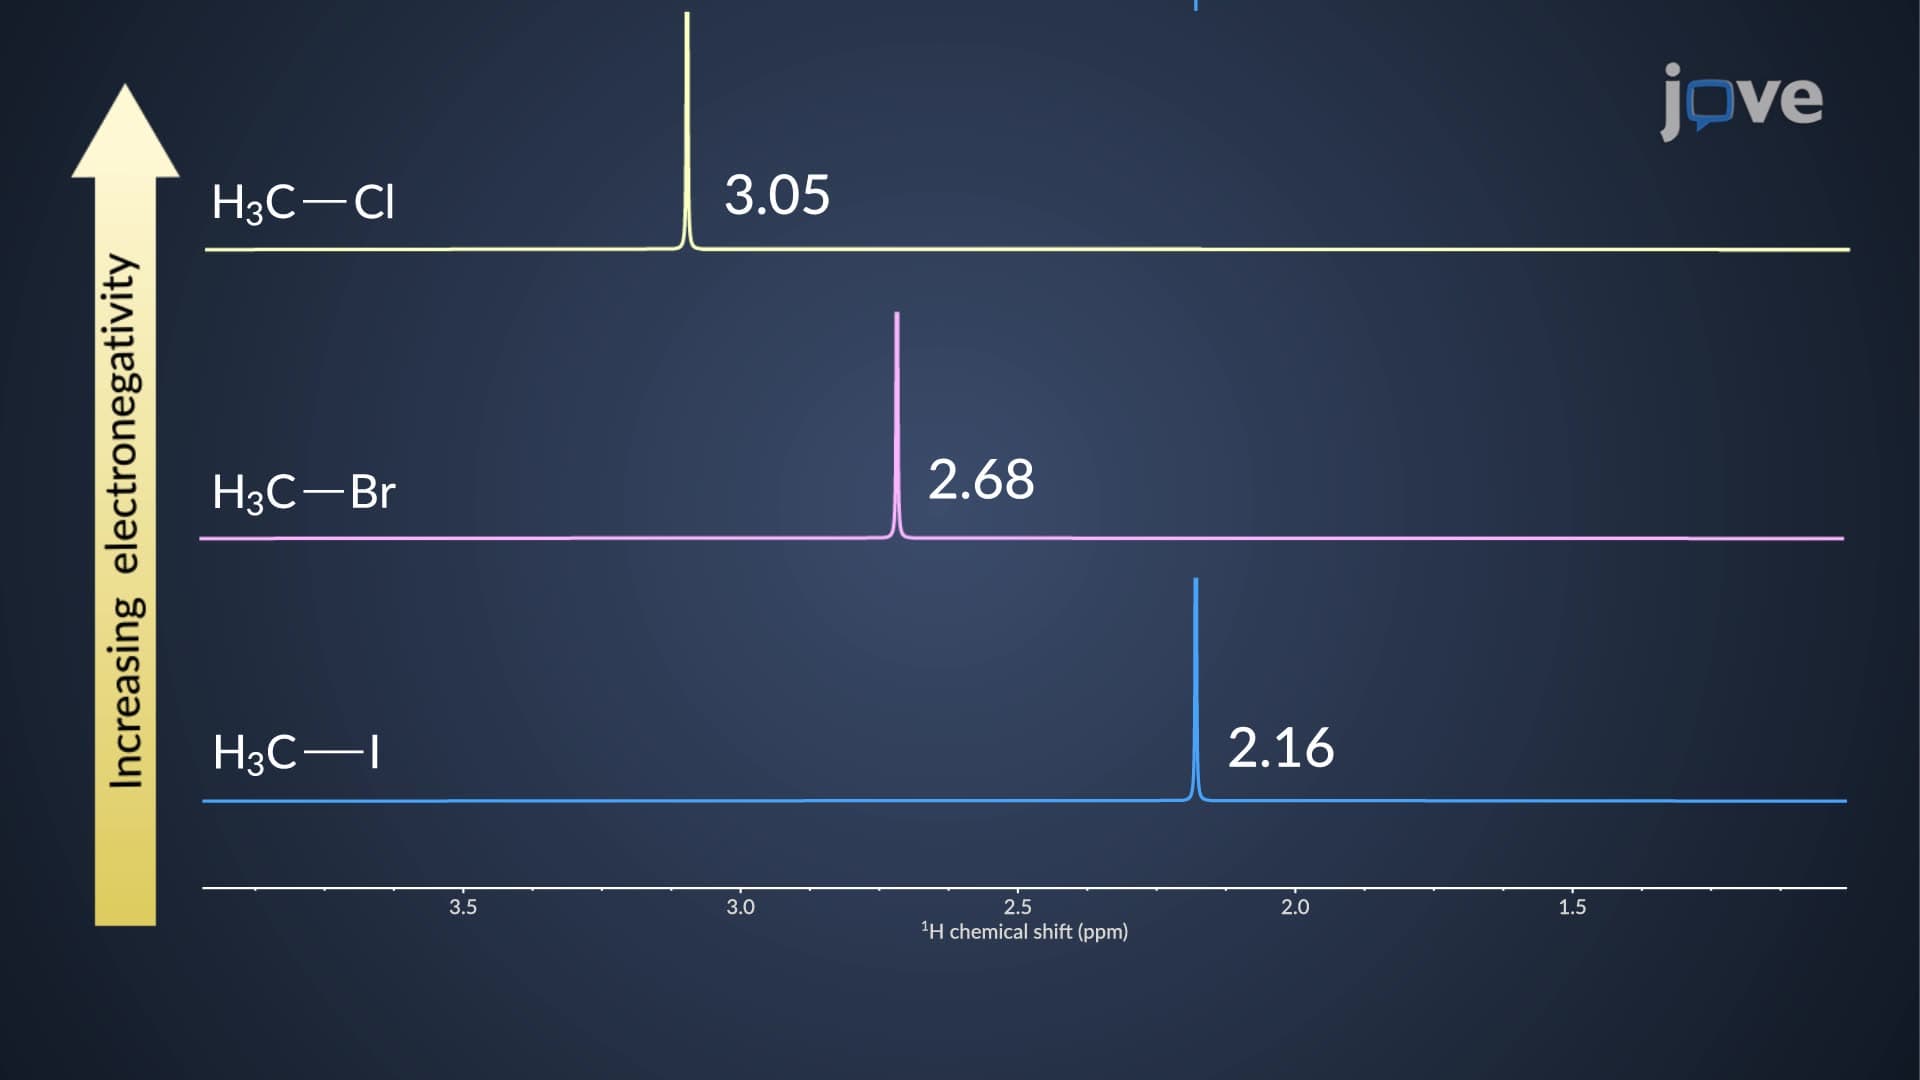
The protons in unsubstituted alkanes are strongly shielded with chemical shifts below 1.8 ppm. Methine, methylene, and methyl protons appear at approximately 1.7, 1.2 and 0.7 ppm, while the proton signal from methane appears at 0.23 ppm. An electronegative substituent, such as chlorine, withdraws the electron density from the protons, increasing their chemical shift. Progressive substitution of the hydrogens in methane by chlorine shifts the proton signals increasingly downfield, to 3.05 ppm in...
π Electron Effects on Chemical Shift: Overview01:27

π Electron Effects on Chemical Shift: Overview
An applied magnetic field causes loosely bound π-electrons in organic molecules to circulate, producing a local or induced diamagnetic field over a large spatial volume. As the molecules tumble in solution, the field generated by π-electrons in spherical substituents results in a zero net field. However, the net field generated by π-electrons in non-spherical substituents is not zero. The effect of this induced field depends on the orientation of the molecule with respect to B0, resulting in...
UV–Vis Spectroscopy: Woodward–Fieser Rules01:29

UV–Vis Spectroscopy: Woodward–Fieser Rules

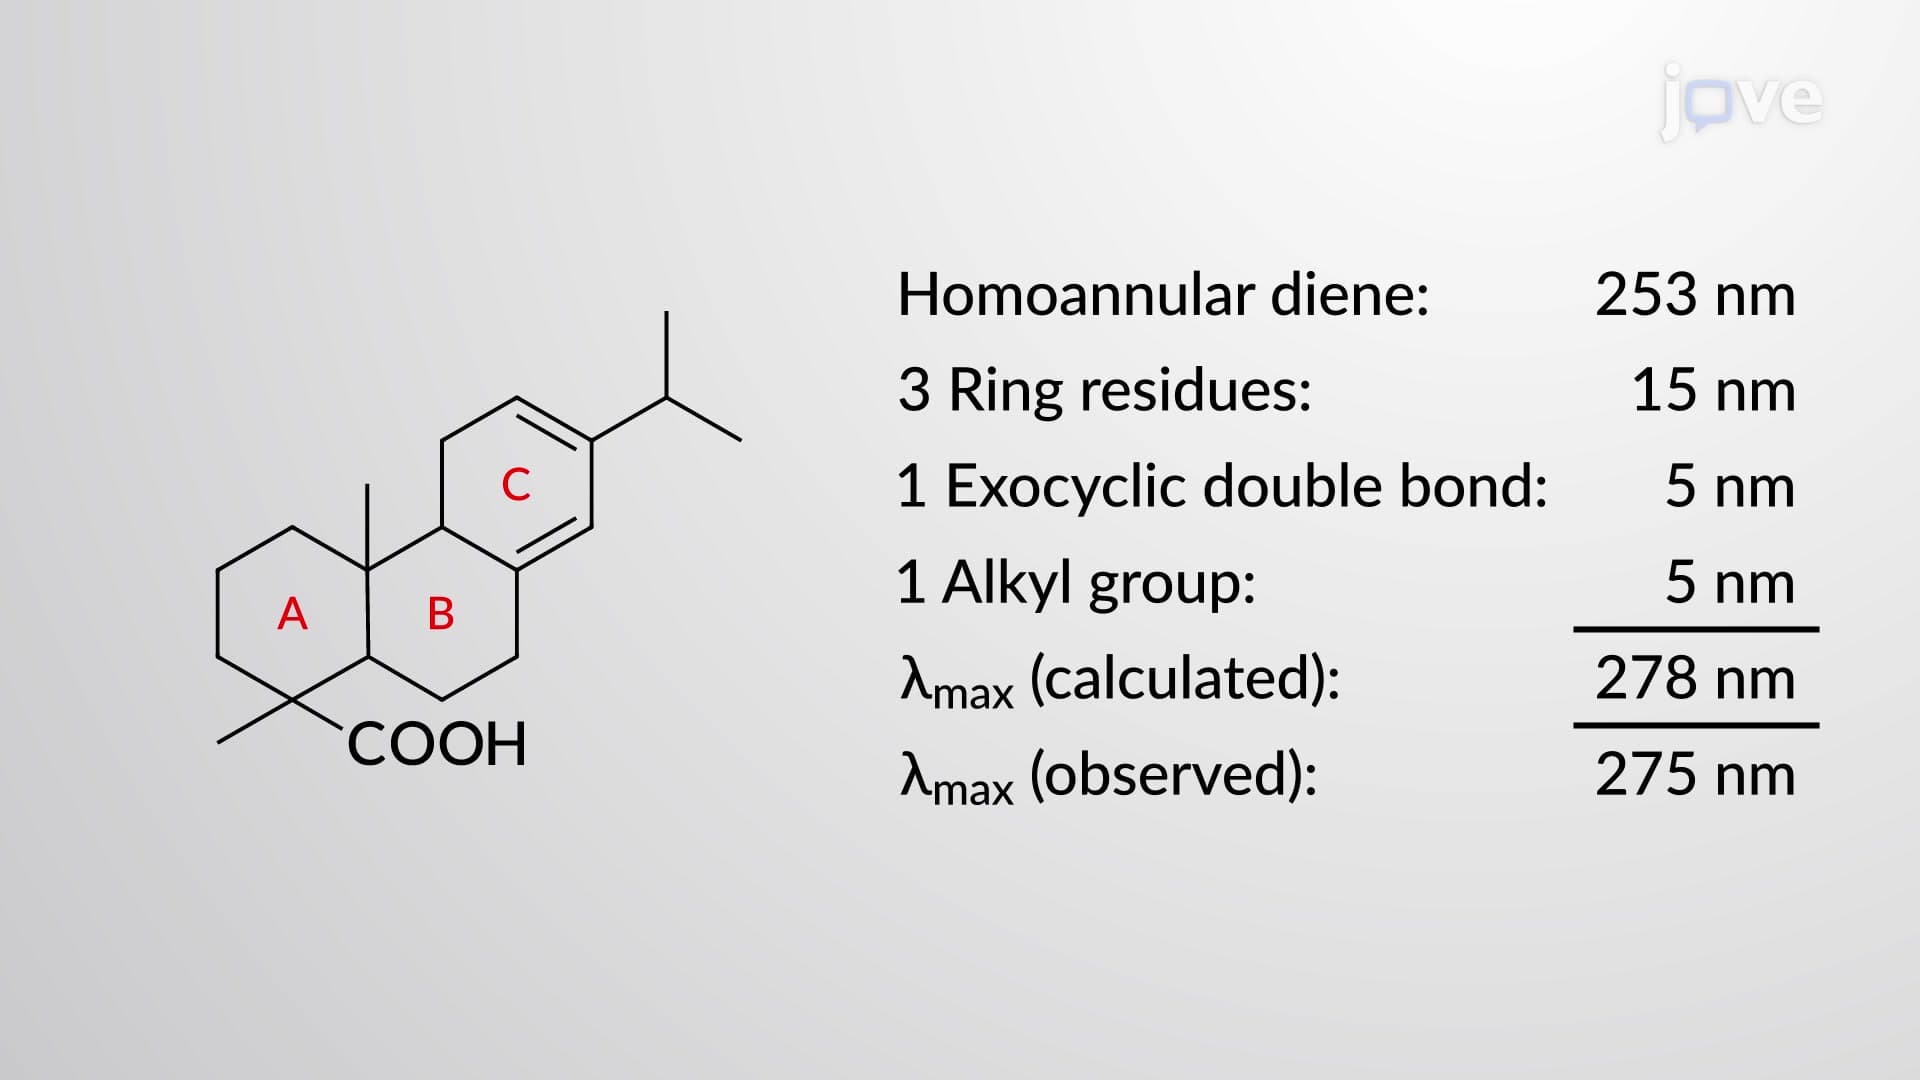
UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given structure by adding the contributions...
IR Absorption Frequency: Delocalization01:04

IR Absorption Frequency: Delocalization

Electron delocalization refers to the distribution of electrons across multiple atoms within a molecule rather than being confined to a single atom or bond. This phenomenon is common in systems with conjugated bonds—structures where alternating single and double bonds allow π-electrons to move freely across the network. The movement of electrons stabilizes the molecule and can affect various chemical properties, including vibrational frequencies observed in IR spectroscopy.
In IR spectroscopy,...
Basicity of Heterocyclic Aromatic Amines01:25

Basicity of Heterocyclic Aromatic Amines
Heterocyclic amines, where the N atom is a part of an alicyclic system, are similar in basicity to alkylamines. Interestingly, the heterocyclic amine having a nitrogen atom as part of an aromatic ring has much less basicity than its corresponding alicyclic counterpart. For this reason, as presented in Figure 1, piperidine (pKb = 2.8) is significantly more basic than pyridine (pKb = 8.8).
Basicity of Aromatic Amines01:18

Basicity of Aromatic Amines

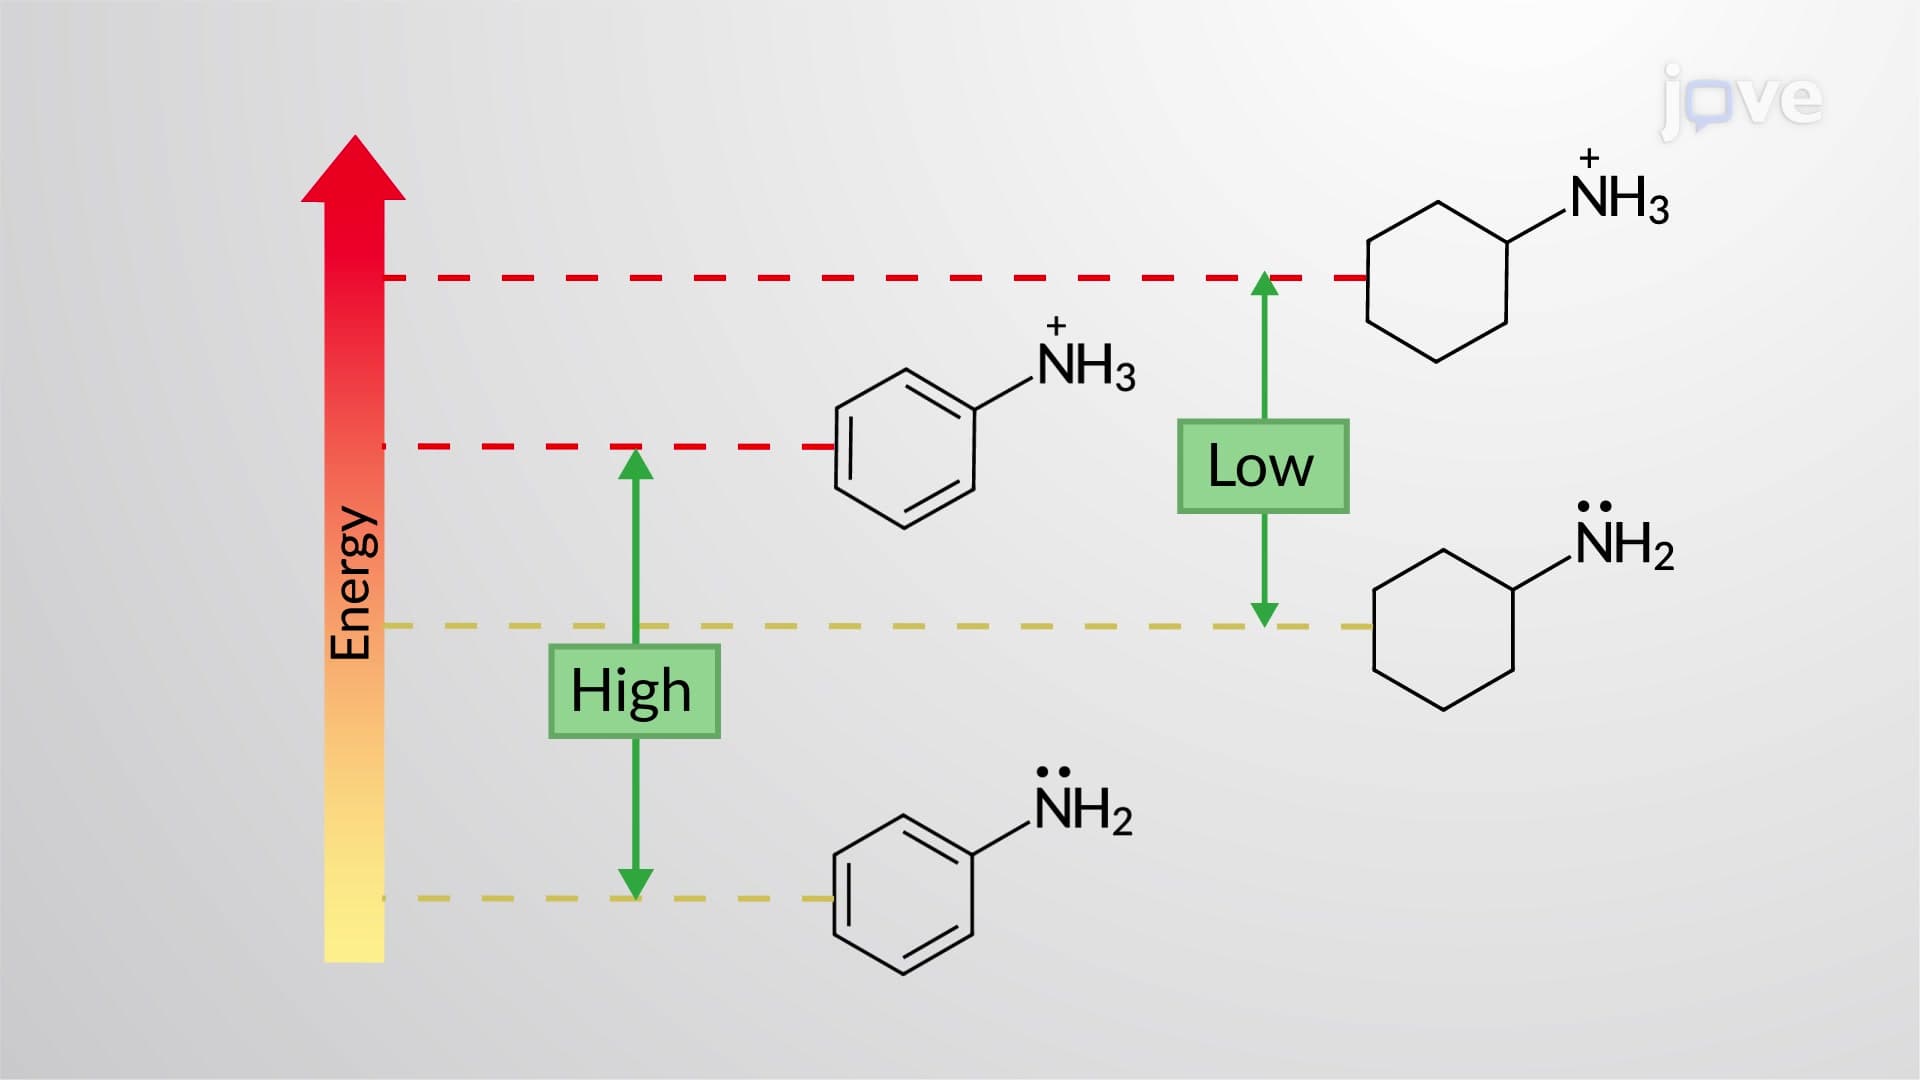
The basicity of aromatic amines is much weaker than that of aliphatic amines due to the involvement of the lone pair of electrons over the N atom in resonance with the aryl rings. Generally, the electron-donating ability of any substituents on the aryl ring of aromatic amines increases the basicity of the amine by increasing electron density, and hence the availability of lone pair on the nitrogen. On the other hand, electron-withdrawing functional groups on the aryl ring of amines decrease the...