Related Experiment Video
Updated: Jun 25, 2026

Tuning Oxide Properties by Oxygen Vacancy Control During Growth and Annealing
Published on: June 9, 2023
Density-functional calculations of the structure of near-surface oxygen vacancies and electron localization on
M Verónica Ganduglia-Pirovano1, Juarez L F Da Silva, Joachim Sauer
1Humboldt-Universität zu Berlin, Unter den Linden 6, 10099 Berlin, Germany.

Abstract:
One of the most topical issues surrounding oxygen vacancies on CeO2(111) is the relative stability of surface and subsurface defects. Using density-functional theory (DFT) with the HSE06 (Heyd-Scuseria-Ernzerhof) hybrid functional as well as the DFT+U approach (where U is a Hubbard-like term describing the on-site Coulomb interactions), we find subsurface vacancies with (2x2) periodicity to be energetically more favorable by 0.45 (HSE06), 0.47 [PBE+U (Perdew-Burke-Ernzerhof functional)], and 0.22 eV [LDA+U (local density approximation)]. The excess electrons localize not on Ce ions which are the nearest neighbor to the defect as priorly suggested, but instead on those that are next-nearest neighbors. The excess-electron distribution and the preference for subsurface vacancies are explained in terms of defect-induced lattice relaxation effects.
More Related Videos
Related Concept Videos
 01:26
Imperfections in Crystal Structure: Stoichiometric Point Defects
 01:29
Imperfections in Crystal Structure: Non-Stoichiometric Defects
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 03:26
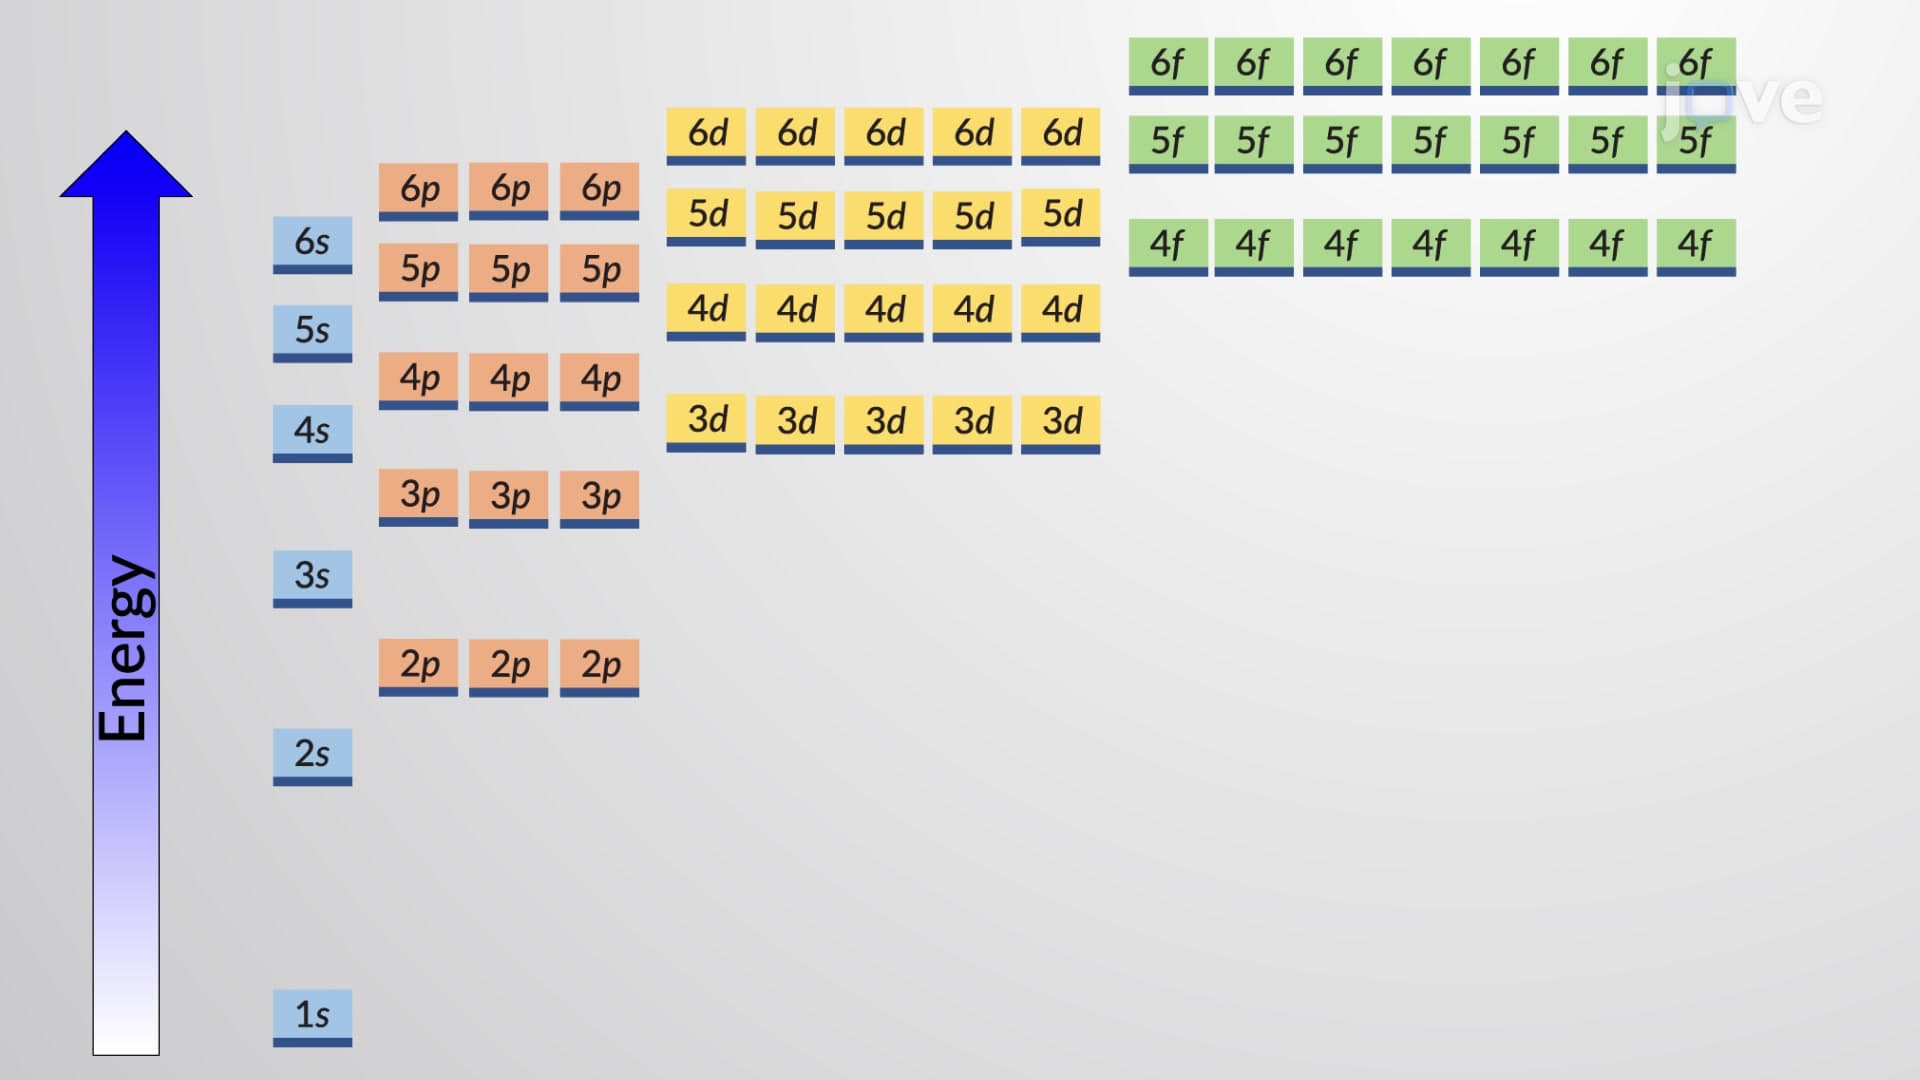
Electron Configuration of Multielectron Atoms
 01:27
Lattice Energies of Ionic Crystals
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...