Related Experiment Video
Updated: Jun 23, 2026
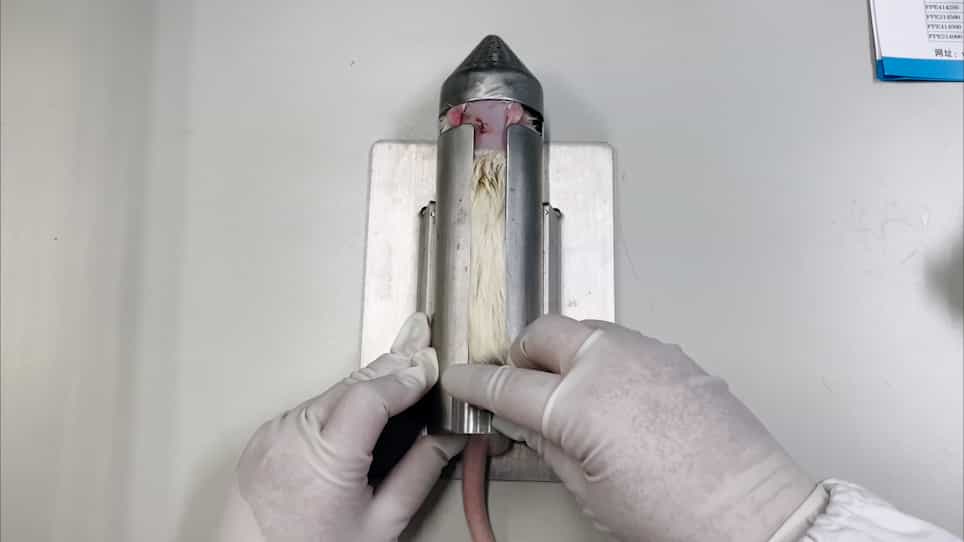

Microsurgical Skills of Establishing Permanent Jugular Vein Cannulation in Rats for Serial Blood Sampling of Orally Administered Drug
Published on: December 14, 2021
Pharmacokinetics of verproside after intravenous and oral administration in rats
Eun Jeong Park1, Hyun Sook Lee, Sei-Ryang Oh
1National Research Laboratory for Drug Metabolism and Bioanalysis, College of Pharmacy, Wonkwang University, Iksan, 570-749, Korea.

Abstract:
Verproside, a catalpol derivative iridoid glucoside isolated from Pseudolysimachion longifolium, is a candidate for anti-asthmatic drug. The dose-dependency of the pharmacokinetics of verproside was evaluated in rats after intravenous and oral administration. After intravenous administration of verproside (2, 5 and 10 mg/kg doses), the systemic clearance (Cl) was significantly reduced and AUC was significantly increased at 10 mg/kg dose compared to 2 and 5 mg/kg doses. The volume of distribution at steady state (V (ss)) remained unchanged as the dose was increased. The extent of urinary excretion was low for both intravenous (3.3-6.2%) and oral (0.01-0.04%) doses. Isovanilloylcatalpol was identified as a metabolite after intravenous administration of verproside and showed the significant decreases in AUC and C (max) at 10 mg/kg verproside dose. The reduced systemic clearance of verproside at high doses appears to be due to the saturable metabolism. Upon oral administration of verproside (20, 50 and 100 mg/kg doses), C (max) was nonlinearly increased. The extent of verproside recovered from the gastrointestinal tract at 24 h after oral administration was 0.01-0.72% for all three doses studied. The absolute oral bioavailability (F) was 0.3 and 0.5% for 50 and 100 mg/kg doses, respectively. Low F appears to be due to first-pass metabolism.
Related Concept Videos
 01:24
Factors Influencing Drug Absorption: Presystemic Elimination
 01:13
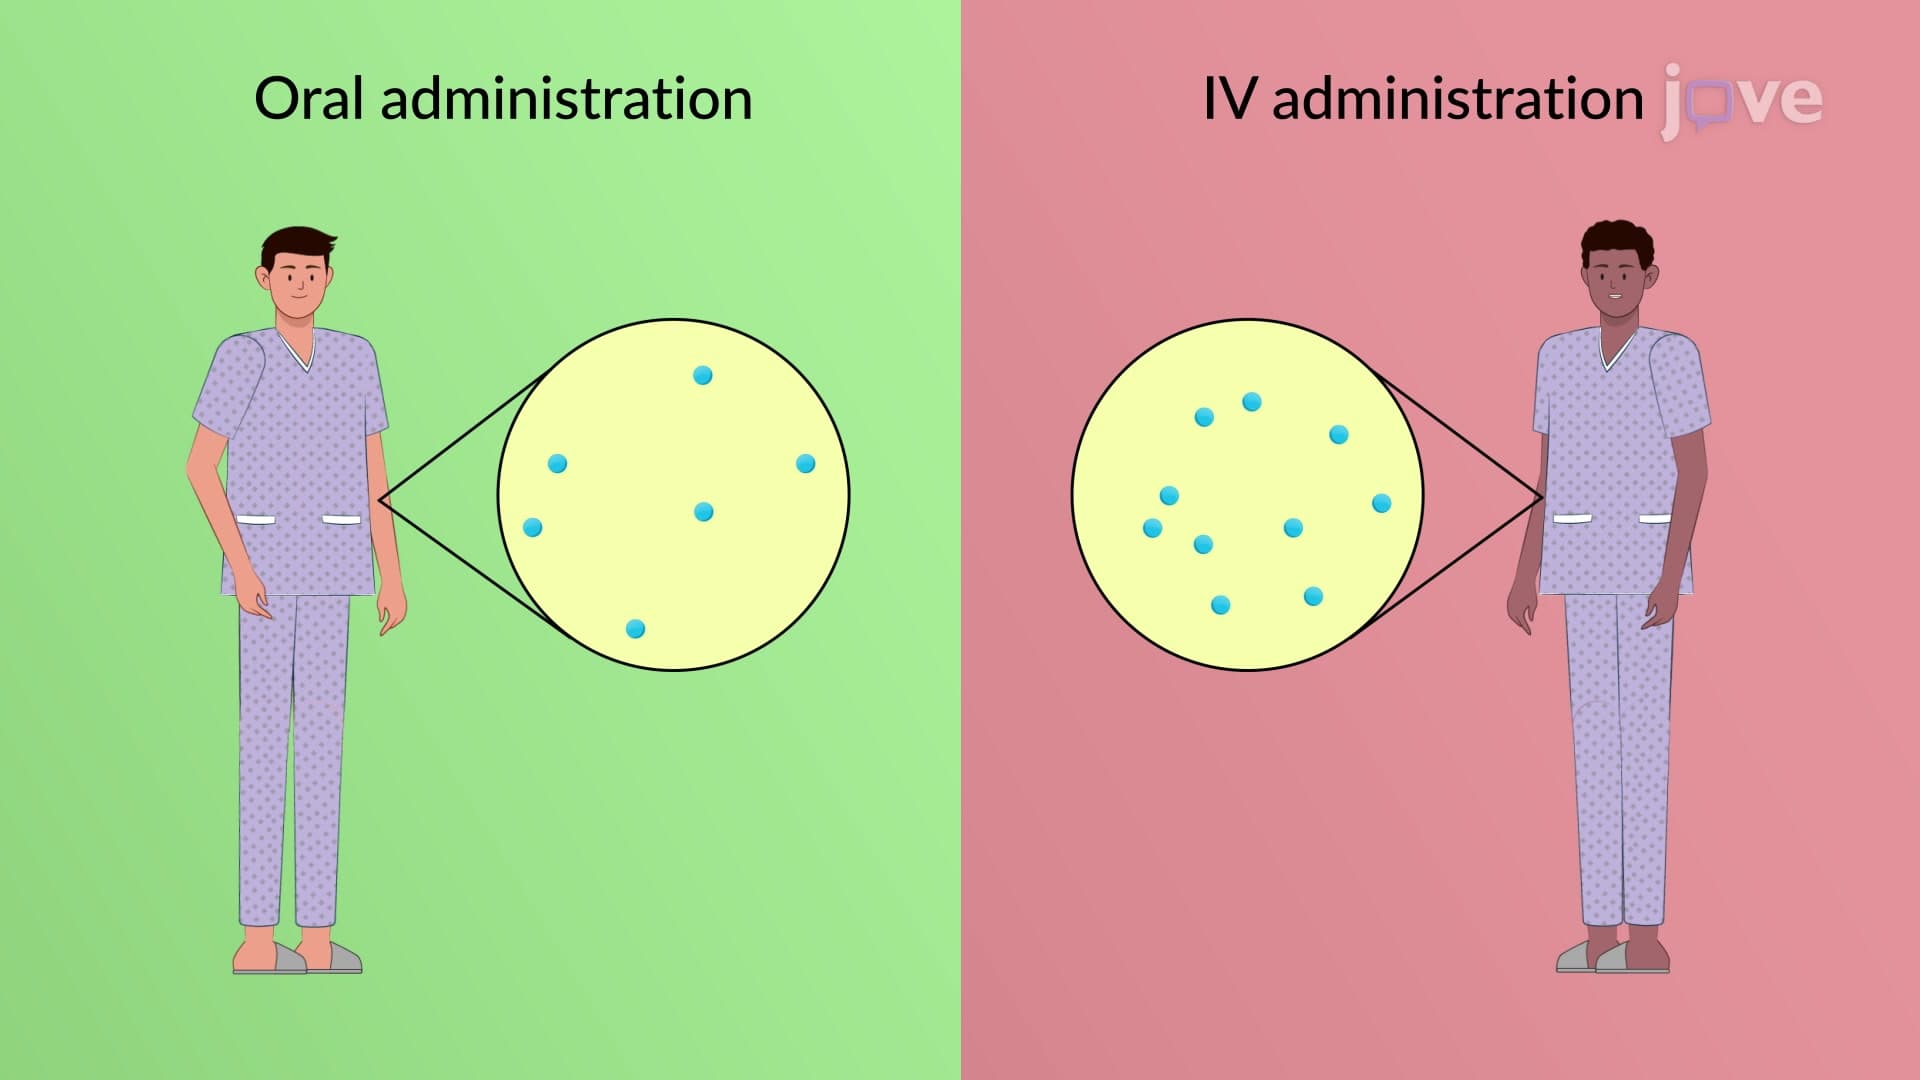
Bioavailability: Overview
 01:17
Bioavailability: Overview
 01:01
Dosage Regimens: Partial Pharmacokinetic Parameters
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...
 01:18
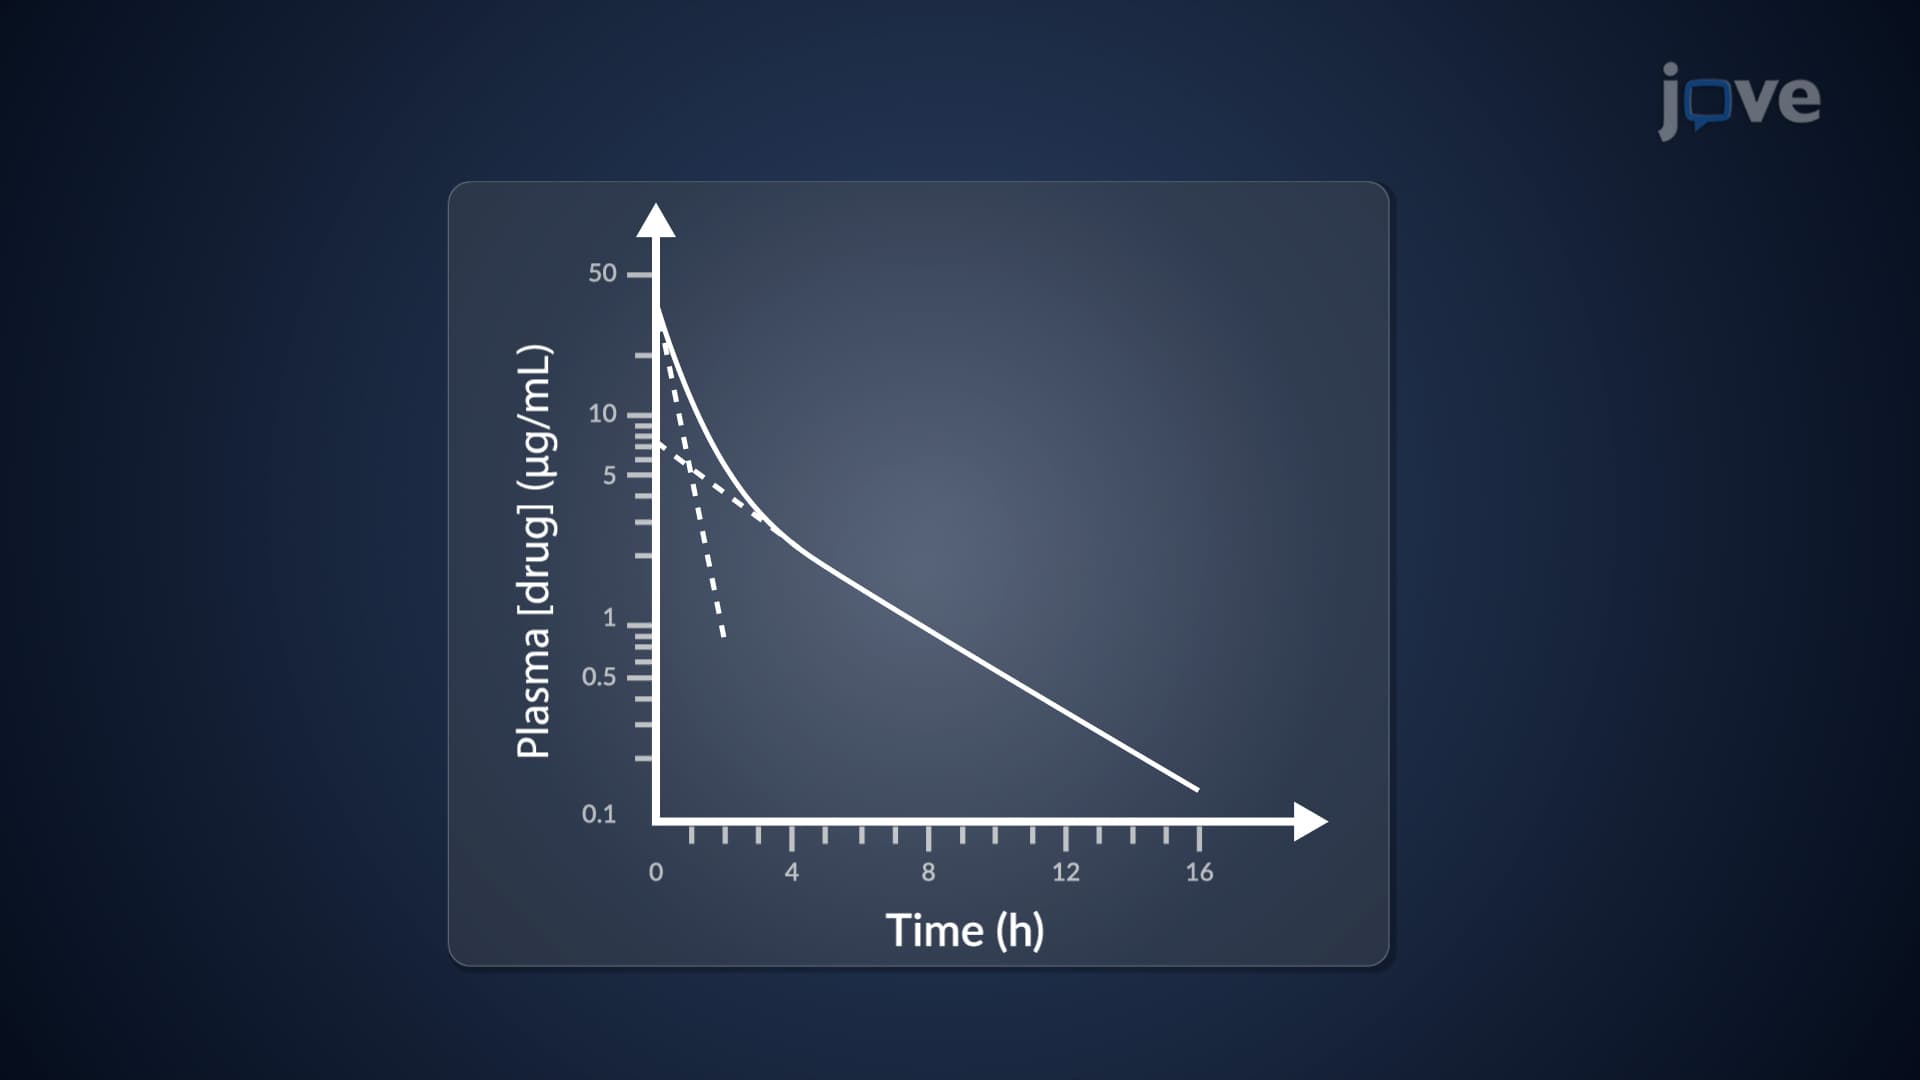
Two-Compartment Open Model: IV Bolus Administration
The disparity between drug input and the sum of drug transfer rates between...