Related Experiment Video
Updated: Jun 21, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Potential energy surface for ground-state H2S via scaling of the external correlation, comparison with extrapolation
Y Z Song1, P J S B Caridade, A J C Varandas
1Departamento de Química, Universidade de Coimbra 3004-535 Coimbra, Portugal.

Abstract:
A global double many-body expansion potential energy surface is reported for the electronic ground state of H2S by fitting accurate ab initio energies calculated at the multireference configuration interaction level with the aug-cc-pVQZ basis set, after slightly correcting semiempirically the dynamical correlation by the double many-body expansion-scaled external correlation method. The function so obtained has been compared in detail with a potential energy surface of the same type recently reported (Song , Y. Z. and Varandas , A. J. C. J. Chem. Phys. 2009 , 130 , 134317.) by extrapolating the calculated raw energies to the complete basis set limit, eschewing any use of information alien to ab initio theory. The new potential energy surface is also used for studying the dynamics and kinetics of the S(1D) + H2/D2/HD reactions.
More Related Videos
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 03:51
Molecular Orbital Theory II
 01:13
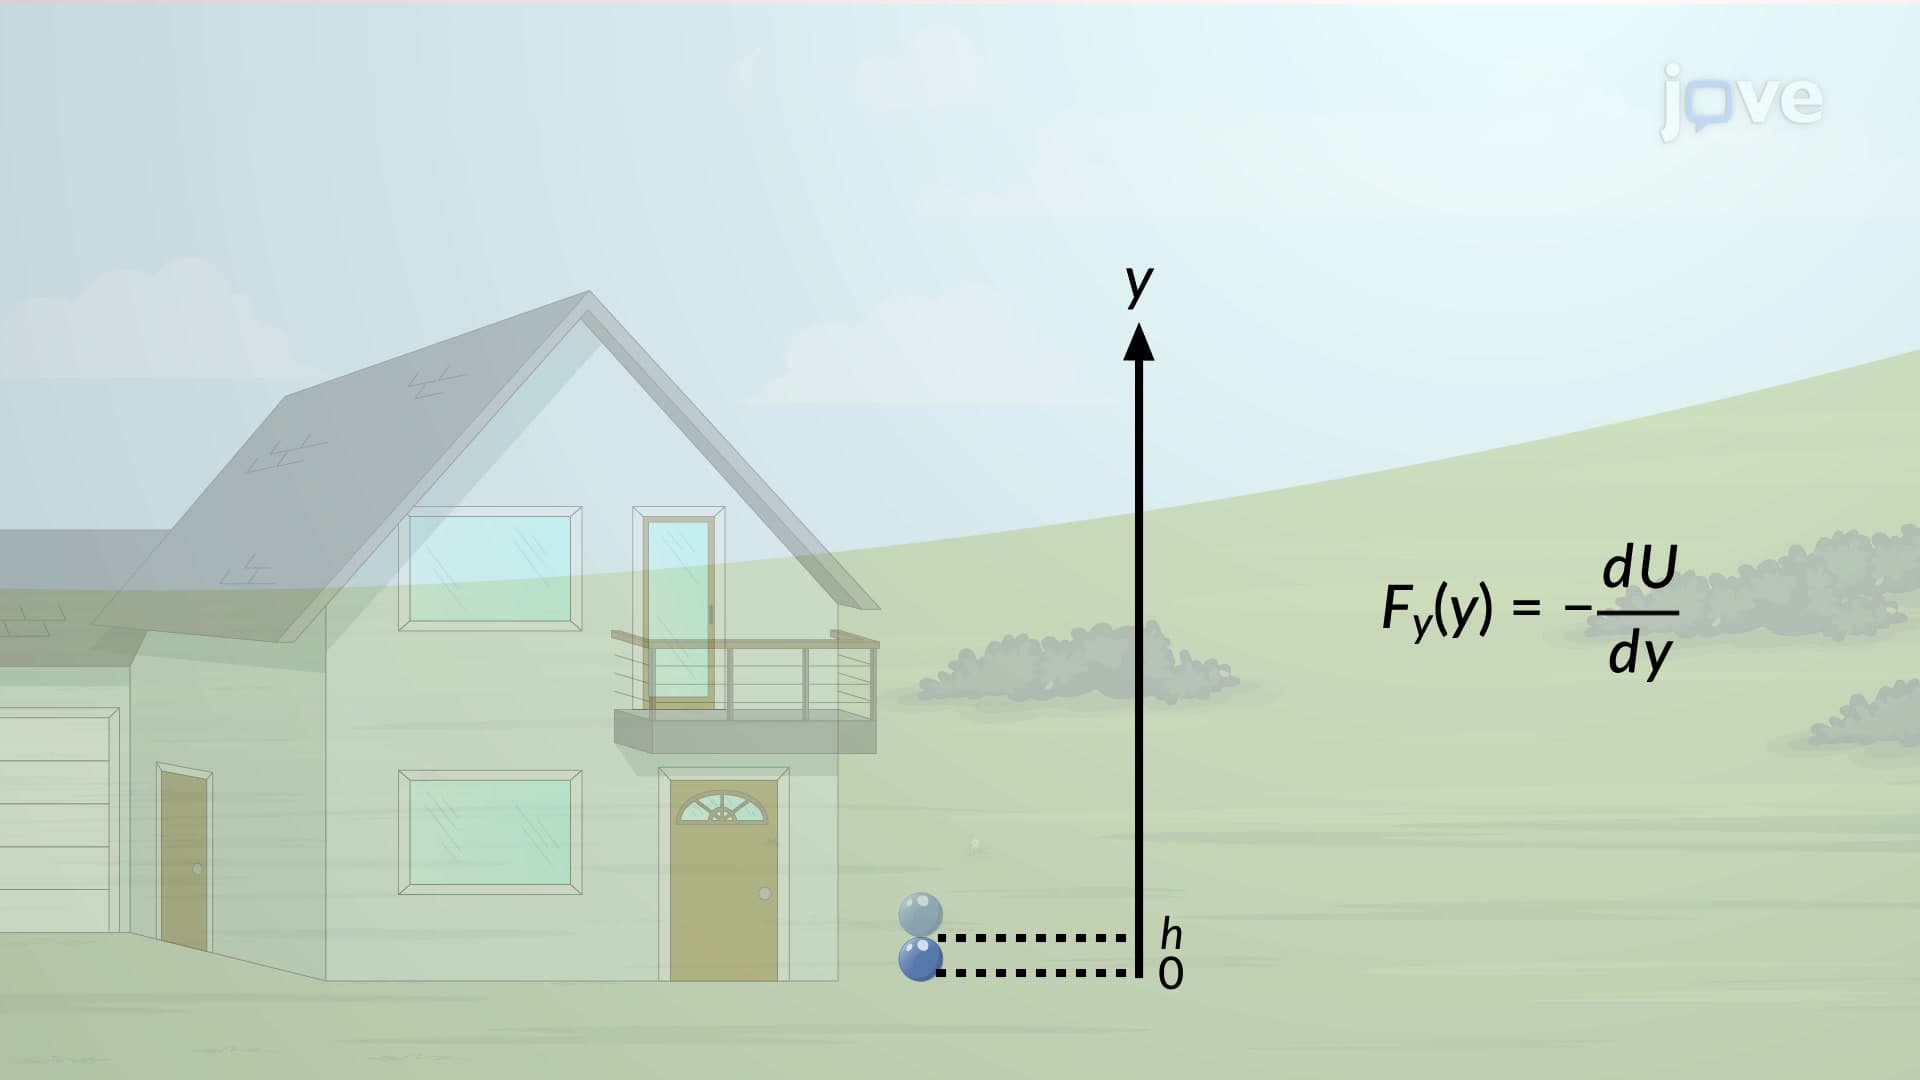
Force and Potential Energy in One Dimension
 03:40
Hess's Law
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
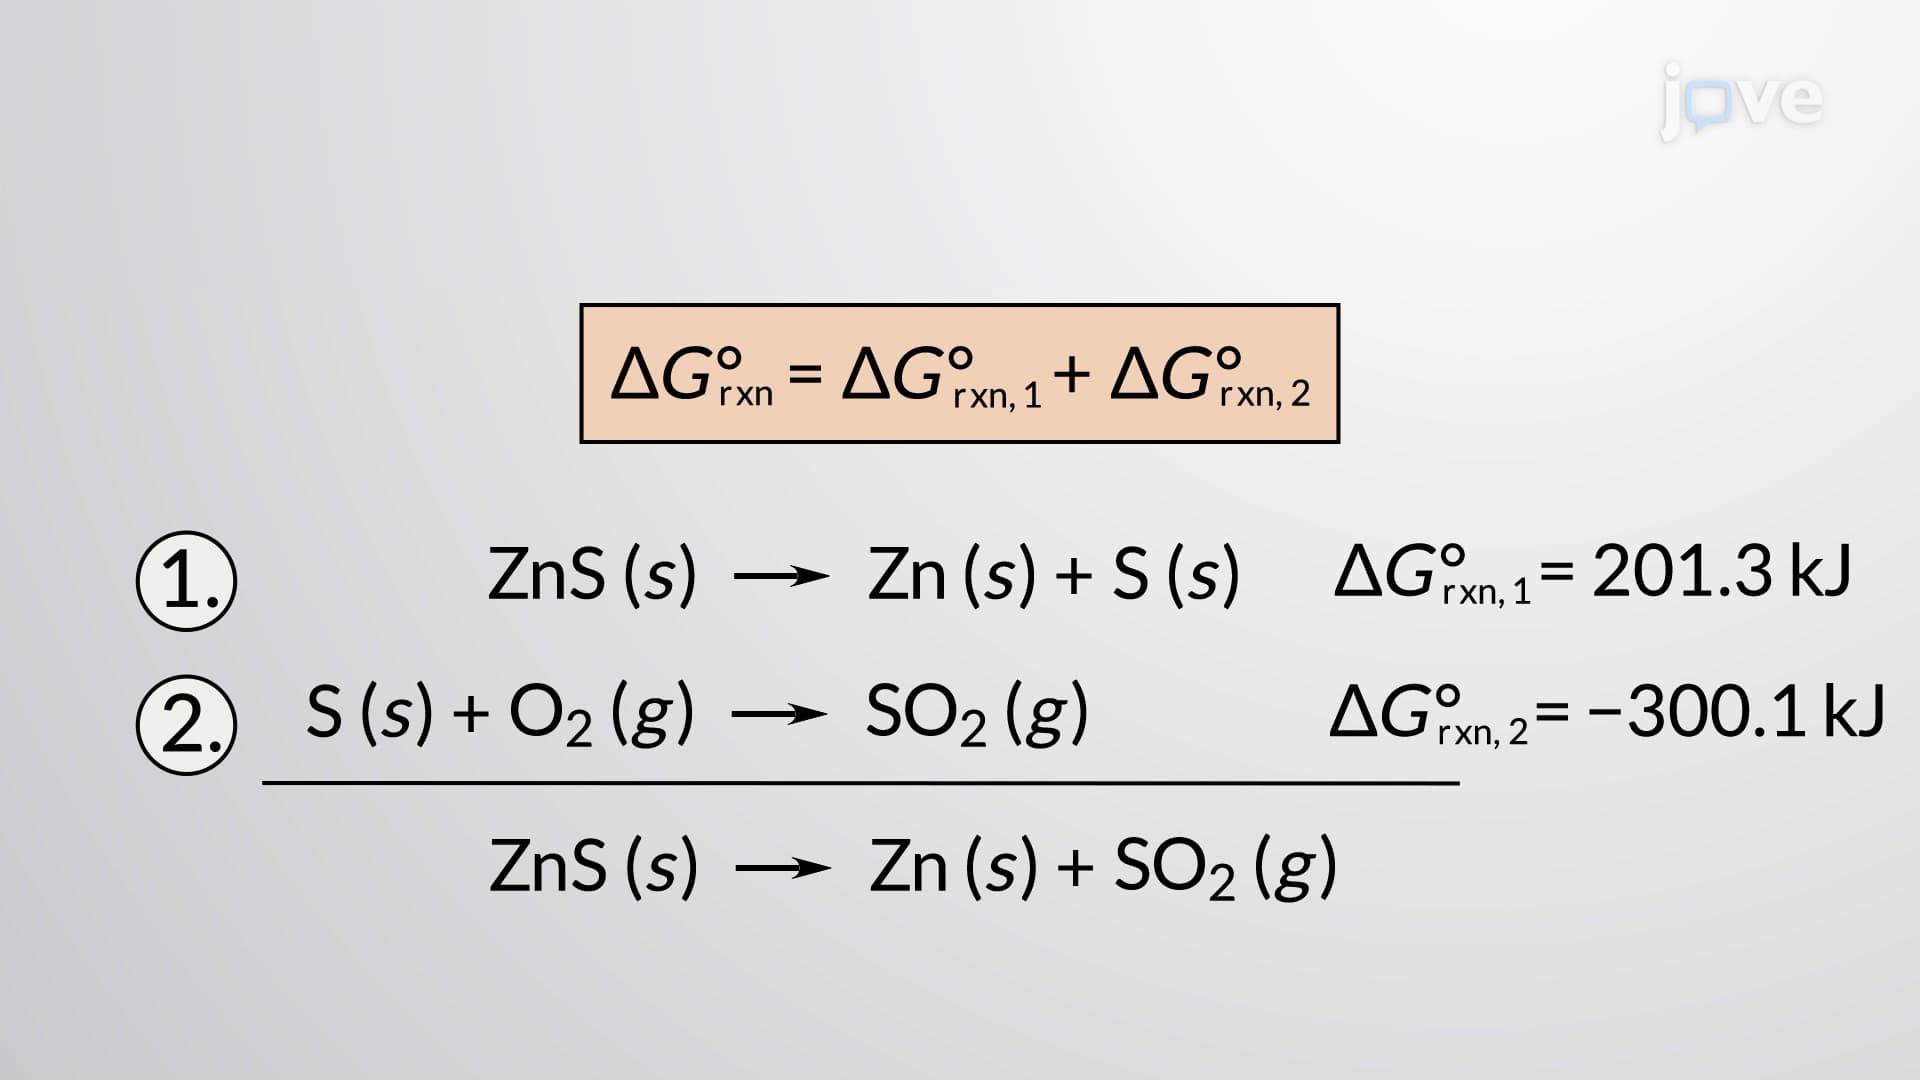 02:49
Calculating Standard Free Energy Changes
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...