N,N-Diethyl-N'-[(E)-4-pyridylmethylene]benzene-1,4-diamine: a combined X-ray and density functional theory study

Rüdiger W Seidel1, William S Sheldrick, Tsonko M Kolev

  • 1Analytische Chemie, Ruhr-Universität Bochum, Universitätsstrasse 150, 44780 Bochum, Germany. ruediger.seidel@rub.de

Related Concept Videos

π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds01:14

π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds

In aromatic compounds, such as benzene, the circulation of (4n + 2) π-electrons sets up a diamagnetic or diatropic ring current around the perimeter of the molecule. This current induces a magnetic field that opposes the external field inside the ring and reinforces it on the outside. The protons in benzene are deshielded and exhibit high chemical shifts in the range 6.5–8.5 ppm. The shielding effect at the center of the ring is evident in complex aromatic molecules, such as annulenes. In...
Structure of Benzene: Molecular Orbital Model01:18

Structure of Benzene: Molecular Orbital Model
According to the molecular orbital (MO) model, benzene has a planar structure with a regular hexagon of six sp2 hybridized carbons. As shown in Figure 1, each carbon is bonded to three other atoms with C–C–C and H–C–C bond angles of 120°. The C–H bond length is 109 pm, and the C–C bond length is 139 pm which is midway between the single bond length of sp3 hybridized carbons (154 pm) and sp2 hybridized carbons (133 pm).
NMR Spectroscopy of Benzene Derivatives01:37

NMR Spectroscopy of Benzene Derivatives

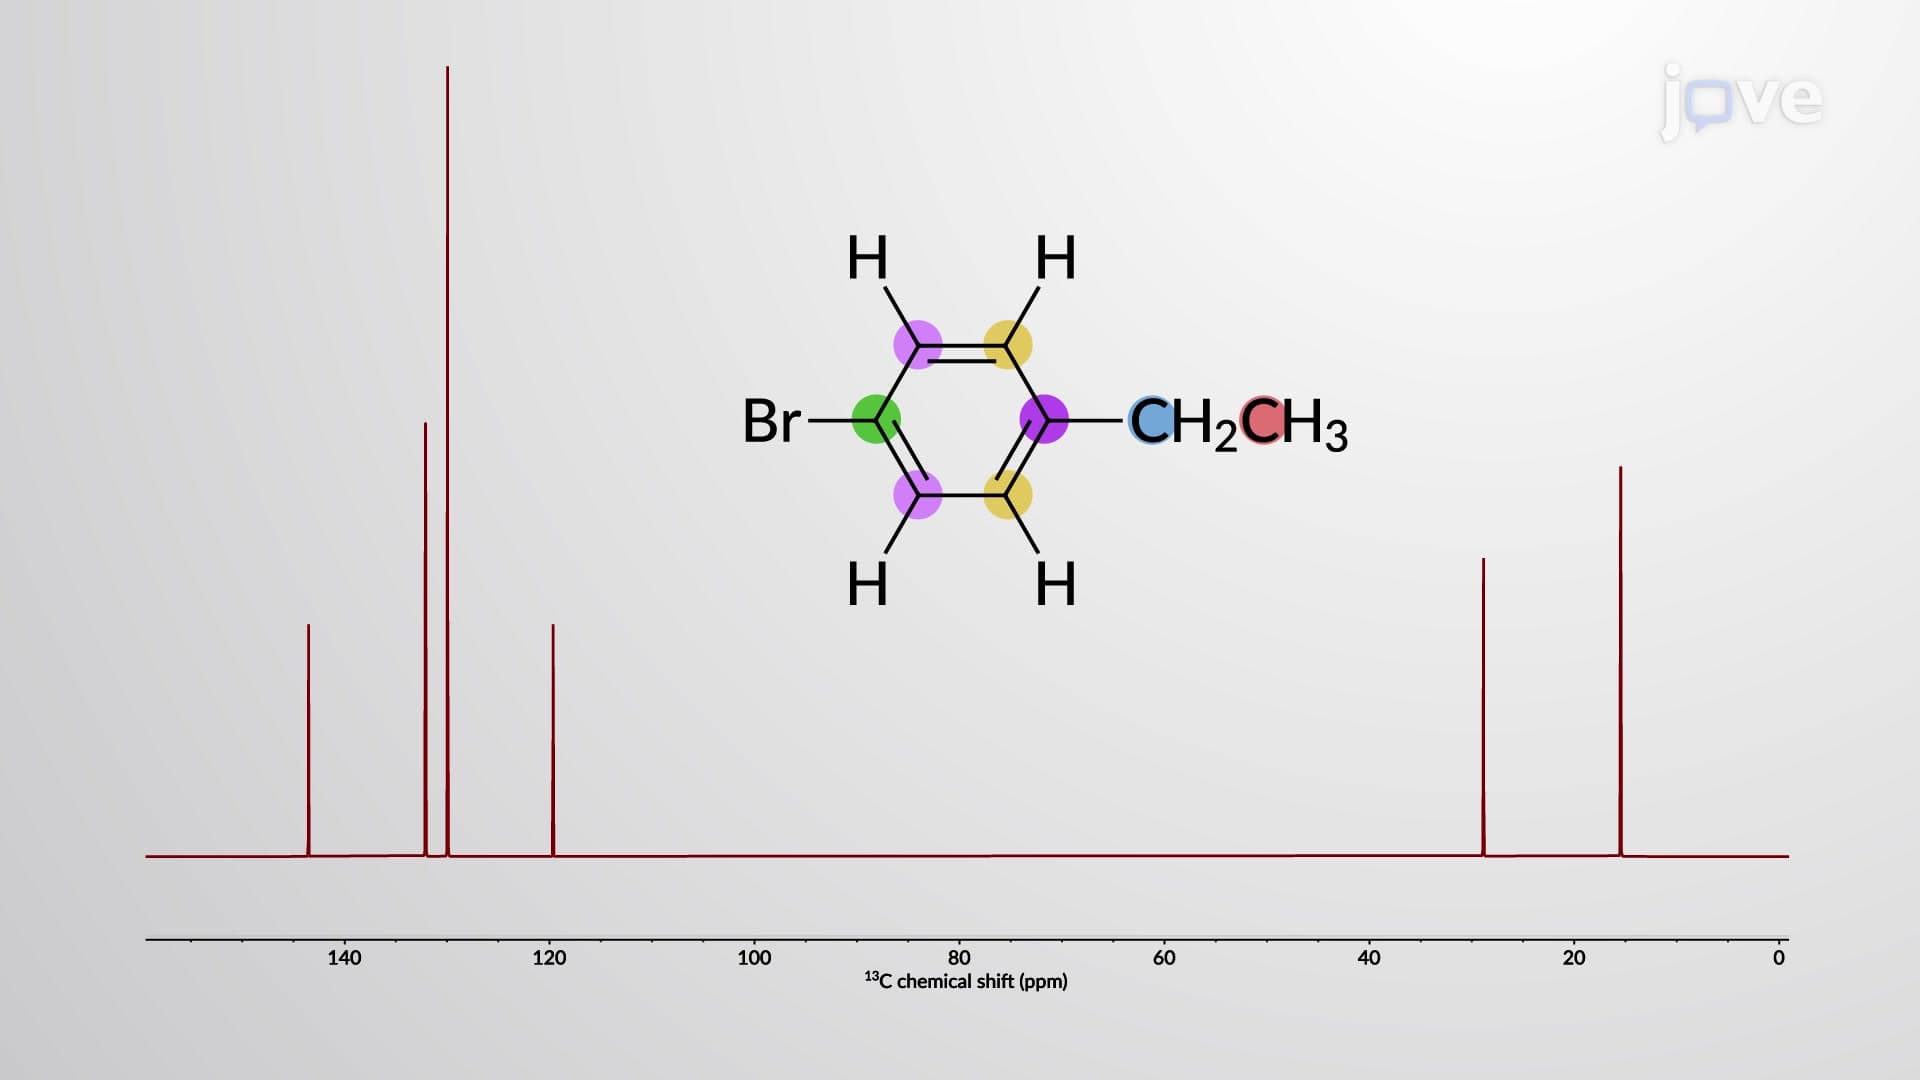
Simple unsubstituted benzene has six aromatic protons, all chemically equivalent. Therefore, benzene exhibits only a singlet peak at δ 7.3 ppm in the 1H NMR spectrum. The observed shift is far downfield because the aromatic ring current strongly deshields the protons. Any substitution on the benzene ring makes the aromatic protons nonequivalent, and the protons split each other. The peak is, therefore, no longer a singlet and the splitting pattern and their associated coupling constants depend...
UV–Vis Spectroscopy: Woodward–Fieser Rules01:29

UV–Vis Spectroscopy: Woodward–Fieser Rules

UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given structure by adding the contributions...
Basicity of Heterocyclic Aromatic Amines01:25

Basicity of Heterocyclic Aromatic Amines

Heterocyclic amines, where the N atom is a part of an alicyclic system, are similar in basicity to alkylamines. Interestingly, the heterocyclic amine having a nitrogen atom as part of an aromatic ring has much less basicity than its corresponding alicyclic counterpart. For this reason, as presented in Figure 1, piperidine (pKb = 2.8) is significantly more basic than pyridine (pKb = 8.8).
π Molecular Orbitals of 1,3-Butadiene01:24

π Molecular Orbitals of 1,3-Butadiene

Conjugated dienes have lower heats of hydrogenation than cumulated and isolated dienes, making them more stable. The enhanced stabilization of conjugated systems can be understood from their π molecular orbitals.
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...