Related Experiment Video
Updated: Jun 20, 2026
Thermochemical Studies of Ni(II) and Zn(II) Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Assessment of the CCSD and CCSD(T) coupled-cluster methods in calculating heats of formation for Zn complexes
Michael N Weaver1, Yue Yang, Kenneth M Merz
1Department of Chemistry, Quantum Theory Project, 2328 New Physics Building, University of Florida, Gainesville, Florida 32611-8435, USA.

Abstract:
Heats of formation were calculated using coupled-cluster methods for a series of zinc complexes. The calculated values were evaluated against previously conducted computational studies using density functional methods as well as experimental values. Heats of formation for nine neutral ZnX(n) complexes [X = -Zn, -H, -O, -F2, -S, -Cl, -Cl2, -CH3, (-CH3)2] were determined at the CCSD and CCSD(T) levels using the 6-31G** and TZVP basis sets as well as the LANL2DZ-6-31G** (LACVP**) and LANL2DZ-TZVP hybrid basis sets. The CCSD(T)/6-31G** level of theory was found to predict the heat of formation for the nonalkyl Zn complexes most accurately. The alkyl Zn species were problematic in that none of the methods that were tested accurately predicted the heat of formation for these complexes. In instances where experimental geometric parameters were available, these were most accurately predicted by the CCSD/6-31G** level of theory; going to CCSD(T) did not improve agreement with the experimental values. Coupled-cluster methods did not offer a systemic improvement over DFT calculations for a given functional/basis set combination. With the exceptions of ZnH and ZnF2, there are multiple density functionals that outperform coupled-cluster calculations with the 6-31G** basis set.
More Related Videos
 09:22
Synthesis and Performance Evaluations of ZnCoS/ZnCdS with Twin Crystal Structure for Multifunctional Redox Photocatalysis in Energy Applications
Published on: July 25, 2025
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:44
The Born-Haber Cycle
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:07
Ladder Diagrams: Complexation Equilibria
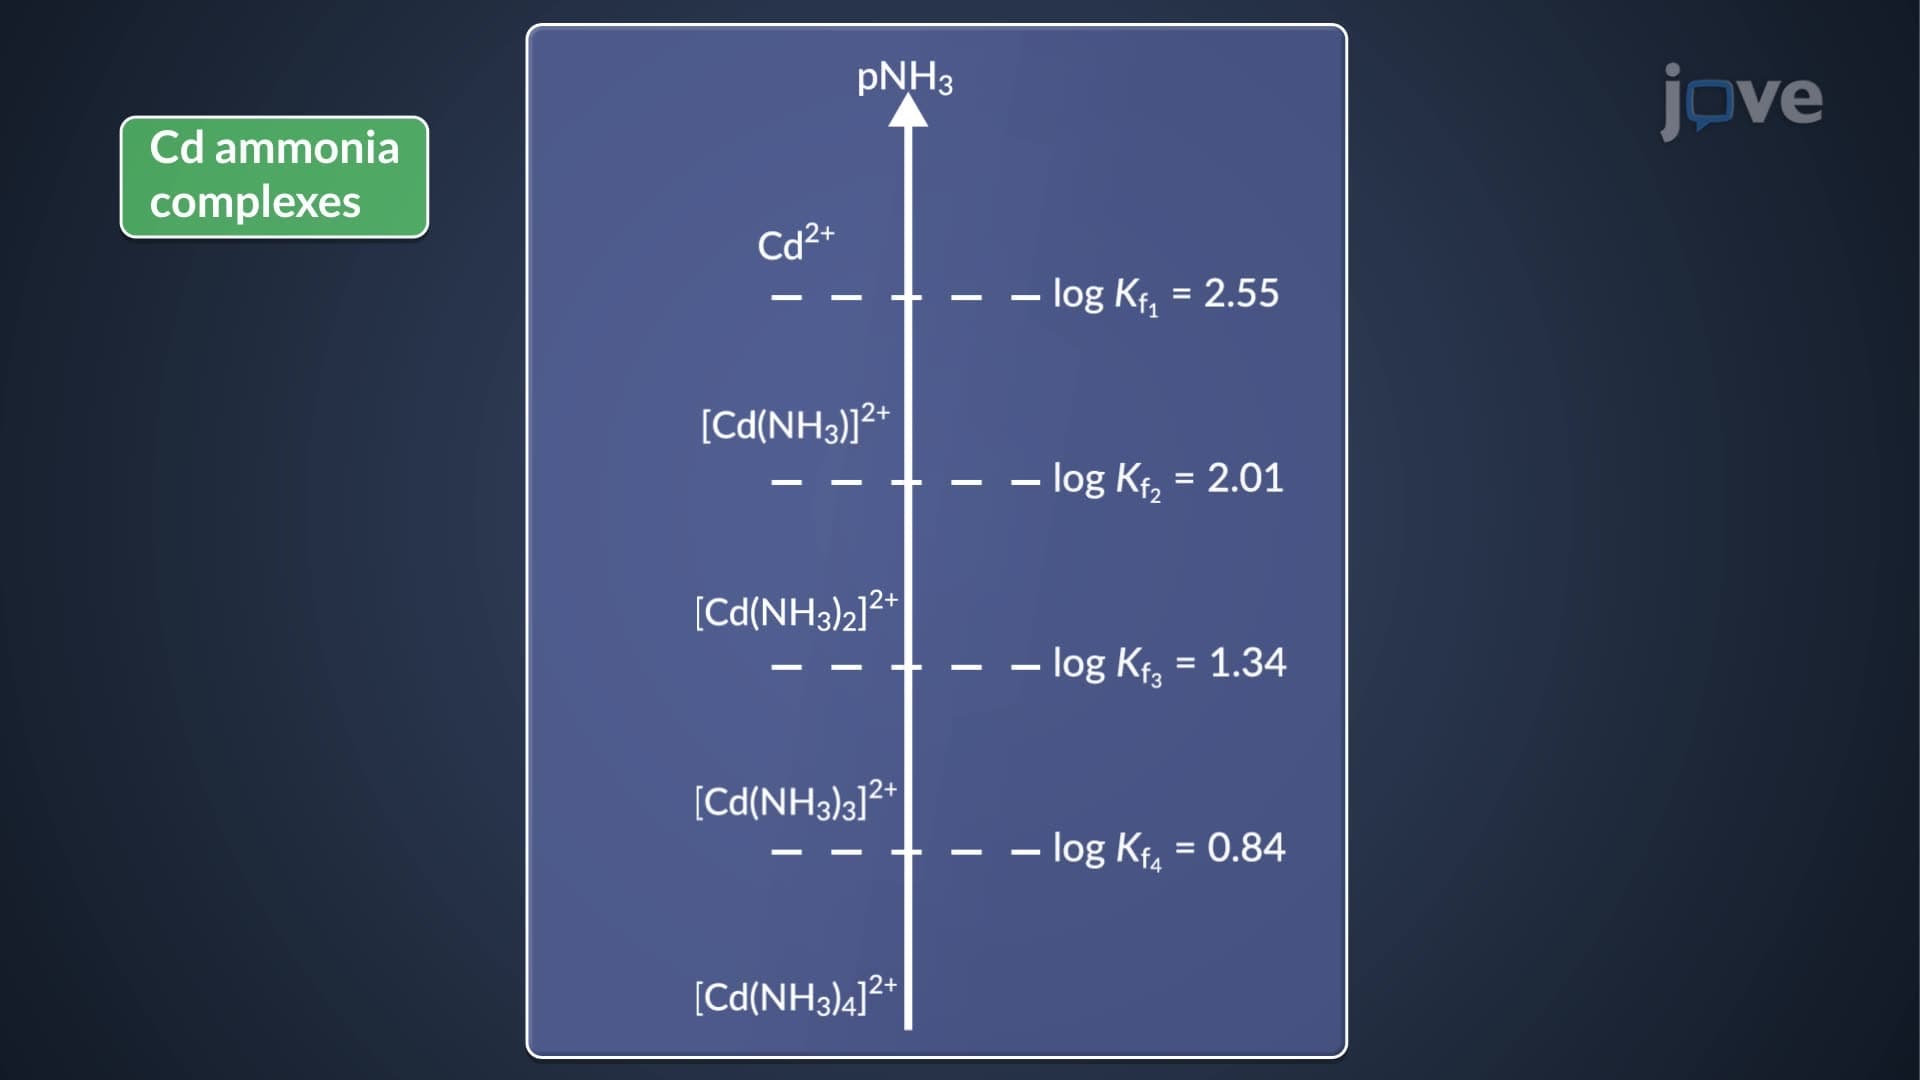
The formation constant, K1, for the formation of Cd(NH3)2+ complex from cadmium and ammonia is 3.55 × 10². Log K1 (i.e. pNH3) is 2.55, and...
 02:37
Standard Enthalpy of Formation
 02:42
Valence Bond Theory