Related Experiment Video
Updated: Jun 19, 2026

Measurements of Long-range Electronic Correlations During Femtosecond Diffraction Experiments Performed on Nanocrystals of Buckminsterfullerene
Published on: August 22, 2017
Intramolecular electronic communication in a dimethylaminoazobenzene-fullerene C60 dyad: an experimental and TD-DFT
K Senthil Kumar1, Archita Patnaik
1Department of Chemistry, Indian Institute of Technology Madras, Chennai 600 036, India.

Abstract:
An electronically push-pull type dimethylaminoazobenzene-fullerene C(60) hybrid was designed and synthesized by tailoring N,N-dimethylaniline as an electron donating auxochrome that intensified charge density on the beta-azonitrogen, and on N-methylfulleropyrrolidine (NMFP) as an electron acceptor at the 4 and 4' positions of the azobenzene moiety, respectively. The absorption and charge transfer behavior of the hybrid donor-bridge-acceptor dyad were studied experimentally and by performing TD-DFT calculations. The TD-DFT predicted charge transfer interactions of the dyad ranging from 747 to 601 nm were experimentally observed in the UV-vis spectra at 721 nm in toluene and dichloromethane. A 149 mV anodic shift in the first reduction potential of the N=N group of the dyad in comparison with the model aminoazobenzene derivative further supported the phenomenon. Analysis of the charge transfer band through the orbital picture revealed charge displacement from the n((N=N)) (nonbonding) and pi ((N=N)) type orbitals centered on the donor part to the purely fullerene centered LUMOs and LUMO+n orbitals, delocalized over the entire molecule. The imposed electronic perturbations on the aminoazobenzene moiety upon coupling it with C(60) were analyzed by comparing the TD-DFT predicted and experimentally observed electronic transition energies of the dyad with the model compounds, NMFP and (E)-N,N-dimethyl-4-(p-tolyldiazenyl)aniline (AZNME). The n((N=N)) --> pi*((N=N)) and pi((N=N)) --> pi*((N=N)) transitions of the dyad were bathochromically shifted with a significant charge transfer character. The shifting of pi((N=N)) --> pi*((N=N)) excitation energy closer to the n --> pi*((N=N)) in comparison with the model aminoazobenzene emphasized the predominant existence of charge separated quinonoid-like ground state electronic structure. Increasing solvent polarity introduced hyperchromic effect in the pi((N=N)) --> pi*((N=N)) electronic transition at the expense of transitions involved with benzenic states, and the extent of intensity borrowing was quantified adopting the Gaussian deconvolution method. On a comparative scale, the predicted excitation energies were in reasonable agreement with the observed values, demonstrating the efficiency of TD-DFT in predicting the localized and the charge transfer nature of transitions involved with large electronically asymmetric molecules with HOMO and LUMO centered on different parts of the molecular framework.
More Related Videos
Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group with both...
 01:18
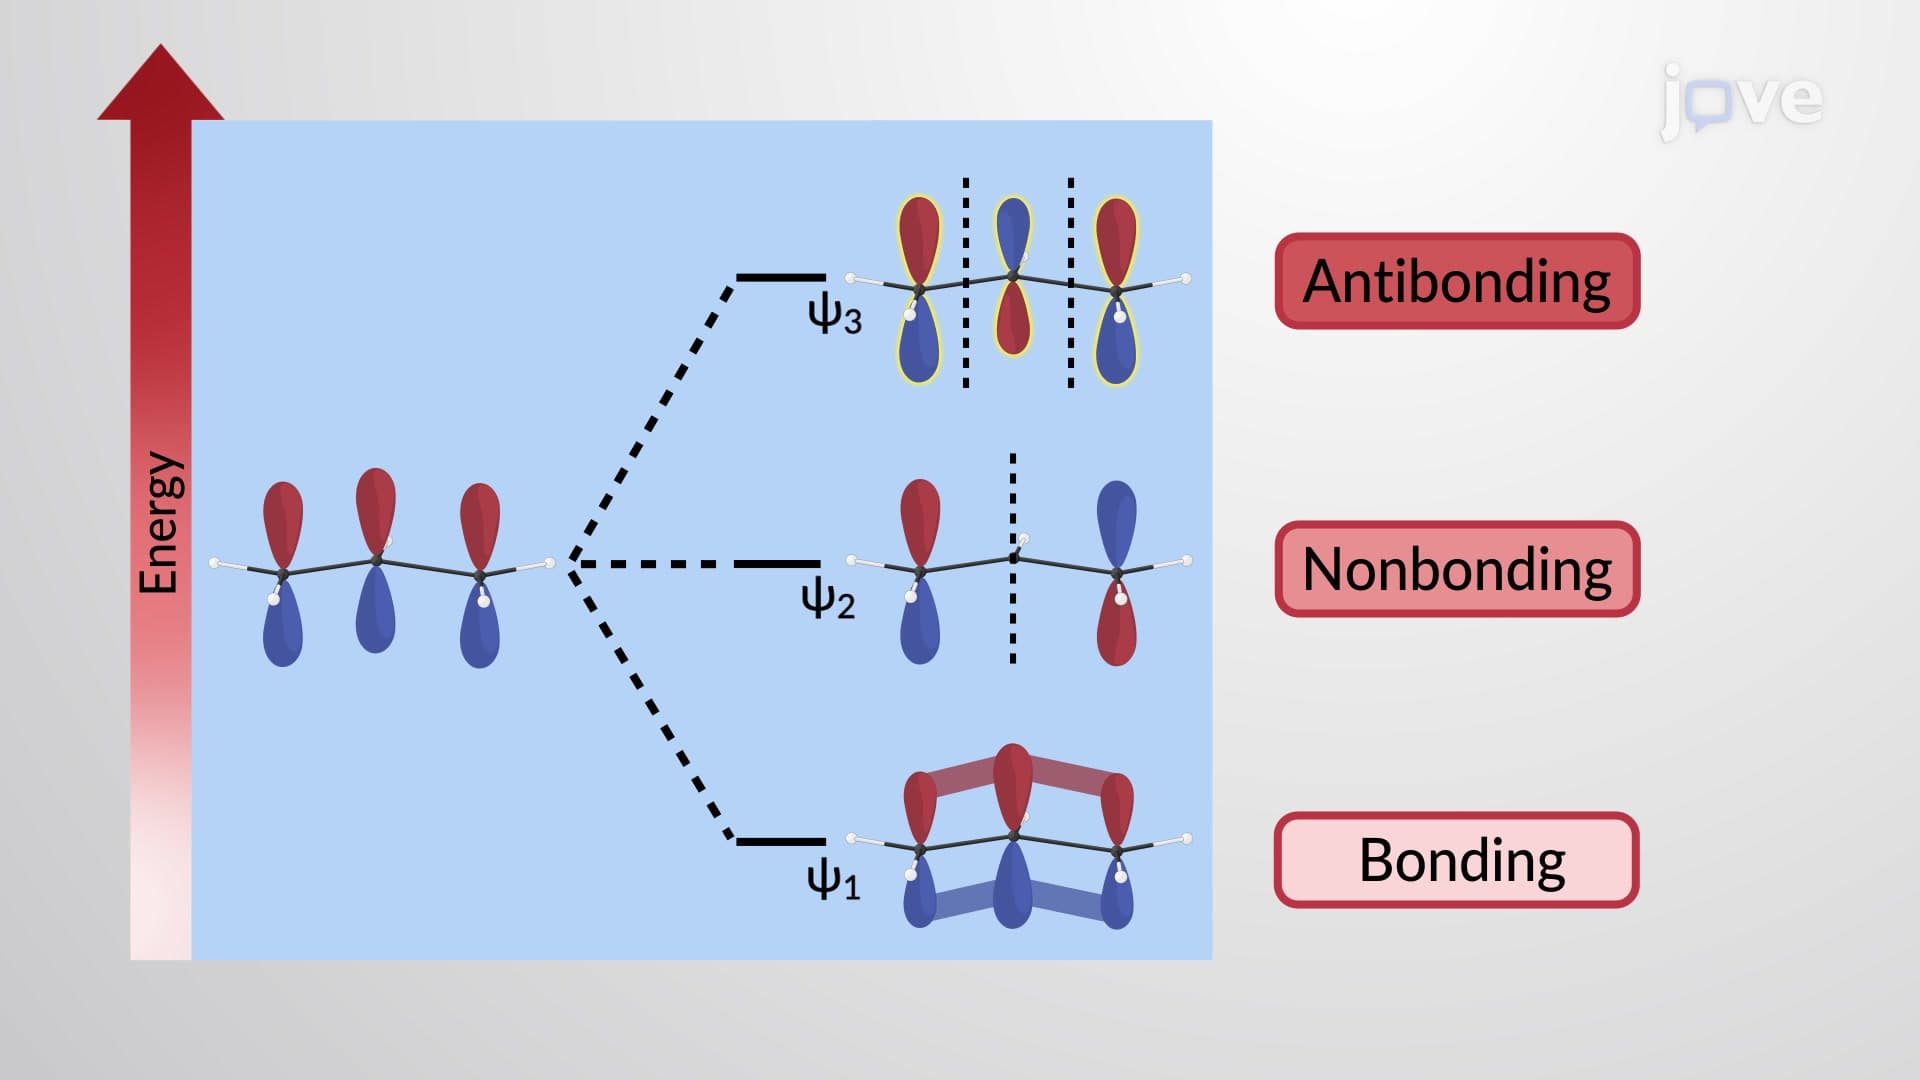
π Molecular Orbitals of the Allyl Cation and Anion
 01:16
[4+2] Cycloaddition of Conjugated Dienes: Diels–Alder Reaction
 01:18
Structure of Benzene: Molecular Orbital Model
 01:16
Structure of Conjugated Dienes
Conjugated dienes are compounds characterized by the presence of alternating double and single bonds. In a conjugated system like 1,3-butadiene, the unhybridized 2p orbital on each carbon overlaps continuously, allowing the π electrons to be delocalized across the entire molecule. In contrast, this type of overlap does not occur in cumulated and isolated dienes, such as 2,3-pentadiene and 1,4-pentadiene, respectively. Instead, the π electrons remain localized between the double...