Related Experiment Video
Updated: Jun 19, 2026
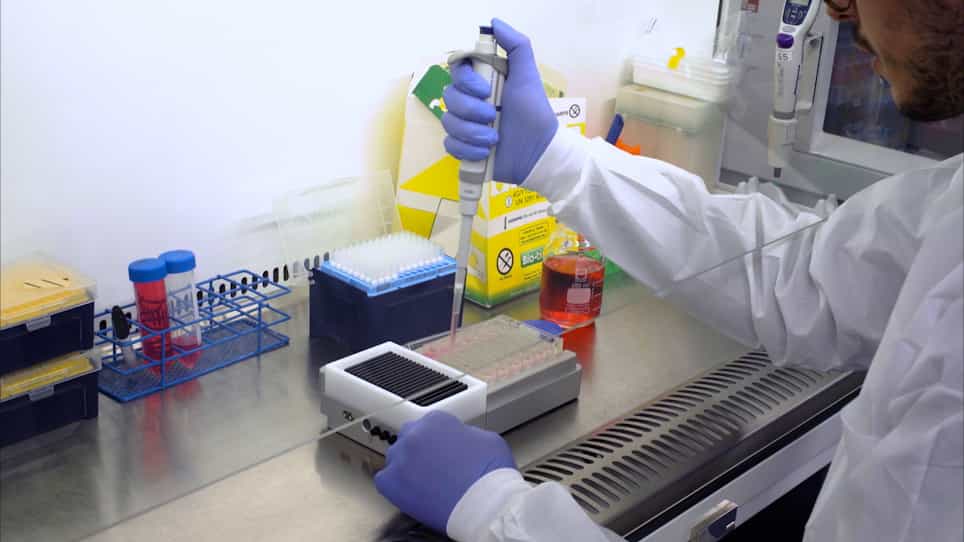

Human Liver Microphysiological System for Assessing Drug-Induced Liver Toxicity In Vitro
Published on: January 31, 2022
Biochemical mechanisms in drug-induced liver injury: certainties and doubts

Abstract:
Drug-induced liver injury is a significant and still unresolved clinical problem. Limitations to knowledge about the mechanisms of toxicity render incomplete the detection of hepatotoxic potential during preclinical development. Several xenobiotics are lipophilic substances and their transformation into hydrophilic compounds by the cytochrome P-450 system results in production of toxic metabolites. Aging, preexisting liver disease, enzyme induction or inhibition, genetic variances, local O(2) supply and, above all, the intrinsic molecular properties of the drug may affect this process. Necrotic death follows antioxidant consumption and oxidation of intracellular proteins, which determine increased permeability of mitochondrial membranes, loss of potential, decreased ATP synthesis, inhibition of Ca(2+)-dependent ATPase, reduced capability to sequester Ca(2+) within mitochondria, and membrane bleb formation. Conversely, activation of nucleases and energetic participation of mitochondria are the main intracellular mechanisms that lead to apoptosis. Non-parenchymal hepatic cells are inducers of hepatocellular injury and targets for damage. Activation of the immune system promotes idiosyncratic reactions that result in hepatic necrosis or cholestasis, in which different HLA genotypes might play a major role. This review focuses on current knowledge of the mechanisms of drug-induced liver injury and recent advances on newly discovered mechanisms of liver damage. Future perspectives including new frontiers for research are discussed.
Insights
Drug-induced liver injury remains a challenge due to incomplete understanding of toxicity mechanisms. This review details known pathways and emerging insights into drug hepatotoxicity.
Area of Science:
- Hepatology
- Toxicology
- Pharmacology
Background:
- Drug-induced liver injury (DILI) is a major clinical issue, hindered by incomplete knowledge of toxicity mechanisms.
- Preclinical detection of hepatotoxic potential is limited, partly due to the complex metabolic activation of xenobiotics by cytochrome P-450 into toxic metabolites.
- Factors like aging, genetic variations, and intrinsic drug properties influence DILI susceptibility.
Discussion:
- Cellular mechanisms of DILI involve both necrosis, characterized by mitochondrial dysfunction and oxidative stress, and apoptosis, involving nucleases and mitochondrial energy production.
- Non-parenchymal liver cells contribute to and are targets of hepatocellular injury.
- Immune system activation can lead to idiosyncratic DILI, with HLA genotypes potentially playing a role in hepatic necrosis or cholestasis.
Key Insights:
- Understanding the metabolic activation of drugs into toxic metabolites is crucial for predicting hepatotoxicity.
- Mitochondrial dysfunction and oxidative stress are key events in drug-induced necrosis.
- Apoptosis pathways are also significantly involved in DILI, highlighting the complexity of cellular responses.
Outlook:
- Future research should focus on novel mechanisms of liver damage and improved preclinical detection methods.
- Investigating the role of genetic factors, such as HLA genotypes, in idiosyncratic DILI is essential.
- Developing strategies to mitigate DILI requires a deeper understanding of its multifaceted pathogenesis.
Related Concept Videos
 01:26
Effect of Hepatic Disease on Pharmacokinetics: Drug Dosing and Hepatic Blood Flow
 01:22
Effect of Hepatic Disease on Pharmacokinetics: Pathophysiologic Assessment and Liver Function Test
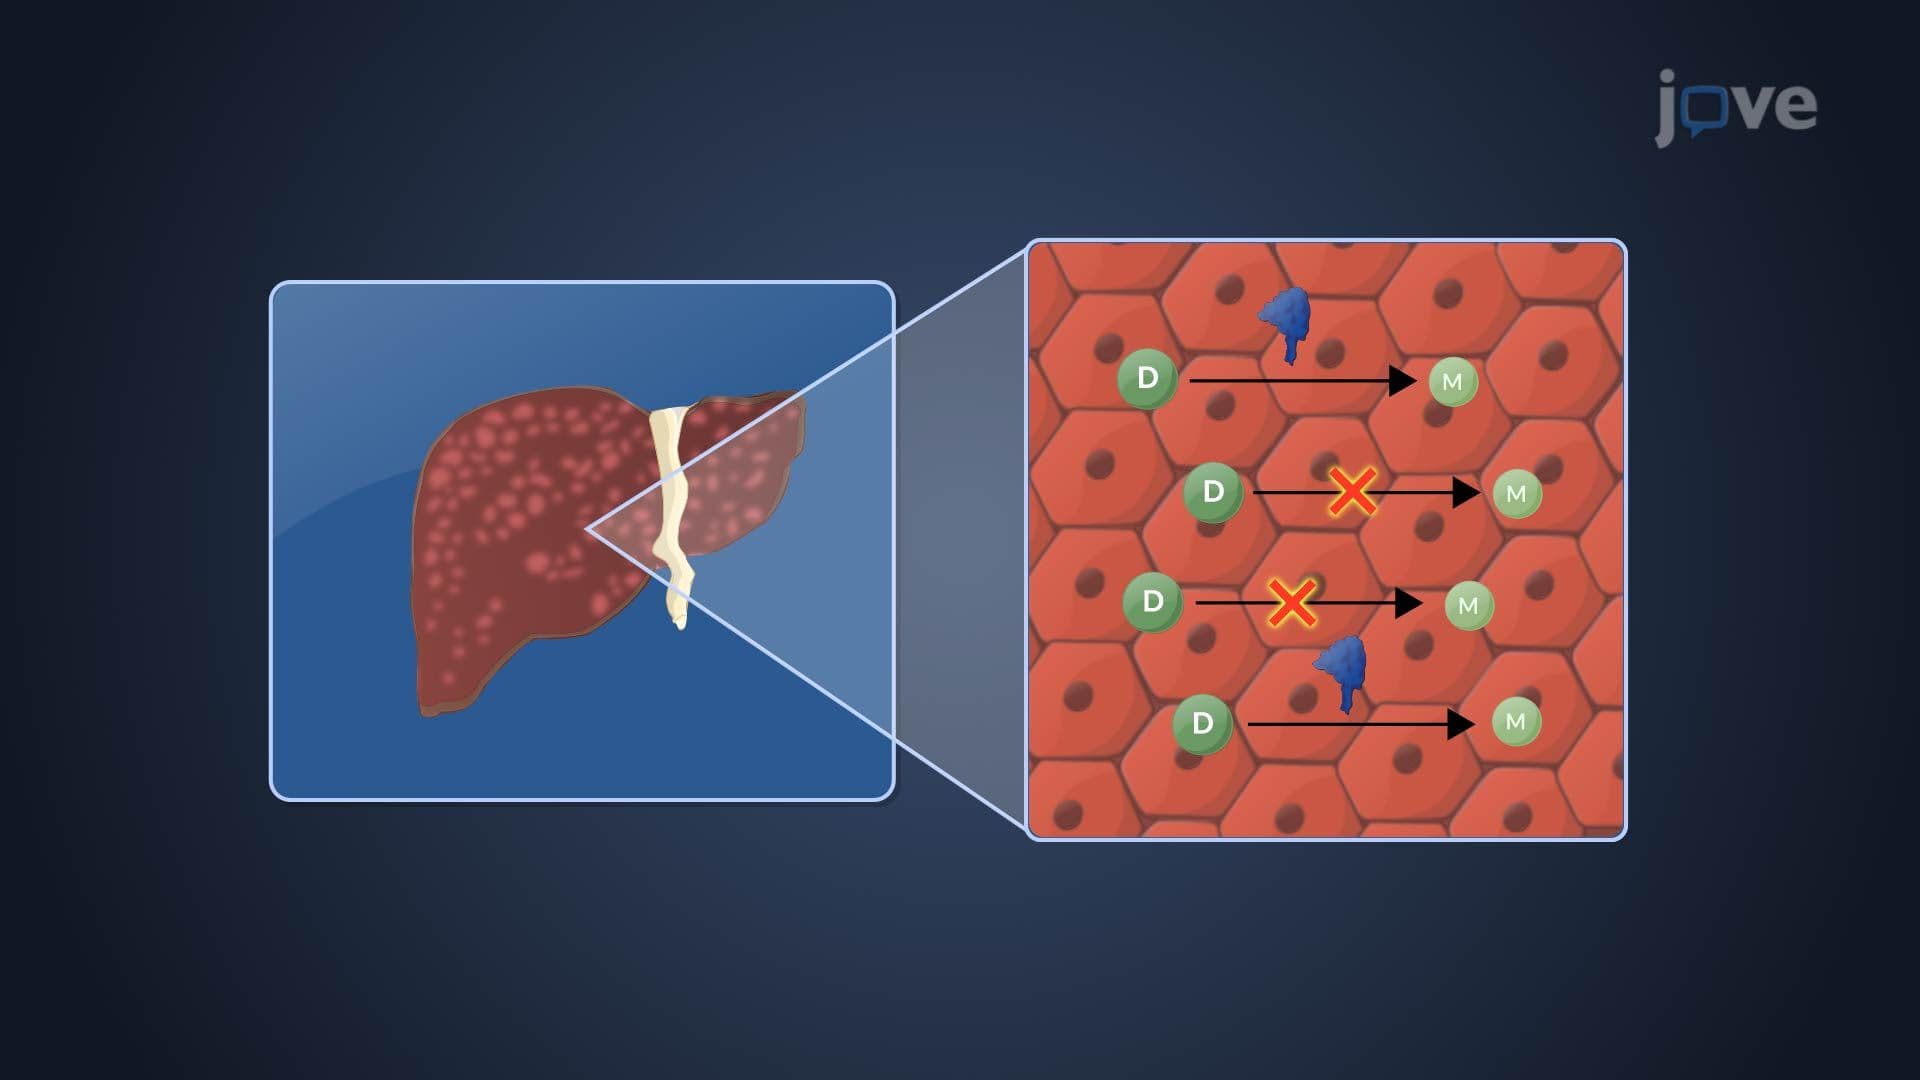 01:14
Effect of Hepatic Disease on Pharmacokinetics: Active Drug, Metabolite and Fraction of Metabolized Drug
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:16
Drug toxicity: Idiosyncratic Reactions
 01:16
Drug Biotransformation: Overview