Related Experiment Video
Updated: Jun 18, 2026
Technical Aspect of the Automated Synthesis and Real-Time Kinetic Evaluation of [11C]SNAP-7941
Published on: April 28, 2019
Prediction of pharmacokinetic profile of valsartan in humans based on in vitro uptake-transport data
Agnès Poirier1, Anne-Christine Cascais, Christoph Funk
1F. Hoffmann-La Roche Ltd., Non-Clinical Development, Drug Safety, CH-4070 Basel. agnes.poirier@roche.com

Abstract:
The aim of this study was to evaluate a physiologically based pharmacokinetic (PBPK) model for predicting PK profiles in humans based on a model refined in rats and humans in vitro uptake-transport data using valsartan as a probe substrate. Valsartan is eliminated unchanged, mostly through biliary excretion, both in humans and rats. It was, therefore, chosen as model compound to predict in vivo elimination based on in vitro hepatic uptake-transport data using a fully mechanistic PBPK model. Plated rat and human hepatocytes, and cell lines overexpressing human OATP1B1 and OATP1B3 were used for in vitro uptake experiments. A mechanistic two-compartment model was used to derive the active and passive transport parameters, namely uptake Michaelis-Menten parameters (V(max) and K(m,u)) together with passive diffusion (P(dif)). These transport parameters were then used as input in a whole body physiologically based pharmacokinetic (PBPK) model. The uptake rate of valsartan was higher for rat hepatocytes (K(m,u)=28.4+/-3.7 microM, V(max)=1320+/-180 pmol/mg/min, and P(dif) =1.21+/-0.42 microl/mg/min) compared to human hepatocytes (K(m,u)=44.4+/-14.6 microM, V(max)=304+/-85 pmol/mg/min, and P(dif)=0.724+/-0.271 microl/mg/min). OATP1B1 and -1B3 parameters were correlated to human hepatocyte data, using experimentally established relative activity factors (RAF). Resulting PBPK simulations were compared for plasma- (humans and rats) and bile- (rats) concentration-time profiles following iv bolus administration of valsartan. Plasma clearances (CL(P)) for rats and humans were predicted within twofold relative to predictions based on respective in vitro data. The simulations were extended to simulate the impact of either OATP1B1 or -1B3 inhibition on plasma profile. The limited data set indicates that the mechanistic model allowed for accurate evaluation of in vitro transport data; and the resulting hepatic uptake transport kinetic parameters enabled the prediction of in vivo PK profiles and plasma clearances, using PBPK modelling. Moreover, the interspecies difference in elimination rate observed in vivo was correctly reflected in the transport parameters determined in vitro.
Related Concept Videos
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:16
Methods for Studying Drug Absorption: In vitro
The diffusion cell method uses a two-compartment cell, including a donor compartment with the drug solution, which simulates the environment where the drug is applied, and a receptor compartment with a buffer solution, which simulates the environment...
 01:30
Measurement of Bioavailability: Pharmacokinetic Methods
 01:30
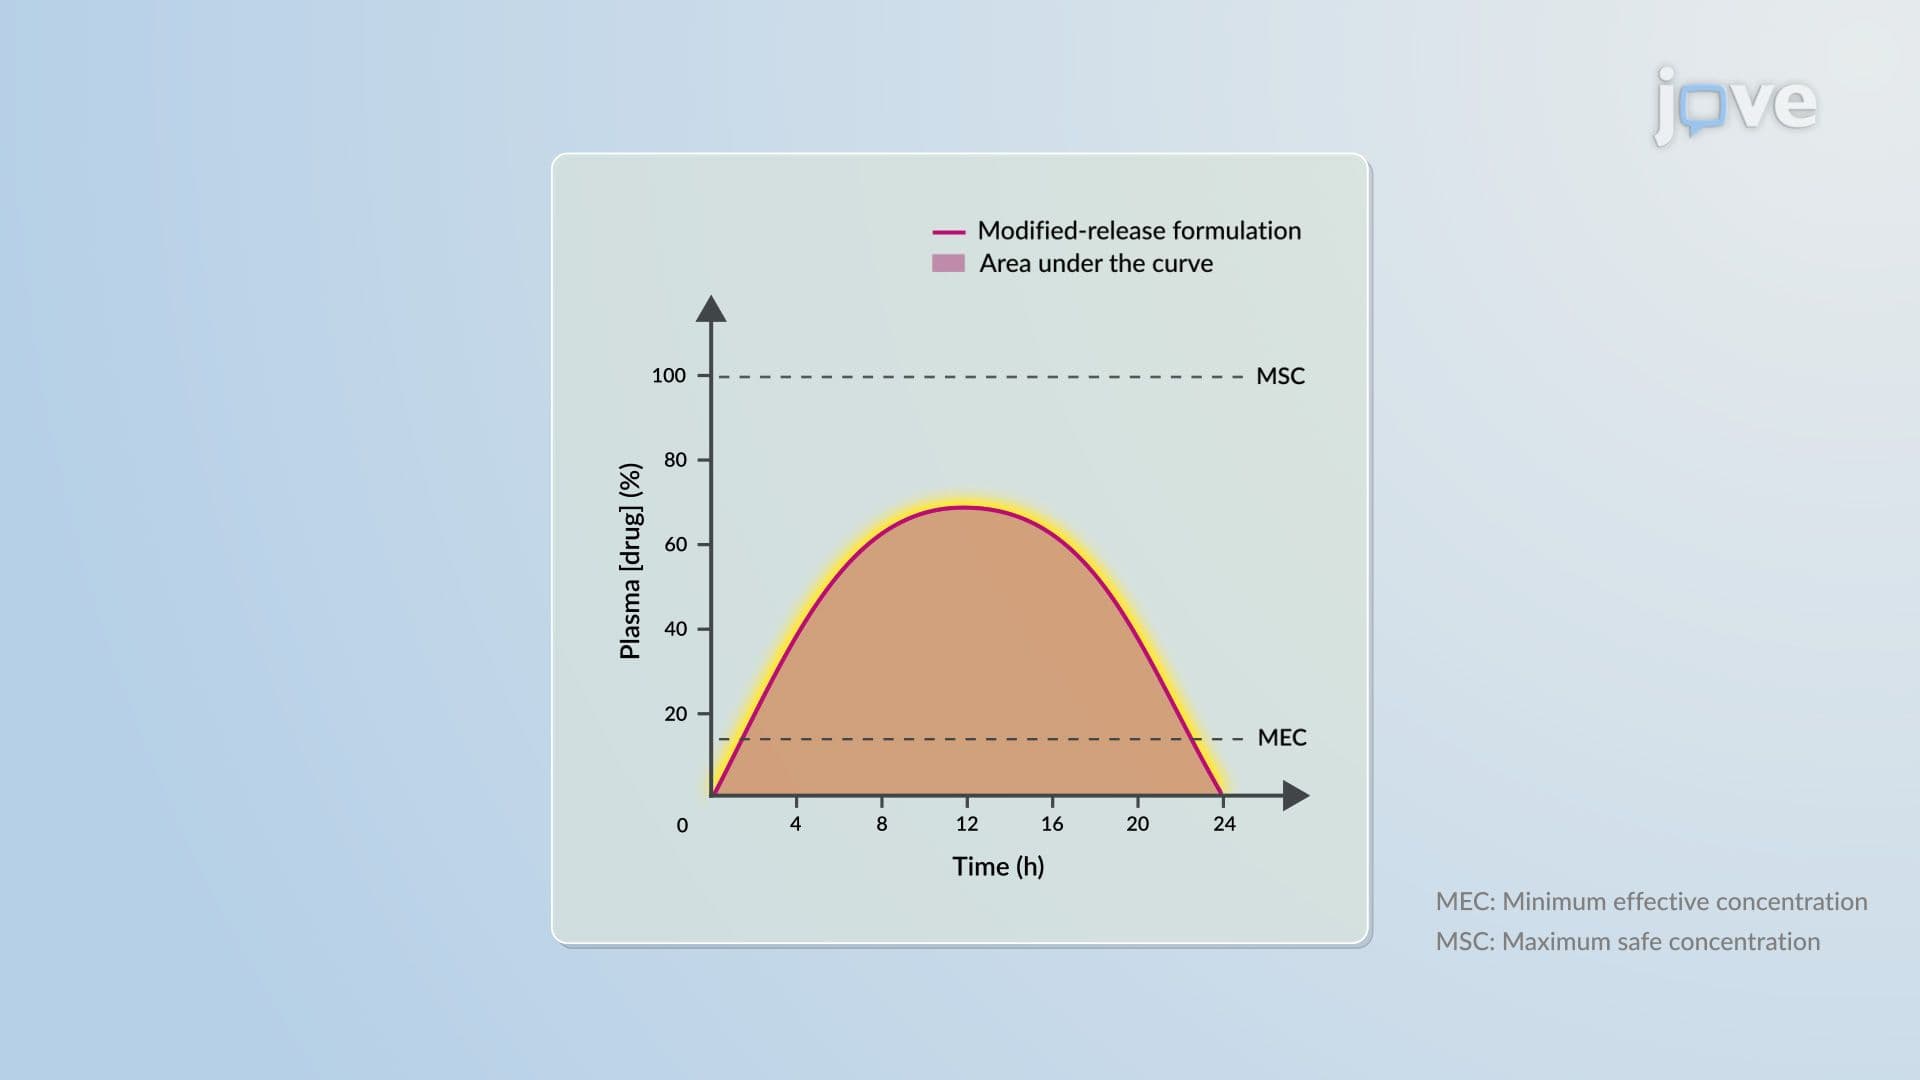
Modified-Release Drug Delivery Systems: Bioavailability
 01:09
Methods for Studying Drug Absorption: In situ
The Doluisio method involves perfusing a prepared segment of a rat's small intestine with a solution of radiolabeled drug and a non-absorbable marker. This helps to differentiate between absorbed and non-absorbed drug concentrations. The intestinal segment is connected at both ends using tubing and syringes,...
 01:01
Dosage Regimens: Partial Pharmacokinetic Parameters