Related Experiment Video
Updated: Jun 17, 2026
Modeling Charcot-Marie-Tooth Disease In Vitro by Transfecting Mouse Primary Motoneurons
Published on: January 7, 2019
The pathogenesis and treatment of acid sphingomyelinase-deficient Niemann-Pick disease
1Department of Genetics & Genomic Sciences, Mount Sinai School of Medicine, New York, NY 10029, USA. edward.schuchman@mssm.edu

Abstract:
Patients with Niemann-Pick disease (NPD) Types A and B have an inherited deficiency of acid sphingomyelinase (ASM) activity. The clinical spectrum of this disorder ranges from the infantile neurological form that results in death by 3 years of age (NPD Type A) to the non-neurological form that is compatible with survival into adulthood (NPD Type B). Intermediate cases have also been reported, and the disease is best thought of as a single entity with a spectrum of phenotypes. ASM deficiency is panethnic, but appears to be more frequent in individuals of Middle Eastern and North African descent. Current estimates of the disease incidence range from 0.5 to 1 per 100,000 births, although these approximations are thought to underestimate the true frequency of the disorder. The gene encoding ASM--SMPD1--has been studied extensively, and over 100 mutations in SMPD1 have been found to cause ASM-deficient NPD. Based on these findings, DNA-based carrier screening has been implemented in the Ashkenazi Jewish community. ASM-knockout mouse models also have been generated and used to investigate disease pathogenesis and treatment with stem cell transplantation, gene therapy and enzyme replacement therapy (ERT). Based on these studies, clinical trials of ERT are underway in patients with non-neurological ASM-deficient NPD.
Related Concept Videos
 01:22
Lysosomal Hydrolases
 01:24
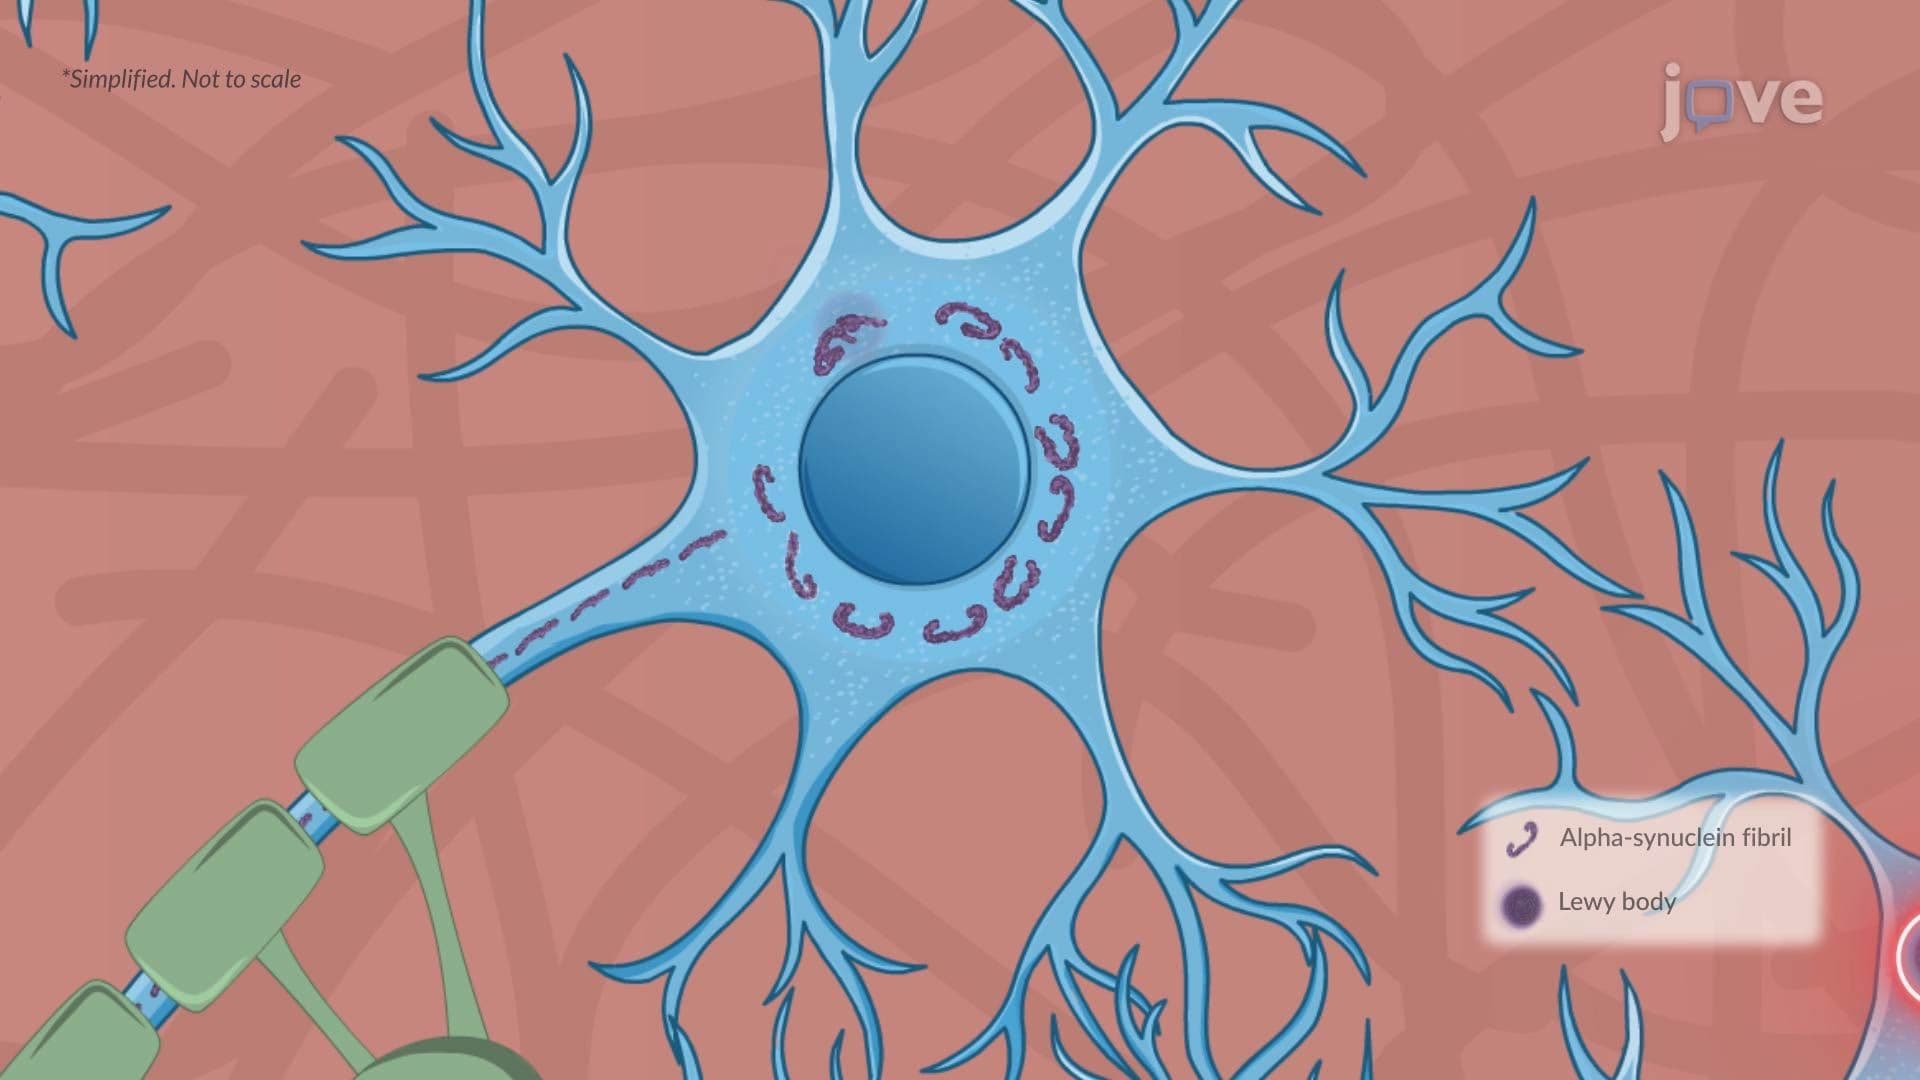
Parkinson Disease ll: Pathophysiology